
Table of Contents
As confidentially submitted to the Securities and Exchange Commission on February 7, 2025
This draft registration statement has not been publicly filed with the Securities and Exchange Commission and all information herein remains strictly confidential.
Registration No. 333-
UNITED STATES
SECURITIES AND EXCHANGE COMMISSION
Washington, D.C. 20549
FORM S-1
REGISTRATION STATEMENT
UNDER
THE SECURITIES ACT OF 1933
CapsoVision, Inc.
(Exact name of registrant as specified in its charter)
| Delaware | 3845 | 20-3369494 | ||
| (State or other jurisdiction of incorporation or organization) |
(Primary Standard Industrial Classification Code Number) |
(I.R.S. Employer Identification Number) |
18805 Cox Avenue, Suite 250
Saratoga, CA 95070
+1-408-624-1488
(Address, including zip code, and telephone number, including area code, of registrant’s principal executive offices)
Kang-Huai (Johnny) Wang
President and Chief Executive Officer
18805 Cox Avenue, Suite 250
Saratoga, CA 95070
+1-408-624-1488
(Name, address, including zip code, and telephone number, including area code, of agent for service)
Copies to:
| Portia Ku, Esq. O’Melveny & Myers LLP JC Plaza, 12th Floor 1225 Nanjing Road West Shanghai 200040 +86-21-2307-7000 |
Kurt Berney, Esq. O’Melveny & Myers LLP Two Embarcadero Center, 28th Floor San Francisco, California 94111 +1-415-984-8700 |
Richard I. Anslow Jonathan Deblinger Joseph A. Smith Ellenoff Grossman & Schole LLP 1345 Avenue of the Americas, 11th Floor New York, New York 10105 +1-212-370-1300 |
Approximate date of commencement of proposed sale to the public: As soon as practicable after the effective date of this registration statement.
If any of the securities being registered on this form are to be offered on a delayed or continuous basis pursuant to Rule 415 under the Securities Act of 1933, check the following box. ☒
If this form is filed to register additional securities for an offering pursuant to Rule 462(b) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this form is a post-effective amendment filed pursuant to Rule 462(c) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this form is a post-effective amendment filed pursuant to Rule 462(d) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, a smaller reporting company, or an emerging growth company. See the definitions of “large accelerated filer,” “accelerated filer,” “smaller reporting company” and “emerging growth company” in Rule 12b-2 of the Exchange Act:
| Large accelerated filer | ☐ | Accelerated filer | ☐ | |||
| Non-accelerated filer | ☒ | Smaller reporting company | ☒ | |||
| Emerging growth company | ☒ | |||||
If an emerging growth company, indicate by check mark if the registrant has elected not to use the extended transition period for complying with any new or revised financial accounting standards provided pursuant to Section 7(a)(2)(B) of the Securities Act. ☐
The registrant hereby amends this registration statement on such date or dates as may be necessary to delay its effective date until the registrant shall file a further amendment which specifically states that this registration statement shall thereafter become effective in accordance with Section 8(a) of the Securities Act of 1933 or until the registration statement shall become effective on such date as the Commission, acting pursuant to said Section 8(a), may determine.
Table of Contents
The information in this preliminary prospectus is not complete and may be changed. These securities may not be sold until the registration statement with the Securities and Exchange Commission is effective. This preliminary prospectus is not an offer to sell, nor does it seek an offer to buy these securities in any jurisdiction where the offer or sale is not permitted.
Subject to Completion Dated , 2025
PRELIMINARY PROSPECTUS
Shares
CapsoVision, Inc.
Common Stock
This is the initial public offering of CapsoVision, Inc. We are offering shares of our common stock, par value $0.001 per share. Prior to this offering, there has been no public market for shares of our common stock. We anticipate the initial public offering price will be between $ and $ per share.
In connection with this offering, we intend to apply to have shares of our common stock listed on the Nasdaq Capital Market (“Nasdaq”) under the symbol “CV”. There can be no assurance that such application will be approved. If shares of our common stock are not approved for listing on Nasdaq, we will not consummate this offering.
We are an “emerging growth company” and a “smaller reporting company” as defined in the federal securities laws and will be subject to reduced public company reporting requirements. See “Prospectus Summary—Implications of Being an Emerging Growth Company” and “Prospectus Summary—Implications of Being a Smaller Reporting Company.”
An investment in shares of our common stock is highly speculative, involves a high degree of risk and should be considered only by persons who can afford the loss of their entire investment, see “Risk Factors” beginning on page 12 of this prospectus before you make your decision to invest in our common stock.
Neither the Securities and Exchange Commission nor any state securities commission has approved or disapproved of these securities or determined if this prospectus is truthful or complete. Any representation to the contrary is a criminal offense.
| Per Share | Total | |||||||
| Initial public offering price(1) |
$ | $ | ||||||
| Underwriting discounts and commissions(2) |
$ | $ | ||||||
| Proceeds to us, before expenses |
$ | $ | ||||||
| (1) | Assuming an initial public offering price is $ , the midpoint of the range set forth on the cover page of this prospectus. |
| (2) | See the section titled “Underwriting” beginning on page 150 for additional information regarding compensation payable to the underwriters. |
In addition to the underwriting discounts and commissions referred to in the table above, we have agreed to issue, upon closing of this offering, warrants to the representative of the underwriters to purchase 3% of the total number of shares of common stock sold in this offering at a per share price equal to 125% of the public offering price (the “Representative’s Warrants”). The registration statement of which this prospectus is a part also covers the Representative’s Warrants and the shares of common stock issuable upon the exercise thereof. See section entitled “Underwriting” on page 150 for more information.
We have granted to the underwriters an option exercisable for a period of 30 days from the closing of this offering to purchase up to additional shares of our common stock from us at the initial public offering price, less the underwriting discounts and commissions, solely to cover over-allotments, if any.
The underwriters expect to deliver the shares of common stock to investors on or about , 2025.
The Benchmark Company
The date of this prospectus is , 2025.
Table of Contents
| Page | ||||
| ii | ||||
| iii | ||||
| 1 | ||||
| 8 | ||||
| 10 | ||||
| 12 | ||||
| 58 | ||||
| 59 | ||||
| 60 | ||||
| 62 | ||||
| Management’s Discussion and Analysis of Financial Condition and Results of Operations |
63 | |||
| 78 | ||||
| 121 | ||||
| 127 | ||||
| 133 | ||||
| Security Ownership of Certain Beneficial Owners and Management |
135 | |||
| 138 | ||||
| 144 | ||||
| Material U.S. Federal Income Tax Consequences for Non-U.S. Holders |
146 | |||
| 150 | ||||
| 162 | ||||
| 163 | ||||
| 164 | ||||
| F-1 | ||||
Table of Contents
No dealer, salesperson or other person is authorized to give any information or to represent anything not contained in this prospectus. You must not rely on any unauthorized information or representations. This prospectus is an offer to sell only the shares offered hereby, but only under circumstances and in jurisdictions where it is lawful to do so. The information contained in this prospectus is current only as of its date.
This prospectus contains estimates, projections, and other information concerning our industry and our business, as well as data regarding market research, estimates, and forecasts prepared by our management or third parties. In particular, there is limited public available data as to the size and growth rates for the broader capsule endoscopy market (including for those medical indications that we seek to address); for the GI-tract medical indications that we and other capsule endoscopy providers seek to target, that portion for which capsule endoscopy is indicated as appropriate for diagnosis and screening; assumptions as to Medicare and other third-party payor reimbursement rates; and the impact of various non-capsule competitive products. Information that is based on estimates, forecasts, projections, market research, or similar methodologies is inherently subject to uncertainties, and actual events or circumstances may differ materially from events and circumstances that are assumed in this information. While we believe the market and industry data included in this prospectus are reliable and are based on reasonable assumptions, these data and the industry in which we operate is subject to a high degree of uncertainty and risk due to a variety of factors, including those described in the section titled “Risk Factors.” These and other factors could cause results to differ materially from those expressed in these estimates, publications, and reports made by third parties or us.
Unless otherwise expressly stated, we obtained such industry, business, market, and other data from reports, research surveys, studies, and similar data prepared by market research firms and other third parties, industry and general publications, government data, and similar sources. In some cases, we do not expressly refer to the sources from which this data is derived. In that regard, when we refer to one or more sources of this type of data in any paragraph, you should assume that other data of this type appearing in the same paragraph is derived from the same sources, unless otherwise expressly stated or the context otherwise requires. The content of these third- party sources, except to the extent specifically set forth in this prospectus, does not constitute a portion of this prospectus and is not incorporated herein.
Forecasts and other forward-looking information with respect to industry, business, market, and other data are subject to the same qualifications and additional uncertainties regarding the other forward-looking statements in this prospectus. See “Cautionary Note Regarding Forward-Looking Statements.”
ii
Table of Contents
CAUTIONARY NOTE REGARDING FORWARD-LOOKING STATEMENTS
This document contains forward-looking statements. In addition, from time to time, we or our representatives may make forward-looking statements orally or in writing. We base these forward-looking statements on our expectations and projections about future events, which we derive from the information currently available to us. Such forward-looking statements relate to future events or our future performance, including: our financial performance and projections; our growth in revenue and earnings; and our business prospects and opportunities. You can identify forward-looking statements by those that are not historical in nature, particularly those that use terminology such as “may,” “should,” “expects,” “anticipates,” “contemplates,” “estimates,” “believes,” “plans,” “projected,” “predicts,” “potential,” or “hopes” or the negative of these or similar terms. In evaluating these forward-looking statements, you should consider various factors, including: our ability to change the direction of the Company; our ability to keep pace with new technology and changing market needs; and the competitive environment of our business. These and other factors may cause our actual results to differ materially from any forward-looking statement. Forward-looking statements are only predictions. The forward-looking events discussed in this document and other statements made from time to time by us or our representatives, may not occur, and actual events and results may differ materially and are subject to risks, uncertainties and assumptions about us. We are not obligated to publicly update or revise any forward-looking statement, whether as a result of uncertainties and assumptions, the forward-looking events discussed in this document and other statements made from time to time by us or our representatives might not occur.
While we believe we have identified material risks, these risks and uncertainties are not exhaustive. Other sections of this prospectus describe additional factors that could adversely impact our business and financial performance. Moreover, we operate in a very competitive and rapidly changing environment. New risks and uncertainties emerge from time to time, and it is not possible to predict all risks and uncertainties, nor can we assess the impact of all factors on our business or the extent to which any factor, or combination of factors, may cause actual results to differ materially from those contained in any forward-looking statements.
Although we believe the expectations reflected in the forward-looking statements are reasonable, we cannot guarantee future results, level of activity, performance or achievements. Moreover, neither we nor any other person assumes responsibility for the accuracy or completeness of any of these forward-looking statements. You should not rely upon forward-looking statements as predictions of future events. We are under no duty to update any of these forward-looking statements after the date of this prospectus to conform our prior statements to actual results or revised expectations, and we do not intend to do so.
Forward-looking statements include, but are not limited to, statements about:
| • | our expectations regarding the acceptance of our products by patients and doctors; |
| • | our expectations regarding the potential market size for our current CapsoCam Plus capsule and CapsoCam Colon (once FDA cleared) and those markets that we may pursue; |
| • | our plans to increase small bowel capsule sales following recent 510(k) clearance for pediatric use and telehealth supervision and related products currently under development; |
| • | our expected receipt of and related timing for FDA 501(k) clearance of our CapsoCam Colon and related sales; |
| • | our plans and efforts to expand into new indications in terms of new GI pathologies and expanded patient populations; |
| • | our plans and efforts to introduce enhancements and improvements to our products and technologies, including the AI capabilities incorporated into our products; |
| • | our commercialization capabilities and strategies, including our plans to increase revenues and sales capabilities in and outside the United States (the “U.S.”); |
iii
Table of Contents
| • | the implementation of our strategic plan for our business and products and technology; |
| • | our relationships with, and capabilities of, our assembly manufacturers and component suppliers; |
| • | the protection of our intellectual property (including our AI capabilities) including through patents and trade secret protections; |
| • | the expected performance of our products; |
| • | our ability to manage our growth; |
| • | the anticipated use of proceeds from this offering; |
| • | our expectations regarding the time during which we will be an emerging growth company under the Jumpstart Our Business Startups Act of 2012 (the “JOBS Act”) or smaller reporting company under U.S. securities laws; |
| • | estimates of our expenses, future revenue, capital requirements, our needs for additional financing, and our ability to obtain additional capital; |
| • | our ability to continue as a going concern; and |
| • | our future financial performance. |
We caution you not to place undue reliance on the forward-looking statements, which speak only as of the date of this prospectus in the case of forward-looking statements contained in this prospectus.
iv
Table of Contents
This summary highlights information contained elsewhere in this prospectus. This summary does not contain all of the information that you should consider before deciding to invest in our common stock. You should read this entire prospectus carefully, including the “Risk Factors” section, our historical financial statements and the notes thereto, each included elsewhere in this prospectus. Unless otherwise indicated or the context requires otherwise, the words “we,” “us,” “our,” the “Company,” or “our Company,” and “CapsoVision” refer to CapsoVision, Inc., a Delaware corporation.
Overview
We are a commercial-stage medical technology company that develops advanced imaging and artificial intelligence (“AI”) technologies, deployed in our capsule endoscopy solutions to identify abnormalities of the gastrointestinal (“GI”) tract for diagnostic and screening purposes.
We developed our first capsule endoscope system, currently comprising the CapsoCam Plus single-use capsule and the CapsoCloud and CapsoView software, to panoramically visualize the small-bowel mucosa to detect abnormalities such as obscure GI bleeding and Crohn’s disease. The capsule acquires and stores video images in onboard memory while moving through the GI tract, and the software component allows healthcare providers to view the video retrieved from the capsule—either by streaming it from the cloud, where it is securely stored, to anywhere, at their convenience, using our CapsoCloud software, or downloading it from the capsule themselves and reviewing it in our CapsoView software. The CapsoCam is a wire-free capsule endoscopy solution, eliminating patient-worn data recorders and providing clinicians a zero-capex, maintenance-free, flexible, and scalable workflow.
We are (i) in the process of updating CapsoCam Plus to add our self-developed AI assisted reading technology and (ii) targeting related FDA 510(k) clearance of the updated capsule for late 2025, with commercialization shortly thereafter. Our AI assisted reading tools detect and highlight suspected abnormalities for a clinician, reducing their time to review the video and making capsule endoscopy more financially attractive to their practice.
We began sales of our small-bowel capsule system to our provider customers (i.e., primarily gastroenterologists practicing in clinics and/or hospitals) both internationally (in 2012) and in the U.S. (in 2017) through our global sales and marketing team. In the U.S., we sell to customers directly. Internationally, we sell both directly and through qualified exclusive distributors in specified regions. Our largest international markets (based on our sales) are Germany, Sweden, and the U.K. In 2023, we established a direct sales team in Germany to better serve our customers and strengthen our market presence in this key market. We plan to (i) further grow our existing sales and marketing team to increase small-bowel-related sales and (ii) leverage our existing sales and marketing team to sell future product additions to our GI-tract capsule endoscopy solution.
Driven by increased acceptance of CapsoCam Plus and growth in the small-bowel capsule endoscopy market, our revenue has increased in each year since we began U.S. direct sales in 2020. Our revenues for the year ended December 31, 2023 and 2024 totaled approximately $9.8 million and $[ ] million, respectively, representing a year-over-year growth of approximately [ ]% from 2023 to 2024. In 2023 and 2024, international sales accounted for 26% and [ ]% of total revenue. As of December 31, 2024, our small bowel capsule has been used by over 120,000 patients worldwide and for 2024 our customer retention rate was approximately 90%.
To expand beyond small-bowel-related sales, we are developing our next pipeline capsule endoscope product, CapsoCam Colon. Our CapsoCam Colon capsule (i) leverages CapsoCam Plus’s existing capsule design
1
Table of Contents
with its panoramic view and (ii) incorporates both our self-developed AI to automatically detect polyps in the video and our polyp-size measurement tool enabled by a 3D sensor in the capsule (polyp size being highly correlated with a polyp’s risk of becoming cancer). Based on our current regulatory development plan, we are targeting CapsoCam Colon revenues beginning in mid-2026 after receiving FDA 510(k) clearance of our second generation of CapsoCam Colon system, designed with a larger field of view and better image quality to improve accuracy.
Longer term, we believe our CapsoCam family of products, incorporating our panoramic imaging solution, can be adapted to address new GI medical indications. Potential new medical indications include pancreatic cancer and esophageal medical conditions (such as esophageal varices and Barrett’s esophagus). In 2025, we plan to commence clinical investigation of CapsoCam’s accuracy in (i) detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) and (ii) screening esophageal varices in cirrhotic patients with portal hypertension.
Market Overview
Overview and Challenges of Visualizing Small Bowel and Detecting Small Bowel Pathologies
Diseases of the small bowel include obscure GI bleeding, chronic iron-deficiency anemia, Crohn’s disease, tumors, and polyposis. Obscure GI bleeding is recurrent or persistent GI bleeding of uncertain origin. 5% of all GI bleeding is obscure GI bleeding. In approximately 80% of obscure GI bleeding cases, the origin is localized to the small bowel. Chronic iron-deficiency anemia is a condition in which blood lacks adequate healthy red blood cells. Crohn’s disease is a type of inflammatory bowel disease that causes swelling and irritation of the tissues, called inflammation, in the digestive tract.
Capsule endoscopy is the first-line modality for imaging the mucosa of the small bowel including the pathologies characterizing these diseases. Various methods of enteroscopy for reaching the entirety of the small bowel, which is approximately 20 feet long, are invasive, time consuming, and require a high level of endoscopy skill. Enteroscopy is still required for biopsy or to provide certain therapies. For diagnostic visualization, however, capsule endoscopy is preferred given its simplicity, non-invasiveness, and relatively low cost.
Our Addressable Market Opportunity in Visualizing Small Bowel and Detecting Small Bowel Pathologies
The U.S. capsule endoscopy market is estimated at approximately $[ ] in [ ] and is forecasted to reach approximately $[ ] in [ ]. The small bowel market is currently the dominant sector within the broader capsule endoscopy market in the U.S.
Overview and Challenges of Detecting Colon Polyps
A colon polyp is a clump of cells that forms on the lining of the colon. Most colon polyps are harmless, but, over time, some colon polyps develop into colorectal cancer (“CRC”). The size of a polyp is highly correlated with its risk of becoming cancerous.
Currently, optical colonoscopy, accompanied by polypectomy and biopsy, is considered the gold-standard for the detection of colorectal polyps and cancers. Each year in the U.S., there are approximately 153,000 new cases of CRC and approximately 53,000 deaths. It is widely accepted that CRC is among the most preventable, yet least-prevented cancers. CRC can take up to 10-15 years to progress from a pre-cancerous lesion to metastatic cancer and death. Patients who are diagnosed early in the progression of the disease—with pre-cancerous lesions or early-stage cancer—are more likely to have a complete recovery and to be treated less expensively.
2
Table of Contents
Our Addressable Market Opportunity in Colon Capsule Endoscopy
We estimate our total addressable market opportunity at over $[ ] annually in the U.S., which represents the potential opportunity from the sale of colon capsule endoscopies to healthcare providers and hospitals. Please see “Business—Market Overview—Our Addressable Market Opportunity in Colon Capsule Endoscopy” for a description of how we calculated this addressable market and related assumptions.
Our Solutions
CapsoCam Plus
Our CapsoCam Plus capsule endoscopy system is intended for visualization of the small bowel mucosa, to detect abnormalities of the small bowel in adults and children aged 2 years and above. We believe that our CapsoCam Plus is a superior capsule endoscopy system compared to competitor systems, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables.
CapsoCam Colon
We have developed our next pipeline product, CapsoCam Colon, for visualization of the colon and detection and measurement of polyps. We believe that our CapsoCam Colon, once FDA cleared, will be a superior capsule endoscopy system compared to competitor systems, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. As described below under “—Our Growth Strategies”, we are conducting relevant clinical development activities (including our recently concluded pivotal study involving approximately 1,200 patients enrolled at 20 sites throughout the U.S.) and product enhancement activities to position us to commercialize CapsoCam Colon by mid-2026.
Our Strengths
We believe the continued growth of our Company will be driven by the following factors:
| • | Sole capsule endoscope with a 360° panoramic view available in the market. Only our CapsoCam Plus boasts a 360° panoramic lateral view. It houses four high-resolution cameras around its circumference, and the images from each are stitched into a single panoramic image. Compared to competitive end-view systems, a 360° panoramic lateral view provides a complete view of the GI mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature, resulting in demonstrated superior diagnostic yield. |
| • | Telemedicine-enabled and zero-capex “wire-free” data collection and remote data analysis. Our CapsoCam is a zero-capex “wire-free” data collection solution for providers, as it stores the entire video in onboard memory. In December 2024, we received FDA 510(k) clearance for telemedicine supervision (i.e., remote ingestion) of our CapsoCam Plus, allowing patients to ingest our capsule in the comfort of their own homes, under the remote supervision of providers. The CapsoCam solution frees up exam-room schedules for providers and provides flexibility to administer capsules any day at any time. A provider’s practice can easily scale to multiple capsules per day with no added cost, and there is no equipment to recover from patients. Our CapsoCam Plus solution includes our cloud-based platform, CapsoCloud, which provides a flexible, trackable, streamlined, and capital-equipment-free workflow for providers in the U.S. It allows clinicians to track procedures and stream in vivo videos anywhere at their convenience, generate reports, store and manage patient data, and transfer data to third-party reading services. We believe at-home procedures and remote analysis via CapsoCloud will be attractive to providers and patients alike, particularly for future screening indications. |
| • | Automated pathology detection due to usage of AI. CapsoCam Colon (subject to FDA clearance) incorporates deep learning AI for automated pathology detection of polyps, a capability that competitive |
3
Table of Contents
| systems lack. In the first half of 2025, we also plan to file for FDA and EU clearance for the use of AI in our CapsoCam Plus capsule. Our AI assisted reading tools detect and highlight suspected abnormalities for a clinician, reducing their time to review the video and making capsule endoscopy more financially attractive to their practice. CapsoCloud continuously acquires consenting patients’ clinical data, enabling our in-house AI experts to develop ever-improving automated lesion detection and classification. |
| • | 3D-sensing technology informs follow-on care decisions. Our CapsoCam Colon (subject to FDA clearance) incorporates our proprietary 3D-sensing technology to more accurately measure polyp sizes. Polyp size is highly correlated with its risk of becoming cancer. No other capsule endoscope currently in the market has 3D-enabled measurement capability. With automated pathology detection and the ability to manually review video frames adjacent to an identified polyp, physicians can more confidently decide that patients with small polyps (e.g., less than 6mm) may forgo a follow-on colonoscopy, increasing the utility of the procedure for healthcare providers and patients alike. |
| • | Experienced leadership team. Our senior management team consists of industry professionals with deep industry expertise across various disciplines, including medical technology, engineering, optics, sales and marketing, finance, operations, data science, AI, and clinical operations and research. |
Our Growth Strategies
Our long-term, lifesaving vision is an ingestible capsule that, in a single convenient non-invasive procedure, screens for multiple cancers—esophageal, gastric, pancreatic, small-bowel, and colorectal—at early and precancerous stages, utilizing AI to analyze thousands of images captured in the GI tract. We are building towards this goal with a planned succession of FDA-cleared indications, targeting existing and nascent markets. Until then, key elements of our nearer-term growth strategy include:
| • | Obtain 510(k) clearance of CapsoCam Colon. We are developing CapsoCam Colon for visualization of the colon and detection and measurement of polyps. In addition to having a 360° panoramic lateral view, CapsoCam Colon incorporates deep learning AI for automated pathology detection of polyps and 3D-sensing technology to more accurately measure polyp sizes. We are currently concluding a pivotal study to support 510(k) clearance of CapsoCam Colon for use by currently indicated patients who are a subset of the colorectal cancer screening and surveillance populations. The pivotal study involved approximately 1,200 patients enrolled at 20 sites throughout the U.S. Based on the clinical results from the pivotal study, we are planning to submit our 510(k) application, with two existing capsule endoscopies as predicate devices (one for the capsule and one for the incorporated AI), to the FDA in the first half of 2025 with a goal of obtaining 510(k) clearance in late 2025. To enhance the sensitivity and specificity of CapsoCam Colon for detecting and measuring polyps, we are developing our second-generation CapsoCam Colon capsule, which will include improvements such as a new lens and illumination optics with an increased field of view and improved image quality, and will extend our pivotal study to include a second arm, which is expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S. We plan to use the clinical results of the second arm to support submission to the FDA of a 510(k) application in late 2025 with a goal of obtaining 510(k) clearance in the first half of 2026. Current indications for colon capsule endoscopy are limited to patients with evidence of lower-GI bleeding (such as a positive stool test) for whom the risk of colonoscopy or moderate sedation is significant and for patients who have had an incomplete colonoscopy, with adequate preparation. As a part of this second 510(k) application to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation. |
| • | Expand clinical use cases and accessories for CapsoCam Plus. In December 2024, we received 510(k) clearance of our CapsoCam Plus for pediatric use in children aged 2 years and above and telemedicine supervision (i.e., remote ingestion). Also, we are currently developing a capsule delivery device |
4
Table of Contents
| (availability expected in 2025) for patients who are unable to swallow the capsule, many of whom are children. The capsule delivery device should enable faster penetration of the newly indicated pediatric market. We are also currently developing a patency capsule (tentative FDA 510(k) submission planned by mid-2025), which is used primarily with Crohn’s disease patients to verify that a capsule endoscope can pass through the GI tract without retention at a stricture, a narrowing of the small bowel which can result from inflammation and scarring associated with Crohn’s disease. |
| • | Continue to improve and innovate our GI-tract capsule endoscopy solution. Our research and development initiatives are focused on introducing enhancements and improvements aimed at increasing the value provided by our GI-tract capsule endoscopy solution. In particular, we are working on improvements to our CapsoCam, including a new lens and illumination optics with an increased field of view, improved image quality and higher peak frame rate. For our AI assisted reading technology, we plan to continue investing to (i) improve the pathology-detection and classification accuracy and the scope of our AI algorithms and (ii) apply AI, including large language models, to streamline the diagnostic and medical-report-generation processes, which in turn improves the efficiency and effectiveness of our healthcare provider customers. We also plan to continue making improvements to our CapsoCloud and CapsoView software. |
| • | Expand into new indications and clinical use cases beyond small bowel and colon. In 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) by visualizing abnormalities of the duodenal papilla. A dilated, or unnaturally opened, duodenal papilla is correlated with GI tract content reflux into the pancreas duct and pancreatic neoplasia and may indicate the presence of, or elevated risk of developing, serious abnormalities like pancreatitis or a tumor of the pancreas. A dilated papilla is frequently not observable by endoscopy due to physical perturbation/disruption in connection with the procedure and heavy patient sedation. Our CapsoCam capsule, with its panoramic view, can observe the papilla in its natural state. Prior studies have demonstrated the CapsoCam’s superior detection of the normal papilla relative to non-panoramic systems. According to the American Cancer Society (the “ACS”), pancreatic cancer is estimated to be diagnosed in over 67,000 patients in the U.S. in 2025, and approximately 52,000 patients are estimated to die from it. The overall 5-year survival rate is approximately 13%, which is the lowest survival rate of all cancer types. On the other hand, the 5-year survival rate is around 44% if the pancreatic cancer is still in the early stages at the time of diagnosis. There is currently no effective screening for pancreatic cancer. Also, in 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in screening esophageal varices in cirrhotic patients with portal hypertension. |
Risks Relating to Our Business
Our business and ability to execute our business strategy are subject to a number of risks of which you should be aware before you decide to buy our securities. In particular, you should consider the following risks, which are discussed more fully in the section entitled “Risk Factors”:
| • | We have a history of net losses and expect to incur additional losses in the foreseeable future. |
| • | Our 2023 audited financial statements include a footnote raising substantial doubt about our ability to continue as a “going concern” and, even if this offering is successful, we may still need to raise additional financing to fund our business and revenue growth plans. |
| • | All of our revenues to date have been, and in the near-term will continue to be, generated from CapsoCam Plus small bowel related sales; and growing our small-bowel-related revenue is subject to our ability to successfully and timely execute related elements of our revenue growth strategy. |
| • | Currently we do not expect CapsoCam Colon-related revenues until mid-2026 given anticipated timing for receiving FDA clearance of our second generation CapsoCam Colon solution. |
5
Table of Contents
| • | Broad adoption of our CapsoCam Colon solution (once cleared) and growing related revenues depend on, among other things, expanding beyond CapsoCam Colon’s initial indicated patient population and a successful completion rate for our capsule colonoscopy, increasing the accuracy of our initial CapsoCam Colon solution; and we may fail to successfully and timely execute related elements of our CapsoCam Colon adoption and revenue growth strategy. |
| • | Longer term efforts to expand our GI-tract capsule endoscopy solutions beyond small bowel and colon pathologies and medical conditions may not succeed. |
| • | We may not obtain or may experience delays in obtaining 510(k) clearance for our initial or planned second generation of our CapsoCam Colon capsule endoscopy solution (with improved optics and other components), which would adversely impact our ability to commercialize this product. |
| • | We may face regulatory challenges and delays in obtaining 510(k) clearance for marketing of those CapsoCam capsules incorporating our self-developed AI assisted reading technology. |
| • | Any failure or defect in our GI-tract capsule endoscopy solution could harm our reputation, expose us to liability, and reduce our sales. |
| • | We rely on various suppliers to assist us in the assembly and manufacture of our GI-tract capsule endoscopy solution and sourcing of critical and other components and many of these suppliers are single-source suppliers located in Asia (particularly Taiwan and Japan); any disruption in our supply chain could adversely affect our ability to meet the demand for our products and fulfil our orders. |
| • | We face substantial geopolitical and trade risks (including a possible trade war and imposition of tariffs and supply chain interruptions) and natural disaster risks associated with our business operations in Asia (particularly Taiwan and Japan). |
| • | If Medicare and other third-party payors do not approve reimbursement for our products at adequate reimbursement rates, we may be unable to successfully commercialize our products. |
| • | If we or our distributors fail to comply with regulatory requirements, we may be subject to stringent penalties and our business may be materially adversely affected. |
| • | Our success will depend on our ability to obtain, maintain, enforce, and protect our intellectual property rights. |
| • | If we fail to identify our patentable inventions or adequately protect our patent rights, the commercial value of our products, services or technologies may be adversely affected and our competitive position may be harmed. |
Implications of Being an Emerging Growth Company
We qualify as an “emerging growth company” under the federal securities laws and, therefore, we may take advantage of certain exemptions from various public company reporting requirements, including:
| • | a requirement to only have two years of audited financial statements and only two years of related selected financial data and management’s discussion and analysis; |
| • | exemption from the auditor attestation requirement on the effectiveness of our internal control over financial reporting; |
| • | election not to adopt new or revised accounting standards until they become effective for private companies; |
| • | reduced disclosure obligations regarding executive compensation; and |
| • | exemptions from the requirements of holding a non-binding advisory stockholder vote on executive compensation and any golden parachute payments. |
6
Table of Contents
We may take advantage of these provisions for up to five years or such earlier time that we are no longer an emerging growth company. We would cease to be an emerging growth company if we have more than $1.235 billion in total annual growth revenues, have issued more than $1 billion of non-convertible debt in the past three years, or if we are deemed to be a large accelerated filer under the rules of the Securities and Exchange Commission (the “SEC”). We may choose to take advantage of some, but not all, of the available benefits available to emerging growth companies. We have taken advantage of some of the reduced reporting requirements in this prospectus. Accordingly, the information contained herein may be different than the information you receive from other public companies in which you hold stock. In addition, an emerging growth company may delay adopting new or revised accounting standards until such time as those standards apply to private companies. We have elected to avail ourselves of this exemption from new or revised accounting standards and, therefore, we will not be subject to the same new or revised accounting standards as other public companies that are not emerging growth companies.
Implications of Being a Smaller Reporting Company
We are a smaller reporting company as defined in the Securities Exchange Act of 1934, as amended (the “Exchange Act”). We may take advantage of certain of the scaled disclosures available to smaller reporting companies and will be able to take advantage of these scaled disclosures for so long as (i) the market value of our voting and non-voting common stock held by non-affiliates is less than $250 million measured on the last business day of our second fiscal quarter or (ii) our annual revenue is less than $100 million during the most recently completed fiscal year and the market value of our voting and non-voting common stock held by non-affiliates is less than $700 million measured on the last business day of our second fiscal quarter. If we are a smaller reporting company at the time we cease to be an emerging growth company, we may continue to rely on exemptions from certain disclosure requirements that are available to smaller reporting companies. Specifically, as a smaller reporting company, we may choose to present only the two most recent fiscal years of audited financial statements in our Annual Report on Form 10-K and we are not required to comply with the auditor attestation requirements of Section 404(b) of the Sarbanes-Oxley Act of 2002, as amended (the “Sarbanes-Oxley Act”), and, similar to emerging growth companies, smaller reporting companies have reduced disclosure obligations regarding executive compensation.
Company Information
We were incorporated under the laws of the State of Delaware on August 1, 2005, under the name “Capso Vision, Inc.” and changed our name to CapsoVision, Inc. on May 31, 2016. Our principal executive office is located at 18805 Cox Ave, Suite 250, Saratoga, CA 95070. Our website address is CapsoVision.com. The information on or accessible through our website is not part of this prospectus.
7
Table of Contents
| Securities we are offering |
shares of common stock. |
| Initial public offering price |
We anticipate the initial public offering price will be between $ and $ per share. |
| Shares of common stock outstanding immediately before this offering |
shares. |
| Shares of common stock outstanding immediately after this offering |
shares. |
| Over-allotment option |
We have granted the underwriters an option, exercisable for 30 days from the closing of this offering, to purchase up to an additional shares of our common stock at the initial public offering price less underwriting discounts and commissions, solely to cover over-allotments, if any. |
| Use of proceeds |
We intend to use the net proceeds of this offering for general corporate purposes. See “Use of Proceeds”. |
| Risk Factors |
See “Risk Factors” on page 12 of this prospectus and other information appearing elsewhere in this prospectus for a discussion of factors you should carefully consider before deciding whether to invest in our securities. |
| Lock-up |
We, our directors, executive officers, and holders (but limited to such holders that hold an aggregate of not more than 95%) of the outstanding shares of our common stock have agreed not to offer, issue, sell, contract to sell, encumber, grant any option for the sale of or otherwise dispose of any of our securities for a period of 6 months after the date of closing of this offering. See “Underwriting”. |
| Representative’s Warrants |
We have agreed to issue to the representative of the underwriters or its designees at the closing of this offering, warrants to purchase the number of shares of our common stock equal to 3% of the aggregate number of shares sold in this offering (the “Representative’s Warrants”). The Representative’s Warrants will be exercisable at any time and from time to time, in whole or in part, during the five (5) year period commencing six (6) months after the closing of this offering and may be exercised on a cashless basis. The exercise price of the Representative’s Warrants will equal 125% of the initial public offering price per share, subject to adjustments. See “Underwriting”. |
| Nasdaq listing and symbol |
In connection with this offering, we intend to apply to have shares of our common stock listed on Nasdaq under the symbol “CV”. No assurance can be given that our application for listing will be approved by Nasdaq or that a trading market will develop for our common stock. We will not proceed with this offering in the event shares of our common stock are not approved for listing on Nasdaq. |
8
Table of Contents
The number of shares of common stock to be outstanding after this offering is based on shares outstanding as of December 31, 2024 and reflects the Preferred Stock Conversion, as defined and described below.
The number of shares of our common stock to be outstanding after this offering does not include:
| • | 50,000 shares of our common stock issuable upon the exercise of outstanding warrants as of December 31, 2024 with a weighted-average exercise price of $1.45 per share; |
| • | shares of our common stock issuable upon the exercise of outstanding stock options as of December 31, 2024, with a weighted-average exercise price of $ per share; |
| • | shares of our common stock issuable upon the exercise of outstanding stock options granted subsequent to December 31, 2024, with a weighted-average exercise price of $ per share; and |
| • | shares of our common stock reserved for future issuance under our 2025 Equity Incentive Plan (the “2025 Plan”), which will become effective as of the date immediately prior to the date our registration statement relating to this offering becomes effective, as well as any future increases in the number of shares of common stock reserved for issuance under the 2025 Plan. |
Unless otherwise indicated, all information contained in this prospectus, including the number of shares of common stock that will be outstanding after this offering, assumes or gives effect to:
| • | the conversion of all the outstanding shares of our Series A, Series B, Series C, Series C-1, Series D, Series D-1, Series D-2, Series E, Series F-1, Series F-2, Series G, Series G-1 and Series H preferred stock into an aggregate of 128,756,397 shares of our common stock, the conversion of which will occur immediately prior to the completion of this offering (the “Preferred Stock Conversion”); |
| • | no exercise of outstanding warrants or options subsequent to December 31, 2024; and |
| • | no exercise by the underwriters of their option to purchase up to additional shares of our common stock and the Representative’s Warrants. |
9
Table of Contents
The following summary statement of operations for the years ended December 31, 2023 and 2024 and summary balance sheet data as of December 31, 2023 and 2024 have been derived from our audited financial statements included elsewhere in this prospectus.
Our financial statements are prepared and presented in accordance with accounting principles generally accepted in the United States, or U.S. GAAP. Our historical results are not necessarily indicative of results expected for future periods. You should read this Summary Financial Data section together with our financial statements and the related notes and “Management’s Discussion and Analysis of Financial Condition and Results of Operations” included elsewhere in this prospectus.
Statement of Operations data:
| For the Year Ended December 31, |
||||||||
| 2023 | 2024 | |||||||
| Revenue |
$ | 9,752,850 | ||||||
| Costs of revenue |
(4,262,373 | ) | ||||||
|
|
|
|
|
|||||
| Gross profit |
5,490,477 | |||||||
| Operating expenses: |
||||||||
| Selling and marketing |
(5,532,852 | ) | ||||||
| Research and development |
(9,332,574 | ) | ||||||
| General and Administrative |
(1,972,274 | ) | ||||||
| Total operating expenses |
(16,837,700 | ) | ||||||
| Operating loss |
(11,347,223 | ) | ||||||
|
|
|
|
|
|||||
| Non-operating income |
||||||||
| Interest income |
49,440 | |||||||
| Other non-operating income, net |
4,371 | |||||||
|
|
|
|
|
|||||
| Total non-operating income, net |
53,811 | |||||||
| Loss before provision for income tax |
(11,293,412 | ) | ||||||
| Provision for income taxes |
11,433 | |||||||
|
|
|
|
|
|||||
| Net loss |
$ | (11,304,845 | ) | |||||
|
|
|
|
|
|||||
| Net loss per share—basic and diluted |
$ | (1.93 | ) | |||||
|
|
|
|
|
|||||
| Weighted average shares—basic and diluted |
5,862,935 | |||||||
|
|
|
|
|
|||||
10
Table of Contents
Balance Sheet data:
| As of December 31, | ||||||||
| 2023 | 2024 | |||||||
| Cash |
$ | 14,559,374 | ||||||
|
|
|
|
|
|||||
| Total assets |
21,526,872 | |||||||
|
|
|
|
|
|||||
| Operating lease liabilities—current and non-current |
1,513,837 | |||||||
|
|
|
|
|
|||||
| Total convertible preferred stock |
128,624,807 | |||||||
| Accumulated deficit |
(110,451,852 | ) | ||||||
|
|
|
|
|
|||||
| Total Liabilities, Convertible Preferred Stock and Stockholders’ Deficit |
$ | 21,526,872 | ||||||
|
|
|
|
|
|||||
11
Table of Contents
Investing in our common stock involves a high degree of risk. You should carefully consider the risks described below, as well as the other information in this prospectus, including our financial statements and the related notes and “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” before deciding whether to invest in our common stock. The occurrence of any of the events or developments described below could have a material adverse effect on our business, financial condition, results of operations, and prospects. In such an event, the market price of our common stock could decline, and you may lose all or part of your investment. Additional risks and uncertainties not presently known to us or that we currently believe are not material may also impair our business, financial condition, results of operations, and prospects.
Risks Relating to Our Business and Industry
We have a history of net losses, and we expect to incur additional losses in the foreseeable future.
We have incurred net losses since inception, and we expect to incur additional losses in the foreseeable future. For the year ended December 31, 2023, we incurred net losses of $11.3 million and expect additional losses in 2024. As of December 31, 2023, we had an accumulated deficit of $110.5 million and we expect to incur additional losses in the foreseeable future. Our accumulated deficit reflects significant front-end spending and investment related to both completed and ongoing key operational milestones, including: (i) the initial and continued development of CapsoCam Plus and CapsoCloud, our cloud-based platform; (ii) development of our next pipeline capsule endoscope, CapsoCam Colon; (iii) initial development and ongoing improvements to our AI assisted reading tools and technologies; and (iv) funding of completed and ongoing related clinical and other studies.
As we execute on our business strategy to grow our business and revenues (including seeking FDA 510(k) clearance for CapsoCam Colon), we will continue to incur development costs and clinical study expenses and will make additional investments. Additionally, as a public company, we will incur significant legal, accounting and other expenses that we did not incur as a private company. Accordingly, we expect to continue to incur losses for at least the near-term and we may never achieve profitability or, if we do achieve profitability, sustain profitability. Our failure to achieve and sustain profitability in the future would make it more difficult to finance our business and accomplish our strategic objectives, which would have a material adverse effect on our business, financial condition and results of operations.
Our audited financial statements for the year ended December 31, 2023 include a footnote raising substantial doubt about our ability to continue as a “going concern” and, even if this offering is successful, we may still need to raise additional financing to fund our business and revenue growth plans.
Our 2023 audited financial statements include a footnote raising substantial doubt about our ability to continue as a going concern. We have funded our historical net losses and negative cash flows through the issuance of convertible preferred stock and common stock. As of December 31, 2024, we had cash of approximately $9 million, including $20.4 million raised in 2023 and $15.0 million raised in 2024 through the issuance of preferred stock to our investors.
Our ability to continue as a going concern depends on a number of factors including the success of this offering, whether we can successfully implement our business and revenue growth plans to generate sufficient cash flow from operating activities, and our ability to raise as needed other equity financing or loan facilities.
Even if this offering is successful, we may need to undertake additional capital-raising activities sufficient to fund our operations and investments to grow our revenues as contemplated by our growth strategy. However, there is no certainty regarding the occurrence, magnitude, or timing of these capital-raising activities. If we raise additional funds through the sale of equity, convertible debt, or other equity-linked securities, our stockholders’
12
Table of Contents
ownership will be diluted. The inability to secure adequate financing on favorable terms, or at all, could have a material adverse effect on our financial position, results of operations, cash flows, and ability to achieve our intended business objectives. See the risk factor titled “We may require additional capital to support business growth, and this capital might not be available on terms favorable to us, or at all, and may dilute existing stockholders’ ownership of our common stock” below.
Given the substantial doubt about our ability to continue as a going concern, potential investors must exercise caution when evaluating our financial condition, results of operations, and business prospects. Our inability to raise sufficient financing or meet our strategic plans could result in a material adverse effect on our financial position, results of operations, cash flows, and ability to achieve our intended business objectives.
All of our revenues to date have been, and in the near-term will continue to be, generated from CapsoCam Plus related sales for the small bowel; and our ability to grow our small-bowel-related revenue is subject to our ability to successfully and timely execute related elements of our revenue growth strategy.
Near term actions to grow our CapsoCam Plus small-bowel-related revenues include those set forth below. We may be unable to timely and effectively execute these or other activities designed to increase our small-bowel-related revenues.
| • | Leveraging the patient and provider benefits of CapsoCam Plus capsule endoscopy solution (which includes the associated software products, CapsoCloud and CapsoView) to effectively compete against competitors, such as (i) Medtronic, with its comparatively greater overall brand recognition, financial resources and other competitive advantages; and (ii) the U.S. subsidiary of a non-U.S. (Chinese) competitor who was the first to use AI-driven lesion detection software to support its small bowel capsule endoscopy system. |
| • | Further penetrating the small bowel market in the U.S. and internationally, including by (i) increasing the size and effectiveness of our U.S. and international sales teams; (ii) pursuing the pediatric market (with children comprising a significant portion of the Crohn’s disease patient population) following FDA clearance in December 2024 for this newly indicated patient population; (iii) introducing complementary products such as our (a) capsule delivery device (availability expected in 2025) and (b) patency capsule (for verifying a capsule endoscope can pass through the bowel without retention prior to an exam) (tentative FDA 510(k) submission planned by mid-2025); and (iv) facilitating increased telemedicine adoption following FDA clearance in December 2024 of remote ingestion of our CapsoCam Plus, allowing patients to ingest our capsules in the comfort of their own homes with remote provider supervision. |
| • | Incorporating our AI assisted reading technology into our CapsoCam Plus capsule. We are currently (i) conducting related clinical studies to demonstrate the benefits of our AI technology as incorporated into CapsoCam Plus; (ii) planning to submit the related 510(k) application to the FDA in the first half of 2025; and (iii) targeting late 2025 for FDA clearance with commercialization planned to begin shortly thereafter. |
Each of these activities is subject to numerous risks. For example, (i) we may be unable to compete effectively against our competitors; (ii) the introduction of our capsule delivery device and patency capsule may be delayed or these products may not be sufficiently adopted by providers and patients initially or in the longer term; and (iii) we may fail to obtain all required FDA and EU approvals for incorporating our AI assisted reading technology into our CapsoCam Plus solution.
For additional information, see the other risks set forth under this “Risk Factors” section including “We operate in a highly competitive industry, and competitive pressures could have a material adverse effect on our business, financial condition, results of operations, and prospects,” “Our longer term efforts to expand our GI-tract capsule endoscopy solution beyond small bowel and colon pathologies and medical conditions may not
13
Table of Contents
succeed,” and “We may face regulatory challenges and delays in obtaining 510(k) clearance for marketing of those CapsoCam capsules incorporating our self-developed AI technology, including our CapsoCam Colon capsule endoscopy solution and CapsoCam Plus capsule endoscopy solution (small bowel).”
CapsoCam Colon is our initial strategic effort to expand revenues for our GI-tract capsule endoscopy solution beyond small-bowel-related sales; we do not currently expect to generate CapsoCam Colon-related revenues until mid-2026 based on the anticipated timing for receiving FDA clearance for the second generation of our CapsoCam Colon solution.
Our CapsoCam Colon capsule endoscopy solution (which includes associated software products, CapsoCloud and CapsoView) is our initial strategic effort to grow our revenues beyond small-bowel-related sales. We do not expect to generate CapsoCam Colon related sales in the U.S. or internationally until after (i) obtaining initial FDA 510(k) or EU clearance, as applicable, for both our first and second generation of our CapsoCam Colon capsule for use by the indicated patients (who are a subset of the colorectal cancer screening and surveillance population) and (ii) satisfying the related conditions for obtaining these clearances.
Both generations of our CapsoCam Colon solution incorporate our AI assisted reading technology. Notably, the planned second-generation product will incorporate improvements—such as a new lens and illumination optics with an increased field of view and improved image quality—designed to increase the accuracy (measured in terms of polyp-detection sensitivity and specificity) and benefits of using our CapsoCam Colon solution to visualize the colon and detect and measure polyps. We plan to focus our regulatory efforts on first obtaining U.S. FDA clearance followed by obtaining EU clearance. We plan to use the clinical results of the second arm of our pivotal study to support submission to the FDA of a 510(k) application in late 2025 with a goal of obtaining FDA 510(k) clearance for the second-generation product in the first half of 2026. Our 510(k) submission and FDA review thereof may be delayed and we may not receive 510(k) clearance from the FDA on a timely basis or at all. Related risks with respect to, among other things, receiving requisite regulatory clearances and the timing thereof and commercializing our CapsoCam Colon capsule endoscopy solution are described further below in these “Risk Factors.”
Broad adoption of our CapsoCam Colon solution (once cleared) and growing related revenues depend on, among other things, expanding beyond CapsoCam Colon’s initial indicated patient population, increasing the accuracy of our initial CapsoCam Colon solution, and achieving and maintaining a satisfactory successful completion rate for capsule endoscopies using our CapsoCam Colon solution; and we may fail to successfully and timely execute related elements of our CapsoCam Colon adoption and revenue growth strategy.
Broader adoption of our CapsoCam Colon capsule endoscopy solution by providers and patients (once cleared) and growing related revenues would involve, among other things:
| • | increasing the indicated patient population for our CapsoCam Colon solution. The initial indicated patient population is a subset of the colorectal cancer screening and surveillance populations. More specifically, GI patients with (i) major risks for colonoscopy or moderate sedation coupled with evidence of GI bleeding of lower GI origin and (ii) an incomplete colonoscopy with adequate bowel preparation. As part of seeking FDA clearance for the planned second generation of our CapsoCam Colon solution, we plan to seek FDA clearance to remove the GI bleeding requirement. Successful removal of the GI bleeding requirement would significantly increase CapsoCam Colon’s addressable market. There is no assurance that the FDA will permit this request or any future request to expand the indicated patient population for our CapsoCam Colon solution on a timely basis or at all. Any such request would involve significant effort and require, among other things, related regulatory submissions and our conducting non-clinical and clinical studies to generate data and evidence to support the request; |
| • | continued enhancements and improvements to our CapsoCam Colon solution to increase its accuracy (measured in terms of polyp-detection sensitivity and specificity) and benefits of using CapsoCam Colon to visualize the colon and detect and measure polyps. Our success in implementing these enhancements |
14
Table of Contents
| and improvements (including obtaining related FDA 510(k) and EU clearances) is critical to (i) our initial and ongoing commercialization efforts (including efforts to increase provider and patient acceptance of our solution); (ii) our ability to increase CapsoCam Colon’s indicated patient populations; and (iii) consequently, increasing that portion of the colorectal cancer screening, surveillance, and diagnostic patient populations for which our CapsoCam Colon may be used. We may be unable to successfully make continued enhancements and improvements to sufficiently increase the accuracy and benefits of using our CapsoCam Colon system on a timely basis or at all; and |
| • | achieving and maintaining a satisfactory and successful examination completion rate for capsule endoscopy using CapsoCam Colon. A successful completion rate for CapsoCam Colon endoscopies much lower than that for traditional colonoscopies may (i) limit the patient population for whom CapsoCam Colon is deemed appropriate; (ii) limit its acceptance by providers and patients; and (iii) result in an unacceptable number of patients requiring a follow-on colonoscopy or other diagnostic test to confirm that they lack significant colon polyps or do not have cancer. A successful colon capsule endoscopy involves successful imaging of either (i) a patient’s entire colon without detecting significant polyps or (ii) a sufficient portion of the patient’s colon to identify the presence of one or more significant polyps (i.e., indicating preventative or treatment driven colonoscopy is recommended). Capsule colonoscopy procedures are unsuccessful most often due to inadequate bowel preparation and/or slow capsule transit through the colon. CapsoCam Colon’s successful completion rate (as demonstrated in our pivotal study) is supported by a physical design that encourages timely transit the GI-tract and a battery lifetime that is currently twice the competition’s. However, the bowel preparation, which is similar to that used for colonoscopy, produces a level of cleansing that is patient specific, and capsule endoscopy does not afford an opportunity, as colonoscopy does, for targeted additional cleansing by the physician during the exam. Although we are positioned to benefit from improvements in bowel preparation medications on the market, these developments are outside our business and difficult to predict. |
We may be unable to timely and effectively drive broad provider and patient adoption of our CapsoCam Colon solution (once cleared) and grow related revenue. Each of the above activities is subject to numerous risks described above or elsewhere in these “Risk Factors.”
Our longer term efforts to expand our GI-tract capsule endoscopy solutions beyond small bowel and colon pathologies and medical conditions may not succeed.
We believe our CapsoCam Plus (small bowel) and CapsoCam Colon solutions, incorporating our panoramic imaging solution, can be adapted to address new GI medical indications. Potential new medical indications include pancreatic cancer and esophageal medical conditions (such as esophageal varices and Barrett’s esophagus) with sales to our customers supported through our existing sales and marketing organization. In 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in (i) detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) and (ii) screening esophageal varices in cirrhotic patients with portal hypertension. These expansions require substantial investment in research and development, rigorous clinical studies or trials, and regulatory clearance, any of which may delay or impede the planned indication expansion for our capsule endoscopy solution. In connection with our efforts to address pancreatic cancer, we may seek an FDA “Breakthrough Device Designation” for our capsule endoscopy solution. A Breakthrough Device Designation prioritizes a device in the FDA’s review queue for all future regulatory submissions and accelerates communications (i.e., negotiations and feedback) with the FDA, thereby expediting the marketing application process. However, our proposed capsule endoscopy solution may not meet the eligibility requirements for this designation and, even if a Breakthough Device Designation is received, it may not receive the FDA authorization required to market the proposed capsule endoscopy solution.
We face significant competition in the capsule endoscopy market for our current and planned medical conditions. Our success in expanding our capsule endoscopy solution is contingent upon our ability to execute on our related strategies, differentiate our technology with initial or improved product accuracy (such as sensitivity
15
Table of Contents
and specificity) and features, and secure acceptance in the GI medical community. Any failure in these areas could significantly impede our growth prospects and adversely affect our financial performance. There is no assurance that our efforts will yield successful outcomes in addressing pancreatic cancer, esophageal medical conditions or other targeted medical conditions.
We may not obtain or may experience delays in obtaining 510(k) clearance for our initial CapsoCam Colon capsule endoscopy solution or our planned second generation of our CapsoCam Colon solution (with improved optics and other components), which would adversely impact our ability to commercialize this product and generate related revenue.
The 510(k) application process can be lengthy. There is no guarantee that the FDA will grant 510(k) clearance for our initial or planned second generation CapsoCam Colon capsule endoscopy solution, and their timeline for doing so may not align with our planned commercialization schedule.
Although our CapsoCam Plus solution, which visualizes the small bowel, has been cleared by the FDA, the CapsoCam Colon capsule endoscopy solution will require a distinct, unrelated regulatory submission, as it is subject to different statutory requirements. This is due to its use in the colon (part of the lower GI-tract) and incorporation of new capabilities (including AI assisted reading technology) to facilitate visualization of the colon and to identify and measure polyps, each of which introduces different safety and effectiveness considerations. As a part of the marketing submission review, the FDA may require additional analysis, data, studies, or modifications to our products or labeling, which could increase our costs, delay our launch, or limit our market opportunity. The FDA may also require other conditions on our products to achieve initial clearance, such as additional studies or clinical trials, labeling restrictions, or post-market surveillance commitments, which could increase our regulatory burden and liability. Risks related to the separate FDA clearance of the AI assisted reading technology incorporated in our CapsoCam Colon capsule endoscopy solution and CapsoCam Plus capsule endoscopy solution are discussed in the risk factor titled “We may face regulatory challenges and delays in obtaining 510(k) clearance for marketing of those CapsoCam capsules incorporating our self-developed AI technology, including our CapsoCam Colon capsule endoscopy solution and CapsoCam Plus capsule endoscopy solution (small bowel)” below.
The FDA is required by statute to review 510(k) applications within defined timelines. For most 510(k) submissions, a 90-day maximum review period is mandated. While this requirement provides predictability for the review process, it also places constraints on available opportunities to address any questions posed by FDA. Factors critical to a timely review and clearance of our 510(k) applications include (i) the quality of our regulatory submission (including both in terms of clarity in presentation and the supportiveness of data) and (ii) the availability of qualified FDA review resources (i.e., individuals possessing relevant technical understanding and requiring a minimal “learning curve”). Although we have (x) engaged with the FDA in early communications (including several pre-submission meetings) regarding our CapsoCam Colon solution and (y) considered FDA feedback during product development, there is no guarantee that the FDA staff members involved in those discussions and/or who are familiar with the CapsoCam technology will be available to participate in the post-submission review of our 510(k) application. The involvement of less experienced FDA staff members may result in a “learning curve” potentially reducing the effective time available to the FDA to critically review our submissions. Likewise, changes in review staff levels or workload—possibly due to changes in FDA funding or operations or the emergence of competing public health priorities (e.g., addressing a device shortage)--may lead to review delay and an inability to complete the FDA review process within the statutorily mandated period, potentially leading to a longer review process for our applications.
Failure to obtain or delays in obtaining 510(k) clearance for our CapsoCam Colon capsule endoscopy solution could result in additional costs or liabilities and adversely impact our reputation, ability to sell our CapsoCam Plus capsule endoscopy solution, and our ability to compete effectively in this or other markets. Any of these outcomes could have a material adverse effect on our business, financial condition, and results of operations.
16
Table of Contents
Our revenue and business growth depends on the success of our CapsoCam capsule endoscopy solution (including our CapsoCam Colon capsule once cleared).
Our ability to capture the market opportunity for our GI-tract capsule endoscopy solution depends on several factors, including the following:
| • | acceptance in the GI medical community; |
| • | the number of patients screened or tested for specific pathology or medical indication and the number of patients who use our CapsoCam capsules for that purpose; |
| • | our ability to introduce enhancements and improvements aimed at increasing the value provided by our GI-tract capsule endoscopy solution; |
| • | our ability to develop and commercialize our CapsoCam capsule endoscopy solution for new indications, patient populations and clinical use cases; |
| • | our ability to successfully complete any required clinical or other studies and obtain and maintain any required regulatory approval or clearances; |
| • | insurer and third-party reimbursement of the costs associated with our GI-tract capsule endoscopy solution; |
| • | successful management of our global supply chain including our component suppliers and assembly manufacturers for our CapsoCam solution, many of which are located in Asia (particularly Taiwan and Japan) and some of which are currently single-source suppliers; |
| • | successful growth and leveraging of our global sales team (including, where appropriate, distributors) and marketing team to sell and market our GI-tract capsule endoscopy solution; and |
| • | the amount and nature of competition from other GI-tract diagnostic products or procedures. |
Our failure to effectively manage our business through the various challenges we face, may result in an inability to execute on our business plan and revenue growth strategies, respond to competitive pressures, take advantage of market opportunities, satisfy customer requirements, or maintain high-quality products, which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We may face regulatory challenges and delays in obtaining 510(k) clearance for marketing of those CapsoCam capsules incorporating our self-developed AI technology, including our CapsoCam Colon capsule endoscopy solution and CapsoCam Plus capsule endoscopy solution (small bowel).
We plan to submit to the FDA the clinical results of the first arm of our CapsoCam Colon pivotal study in a 510(k) submission in the first half of 2025. This submission will seek FDA clearance of our initial CapsoCam Colon capsule endoscopy solution which incorporates our self-developed AI technology. Similarly, in the first half of 2025, we plan to seek FDA 510(k) clearance for the use of AI in our small bowel capsule, CapsoCam Plus.
The incorporation of AI technology into our GI-tract capsule endoscopy solution (including the associated software products, CapsoCloud and CapsoView) requires appropriate FDA regulatory authorization, supported by requisite clinical and other studies, prior to its commercialization in the U.S. Under the FDA’s current regulatory framework, our capsule endoscopy solution and the AI technology are assessed separately, with each component falling under a different regulatory classification and subject to distinct regulatory requirements. This bifurcated approach, set by precedent, ensures that each component meets the necessary safety, effectiveness, and performance expectations.
In the case of CapsoCam Colon, as of the date of this prospectus, the FDA has only authorized one colon capsule imaging system, the Medtronic PillCam COLON 2, introduced in 2014, which does not include any
17
Table of Contents
AI-driven analytics; and one AI-driven capsule endoscopy analysis software, the AnX Robotics NaviCam ProScan, introduced in 2023, which exclusively supports AnX Robotics’ small bowel capsule system. These devices will respectively serve as predicate devices for our CapsoCam Colon 510(k) submission. In particular, to receive initial FDA clearance for our first generation CapsoCam Colon, the clinical results must demonstrate that:
| • | CapsoCam Colon, considered without the integration of AI, demonstrates a polyp-detection accuracy that is, at a minimum, comparable to the performance of the predicate device (i.e., Medtronic PillCam COLON 2); and |
| • | our AI technology can (i) reliably and accurately identify and analyze images and video of the colon to detect abnormalities as quantified by diagnostic accuracy measures such as sensitivity and specificity, and, in doing so, (ii) aid qualified physicians in achieving improved diagnostic performance relative to not using AI. |
Clearance of the incorporated AI technology is unlikely to occur if the CapsoCam Colon solution, absent AI, fails to meet the applicable regulatory requirements. In connection with FDA clearance of our planned second generation of CapsoCam Colon, which will incorporate various hardware and image processing improvements, we will need to demonstrate that the incorporated AI technology is not compromised by these modifications.
The FDA may not agree that our AI technology meets these evidentiary requirements, or it may require us to submit additional data or information over the course of regulatory review to support the safety and effectiveness of CapsoCam Colon (including the incorporated AI technology). Also, the FDA may determine that CapsoCam Colon (including the incorporated AI technology) is not eligible for the 510(k) pathway and that we must utilize a different, more rigorous and costly regulatory pathway, such as a PMA or a De Novo request, which could significantly delay our ability to market CapsoCam Colon (including the incorporated AI technology), increase our development costs and reduce our competitive advantage. The FDA recognizes that there are challenges within the current regulatory framework for medical devices incorporating artificial intelligence and is actively pursuing regulatory reforms to better foster innovation while maintaining patient safety. We proactively monitor new FDA policies and programs and will update our regulatory strategy to reduce regulatory uncertainty and burden where applicable.
In seeking 510(k) clearance for our AI technology as incorporated into CapsoCam Plus we are (i) utilizing the NaviCam ProScan as the predicate device (confirmed with the FDA) and (ii) currently conducting a retrospective clinical study of the CapsoCam Plus solution with the AI technology incorporated to analyze in vivo videos from completed, real-world clinical cases, to assess the performance of the AI technology for small bowel. Similar to the first arm of our CapsoCam Colon pivotal study, the study seeks to demonstrate that the AI technology can (i) reliably and accurately identify and analyze images and video of the small bowel to detect abnormalities as quantified by diagnostic accuracy measures such as sensitivity and specificity, and, in doing so, (ii) aid qualified physicians in achieving improved diagnostic performance relative to not using AI.
These regulatory considerations may apply to updates or improvements to previously cleared AI technology, such as major changes in algorithm design, significant changes in clinical performance, or the addition of new functions (e.g., detection of new types of abnormalities). In these scenarios, we will be expected to demonstrate the safety and performance of the updated technology through the appropriate studies and pursue regulatory clearance through additional FDA filings prior to commercial distribution. Also, changes to the device hardware or imaging processing methods contained in CapsoCam capsules incorporating our AI technology may affect the performance of the incorporated AI technology, even if the related algorithm is not directly changed. As a result, we may be required to demonstrate that performance of the incorporated AI technology is not degraded by those critical capsule component changes and obtain regulatory clearance prior to distribution.
18
Table of Contents
Any failure or defect in our GI-tract capsule endoscopy solution could harm our reputation, expose us to liability, and reduce our sales.
Our GI-tract capsule endoscopy solution comprises our CapsoCam capsules and the associated software (CapsoCloud and CapsoView), including their related components. We may encounter technical, operational, or regulatory challenges or limitations in developing, testing, validating, or implementing improvements to our GI-tract capsule endoscopy solution, which could delay or prevent us from achieving our desired outcomes. Moreover, related components (including the lens module, image senser, processor and storage module) and incorporated AI technology may fail or malfunction due to defects, errors, bugs, viruses, cyberattacks, or other causes, which could compromise the quality, accuracy, or reliability of our products and the images and analyses they produce. Any failure or defect in our GI-tract capsule endoscopy solution (including capsule components and the associated software, CapsoCloud and CapsoView) could harm our reputation, expose us to liability, and reduce our sales. Any of these outcomes could have a material adverse effect on our business, financial condition, and results of operations.
We rely on various suppliers to assist us in the assembly and manufacture of our GI-tract capsule endoscopy solution and sourcing of critical and other components and many of these suppliers are single source suppliers located in Asia (particularly Taiwan and Japan); any disruption in our supply chain could adversely affect our ability to meet the demand for our products and fulfil our orders.
To assist us in manufacturing our GI-tract capsule endoscopy products we rely on component suppliers and assembly manufacturers based in Asia (particularly Taiwan and Japan). Critical components found in our CapsoCam capsules include lens modules from Largan, complementary metal oxide semiconductor (“CMOS”) image sensors from Toshiba, and application-specific integrated circuits (“ASICs”) from Moai Electronics Corporation. Many of our suppliers/manufacturers are single source and located in Asia. For example, the single source suppliers of our lens modules and ASICs are located in Taiwan and the single source supplier of our CMOS image sensors is located in Japan. Currently, assembled CapsoCam capsules are shipped by Largan from Taiwan to our U.S. facility where we complete the manufacturing process before distributing the capsules to our distribution network. Any disruption to our supply chain could significantly harm our ability to effectively manufacture and deliver our CapsoCam capsules and, in turn materially harm, our financial results.
The numerous supply chain related risks we face that could adversely impact our ability to manufacture and sell our CapsoCam capsules include:
| • | our supplier may be unable to meet our demand requirements, including demands for desired improvements or upgrades, or may prioritize the orders of its other customers; |
| • | our supplier may be unable to meet our quality standards, specifications, or requirements for critical components, or may experience defects, errors, or failures in these components, which could compromise the quality, performance, or safety of our CapsoCam capsules and the images and analyses they produce; |
| • | our supplier may increase or seek to increase prices, or impose unfavorable purchase terms or conditions, which could increase our costs and reduce our margins; |
| • | our supplier may discontinue the production of critical components, or cease to do business with us, for any reason; |
| • | it may be difficult for us to find an alternative or second supply source, which may not be available, affordable, or compatible with our CapsoCam capsules and, for commercial reasons, we may prefer or be required to work on an exclusive basis with our supplier; and |
| • | supplier concentration in Asia (particularly Taiwan and Japan) subjects us to heightened risks related to geopolitical tensions (particularly between China and Taiwan), potential trade disputes and imposition of tariffs or threats of tariffs, transportation interruptions, natural disasters, and manufacturing constraints, any of which can affect the manufacture and delivery of our CapsoCam capsules and lead to delays, |
19
Table of Contents
| increased costs, and potential shortages of our CapsoCam capsules and critical components. For additional information, see the risk factor titled “We face substantial geopolitical and trade risks (including a possible trade war and imposition of tariffs and supply chain interruptions) and natural disaster risks associated with our business operations in Asia (particularly Taiwan and Japan)” below. |
Although we are in the process of planning or implementing various mitigation measures to address supply chain risks (including qualifying a backup supplier for certain critical components and looking to build reserve supplies of capsules and critical components to address unanticipated delays), these efforts may not be successful. For example, implementation of mitigation measures to address the supply chain disruption of CMOS image sensors would be complicated, and resolving an industry-wide issue relating to CMOS image sensors would require a significant amount of time and coordinated efforts across multiple industry participants.
In addition, our operations depend heavily on expedited and reliable shipping services to ensure the secure and timely delivery of our products to customers. Any delivery performance issues, such as loss, damage, or destruction of our CapsoCam capsules or the components thereof, would incur substantial costs for timely replacement and could tarnish our reputation, leading to decreased demand for our products and increased operational expenses. Additionally, significant increases in shipping rates could negatively affect our operating margins and overall financial performance. Service interruptions caused by strikes, severe weather, natural disasters, or other disruptions could further impede our ability to process orders promptly. These factors collectively pose a risk to our business, financial condition, and operational results, and can potentially damage our reputation.
We face substantial geopolitical and trade risks (including a possible trade war and imposition of tariffs and supply chain interruptions) and natural disaster risks associated with our business operations in Asia (particularly Taiwan and Japan).
We face substantial geopolitical risks due to our business operations in Asia (particularly Taiwan and Japan). Our major assembly manufacturers are located in Taiwan. Most of our single-source suppliers of critical components of our products are located primarily in Taiwan and Japan. Political events, trade and other international disputes, geopolitical tensions, conflict, terrorism, public health issues, industrial accidents and other business interruptions in these areas can have a material adverse effect on our business operations. Restrictions on international trade, such as tariffs and other controls on imports or exports of goods, technology or data, can materially adversely affect our business and supply chain. Restrictive measures can increase the cost of our products, and can require us to take various actions, including changing suppliers and restructuring our supply chain, business relationships and operations. Changing our business and supply chain in accordance with new or changed restrictions on international trade can be expensive, time-consuming and disruptive to our operations. Such restrictions can be announced with little or no advance notice, which can create uncertainty, and we may not be able to effectively mitigate all adverse impacts from such measures. For example, an increase in the recent tensions between mainland China and Taiwan and the possibility of instability and uncertainty caused by prolonged or regular military drills in the Taiwan Strait may result in disruptions in the overall trading environment, and in turn result in higher transportation cost and interruption in delivery of our products or its critical components from our single-source suppliers in Taiwan, adversely affecting our business and financial condition.
In addition to political risks, Taiwan and Japan are susceptible to natural disasters, such as earthquakes and typhoons, that could disrupt the normal operations of our business and adversely affect earnings. There can be no assurance that future natural disasters will not occur and result in major damages to our component suppliers and assembly manufacturers (many of whom are single-source and located in Taiwan and Japan), which could have a material adverse effect on our business, financial condition and results of operations.
20
Table of Contents
If Medicare and other third-party payors, including managed care organizations, do not approve reimbursement for our products at adequate reimbursement rates, we may be unable to successfully commercialize our products which would likely have a material adverse effect on our business.
Our ability to increase sales of our products depends, in significant part, on the availability of adequate coverage and reimbursement from third-party payers, including governmental payers, managed care organizations, and private health insurers. If Medicare and other third-party payors do not approve reimbursement for our CapsoCam capsules at adequate reimbursement rates, we may be unable to successfully commercialize our products which would have a material adverse effect on our business.
In particular, we believe that obtaining a positive national coverage decision and favorable reimbursement rate from the Centers for Medicare and Medicaid Services (“CMS”) for our CapsoCam capsules will be a necessary element in achieving material commercial success. These third-party payors are increasingly attempting to contain healthcare costs by limiting both coverage and the level of reimbursement for new healthcare products approved for marketing by the FDA. As a result, there is significant uncertainty surrounding whether the use of capsules will be eligible for coverage by third-party payors or, if eligible for coverage, what the reimbursement rates will be for those products.
CMS currently reimburses properly submitted reimbursement claims for most capsule endoscopies utilizing our small bowel CapsoCam Plus capsule. For CapsoCam Colon, we expect to qualify for CMS reimbursement for our initial indicated patient population. CMS currently provides reimbursement for capsule endoscopy procedures for patients with major risks for colonoscopy or moderate sedation with evidence of GI bleeding of lower GI origin and patients who have had an incomplete colonoscopy with adequate preparation. As a part of our 510(k) FDA application for our second generation CapsoCam Colon capsule and to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation. Successfully growing our CapsoCam Colon revenues will depend on our success in timely securing CMS reimbursement for this and any newly indicated patient population.
Reimbursement of capsule endoscopy by a third-party payor may depend on a number of factors, including a payor’s determination that products using our technologies are: effective for detecting or diagnosing gastrointestinal disorders; not experimental or investigational; approved by the major guidelines organizations; reliable, safe and minimally invasive; medically necessary; appropriate for the specific patient; and cost-effective.
If we are unable to obtain positive policy decisions from third-party payors, including Medicare and managed care organizations, approving reimbursement for our CapsoCam capsules, including our CapsoCam Colon, at adequate levels, the commercial success of these products would be compromised and our revenues would be significantly limited. Moreover, coverage policies and reimbursement rates are subject to change and we cannot guarantee that even if we initially achieve adequate coverage and reimbursement rates that they will be applicable to our products in the future. Any failure to obtain or maintain favorable reimbursement could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We may face expansion risks when expanding in existing and entering into new foreign markets.
Our growth strategy includes increasing our international revenues and potential entry into new international markets. In 2023 and 2024, international sales accounted for approximately 26% and % of our revenues, respectively. We have sales in each of the EU 7 countries with Germany (our largest international market), Sweden and the U.K. being our largest international markets. Some of our existing and new international markets may be highly regulated and competitive. However, we may face significant challenges and risks in expanding in
21
Table of Contents
existing and entering into new international markets (including risks related to expanding our market share and customer base), such as the following:
| • | The need to grow our sales and marketing team (including potentially through qualified distributors) in these markets. |
| • | The need to comply with complex and evolving regulatory requirements, quality standards, and post-market surveillance obligations, which may entail significant time, cost, and resources, and may subject us to regulatory actions, penalties, or product recalls if we fail to meet them. |
| • | The need to compete with established and emerging players in the capsule endoscopy market, which may have greater financial capabilities, brand recognition, market access, distribution networks, or technological capabilities, and which may offer lower prices, superior features, or better customer service. |
| • | The need to secure adequate reimbursement and coverage policies from public and private payers, which may affect the affordability, accessibility, and adoption of our CapsoCam capsule endoscopy solution, and which may be subject to changes, limitations, or uncertainties due to budget constraints, policy reforms, or competitive pressures. |
We may fail to successfully execute on our growth strategy, overcome these challenges and risks, or achieve our expected returns on our investments in these foreign markets. In addition, we may encounter unforeseen difficulties or liabilities related to our operations, supply chain, intellectual property, or litigation in these foreign markets. Any of these factors could negatively impact our sales, margins, reputation, or market position in new foreign markets, and as a result, our business, results of operations and financial condition.
The clinical results of our various clinical studies (including those for our CapsoCam Colon capsule) may not be released in any peer-reviewed publications.
We are currently evaluating the results of the first arm of our pivotal study for CapsoCam Colon and plan to submit these results to the FDA as part of an initial 510(k) clearance submission for our “first generation” product. We will do the same with the clinical results of the second arm of the pivotal study to evaluate our second generation CapsoCam Colon capsule (incorporating improvements such as a new lens and illumination optics with an increased field of view and improved image quality).
From time to time, we may publicly disclose interim, top-line or preliminary data from our pivotal studies (including those related to our CapsoCam Colon pivotal study included in this prospectus), which are based on a preliminary analysis of then-available data, and the results and related findings and conclusions are subject to change following a more comprehensive review of the data related to the pivotal study or additional data collected at a later time. We also make assumptions, estimations, calculations, and conclusions as part of our analyses of data, and we may not have received or had the opportunity to fully and carefully evaluate all data. The data from a study may not be released in any peer-reviewed publication, either initially or in the future. As a result, the interim, top-line, or preliminary results that we report may differ from future results of the same trial, or different conclusions or considerations may qualify such results, once additional data have been received and fully evaluated. The FDA may not accept or agree with our assumptions, estimates, calculations, conclusions, or analyses or may interpret or weigh the importance of data differently, which could impact the value of our study (including our CapsoCam Colon pivotal study), or the approvability or potential for commercialization.
We do not plan to release the clinical results from our various clinical studies (including the first arm or second arm of our CapsoCam Colon clinical study and our planned CapsoCam Plus clinical study (AI)) before submitting, as applicable, the related 510(k) clearance submission to the FDA. Typically, the submission of study results for publication in a peer-reviewed journal occurs after its use for an FDA submission. The initial or future absence of peer-reviewed research could hinder the acceptance of our GI-tract capsule endoscopy solution products within the medical community, potentially affecting its credibility and market adoption.
22
Table of Contents
Furthermore, the results of the utilization of our GI-tract capsule endoscopy solution (including CapsoCam Colon) may not be accepted or replicated by the scientific or medical community, or may be challenged or disputed by our competitors or other third parties. Physicians and third-party payors may be skeptical of new technologies and may require substantial evidence of clinical efficacy and cost-effectiveness before adopting new products. If the results of our research and clinical studies and our sales and marketing activities relating to communication of these results do not convince thought-leading gastroenterologists, guidelines organizations, primary care physicians, third-party payors, and patients that our GI-tract capsule endoscopy solution (including CapsoCam Colon) is reliable, effective, and superior to existing screening methods, we may not achieve the necessary market acceptance. The publication of negative or inconclusive results, or the emergence of new or conflicting data, could undermine the validity or reliability of our study and our GI-tract capsule endoscopy solution, and could reduce the demand for or acceptance of our CapsoCam capsules by physicians, patients, payors, or other stakeholders. Any of these outcomes could have a material adverse effect on our business, financial condition, and results of operations.
We may face risks associated with our use and development of artificial intelligence, including our AI technology.
Clinicians are subject to fatigue and distraction and have varying degrees of expertise in identifying GI-tract abnormalities. Our AI assisted reading tool is designed to (i) reduce viewing times for clinicians by highlighting the areas of interest (via a bonding box placed around suspected abnormalities) and (ii) improve diagnostic yield and provide more consistent accuracy by providing suggestions or recommendations. However, risks related to the use of our AI technology include:
| • | our AI assisted reading technology may fail to accurately identify all or a sufficient portion of the patient’s GI-tract abnormalities (measured in terms of sensitivity and specificity); |
| • | a clinician may (i) place undue reliance on our AI assisted reading technology to identify GI-tract abnormalities (e.g., polyps in colon) or (ii) fail to fully utilize product features designed to supplement a clinician’s review (such as the ability to review frames adjacent to an AI bonded frame identifying a suspected GI-tract abnormality or view the video in non-AI mode); |
| • | our AI technology may not prevent or reduce the occurrence or impact of clinician alertness fatigue; and |
| • | product enhancements and improvements (including improvements to our AI algorithms) may fail to deliver improved viewing, accuracy improvements (including as measured in terms of sensitivity and specificity) and other expected benefits. |
We plan to further invest in AI to improve our products. However, we may not achieve the expected benefits from our AI investments, or we may face technical, regulatory, ethical or legal challenges that could limit or delay the development, deployment or adoption of our AI technology. Additionally, we may encounter difficulties in integrating our AI technology with our existing or future products, software or platforms, or in ensuring the compatibility, interoperability or reliability of our AI technology.
Our AI technology may introduce errors or biases that could compromise the quality or validity of the clinician’s viewing. In particular, for data collected by CapsoCloud to serve as the clinically derived reference standard (“ground truth”) for enhancing AI capabilities, it is imperative that the collected data be meticulously labeled. This labeling process is resource-intensive and necessitates the involvement of qualified gastroenterologists who meticulously review and annotate the images. The accuracy of these labels is crucial, as it directly impacts the AI’s ability to make precise and reliable predictions based on the data. If the models underlying our AI technologies are incorrectly designed or implemented; trained or reliant on incomplete, inadequate, inaccurate, biased, or otherwise poor quality data (including as a result of the labeling process described above), or on data to which we do not have sufficient rights or in relation to which we and/or the providers of such data have not implemented sufficient legal compliance measures; used without sufficient oversight and governance to ensure their responsible use; misused or used outside of scope of applicable
23
Table of Contents
regulatory authorizations; and/or adversely impacted by unforeseen defects, technical challenges, cybersecurity threats, or material performance issues, the performance of our products, as well as our reputation, could suffer or we could incur liability resulting from the violation of laws or contracts to which we are a party, regulatory enforcement actions, or civil claims.
The performance of any algorithm incorporated in our AI technology will generally be assessed by comparing the output of the algorithm against a ground truth for a specified dataset. This applies to internal evaluation of an algorithm’s performance, supporting external presentations and publications, and testing to support regulatory submissions. Our algorithm output will not always agree with the opinion of a qualified gastroenterologist, and in some cases multiple qualified gastroenterologists may not agree with each other. While we constantly work to improve our GI-tract capsule endoscopy solution, our AI technology is novel and complex, and our AI technology may not perform as intended under all circumstances.
CapsoCloud is designed to automatically accumulate patient data critical to develop and train AI-based GI-tract abnormality detection and classification capabilities. If we are unable to obtain sufficient rights to use such data under applicable regulatory frameworks or our patients were to withdraw or withhold their consent, our ability to continue to develop new or improved CapsoCam capsule endoscopy products, and our revenue prospects, could be materially adversely impacted.
The regulatory landscape for AI technologies is rapidly changing as various federal, state, and international government bodies introduce or consider new laws and regulations. The FDA has issued guidance on incorporating AI into medical devices, and existing laws may be interpreted in ways that impact AI operations. This creates uncertainty around implementation standards and enforcement practices, making it difficult to predict the future impact on businesses. The Biden Administration’s Executive Order on the Safe, Secure and Trustworthy Development and Use of Artificial Intelligence outlines principles for AI design and deployment, signaling increased governmental regulation. This order established new requirements for AI model training, testing, and cybersecurity, and directed federal agencies to develop additional regulations. Agencies like the Department of Commerce and the FTC have proposed rules, and state-level legislation, such as California’s CCPA regulations on automated decision-making, is also advancing.
New laws and regulations are expected in the U.S. and other jurisdictions, potentially limiting the use of AI technologies or requiring changes that could negatively affect business performance. Compliance with these laws may necessitate significant resource expenditure, especially if regulations vary across jurisdictions. The cost of compliance could increase operating expenses due to additional reporting obligations. Any failure, actual or perceived, to comply with these laws and regulations could materially and adversely impact our business operations, financial condition, and future prospects.
Actual or perceived failures to comply with applicable data privacy and security laws, regulations, standards and other requirements could adversely affect our business, financial condition, results of operations, and prospects.
We collect, store, process and analyze large amounts of data for GI tract from our products, customers, patients, hospitals and other sources, to support our AI technology. With CapsoCloud, we have access to a rich and growing source of clinical data from patients, which enables us to develop ever-improving automated detection and classification of GI-tract abnormalities. However, the collection and use of such data involves significant risks and challenges, such as data breaches, cyberattacks, unauthorized access, theft, loss, corruption, misuse, manipulation, infringement, disclosure or destruction. Any of these events could compromise the security, privacy, integrity, availability or accuracy of our data, and expose us to legal, regulatory, reputational or financial liabilities, as well as potential claims, fines, penalties, sanctions, litigation or investigations.
We must comply with various U.S. laws and regulations that protect personal information, especially health-related information, such as the Health Insurance Portability and Accountability Act (HIPAA), which imposes
24
Table of Contents
privacy, security, and breach notification obligations and penalties on entities that handle such information. Some states, such as California, have also enacted their own privacy and security laws, which may differ from or conflict with federal laws and require additional disclosures, responses, and contracts. These laws and regulations are subject to interpretation and change, and may increase our compliance costs and risks.
We may also face data protection laws and regulations in foreign jurisdictions, such as the General Data Protection Regulation, which applies to personal data of individuals in the European Economic Area and imposes stringent requirements and fines on data processors and controllers, including for transfers of such data to third countries like the U.S. These laws and regulations may affect how we conduct business, transfer data, and provide services, and may expose us to additional costs, complaints, investigations, or fines. The Federal Trade Commission may also take action against us for misleading, deceptive, or unfair practices related to privacy and data security.
Although we work to comply with applicable laws, regulations and standards, our contractual obligations, research protocols, and other obligations, any actual or perceived failure by us or our employees, representatives, contractors, consultants, or other third parties to comply with such requirements or adequately address data privacy and security concerns, even if unfounded, could result in, among other adverse impacts, damage to our reputation, loss of customer confidence in our security measures, withdrawal or withholding of customer consent for using patient data, government investigations, and enforcement actions and litigation and claims by third parties, any of which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We operate in a highly competitive industry, and competitive pressures could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We operate in a highly competitive industry, and competitive pressures could have a material adverse effect on our business, financial condition, results of operations, and prospects. The medical device market, particularly in the area of diagnostic imaging and endoscopic procedures, is characterized by rapid technological advancements and the continuous introduction of new products.
The competition that we face for our small bowel CapsoCam Plus is primarily from traditional enteroscopy procedures performed by trained physicians in hospital or clinical settings and other capsule-based imaging solutions manufactured by companies such as Medtronic, IntroMedic, JinShan and Ankon. Those competitors include well-established companies with significant resources and brand recognition such as Medtronic, that are constantly developing and marketing innovative products that may offer superior features or lower costs. With respect to Medtronic, it also enjoys other competitive advantages including (i) a “first mover advantage” as the first manufacturer of a small bowel capsule endoscopy and a colon capsule endoscopy (as described below); (ii) exclusive supply arrangements (sometimes up to three years) with some of the larger GI practices and hospitals (particularly in the Northeast region of the U.S.) which our sales team also targets; and (iii) greater brand recognition and financial resources.
Notably, one competitor, Ankon (through its affiliate AnX Robotics), has already established a market presence with their FDA-cleared AI product for the small bowel in the U.S. and various other markets outside the U.S. This puts us at a market disadvantage until we can launch our own AI product for the small bowel, which is anticipated to occur in late 2025, subject to FDA clearance. The delay in our product launch could result in a loss of market share and reduced revenue opportunities, as potential customers may opt for the already available and proven solutions from our competitors. Additionally, the competitive landscape is further impacted by the pricing strategies of Asia-based capsule endoscopy companies, such as Ankon (China), Jinshan (China) and IntroMedic (South Korea), who are offering their products at lower than average prices. This aggressive pricing approach could pressure us to lower our prices to remain competitive, potentially impacting our profit margins.
The competition that we face for CapsoCam Colon is primarily from (i) procedure-based detection technologies such as optical colonoscopy, flexible sigmoidoscopy and CTC (or “virtual” colonoscopy); (ii) stool-
25
Table of Contents
based DNA tests such as Cologuard (initial FDA clearance in 2014); and (iii) other capsule-based imaging solutions like PillCam COLON 2 (initial FDA clearance in 2014). Other sources of competition include (a) common CRC screening tests, such as the fecal occult blood test (“FOBT”) and the fecal immunochemical test (“FIT”), and (b) screening technologies including liquid biopsy tests, such as Epi proColon (FDA approval in 2016) and C-Scan (CE Mark obtained in 2019). Those competitors include well-established companies with significant resources and brand recognition such as Medtronic, that are constantly developing and marketing innovative products that may offer superior features or lower costs. For instance, the Medtronic PillCam COLON 2 is the only U.S. FDA approved colon capsule endoscopy and is prominent in the market outside the U.S.
The combination of these factors could significantly affect our ability to attract and retain customers, thereby adversely impacting our overall business performance and future growth prospects.
Cost-containment efforts of our customers and purchasing groups could have a material adverse effect on our sales and results of operations. Consolidation in the healthcare industry or group purchasing organizations could lead to demands for price concessions, which may affect our ability to sell our products at prices necessary to support our current business strategies.
The cost-containment efforts of our customers and integrated group purchasing organizations (“GPOs”) have a material adverse effect on our sales and results of operations. In the United States, many clinics and hospitals, including some of our customers, are members of the GPOs, which are organizations that leverage the collective purchasing power of their members, such as hospitals, clinics, and other healthcare providers, to negotiate favorable pricing and terms for medical products and services. These entities negotiate pricing arrangements with medical device companies and distributors, offering these negotiated prices to affiliated hospitals and other members. GPOs typically award contracts on a category-by-category basis through a competitive bidding process, soliciting bids from multiple providers to drive down pricing or reduce the number of vendors. Due to the highly competitive nature of these contracting processes, we may face challenges in obtaining new or maintaining existing contract positions with major GPOs. The increasing leverage of the GPOs may reduce market prices for our products, thereby impacting our revenue and margins.
While securing a contract with a GPO can facilitate sales to their members and enhance our ability to meet the stringent requirements set by their members, it does not guarantee any level of sales, as purchases are typically made through individual purchase orders. Even as a sole contracted supplier for a certain product category, members of the GPO are generally free to purchase from other suppliers. Consequently, members of the GPOs may opt for alternative products based on price or quality offered by other companies, potentially leading to a decline in our sales volumes and revenue.
The rising healthcare costs over the past decade have led to numerous cost reform initiatives by legislators, regulators, and third-party payors, triggering a consolidation trend in the healthcare industry to aggregate purchasing power. This trend may result in more requests for pricing concessions in the future. We anticipate that market demand, government regulation, third-party coverage and reimbursement policies, and societal pressures will continue to evolve the healthcare industry, leading to further business consolidations and alliances among our customers. These changes may exert additional downward pressure on the prices of our products, affecting our results of operations and our ability to support our current business strategies.
The effectiveness of our CapsoCam capsules heavily relies on patients adhering to our use protocols. Other companies or institutions may develop and market novel or improved methods that may make our technologies less competitive or obsolete.
The effectiveness of our CapsoCam capsule endoscopy relies on patients adhering to proper dietary restriction and bowel cleansing protocols prior to the screening procedure. Inadequate preparation can result in suboptimal imaging, which may lead to missed screenings and diagnoses or the need for repeat procedures or alternative procedure such as colonoscopy. This dependence on patient compliance introduces a variable that is
26
Table of Contents
beyond our direct control, potentially impacting the reliability and accuracy of our diagnostic results. If patients do not follow the prescribed fasting and cleansing regimen, the performance of our CapsoCam capsules may be compromised, leading to decreased diagnostic accuracy and patient dissatisfaction.
The use of our CapsoCam capsule also requires manual retrieval of the capsule by the patient, who must send it back to our download center for data collection and analysis. This manual retrieval process, while designed to be as convenient as possible, may not meet the expectations of all patients and could lead to potential errors as patients are required to diligently monitor their bowel movements to ensure the capsule is retrieved, which may be inconvenient and burdensome for them. The capsules are transported by shipping companies. There have been instances where the shipping company has either lost or damaged the capsules, leading to the loss of critical patient data. This compromises the integrity of the patient data and disrupts the entire procedure.
If other companies or institutions develop or market methods that do not require fasting or bowel cleansing or require less stringent preparation, our products may become less attractive to patients. For example, non-invasive stool-based DNA tests generally requires no specific dietary restrictions or bowel preparation and enables patients to collect a stool sample at home using the provided kit and send it to the lab for analysis. Such test may be more attractive to low-risk or average-risk patients who require more convenient options for GI-tract screening. Also, recent advancements in medical research have opened up new possibilities for early detection and screening of colorectal cancer by leveraging key metabolites and early diagnostic biomarkers. Advances in technology or alternative screening methods that eliminate or reduce the need for fasting or bowel cleansing could provide a more convenient and patient-friendly option, potentially making our CapsoCam capsules less competitive. Should such alternatives gain acceptance in the market, we may experience a decline in demand for our products, which could materially and adversely affect our business, financial condition, and results of operations. Additionally, our ability to compete effectively may be further challenged if these new methods are protected by patents or other intellectual property rights that we cannot circumvent.
There are risks associated with use of our CapsoCam capsules, including capsule retention, component failures and aspiration.
Our CapsoCam capsule endoscopy is generally a safe procedure providing non-invasive visualization of the entire small bowel and colon (once cleared). However, there are risks associated with this procedure, including capsule retention, component failures and, in rare cases, aspiration.
For 2024, we recorded a provider complaint rate of approximately 2.4% (based on number of capsules sold in 2024). The bulk of recorded complaints related to patient failure to (i) timely retrieve the capsule following completion of the procedure and (ii) enter the correct serial number or use the correct shipping label when returning the capsule for download. In addition, a small number of the recorded complaints related to defective battery packs. Although small, these failures indicate potential issues with the usability or clarity of instructions provided to patients by providers or us and shipment and inventory management issues. These issues, in turn, can significantly impact customer and patient satisfaction and our reputation and potential sales.
Capsule retention by a patient refers to the capsule getting stuck in the GI tract, which can occur due to various reasons such as strictures, tumors, or inflammatory bowel diseases. We believe the industry retention rate may be as high as 2%. Based on (i) incidences reported to us for all CapsoCam capsule patients in 2024 and (ii) total CapsoCam capsules sold by us in 2024, our 2024 retention rate was less than 1/10 of 1%. However, we believe our retention rate is understated due to (i) underreporting by providers and patients and, relatedly, (ii) the fact that most retained capsules are eventually excreted without the need for invasive intervention and medication may be used to encourage this process. In some instances, endoscopic retrieval or surgical intervention may be necessary to remove the retained capsule. Aspiration is rare but potentially fatal where the capsule is accidentally inhaled into the respiratory tract instead of being swallowed into the esophagus. As of the date of this prospectus, there have been no complaints regarding any aspiration incident involving a CapsoCam capsule.
27
Table of Contents
While we strive to ensure the highest safety standards and patient experience for our CapsoCam capsule, we cannot guarantee that it is entirely free of safety issues, component failures or errors. Despite rigorous testing and quality control measures, unforeseen complications may arise during the use of our GI-tract capsule endoscopy solution. These could include, but are not limited to, adverse reactions, product malfunction, or unforeseen interactions with the patient’s medical condition.
We may need additional capital to execute our business plan, and we may be unable to raise additional capital on acceptable terms.
Executing our business plan will require additional capital. We may need to raise funds to support research and development, and expand our sales and marketing efforts. If we are unable to raise additional capital on acceptable terms, we may not be able to implement our business plan effectively, which could adversely affect our growth prospects and financial condition. Furthermore, market conditions and other factors outside our control could make it difficult to raise capital when needed. If we are unable to secure additional financing, we may have to delay, reduce, or eliminate certain aspects of our business plan, which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
Our existing sales and marketing team may face challenges in selling additional products and features added to our GI-tract capsule endoscopy solution.
We may face unique challenges in leveraging our existing sales channels to sell and market product and service additions to our GI-tract capsule endoscopy solution (including CapsoCam Colon once cleared). For example, the primary difference between small-bowel and colon diagnostic visualization lies in their distinct medical purposes, preparation requirements, and market dynamics. A small bowel capsule endoscopy is typically conducted to diagnose conditions such as OGIB, Crohn’s disease, or small bowel tumors and a different cleansing preparation compared to a colon capsule endoscopy which requires more rigorous bowel preparation. Additionally, the market for colorectal cancer screening is highly competitive, with established products such as traditional colonoscopies, other imaging technologies (mainly CT colonography and to a decreasing extent sigmoidoscopy) and stool-based screening products already in widespread use. These and other procedure and market differences and dynamics will require additional salesforce training and may necessitate tailored marketing strategies and sales approaches, which could complicate our efforts to utilize our current sales teams for products beyond small bowel.
If we or our distributors fail to comply with regulatory requirements, we may be subject to stringent penalties and our business may be materially adversely affected.
The marketing and sale of our CapsoCam capsules are subject to various state, federal, and foreign regulations. Compliance with these regulatory requirements is essential to avoid significant penalties and ensure the continued commercialization of our products. However, the regulatory landscape is complex and constantly evolving, making it challenging to stay compliant. Any failure by us or our distributors to adhere to applicable regulations could result in substantial fines, sanctions, or other enforcement actions and damage to our reputation.
Healthcare policy changes and increased scrutiny of marketing practices in the medical device industry add another layer of risk. New regulations or changes to existing laws could impose additional burdens on our business, affecting our ability to operate effectively. Ensuring ongoing compliance with regulatory requirements is critical to maintaining our market position and avoiding adverse impacts on our business.
The success of our business is substantially dependent upon the efforts of our senior management team and certain other key personnel and our ability to retain these individuals and to attract additional key personnel.
Our success depends in large part on our ability to attract and retain managerial and key personnel, including our senior executive officers and various highly skilled employees. If we were to lose any of our senior
28
Table of Contents
management team or these key personnel, we may experience difficulties in competing effectively, developing our technologies, and implementing our business strategies. Competition for desirable personnel is intense, and there can be no assurance that we will be able to attract and retain the necessary staff. The failure to maintain management or to attract sales and key personnel as we move towards the commercialization of our CapsoCam capsules could materially adversely affect our business, financial condition, and results of operations.
Product and professional liability suits against us could result in expensive and time-consuming litigation, payment of substantial damages and increases in our insurance rates.
Our CapsoCam capsules are cleared or authorized by the FDA for specific intended uses. However, if physicians elect to use our products in manners outside of these intended uses, such use may result in adverse outcomes and may pose significant risks, especially for individuals with known or suspected gastrointestinal obstructions, strictures, or fistulas, as the capsule may become lodged, potentially leading to an obstruction that requires urgent medical or surgical intervention. We cannot prevent physicians from using our products off-label or from using non-our products in conjunction with our products. The risk of injury to patients may increase if physicians attempt off-label use. Additionally, we cannot guarantee that physicians are adequately trained by us or their peers before using our products. Complications from off-label use or use by untrained physicians may not effectively treat conditions and may expose us to product liability claims or litigation, harming our reputation.
The sale and use of our CapsoCam capsules could lead to product or professional liability claims based on allegations that one of our CapsoCam capsules contained a design or manufacturing defect, which resulted in the failure to detect the disease for which it was designed. A product or professional liability claim could result in substantial damages and be costly and time-consuming to defend, either of which could materially harm our business or financial condition. We cannot assure you that our liability insurance would protect our assets from the financial impact of defending a liability claim.
Any claim brought against us, with or without merit, could increase our liability insurance rates or prevent us from securing insurance coverage in the future. Furthermore, product liability claims could result in negative publicity, injury to our reputation, and loss of revenue. Even successful defense would require significant financial and management resources, which could adversely affect our business, financial condition, results of operations, and prospects.
We may engage in acquisitions that could disrupt our business, cause dilution to our stockholders and reduce our financial resources.
In the future, we may enter into transactions to acquire other businesses, products, or technologies. Because we have not made any acquisitions to date, our ability to do so successfully is unproven. If we do identify suitable candidates, we may not be able to make such acquisitions on favorable terms or at all. Any acquisitions we make may not strengthen our competitive position, and these transactions may be viewed negatively by customers or investors.
We may decide to incur debt in connection with an acquisition or issue our common stock or other securities to the stockholders of the acquired company, which would reduce the percentage ownership of our existing stockholders. We could incur losses resulting from undiscovered liabilities of the acquired business that are not covered by the indemnification we may obtain from the seller. In addition, we may not be able to successfully integrate the acquired personnel, technologies, and operations into our existing business in an effective, timely, and non-disruptive manner.
Macroeconomic conditions could materially adversely affect our business, financial condition, results of operations, and prospects.
Macroeconomic conditions, such as high inflationary pressure, potential military conflicts, possible trade wars, introduction of or changes in tariffs or trade barriers, changes to monetary policy, high interest rates,
29
Table of Contents
volatile currency exchange rates, credit and debt concerns, decreasing consumer confidence and spending, including capital spending, concerns about the stability and liquidity of certain financial institutions, and global recessions can adversely impact demand for our products, which could negatively impact our business, financial condition, results of operations, and prospects. Recent macroeconomic conditions have been adversely impacted by geopolitical instability and military hostilities in multiple geographies and monetary and financial uncertainties.
The impacts of these macroeconomic conditions, and the actions taken by governments, central banks, companies, and consumers in response, have resulted in, and may continue to result in, higher inflation in the United States and globally, which is likely, in turn, to lead to an increase in costs and may cause changes in fiscal and monetary policy, including additional increases in interest rates. Other adverse impacts of recent macroeconomic conditions have been, and may continue to be, supply chain constraints, logistics challenges, liquidity concerns in the broader financial services industry, and fluctuations in labor availability. In a higher inflationary environment, we may be unable to raise the prices of our products sufficiently to keep up with the rate of inflation.
Healthcare policy changes could have a material adverse effect on our business, financial condition, results of operations, and prospects.
In recent years, the healthcare industry in the United States has experienced significant legislative and regulatory changes aimed at reducing costs, increasing access to health insurance, and improving the overall efficiency of healthcare delivery. The Affordable Care Act (the “ACA”) has been a cornerstone of these efforts, expanding health insurance coverage through public program expansion and private sector reforms. Additionally, the American Rescue Plan Act and the Inflation Reduction Act have introduced enhanced subsidies to make health insurance more affordable. However, these changes bring uncertainty regarding the number of individuals who will obtain public or private health insurance and the scope of such coverage. This uncertainty can impact our business by potentially reducing the number of insured individuals or altering the scope of insurance coverage, which may affect our revenue streams and financial stability.
The ACA has been subject to continuous legislative, regulatory, and judicial challenges, creating an unpredictable environment for healthcare providers. Changes in the interpretation or implementation of the ACA could eliminate or modify provisions that are beneficial to our business while maintaining or introducing provisions that reduce our reimbursement rates or otherwise negatively impact our operations. For instance, if certain beneficial provisions are repealed or altered, we may face increased financial pressure due to reduced reimbursement rates or additional regulatory compliance costs. This uncertainty makes it challenging to plan for the future and could lead to financial instability if adverse changes are implemented.
Moreover, other health reform initiatives at both federal and state levels add another layer of complexity. These initiatives could result in coverage reductions or decreased enrollment in Medicaid, further impacting our revenue. Additionally, recent reforms focused on price transparency and out-of-network charges, such as the No Surprises Act, may limit our ability to set and negotiate prices, affecting our relationships with insurers and patients. These factors collectively create a challenging environment that could adversely affect our financial condition, results of operations, and cash flows.
Adverse developments affecting the financial services industry could adversely affect our current and projected business operations, financial condition and results of operations.
Adverse developments that affect financial institutions, such as events involving liquidity that are rumored or actual, have in the past and may in the future lead to bank failures and market-wide liquidity problems. We are exposed to concentrations of credit risk through our financial instruments, which primarily include demand deposits held at reputable financial institutions and accounts receivable. These financial instruments are subject to potential credit risk, as we rely on the financial stability and creditworthiness of these institutions and its
30
Table of Contents
customers. While we take measures to mitigate these risks, such as conducting thorough credit evaluations and maintaining relationships with well-established financial institutions, there remains an inherent risk of financial loss if these counterparties fail to meet their obligations.
Additionally, our cash deposits may occasionally exceed the limits insured by the U.S. Federal Deposit Insurance Corporation. This situation can arise during periods of high cash flow or when large payments are received. When cash deposits exceed FDIC insurance limits, we are exposed to the risk of loss in the event of a bank failure. Although we strive to manage this risk by spreading out our cash deposits in accounts with multiple reputable and financially stable banks, the potential for uninsured losses remains a significant risk factor that could adversely impact our financial position and operational stability.
Changes to U.S. tariff measures and other potential changes in international trade relations implemented by the U.S. could have a material adverse effect on our business, financial condition, cash flows and results of operations.
Our supply chain is heavily reliant on products and components manufactured and assembled in various Asian countries (particularly Taiwan and Japan). These import operations are subject to tariff and other international trade regulations. When imported into the U.S., such products and components are subject to applicable rates of duty. The U.S. has recently enacted and proposed to enact significant new tariffs. Additionally, President Trump has directed various federal agencies to further evaluate key aspects of U.S. trade policy and there has been ongoing discussion and commentary regarding potential significant changes to U.S. trade policies, treaties and tariffs. Such changes to trade policies, treaties and tariffs could adversely impact the price and demand for our GI-tract capsule endoscopy solution and increase our costs. There continues to exist significant uncertainty about the future relationship between the U.S. and other countries with respect to such trade policies, treaties and tariffs. These developments, or the perception that any of them could occur, may have a material adverse effect on global economic conditions and the stability of global financial markets, and may significantly reduce global trade and, in particular, trade between the impacted nations and the U.S. Any of these factors could depress economic activity and adversely impact the price and demand for our products, increase our costs, and affect our customers and suppliers and have a material adverse effect on our business, financial condition and results of operations.
We leverage cloud-based CapsoCloud for easy access to patients’ in vivo videos, however use of CapsoCloud faces complicated compliance requirements.
Utilizing our cloud-based platform, CapsoCloud, physicians can remotely access and analyze their patients’ in vivo videos and other exam data. However, the regulatory environment for cloud-based platforms is complex and varies across different jurisdictions, and as described below, some states or countries may impose restrictions or prohibitions on the use of cloud-based platforms or the transmission of patient data across borders. Initially, following the commercial launch of our CapsoCam Colon solution, providers will utilize CapsoCloud to download in vivo videos for remote review. Within one year of commercial launch, we plan to introduce user-friendly streaming functions to facilitate via CapsoCloud remote in vivo video review, procedure report generation and image annotation.
Outside the U.S., CapsoCloud also faces significant operational and compliance issues due to stringent data privacy and security regulations in various countries. This necessitates the development of country-specific cloud infrastructures for CapsoCloud, which can lead to increased costs, complexity, and potential delays in service delivery. Additionally, for certain U.S. government-related customers, such as the U.S. Department of Defense and Veterans Administration associated hospitals, CapsoCloud must implement additional layers of IT security to meet their stringent security requirements before these customers can utilize CapsoCloud. This could further complicate our cloud infrastructure, increase the risk of security breaches if not properly managed, and require substantial investment in specialized security measures and compliance protocols.
31
Table of Contents
If we are unable to comply with these regulations or obtain the necessary approvals or licenses, we may face legal actions, fines, penalties, or loss of business, which could adversely affect our reputation, operations, and financial results. Additionally, we may incur significant costs and resources to adapt CapsoCloud to meet the changing regulatory requirements or customer preferences in different markets and countries.
We depend on our authorized agent and importer for a significant portion of our revenue.
We depend on Aureliance, our authorized agent and importer in the EU and EEA for a significant portion of our revenue. It has historically contributed a significant percentage of our revenue generated from EU countries, such that if it were to materially reduce or terminate its business with us, our revenue generated from such countries would suffer. For the years ended, and as of December 31, 2023 and 2024, it represented approximately 12% and [ ]% of our revenue and 28% and [ ]% of our accounts receivable, respectively. The loss of it or a significant reduction in its business with us could have a material adverse effect on our financial condition and results of operations.
Our ability to utilize our net operating loss carryforwards and research and development tax credits may be limited.
We have experienced net operating losses (“NOLs”) for tax purposes since our inception. As of December 31, 2023, we have total available gross (pre-tax) Federal and U.S. state NOL carryforwards of $91 million and $69 million, respectively. Under the Federal Tax Cuts and Jobs Act of 2017, NOLs incurred in tax years beginning on or after January 1, 2018, are carried forward indefinitely and are subject to a usage limitation of 80% of taxable income. NOLs incurred in tax years prior to January 1, 2018, are subject to a twenty-year carryforward period before expiring but are not subject to a usage limitation based on taxable income.
Utilization of NOL carryforwards and research and development tax credit carryforwards may be subject to substantial annual limitations under Section 382 of the Internal Revenue Code of 1986, as amended (“Section 382”), or, for states, under state laws, due to ownership changes that have occurred previously or that could occur in the future. These ownership changes may limit the amount of carryforwards that can be utilized annually to offset future taxable income. In general, an ownership change, as defined by Section 382, results from transactions increasing the ownership of certain stockholders or public groups in the stock of a corporation by more than 50% over a three-year period. We have not conducted a study to assess whether an ownership change has occurred or whether there have been multiple ownership changes since inception. If we have experienced an ownership change, as defined by Section 382, at any time since inception, utilization of the NOL carryforwards or research and development tax credit carryforwards would be subject to an annual limitation under Section 382. Any limitation may result in the expiration of a portion of the NOL carryforwards or research and development tax credit carryforwards before utilization.
The inability to fully utilize the NOL carryforwards and research and development tax credits could adversely affect our financial position and results of operations. If we are unable to generate sufficient taxable income in future periods to utilize the NOL carryforwards and research and development tax credit carryforwards before they expire, we may incur higher tax liabilities, which could have a material adverse effect on our business, financial condition, and results of operations.
Our relationships with contracted physicians are subject to various state laws and there is no assurance that our current practices will remain compliant.
We provide remote capsule endoscopy reading services through contracted physicians to our customers. Our relationships with contracted physicians are subject to various state laws that prohibit the corporate practice of medicine and fee-splitting. The interpretation and enforcement of these laws vary significantly across states, and there is no assurance that our current practices will remain compliant. If regulatory authorities determine that our
32
Table of Contents
arrangements with physicians violate these laws, we may be forced to restructure or terminate these relationships, leading to potential disciplinary actions, penalties, and a loss of revenue.
Our products must be manufactured in accordance with applicable laws and regulations, and we could be forced to recall our products or terminate production if we fail to comply with these regulations.
Our CapsoCam capsules are regulated as medical devices in the United States and must be manufactured in compliance with the FDA’s current good manufacturing practices (cGMPs) for medical devices, known as the Quality System Regulation (“QSR”). The FDA enforces the QSR through periodic announced or unannounced inspections of medical device manufacturing facilities, which may include the facilities of subcontractors. Failure to comply with applicable FDA requirements or later discovery of previously unknown problems with our products or manufacturing processes could result in, among other things, warning letters, fines, injunctions, civil penalties, suspension or withdrawal of marketing authorizations, product recalls, and total or partial suspension of production or distribution.
In addition, our products are subject to similar state regulations governing manufacturing. Our third-party manufacturers may not take the necessary steps to comply with applicable regulations, which could cause delays in the delivery of our medical devices. If we or our third-party manufacturers fail to comply with applicable laws and regulations, we could be forced to recall our products or terminate production, which could have a material adverse effect on our business, financial condition, results of operations, and prospects.
Some of our activities may subject us to risks under federal and state laws prohibiting “kickbacks” and false or fraudulent claims.
Our business practices and relationships with healthcare providers are subject to scrutiny under various federal and state laws designed to prevent fraud and abuse, including the federal Physician Payments Sunshine Act, Anti-Kickback Statute and the False Claims Act. The Physician Payments Sunshine Act increases the transparency of financial relationships between medical device manufacturers (etc.) and healthcare providers in order to uncover potential conflicts of interest that could compromise treatment decisions and medical research, or that could increase the cost of healthcare services billed to federal health programs. The Anti-Kickback Statute prohibits the exchange of remuneration to induce or reward the referral of business reimbursable under federal healthcare programs. The False Claims Act imposes liability on individuals and companies who submit false claims under federal health care programs. Violations of these laws can result in severe penalties, including criminal and civil fines, exclusion from participation in federal healthcare programs, and significant reputational harm.
Additionally, state laws often mirror or expand upon federal prohibitions, applying to all payers, not just federal programs. Compliance with these laws requires substantial resources and ongoing monitoring of our business practices. Any failure to comply with these laws, whether intentional or inadvertent, could lead to investigations, legal actions, and significant financial penalties, which could materially and adversely affect our business, financial condition, results of operations, and prospects.
Our independent sales representatives earn commissions for capsule sales in their territories, with potential additional commissions for reorders from new accounts and quarterly bonuses. Despite our efforts to provide compliance training and integrate some of them into our direct sales team to improve regulatory oversight, there remains a risk that they might inadvertently breach laws or ethical guidelines in their pursuit of higher earnings, which is not entirely within our control. Independent sales representatives, motivated by commissions and bonuses, might engage in practices that could be construed as kickbacks. If our independent sales representatives engage in unethical or illegal practices to boost sales, it could lead to the submission of false or fraudulent claims for payment submitted to the government. Their malfeasance may expose us to legal and financial liabilities under the federal and state laws.
33
Table of Contents
Failure to comply with the Foreign Corrupt Practices Act (the “FCPA”), economic and trade sanctions regulations, and similar laws could subject us to penalties and other adverse consequences.
We are subject to the FCPA and other laws that prohibit improper payments or offers of payments to foreign governments and their officials for the purpose of obtaining or retaining business. Our operations in countries with a high risk of corruption expose us to the risk that our employees, contractors, or agents may engage in activities that violate the FCPA or similar laws, despite our policies and procedures designed to prevent such conduct. Violations of these laws can result in severe penalties, including substantial fines, criminal sanctions, and reputational damage.
Additionally, we are subject to economic and trade sanctions regulations administered by the U.S. Department of the Treasury’s Office of Foreign Assets Control (OFAC), which restrict our ability to engage in transactions with certain countries, entities, and individuals. Non-compliance with these regulations can lead to significant financial penalties, restrictions on our business operations, and adverse effects on our reputation. Ensuring compliance with these complex and evolving regulations requires significant resources and ongoing vigilance.
We sell our GI-tract capsule endoscopy products in the international market through a combination of our direct sales team and third-party distributors. Additionally, the bulk of our component suppliers and assembly manufacturers are currently located outside the U.S., and we anticipate increasing our international activities in the future. Despite our implementation of policies, internal controls, and other measures designed to ensure compliance with applicable anti-corruption and anti-bribery laws and regulations, as well as U.S. trade sanctions laws, there remains a risk that our employees or agents may engage in improper conduct for which we could be held accountable. Any violations, or even allegations of violations, of these laws can result in investigations and enforcement actions that may disrupt our operations, divert significant management attention, and incur substantial costs and expenses, including legal fees. If we, our sales representatives or distributors are found to have violated these laws, we could face severe penalties, including fines, profit disgorgement, injunctions, securities litigation, bans on government business transactions, delisting from securities exchanges, and other consequences that could materially and adversely affect our business, financial condition, and results of operations. Furthermore, our reputation, net sales, or stock price could suffer if we become the subject of negative publicity related to actual or potential violations of anti-corruption, anti-bribery, or trade sanctions laws and regulations.
Our business and operations may suffer in the event of information technology system failures, cyberattacks, or deficiencies in our cybersecurity.
We rely heavily on information technology systems and infrastructure to operate our business, including the collection, storage, and transmission of sensitive patient exam data. Our systems, including CapsoCloud, and those of our third-party service providers, are vulnerable to cyberattacks, data breaches, and other security incidents that could compromise the confidentiality, integrity, and availability of our data. Such incidents could result in significant disruptions to our operations, financial losses, and damage to our reputation.
We seek to protect against cybersecurity threats. Among other measures, our CapsoCloud systems are HIPAA-compliant and operate in an ISO 27001 certified cloud-computing environment, we have adopted various cybersecurity controls and procedures and we conduct cybersecurity related trainings. However, there is no guarantee that our systems will be immune to cyberthreats. For example, in October 2019, we observed and investigated a cyberattack that breached our Amazon Web Services (AWS) account hosting the CapsoCloud service. In connection with this investigation, we determined no data was compromised during this incident and, in addition to related employee trainings, we adopted additional procedures and controls designed to protect against these types of incidents. Any failure to adequately protect our IT systems and sensitive patient exam data collected, stored and transmitted on or through our systems could lead to regulatory investigations, legal actions, and substantial costs associated with remediation and notification efforts, which could materially and adversely affect our business, financial condition, results of operations, and prospects.
34
Table of Contents
We have identified two material weaknesses in our internal control over financial reporting. These material weaknesses could adversely affect our ability to report our results of operations and financial condition accurately and in a timely manner.
Our management is responsible for establishing and maintaining adequate internal control over financial reporting designed to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements for external purposes in accordance with U.S. GAAP. Our management is likewise required, on a quarterly basis, to evaluate the effectiveness of our internal controls and to disclose any changes and material weaknesses identified through such evaluation in those internal controls. A material weakness is a deficiency, or a combination of deficiencies, in internal control over financial reporting, such that there is a reasonable possibility that a material misstatement of our annual or interim financial statements will not be prevented or detected on a timely basis.
In connection with the preparation and audit of our 2023 financial statements in accordance with the standards of the U.S. Public Company Accounting Oversight Board (“PCAOB”), we identified two material weaknesses in our internal control over financial reporting.
The first material weakness relates to our failure to design or maintain sufficient controls over implementation of information technology general controls or complementary user entity controls for applications (such as our enterprise resource planning (“ERP”), payroll and stock option management IT systems) used in the preparation of our financial statements. More specifically, we did not design or maintain sufficient controls related to user access provisioning and monitoring, change management, program development and data management.
The second material weakness relates to a lack of segregation of duties in the financial reporting function due to a limited number of staff performing the financial reporting function. More specifically, there is a limited level of multiple reviews among those tasked with preparing our financial records and with respect to our existing ERP system proper segregation of duties was not enforced (i.e., for journal entries we did not always different individuals responsible for the entry process and another approving the journal entry).
To respond to these material weaknesses, we have devoted, and plan to continue to devote, significant effort and resources to the remediation and improvement of our internal control over financial reporting. For example, in addition to hiring a Corporate Controller in Q1 2025, we are also evaluating the current and future headcount and other needs of the accounting department to ensure proper segregation of duties. We are also evaluating an upgrade to or replacement of our existing ERP system which would help address many of the control issues contributing to these material weaknesses.
Any failure to remediate these material weaknesses could adversely impact our ability to report our financial position and results from operations on a timely and accurate basis. If our financial statements are not accurate, investors may not have a complete understanding of our operations. Likewise, if our financial statements are not filed on a timely basis, we could be subject to sanctions or investigations by the stock exchange on which our common stock is listed, the SEC or other regulatory authorities. Either of the foregoing could have a material adverse effect on our business. Ineffective internal controls could also cause investors to lose confidence in our reported financial information, which could have a negative effect on the trading price of our securities.
We can give no assurance that the measures we have taken and plan to take in the future will remediate the material weaknesses identified or that any additional material weaknesses or restatements of financial results will not arise in the future due to a failure to implement and maintain adequate internal control over financial reporting or circumvention of these controls. In addition, even if we are successful in strengthening our controls and procedures, in the future those controls and procedures may not be adequate to prevent or identify irregularities or errors or to facilitate the fair presentation of our financial statements.
35
Table of Contents
We may face litigation and other risks as a result of the material weaknesses in our internal control over financial reporting.
As a result of such material weaknesses and other matters raised or that may in the future be raised by the SEC, we face potential for litigation or other disputes which may include, among others, claims invoking the federal and state securities laws, contractual claims or other claims arising from the restatement and material weaknesses in our internal control over financial reporting and the preparation of our financial statements. Any such litigation or dispute, whether successful or not, could have a material adverse effect on our business, results of operations and financial condition.
Risks Relating to Our Intellectual Property
Our success will depend on our ability to obtain, maintain, enforce, and protect our intellectual property rights.
Our success and ability to compete depends in part on our ability to obtain, maintain, enforce, and protect issued patents, trademarks, trade secret, and other intellectual property rights and proprietary technology in the U.S. and elsewhere. If we cannot adequately obtain, maintain, and enforce our intellectual property rights and proprietary technology, competitors may be able to use our technologies or the goodwill we have acquired in the marketplace and erode or negate any competitive advantage we may have and our ability to compete, which could harm our business and ability to achieve profitability and/or cause us to incur significant expenses. We generally seek to protect our proprietary position by filing patent applications that are important to our business. We also seek to protect our proprietary position by acquiring or in-licensing relevant issued patents or pending patent applications or other intellectual property or proprietary rights from third parties. If we are unable to obtain or maintain patent protection with respect to any proprietary technology, our business, financial condition, results of operations, and prospects could be materially harmed.
We rely on a combination of contractual provisions, confidentiality procedures, and patent, trademark, copyright, trade secret and other intellectual property laws to protect the proprietary aspects of our brands, technologies, trade secrets, know-how, and data. These legal measures afford only limited protection, and competitors or others may gain access to or use our intellectual property rights and proprietary information. In addition, patents have a limited lifespan. In the U.S., for example, the natural expiration of a utility patent is generally 20 years from the earliest effective non-provisional filing date. Our success will depend, in part, on preserving our trade secrets, maintaining the security of our data and know-how, and obtaining, maintaining, and enforcing other intellectual property rights. We may not be able to obtain, maintain, and/or enforce our intellectual property or other proprietary rights necessary to our business or in a form that provides us with a competitive advantage.
The patent prosecution process and enforcement of any resulting patents are expensive, time-consuming, and complex, and we may not be able to file, prosecute, maintain, enforce, defend, or license all necessary or desirable patents or patent applications at a reasonable cost or in a timely manner, or in all jurisdictions. Moreover, pending patent applications cannot be enforced against third parties practicing the technology claimed in such applications unless, and until, patents issue from such applications, and then only to the extent the issued claims cover relevant product, service, or the technology. There can be no assurance that our current or future patent applications will result in patents being issued or that our issued patents will afford sufficient protection against competitors or other third parties with similar products, services or technologies competitive with ours, nor can there be any assurance that the patents issued will not be infringed, designed around, or invalidated by third parties.
Even our issued patents may later be found invalid or may be modified or revoked in proceedings instituted by third parties before various patent offices or in courts. In addition, our issued patents may later be found to be unenforceable for a number of possible reasons, such as for a failure to properly identify material prior art or
36
Table of Contents
other references to the U.S. Patent and Trademark Office (the “USPTO”). The degree of future protection for our intellectual property or other proprietary rights is uncertain. Only limited protection may be available and may not adequately protect our rights or permit us to gain or keep any competitive advantage. These uncertainties and/or limitations in our ability to properly protect or be able to enforce the intellectual property or other proprietary rights relating to our products, services and technologies could have a material adverse effect on our business, financial condition, results of operations, and prospects.
We cannot be certain that the claims in our U.S. pending patent applications, corresponding international patent applications and patent applications in certain foreign territories will be considered patentable by the USPTO, courts in the U.S., or by the patent offices and courts in foreign countries, nor can we be certain that the claims in our future issued patents will not be found invalid or unenforceable if challenged. Our ability to obtain and maintain valid and enforceable patents depends in part on whether the differences between our inventions and the prior art allow our inventions to be patentable over the prior art. Additionally, regardless of when filed, we may fail to identify relevant third-party patents or patent applications, or we may incorrectly conclude that a third-party patent is invalid or not infringed by our products, services, technologies, or activities. Therefore, we cannot be certain that we were the first to make the inventions claimed in any of our patents or pending patent applications, or that we were the first to file for patent protection of such inventions.
Failure to obtain, maintain, and/or enforce intellectual property rights necessary to our business and failure to protect, monitor and control the use of our intellectual property rights could negatively impact our ability to compete and cause us to incur significant expenses. The intellectual property laws and other statutory and contractual arrangements in the U.S. and other jurisdictions we depend upon may not provide sufficient protection in the future to prevent the infringement, use, violation or misappropriation of our patents, trademarks, data, technology, and other intellectual property rights by others, and may not provide an adequate remedy if our intellectual property rights are infringed, misappropriated, or otherwise violated by others.
The degree of future protection for our intellectual property rights is uncertain. Therefore, we cannot ensure that others will not develop, manufacture and/or commercialize similar or alternative products, services, or technologies that do not infringe, misappropriate, or violate any patents or other intellectual property rights that we own or have rights to. We cannot ensure that any patents issued to us will provide a basis for an exclusive market for our products, services, or technologies, will provide us with any competitive advantages or will not be challenged, invalidated, modified, revoked, or circumvented by third parties. We cannot ensure that any of our challenged patents will be found to ultimately be valid and enforceable. We cannot ensure that any of our patents, or any of our pending patent applications, if issued, will include claims having a scope sufficient to protect our products, services, or technologies. We cannot ensure that any of our pending patent applications will issue as patents, or even if issued, will include claims with a scope sufficient to protect our products, services, or technologies. We cannot ensure that we will be able to successfully develop, manufacture, and commercialize our products, services, or technologies on a substantial scale before relevant patents we may have expire. We cannot ensure that we were the first to make the inventions covered by each of our patents and pending patent applications or we were the first to file patent applications for such inventions. We cannot ensure that we will develop additional proprietary inventions, products, services, or technologies that are separately patentable. In addition, we cannot ensure that our commercial activities, products, services, or technologies will not infringe upon the patents of others. Defending against any such infringement claims is also expensive, time-consuming, complex, and can distract us from pursuing commercial activities.
If we fail to identify our patentable inventions or adequately protect our patent rights, the commercial value of our products, services or technologies may be adversely affected and our competitive position may be harmed.
We rely in part on our portfolio of issued patents and pending patent applications in the U.S. and other countries to protect our intellectual property and competitive position. However, it is also possible that we may fail to identify patentable aspects of inventions made in the course of the development, manufacture, and commercial activities conducted by or on behalf of us before it is too late to obtain patent protection on such
37
Table of Contents
inventions. If we fail to timely file for patent protection in any jurisdiction, we may be precluded from doing so at a later date. Although we enter into non-disclosure and confidentiality agreements with parties who have access to patentable aspects of our research and development output, such as our employees, outside scientific collaborators, suppliers, consultants, advisors, and other third parties, any of these parties may breach the agreements and disclose such output before a patent application is filed, thereby jeopardizing our ability to seek patent protection, and any of these parties may decide to independently file for intellectual property without naming us as owners, inventors, or co-owners for the intellectual property. Furthermore, publications of discoveries in the scientific literature often lag behind the actual discoveries, and patent applications in the U.S. and other jurisdictions are typically not published until 18 months after filing, or in some cases not at all. Therefore, we cannot be certain that we were the first to make the inventions claimed in any of our patents or pending patent applications, or that we were the first to file for patent protection of such inventions. Moreover, should we become a licensee or co-owner of a third party’s patents or patent applications, depending on the terms of any future in-licenses or co-ownership agreement to which we may become a party, we may not have the right to control the preparation, filing, and prosecution of patent applications, or to maintain or enforce the patents, covering technology in-licensed from third parties. Therefore, these patents and patent applications may not be prosecuted, maintained, and/or enforced in a manner consistent with the best interests of our business. While we may apply for patents in some countries outside of the U.S., we may not accurately predict all of the countries where patent protection will ultimately be desirable. Furthermore, the issuance of a patent does not give us the right to practice the patented invention. Third parties may have blocking patents that could prevent us from importing, using, manufacturing, and/or commercializing our own products or services, or otherwise practicing our own technology. Any of these outcomes could impair our ability to prevent competition from third parties, which may have an adverse impact on our business.
The patent positions of companies, including our patent position, may involve complex legal and factual questions that have been the subject of much litigation in recent years, and, therefore, the scope of any patent claims that we have or may obtain cannot be predicted with certainty. Accordingly, we cannot provide any assurances about which of our patent applications will issue, the breadth of any resulting patent, whether any of the issued patents will be found to be infringed, invalid, or unenforceable or will be threatened or challenged by third parties, that any of our issued patents have, or that any of our currently pending or future patent applications that mature into issued patents will include claims with a scope sufficient to protect our products, services, or technology. Our pending and future patent applications may not result in the issuance of patents or, if issued, may not issue in a form that will be advantageous to us.
The coverage claimed in a patent application can be significantly reduced before the patent is issued, and its scope can be reinterpreted after issuance. We cannot offer any assurances that the breadth of our issued patents will be sufficient to stop a competitor from developing, manufacturing, and commercializing one or more products, services, or technologies in a non-infringing manner that would be competitive with one or more of our products, services, or technologies, or otherwise provide us with any competitive advantage. Furthermore, any successful challenge to these patents or any other patents owned by us after patent issuance could deprive us of rights necessary for our commercial success. Further, there can be no assurance that we will have adequate resources to enforce our patents.
Though an issued patent is presumed valid and enforceable, its issuance is not conclusive as to its validity or its enforceability and it may not provide us with adequate proprietary protection or competitive advantages against competitors with similar products or services. Patents, if issued, may be challenged, deemed unenforceable, invalidated, narrowed, or circumvented. Proceedings challenging our patents or patent applications could result in either loss of the patent, or denial of the patent application or loss or reduction in the scope of one or more of the claims of the patent or patent application. Any successful challenge to our patents and patent applications could deprive us of exclusive rights necessary for our commercial success. In addition, defending such challenges in such proceedings may be costly. Thus, any patents that we own may not provide the anticipated level of, or any, protection against competitors. Furthermore, an adverse decision may result in a third
38
Table of Contents
party receiving a patent right sought by us, which in turn could affect our ability to develop, manufacture, commercialize, import, or otherwise use our products, services, or technologies.
Some of our patents and patent applications are or may in the future be co-owned with third parties. If we are unable to obtain an exclusive license to any such third- party co-owners’ interest in such patents or patent applications, such co-owners may be able to license their rights to other third parties, including our competitors, and our competitors could market competing products, services, or technologies. In addition, we may need the cooperation of any such co-owners of our patents to enforce such patents against third parties, and such cooperation may not be provided to us.
Obtaining and maintaining patent protection depends on compliance with various procedural, document submission, fee payment, and other requirements imposed by governmental patent agencies, and our patent protection could be reduced or eliminated for non-compliance with these requirements.
The USPTO, and various foreign governmental patent agencies require compliance with a number of procedural, documentary, fee payment, and other similar provisions during the patent application process. In addition, periodic maintenance fees, renewal fees, annuity fees, and various other government fees on issued patents often must be paid to the USPTO and foreign patent agencies over the lifetime of the patent and/or applications and any patent rights we may obtain in the future. While an unintentional lapse of a patent or patent application can in many cases be cured by payment of a late fee or by other means in accordance with the applicable rules, there are situations in which non-compliance can result in abandonment or lapse of the patent or patent application, resulting in partial or complete loss of patent rights in the relevant jurisdiction. Non-compliance events that could result in abandonment or lapse of a patent or patent application include, but are not limited to, failure to respond to official actions within prescribed time limits, non-payment of fees, and failure to properly legalize and submit formal documents. If we fail to maintain our patents and patent applications, we may not be able to stop a competitor from marketing products, services, or technologies that are the same as or similar to our products, services, or technologies, which would have a material adverse effect on our business, financial condition, results of operations, and prospects.
Our patents will eventually expire, and our patent protection will be reduced or eliminated by expiration of patents in our patent portfolio.
We have prosecuted and obtained a variety of patents to protect the intellectual property rights to our products and services. These patents protect our inventions relating to camera, image sensor, display and imaging system, capsule detection, power source control, and other key and ancillary components of our products. We rely on these patents to prevent others from using, developing, manufacturing and selling competing products or solutions or infringing on our proprietary rights.
However, patents have a limited lifespan, with the term of individual patents depends upon the date of filing of the patent application, the date of patent issuance and the legal term of patents in jurisdictions in which they are obtained. In most countries, including the U.S., issued patents are granted a term of 20 years from the earliest effective non-provisional filing date. We have patents that are anticipated to expire as indicated in “Business—Intellectual Property.” After patents in our portfolio begin to expire, we cannot guarantee that our remaining patents will be sufficient to maintain our competitive advantage or prevent competitors from developing similar or superior products. Once our patents expire, we may lose the exclusive right to prevent others from using, developing, manufacturing or selling products that incorporate our patented technology that are claimed in the expired patents. If third parties use our technology within the scope of such expired patents, we may not be able to sue them for infringement or recover any damages. This could result in increased competition, reduced market share, lower revenues and margins, and harm to our reputation and brand.
In certain instances, a patent term of a U.S. patent may be adjusted to recapture a portion of delay by the USPTO in examining the patent application or extended to account for term effectively lost as a result of the
39
Table of Contents
FDA regulatory review period, or both. While the adjustment of the patent term based upon delay by the USPTO is automatically calculated by the USPTO, it is possible that this patent term adjustment is improperly calculated by the USPTO and thus does not accurately reflect the true amount of adjustment that is entitled to a patent holder. Indeed, the USPTO has publicly acknowledged in 2024 that numerous errors have possibly occurred for patent term adjustments that have been calculated by the USPTO for previously issued patents. Moreover, it is possible that actions we have taken to prosecute a patent application will be considered applicant delay and will reduce the amount of patent term adjustment that is granted to a patent. For a patent term extension based upon the FDA regulatory review period, the period of extension may be up to five years, but cannot extend the remaining term of a patent beyond a total of 14 years from the date of approval. Only one patent among those eligible for an extension may be extended. However, there is no guarantee that the applicable authorities, including the FDA, will agree with our assessment of whether such extensions should be granted, and even if granted, the length of such extensions may be less than the maximum extension available. In addition, this type of patent term extension will only be granted if requested within a designated timeframe after FDA approval. If the applicable authorities do not approve the extension or if we do not make a timely application for extension, then it is possible that a term extension will not be granted for a given patent in our patent portfolio, thereby reducing the amount of enforceable term for the patent.
Changes in U.S. or foreign patent laws or their interpretations could diminish the value of our patents in general, thereby impairing our ability to protect our current and future products, services, or technologies, and could increase the uncertainties and costs surrounding the prosecution of our patent applications and the enforcement or defense of our current or future patents.
Our ability to obtain patents and the breadth of any patents obtained is uncertain in part because, to date, some legal principles remain unresolved, and there has not been a consistent policy regarding the breadth or interpretation of claims allowed in patents in the U.S. and other countries. Changes in either patent laws or in interpretations of patent laws in the U.S. and other countries may diminish the value of our intellectual property rights or narrow the scope of our patent protection, which in turn could diminish the commercial value of our products, services, and technologies.
Patent reform legislation may pass in the future that could lead to additional uncertainties and increased costs surrounding the prosecution, enforcement, and defense of our patents and applications. Furthermore, the U.S. Supreme Court and the U.S. Court of Appeals for the Federal Circuit have made, and will likely continue to make, changes in how the patent laws of the U.S. are interpreted. The U.S. Supreme Court has ruled on several patent cases in recent years, either narrowing the scope of patent protection available in certain circumstances or weakening the rights of patent owners in certain situations.
In addition to increasing uncertainty with regard to our ability to obtain patents in the future, this combination of events has created uncertainty with respect to the value of patents, once obtained. Depending on actions by the U.S. Congress, the federal courts and the USPTO, the laws and regulations governing patents could change in unpredictable ways that would weaken our ability to obtain new patents or to enforce patents that we own or that we might obtain or license in the future. An inability to obtain, enforce, and defend patents covering our proprietary technologies would materially and adversely affect our business, financial condition, results of operations, and prospects.
Similarly, foreign courts have made, and will likely continue to make, changes in how the patent laws in their respective jurisdictions are interpreted. Changes in patent laws and regulations in other countries or jurisdictions, changes in the governmental bodies that enforce them, or changes in how the relevant governmental authority enforces patent laws or regulations may weaken our ability to obtain new patents or to enforce patents that we own or may obtain in the future. Further, the laws of some foreign countries do not protect proprietary rights to the same extent or in the same manner as the laws of the U.S. In addition, any protection afforded by foreign patents may be more limited than that provided under U.S. patent and intellectual property laws. We may encounter significant problems in enforcing and defending our intellectual property both
40
Table of Contents
in the U.S. and abroad. For example, if the issuance in a given country of a patent covering an invention is not followed by the issuance in other countries of patents covering the same invention, or if any judicial interpretation of the validity, enforceability or scope of the claims or the written description or enablement in a patent issued in one country is not similar to the interpretation given to the corresponding patent issued in other countries, our ability to protect our intellectual property rights in those countries may be limited. We cannot predict future changes in the interpretation of patent laws in the U.S. and other countries or changes to patent laws that might be enacted into law by U.S. and foreign legislative bodies. Those changes may materially affect our patents or patent applications and our ability to obtain additional patent protection in the future. Any of the foregoing could have a material adverse effect on our competitive position, business, financial condition, results of operations, and prospects.
In June 2023, the European Unitary Patent system and the European Unified Patent Court (“UPC”) were launched. European patent applications now have the option, upon grant of a patent, of becoming a Unitary Patent which is subject to the jurisdiction of the UPC. In addition, conventional European patents, both already granted at the time the new system began and granted thereafter, are subject to the jurisdiction of the UPC, unless actively opted out. This was a significant change in European patent practice, and deciding whether to opt-in or opt-out of Unitary Patent practice entail strategic and cost considerations. The UPC provides third parties with a new forum to centrally revoke our European patents and makes it possible for a third party to obtain pan-European injunctions against us. It will be several years before we will understand the scope of patent rights that will be recognized and the strength of patent remedies that will be provided by the UPC. While we have the right to opt our patents out of the UPC over the first seven years of the court’s existence, doing so may preclude us from realizing the benefits of the UPC. Moreover, the decision whether to opt-in or opt-out of Unitary Patent status will require coordinating with co-applicants, if any, adding complexity to any such decision.
The legal systems in certain countries may also favor state-sponsored or companies headquartered in particular jurisdictions over our first-in-time patents and other intellectual property protection. We are aware of incidents where such entities have stolen the intellectual property of other companies in order to create competing products. For example, through its “Annual Special 301 Report on Intellectual Property,” the Office of the United States Trade Representative has been reporting on the adequacy and effectiveness of intellectual property protection in a number of foreign countries that are U.S. trading partners and their protection and enforcement of intellectual property rights. Placement of a country on the Priority Watch List indicates that particular problems exist in that country with respect to intellectual property protection, enforcement, or market access for persons relying on intellectual property rights. Countries placed on the Priority Watch List are the focus of increased bilateral attention concerning the specific problem areas. It is possible that we will not be able to enforce our intellectual property rights against third parties that misappropriate our proprietary technology in those countries.
We may not be able to protect our intellectual property rights throughout the world.
Filing, prosecuting, and defending patents on our products, services, and technologies in all countries throughout the world would be prohibitively expensive, and the laws of foreign countries may not protect our rights to the same extent as the laws of the U.S. The requirements for patentability may differ in certain countries, particularly in developing countries. In addition, the laws of some foreign countries do not protect intellectual property rights to the same extent as federal and state laws in the U.S. Consequently, we may not be able to prevent third-parties from practicing our inventions in all countries outside the U.S. or from selling or importing products made using our inventions in and into the U.S. or other jurisdictions. Competitors may use our technologies in jurisdictions where we have not obtained patent protection to develop their own products, services, or technologies and, further, may export otherwise infringing products, services, or technologies to territories where we have patent protection, but enforcement is not as strong as that in the U.S. These products, services, or technologies may compete with our products, services, or technologies, and our patents or other intellectual property rights may not be effective or sufficient to prevent such competition.
41
Table of Contents
Various companies have encountered significant problems in protecting and defending intellectual property rights in certain foreign jurisdictions. The legal systems of certain countries may not favor the enforcement of patents and other intellectual property protection, particularly those relating to medical devices and related services and technologies, which could make it difficult for us to stop the infringement of our patents or marketing of competing products, services, and technologies in violation of our intellectual property and proprietary rights. In addition, some jurisdictions, such as Europe, Japan, and China, may have a higher standard for patentability than in the U.S., including, for example, imposing a high standard for making claim amendments and for the submission of supplemental experimental data during patent examination. Under those heightened patentability requirements, we may not be able to obtain sufficient patent protection in certain jurisdictions even though the same or similar patent protection can be secured in the U.S. and other jurisdictions.
Proceedings to enforce our intellectual property and proprietary rights in foreign jurisdictions could result in substantial costs and divert our efforts and attention from other aspects of our business, could put our patent rights at risk of being invalidated or interpreted narrowly, could put our owned patent applications at risk of not issuing and could provoke third parties to assert claims against us. We may not prevail in any lawsuits that we initiate, and the damages or other remedies awarded, if any, may not be commercially meaningful. Accordingly, our efforts to enforce our intellectual property and proprietary rights around the world may be inadequate to obtain a significant commercial advantage from the intellectual property that we develop, own or license. Various countries outside the U.S., including certain countries in Europe, India, and China, have compulsory licensing laws under which a patent owner may be compelled to grant licenses to third parties. If we are forced to grant a license to third parties with respect to any patents relevant to our business, our competitive position may be impaired, and our business, financial condition, results of operations, and prospects may be adversely affected. In addition, many countries limit the enforceability of patents against government agencies or government contractors. As a result, a patent owner in such countries may have limited remedies in certain circumstances, which could materially diminish the value of such patent. Accordingly, our efforts to enforce our intellectual property rights around the world may be inadequate to obtain a significant commercial advantage from our intellectual property.
Further, the standards applied by the USPTO and foreign patent offices in granting patents are not always applied predictably. As such, we do not know the degree of world-wide uniform protection that we will have on our technologies and products in the future.
If we cannot successfully enforce our intellectual property rights, the commercial value of our products, services, or technologies may be adversely affected and our competitive position may be harmed.
Third parties, including our competitors, may currently, or in the future, infringe, misappropriate, or otherwise violate our issued patents or other intellectual property rights, and we may file lawsuits or initiate other proceedings to protect or enforce our patents or other intellectual property rights, which could be expensive, time-consuming, and unsuccessful. We regularly monitor for unauthorized use of our intellectual property rights and, from time to time, analyze whether to seek to enforce our rights against potential infringement, misappropriation, or violation of our intellectual property rights. However, the steps we have taken, and are taking, to protect our proprietary rights may not be adequate to enforce our rights as against such infringement, misappropriation, or violation of our intellectual property rights. In certain circumstances it may not be practicable or cost-effective for us to enforce our intellectual property rights fully, particularly in certain developing countries or where the initiation of a claim might harm our business relationships. We may also be hindered or prevented from enforcing our rights with respect to a government entity or instrumentality because of the doctrine of sovereign immunity. Our ability to enforce our patent or other intellectual property rights depends on our ability to detect infringement. It may be difficult to detect infringers who do not advertise the components or methods that are used in connection with their products, services, or technologies. Moreover, it may be difficult or impossible to obtain evidence of infringement in a competitor’s or potential competitor’s product, services, or technologies. Thus, we may not be able to detect unauthorized use of, or take appropriate steps to enforce, our intellectual property rights. Any inability to meaningfully enforce our intellectual property rights
42
Table of Contents
could harm our ability to compete and reduce demand for our products, services, and technologies. We may in the future become involved in lawsuits to protect or enforce our intellectual property rights. An adverse result in any litigation proceeding could harm our business. In any lawsuit we bring to enforce our intellectual property rights, a court may refuse to stop the other party from manufacturing, commercializing, using or importing the product, service, offering or technology at issue on grounds that our intellectual property rights do not cover, and the other party is not infringing, violating or otherwise misappropriating our intellectual property, through the manufacture, commercialization, use or importation of the product, service, offering or technology in question. Any claims we assert against perceived infringers could also provoke these parties to assert counterclaims against us alleging that we infringe, misappropriate, or otherwise violate their intellectual property rights. If we initiate legal proceedings against a third party to enforce a patent covering a product, service, offering or technology, the defendant could counterclaim that such patent is invalid or unenforceable. In patent litigation in the U.S., defendant counterclaims alleging invalidity or unenforceability are common, and there are numerous grounds upon which a third party can assert invalidity or unenforceability of a patent. Grounds for a validity challenge could be an alleged failure to meet any of several statutory requirements, including lack of patentable subject matter, novelty, obviousness, or non-enablement. Grounds for an unenforceability assertion could be an allegation that someone connected with prosecution of the patent withheld relevant information from USPTO, or made a misleading statement, during prosecution. Mechanisms for such challenges include re-examination, post-grant review, inter partes review, interference proceedings, derivation proceedings, and equivalent proceedings in foreign jurisdictions (e.g., opposition proceedings). In a patent or other intellectual property proceeding, a court may decide that a patent or other intellectual property right of ours is invalid or unenforceable, in whole or in part, construe the patent’s claims or other intellectual property narrowly or refuse to stop the other party from manufacturing, commercializing, using or importing the product, service, offering, or technology at issue on the grounds that our patents or other intellectual property do not cover the manufacture, commercialization, use, or importation of the product, service, offering, or technology in question. Furthermore, even if our patents or other intellectual property rights are found to be valid and infringed, a court may refuse to grant injunctive relief against the infringer and instead grant us monetary damages and/or ongoing royalties. Such monetary compensation may be insufficient to adequately offset the damage to our business caused by the infringer’s competition in the market. An adverse result in any litigation or administrative proceeding could put one or more of our patents or other intellectual property rights at risk of being invalidated or interpreted narrowly, which could adversely affect our competitive business, financial condition, results of operations and prospects. Moreover, even if we are successful in any litigation, we may incur significant expense in connection with such proceedings, and the amount of any monetary damages may be inadequate to compensate us for damage as a result of the infringement and the proceedings.
We may become a party to intellectual property litigation or administrative proceedings that could be expensive, time-consuming, and unsuccessful, and could interfere with our ability to develop, manufacture, commercialize, import, or otherwise use our products, services, or technologies.
Our commercial success depends, in part, on our ability to develop, manufacture, commercialize, import, or use our products, services, and technologies without infringing, misappropriating, or otherwise violating the intellectual property rights of third parties. Our industry has been characterized by extensive litigation regarding patents, trademarks, trade secrets, and other intellectual property rights, and companies in the industry have used intellectual property litigation to gain a competitive advantage. While we do not intend to infringe upon, misappropriate, or otherwise violate the intellectual property rights of others, there may be pertinent rights of which we are presently unaware.
Third parties may initiate legal proceedings alleging that we are infringing, misappropriating, or otherwise violating their intellectual property rights. The outcomes of such proceedings are uncertain and could have a negative impact on the success of our business. It is possible that U.S. and foreign patents and pending patent applications controlled by third parties may be alleged to cover our products, services, and technologies, or that we may be accused of misappropriating third parties’ trade secrets or infringing third parties’ trademarks. We may in the future become party to, or be threatened with, adversarial proceedings or litigation regarding
43
Table of Contents
intellectual property rights with respect to our products, services, or technologies, including interference proceedings, post grant review, and inter partes review before the USPTO or equivalent foreign regulatory authority. Furthermore, we may also become involved in other proceedings, such as reexamination, derivation, or opposition proceedings before the USPTO or other jurisdictional body relating to our intellectual property rights or the intellectual property rights of others. Third parties may assert infringement claims against us based on existing patents or patents that may be granted in the future, regardless of their merit. Because patent applications can take many years to issue and because publication schedules for pending applications vary by jurisdiction, there may be applications now pending of which we are unaware and which may result in issued patents, which our current or future products, services, or technologies infringe. Also, because the claims of published patent applications can change between publication and patent grant, there may be published patent applications that may ultimately issue with claims that we infringe. There is a risk that third parties may choose to engage in litigation with us to enforce or to otherwise assert their patent rights against us. Even if we believe such claims are without merit, a court of competent jurisdiction could hold that these third-party patents are valid and enforceable, and infringed by the use of our products, services, or technologies, which could have a negative impact on the commercial success of our current and any future products, services, or technologies. If we were to challenge the validity of any such third-party U.S. patent in federal court, we would need to overcome a presumption of validity. As this burden is a high one requiring us to present clear and convincing evidence as to the invalidity of any such U.S. patent claim, there is no assurance that a court of competent jurisdiction would invalidate the claims of any such U.S. patent. We will have similar burdens to overcome in foreign courts in order to successfully challenge a third-party claim of patent infringement.
Our defense of any litigation or interference proceedings may fail and, even if successful, defending such claims brought against us would cause us to incur substantial expenses and distract our management and other employees. If such claims are successfully asserted against us, we could be forced to pay substantial damages. Further, if a patent infringement or other intellectual property rights-related lawsuit were brought against us, we could be forced, including by court order, to cease developing, manufacturing, commercializing, importing, or using the infringing product, service, or technology. In addition, we could be found liable for monetary damages, including treble damages and attorneys’ fees, if we are found to have willfully infringed a patent or other intellectual property right. Although patent, trademark, trade secret, and other intellectual property disputes have often been settled through licensing or similar arrangements, costs associated with such arrangements may be substantial and could include ongoing royalties. We may not be able to obtain licenses on commercially reasonable terms or at all, in which event our business would be materially and adversely affected. Even if we were able to obtain a license, the rights may be non-exclusive, which could result in our competitors and other third parties gaining access to the same intellectual property. Ultimately, if we are unable to obtain such licenses or make any necessary changes to our products, services, or technologies, we could be forced to cease some aspect of our business operations, which could harm our business significantly.
A finding of infringement or an unfavorable interference or derivation proceedings outcome could prevent us from developing, manufacturing, commercializing, importing, or using our products, services, or technologies, or force us to cease some or all of our business operations, which could materially harm our business. Claims that we have misappropriated the confidential information or trade secrets of third parties could have a similar negative impact on our business, financial condition, results of operations, and prospects. We may not have sufficient financial or other resources to conduct such litigation or proceedings adequately. Some of our competitors may be able to sustain the costs of litigation or administrative proceedings more effectively than we can because of greater financial resources and more mature and developed intellectual property portfolios. We could encounter delays in product introductions while we attempt to develop alternative products or technologies.
If third parties assert infringement, misappropriation, or other claims against our customers, these claims may require us to initiate or defend protracted and costly litigation on behalf of our customers, regardless of the merits of these claims. If any of these claims succeed or settle, we may be forced to pay damages or settlement payments on behalf of our customers or may be required to obtain licenses for the products, services, or
44
Table of Contents
technologies they use. If we cannot obtain all necessary licenses on commercially reasonable terms, our customers may be forced to stop using our products, services, or technologies.
Our competitors may have substantially greater resources and have made substantial investments in patent portfolios, trade secrets, trademarks, and competing technologies, and may have applied for or obtained, or may in the future apply for or obtain, patents or trademarks that will prevent, limit, or otherwise interfere with our ability to make, use, sell, import, and/or export our products, services, or technologies. As the number of competitors in our market grows and the number of patents issued in this area increases, the possibility of patent infringement claims against us may increase. Moreover, individuals and groups that are non-practicing entities, commonly referred to as “patent trolls,” purchase patents, and other intellectual property assets for the purpose of making claims of infringement in order to extract settlements. From time to time, we may receive threatening letters, notices or “invitations to license,” or may be the subject of claims that our products, services, or technologies and business operations infringe, misappropriate, or otherwise violate the intellectual property rights of others. These matters can be time-consuming, costly to defend in litigation, divert management’s attention and resources, damage our reputation and brand, and cause us to incur significant expenses or make substantial payments. In addition, we purchase product components, including hardware and software, from suppliers, and the design of these components may be outside of our direct control. These suppliers may not indemnify us in the event that a third party alleges the use of such components infringes its intellectual property rights.
Any lawsuits relating to intellectual property rights could subject us to significant liability for damages and invalidate our intellectual property. Any potential intellectual property litigation also could force us to do one or more of the following: (a) stop developing, making, selling, importing, or using products, services, or technologies that allegedly infringe, misappropriate, or otherwise violate the asserted intellectual property right; (b) pay substantial damages or royalties to the party whose intellectual property rights we may be found to be infringing, misappropriating, or otherwise violating; (c) redesign those products, services, or technologies that contain the allegedly infringing intellectual property, which could be costly, disruptive, and infeasible; and attempt to obtain a license to the relevant intellectual property rights from third parties, which may not be available on commercially reasonable terms or at all, or from third parties who may attempt to license rights that they do not have; (d) lose the opportunity to license our intellectual property rights to others or to collect royalty payments based upon successful protection and assertion of our intellectual property rights against others; (e) incur significant legal expenses; or (f) pay the attorney’s fees and costs of litigation to the party whose intellectual property rights we may be found to be infringing, misappropriating, or otherwise violating.
Third parties may also raise similar claims before administrative bodies in the U.S. or abroad, even outside the context of litigation. Such mechanisms include re-examination, post grant review, inter partes review, and equivalent proceedings in foreign jurisdictions (for example, opposition proceedings). Such proceedings could result in revocation of or amendment to our patents in such a way that they no longer cover our products, services, or technologies. With respect to the validity question, for example, we cannot be certain that there is no invalidating prior art, of which we, our patent counsel, and the patent examiner were unaware during prosecution. If a third party were to prevail on a legal assertion of invalidity and/or unenforceability, we may lose at least part, and perhaps all, of the patent protection on our products, services, or technologies. Such a loss of patent protection would have a material adverse impact on our business, financial condition, results of operations, and prospects.
Because of the substantial amount of discovery required in connection with intellectual property litigation, there is a risk that some of our confidential information could be compromised by disclosure during litigation. There could also be public announcements of the results of hearing, motions, or other interim developments. If securities analysts or investors perceive these results to be negative, it could have a material adverse effect on the price of shares of our common stock. Even if we ultimately prevail, a court may decide not to grant an injunction against further infringing activity and instead award only monetary damages, which may not be an adequate remedy. Furthermore, even if resolved in our favor, the monetary cost of such litigation and the diversion of the attention of our management could outweigh any benefit we receive as a result of the proceedings. Uncertainties
45
Table of Contents
resulting from the initiation and continuation of patent litigation or other proceedings could have a material adverse effect on our business. Any of the foregoing may cause us to incur substantial costs and could place a significant strain on our financial resources, divert the attention of management from our core business, and harm our reputation.
We may be subject to claims challenging the inventorship or ownership of our patents and other intellectual property rights.
We may also be subject to claims that our current or former employees, contractors, partners, vendors or other third parties have an ownership interest in our current or future patents, patent applications, or other intellectual property rights, including as an inventor, co-inventor, or co-owner. We may be subject to ownership or inventorship disputes in the future arising, for example, from conflicting obligations of employees, consultants, partners, vendors, or others who were or are involved in developing our products, services, or technologies. Although it is our policy to require those who may be involved in the conception or development of inventions to execute agreements assigning such inventions and intellectual property rights therein to us, we may be unsuccessful in executing such an agreement with each party who, in fact, conceives or develops inventions that we regard as our own, and we cannot be certain that our agreements with such parties will be upheld in the face of a potential challenge, or that they will not be breached, for which we may not have an adequate remedy. The assignment of inventions may not be self-executing, or the assignment agreements may be breached, and litigation may be necessary to defend against these and other claims challenging inventorship or ownership of inventorship. If we fail in defending any such claims, in addition to paying monetary damages, we may lose valuable intellectual property rights, such as exclusive ownership of, or the right to use, valuable intellectual property rights, and other owners may be able to license their interest in such intellectual property rights to other third parties, including our competitors. Such an outcome could have a material adverse effect on our business. Even if we are successful in defending against such claims, litigation could result in substantial costs and be a distraction to management and other employees.
In addition, we may be subject to claims from third parties challenging inventorship or ownership of intellectual property rights we regard as our own, based on claims that our agreements with employees, consultants, partners, vendors, or others who were or are involved in developing our products, services, or technologies obligating them to assign their inventions and intellectual property rights therein to us are ineffective or in conflict with prior or competing contractual obligations to assign inventions and intellectual property rights therein to another employer, to a former employer, or to another person or entity. Many of our current and former employees and consultants were previously employed at or engaged by other medical device companies, including our competitors or potential competitors. Some of these employees and consultants have executed with such previous employment or engagements confidential information non-disclosure and non-use agreements and inventions assignment agreements, which may have included non-competition provisions. Although we try to ensure that such employees and consultants do not use or otherwise disclose confidential information or intellectual property rights of others in their work for us without such other person’s consent, we may be subject to claims that we or our current or former employees or consultants have, inadvertently or otherwise, infringed, violated, or otherwise misappropriated the confidential information or the intellectual property rights of these former employers, clients, or other third parties. To the extent that our current or former employees or consultants disclose or use confidential information or intellectual property rights owned by others in their work for us, disputes may arise as to the rights in any related or resulting inventions and litigation may be necessary to defend against these claims. It may also be necessary or we may desire to obtain a license to such third party’s intellectual property rights to settle any such claim; however, there can be no assurance that we would be able to obtain such license on commercially reasonable terms, if at all. If our defense to those claims fails, in addition to paying monetary damages or a settlement payment, a court could prohibit us from manufacturing, commercializing, using or importing the product, service, or technology features or practicing other intellectual property rights that are essential to our business, which could have a material adverse effect on our competitive position as well as our business, financial condition, results of operations, and prospects. In addition, we may lose valuable intellectual property rights or personnel. Even if we are successful in defending
46
Table of Contents
against these claims, litigation could result in substantial costs and could be a distraction to management and our employees. Any litigation or the threat thereof may adversely affect our ability to hire employees or contract with collaborators, partners, services provides, or contractors. A loss of key personnel or their work product could hamper or prevent our ability to develop, manufacture, commercialize, import, or use our products, services, or technologies, which could materially and adversely affect our business, financial condition, results of operations, and prospects.
If we are unable to protect the disclosure and use of our confidential information and trade secrets, the value of our products, services, and technologies and our business and competitive position could be harmed.
In addition to patent protection, we also rely on other intellectual property rights, including trade secrets, know-how, and/or other proprietary information that is not patentable or that we elect not to patent. However, trade secrets can be difficult to protect, and some courts are less willing or unwilling to protect trade secrets. To protect and maintain the confidentiality of our trade secrets and other proprietary information, we rely heavily on confidentiality provisions that we have in contracts with our employees, consultants, collaborators, and other third parties. We generally enter into confidentiality and inventions assignment agreements with our employees, consultants, and applicable third parties upon their commencement of a relationship with us. However, we cannot guarantee that we have entered into such agreements with each party that may have or have had access to our trade secrets or proprietary technology and processes, and we may not enter into such agreements with all employees, consultants, and third parties who have been involved in the development of our inventions. Although we generally require all of our employees, consultants, advisors, and any third parties who have access to our proprietary know-how, information, or technology to enter into confidentiality agreements, we cannot provide any assurances that all such agreements have been duly executed, and any of these parties may breach the agreements and disclose our proprietary information, including our trade secrets.
In addition, despite the protections we place on our intellectual property and our other proprietary rights, monitoring unauthorized use and disclosure by employees, consultants, and other third parties who have access to such intellectual property or other proprietary rights is difficult, and we do not know whether the steps we have taken to protect our intellectual property or other proprietary rights will be adequate. Therefore, we may not be able to prevent the unauthorized disclosure or use of our technical knowledge or other trade secrets by such employees, consultants, advisors, or third parties, despite the existence of our protections, including non-disclosure and use restrictions. These agreements may not provide meaningful protection against the unauthorized disclosure or use or of our trade secrets, know-how, or other proprietary information in the event the unwanted use is outside the scope of the provisions of the contracts or in the event of any unauthorized use, misappropriation, or disclosure of such trade secrets, know-how or other proprietary information that we fail to detect. There can be no assurances that such employees, consultants, advisors, or third parties will not intentionally or unintentionally breach their agreements with us, that we will have adequate remedies for any breach, or that our trade secrets will not otherwise become known or independently developed by third parties, including our competitors. If any of our trade secrets were to be lawfully obtained or independently developed by a competitor, we would have no right to prevent them, or those to whom they communicate it, from using that information to compete with us. In addition to contractual measures, we try to protect the confidential nature of our proprietary information by maintaining physical security of our premises and electronic security of our information technology systems. Such security measures may not, for example, in the case of misappropriation of a trade secret by an employee, consultant or other third party with authorized access, provide adequate protection for our proprietary information. Our security measures may not prevent an employee, consultant, or other third party from misappropriating our trade secrets and providing them to a competitor, and recourse we take against such misconduct may not provide an adequate remedy to protect our interests fully.
If any of our trade secrets were to be disclosed to or independently developed by a competitor, our competitive position would be harmed. The exposure of our trade secrets and other proprietary information would impair our competitive advantages and could have a material adverse effect on our business, financial condition, results of operations, and prospects. In particular, a failure to protect our proprietary rights may allow
47
Table of Contents
competitors to copy our products, services, or technologies, which could adversely affect our pricing and market share. Unauthorized parties may also attempt to copy or reverse engineer certain aspects of our products, services, or technologies that we consider proprietary. Costly and time-consuming litigation could be necessary to enforce and determine the scope of our trade secret rights and related confidentiality, non-disclosure, and non-use provisions, and outcomes of such litigation are unpredictable. Enforcing a claim that a party illegally disclosed, used or misappropriated a trade secret can be difficult, expensive, and time-consuming, and the outcome is unpredictable. While we use commonly accepted security measures, trade secret violations are often a combination of federal and state law in the U.S., and the criteria for protection of trade secrets can vary among different jurisdictions. If the steps we have taken to maintain our trade secrets are deemed inadequate, we may have insufficient recourse against third parties for misappropriating the trade secret. In addition, some courts are less willing or unwilling to protect trade secrets and agreement terms that address non-competition are difficult to enforce in many jurisdictions and might not be enforceable in certain cases. Finally, even if we were to be successful on the enforcement of our claims, we may not be able to obtain adequate remedies.
It is also possible that others may independently develop information or technologies that are the same as or similar to our trade secrets or other proprietary technologies and develop products, services, or technologies without obtaining access to our trade secrets or other proprietary information in which case we could not assert any intellectual property rights, including trade secret rights, against such parties in a manner that could prevent legal recourse by us. If we fail to obtain or maintain trade secret protection, or if any of our confidential or proprietary information, such as our trade secrets, were to be disclosed or used by others without our consent or otherwise misappropriated, or if any such information was independently developed by a competitor, or if our competitors obtain our trade secrets or independently develop products, services, or technologies that are the same as or similar to ours, our competitive market position could be materially and adversely harmed.
If our trademarks and trade names are not adequately protected, we may not be able to build brand name recognition in our markets of interest and our competitive position may be harmed.
Our trademarks could be challenged, opposed, invalidated, infringed, and circumvented by third parties, and our trademarks could also be diluted, declared generic or descriptive, or found to be infringing on other marks. If any of the foregoing occurs, we could be forced to re-brand our Company, products, services, or technologies, resulting in loss of brand recognition and requiring us to devote resources to advertising and marketing new brands, and suffer other competitive harm. Third parties may also adopt trademarks similar to ours, which could harm our brand identity and lead to market confusion. Further, there can be no assurance that competitors will not infringe our trademarks or that we will have adequate resources to enforce our trademarks. At times, competitors may adopt trade names or trademarks similar to ours, thereby impeding our ability to build brand identity and possibly leading to market confusion. Certain of our current or future trademarks may become so well known by the public that their use becomes generic and they lose trademark protection. Over the long term, if we are unable to establish name recognition based on our trademarks and trade names, then we may not be able to compete effectively. Any of the foregoing could have a material adverse effect on our competitive position, business, financial condition, results of operations, and prospects.
We rely on our trademarks, trade names, and brand names to distinguish our products, services, and technologies from the products, services, and technologies of our competitors, and have registered or applied to register many of these trademarks in the U.S. and certain countries outside the U.S.; however, we have not yet registered all of our trademarks in all of our current and potential markets. There can be no assurance that all of our trademark applications will be approved for registration. During trademark registration proceedings, we may receive rejections. Although we are given an opportunity to respond to those rejections, we may be unable to overcome such rejections. In addition, in proceedings before the USPTO and comparable agencies in many foreign jurisdictions, third parties have opposed and may oppose in the future further our trademark applications and may seek to cancel trademark registrations or otherwise challenge our use of the trademarks. Opposition or cancellation proceedings may be filed against our trademark filings in these agencies, and such filings may not survive such proceedings. While we may be able to continue the use of our trademarks in the event registration is
48
Table of Contents
not available, particularly in the U.S., where trademark rights are acquired based on use and not registration, third parties may be able to enjoin the continued use of our trademarks if such parties are able to successfully claim infringement in court. In addition, opposition or cancellation proceedings may be filed against our trademark applications and registrations and our trademarks may not survive such proceedings. If we do not secure registrations for our trademarks, we may encounter more difficulty in enforcing them against third parties than we otherwise would. Our trademarks or trade names may be infringed, circumvented, declared generic, or determined to be violating or infringing on other marks.
Our products contain third-party open source software components and failure to comply with the terms of the underlying open source software licenses could restrict our ability to sell our products, affect our ability to protect our proprietary information, and subject us to possible litigation.
Our products contain software tools licensed by third parties under open source software licenses. Use and distribution of open source software may entail greater risks than use of third-party commercial software, as open source software licensors generally do not provide warranties or other contractual protections regarding infringement claims or the quality of the code. Some open source software licenses contain requirements that the licensee make its source code publicly available if the licensee creates modifications or derivative works using such open source software, depending on the type of open source software the licensee uses and how the licensee uses it. If we combine our proprietary software with open source software in a certain manner, we could, under certain open source software licenses, be required to make available the source code of certain of our proprietary software to the public for free. This could allow our competitors to create similar products with less development effort and time and ultimately could result in a loss of product sales and revenue. In addition, some companies that use third-party open source software have faced claims challenging their use of such open source software and their compliance with the terms of the applicable open source license. We may be subject to suits by third parties claiming ownership of what we believe to be open source software or claiming non-compliance with the applicable open source licensing terms. Use of open source software may also present additional security risks because the public availability of such software may make it easier for hackers and other third parties to compromise or attempt to compromise our technology platform and systems.
Although we typically review our use of open source software to avoid subjecting our products, services or technology to conditions we do not intend, the terms of many open source software licenses have not been interpreted by U.S. courts, and there is a risk that these licenses could be construed in a way that could impose unanticipated conditions or restrictions on our ability to commercialize our products, services or technology. Moreover, our processes for monitoring and controlling our use of open source software in our products, services or technology may not be effective. If we are held to have breached the terms of an open source software license, we could be required to seek licenses from third parties to continue offering our solutions on terms that are not economically feasible, to re-engineer our products, services, or technology, to discontinue the sale of our products, services, or technology if re-engineering could not be accomplished on a timely basis, to pay statutory or other damages to the license holder, or to make generally available, in source code form, our proprietary code, any of which could materially adversely affect our business, financial condition, results of operations, and prospects.
Risks Relating to Our Common Stock and this Offering
There may not be an active trading market for our common stock, which may cause shares of our common stock to trade at a discount from the initial public offering price and make it difficult to sell the shares of common stock you purchase.
Prior to this offering, there has been no public market for our common stock. It is possible that after this offering, an active trading market will not develop or, if developed, that any market will not be sustained, which would make it difficult for you to sell your shares of common stock at an attractive price or at all. The initial public offering price per share of common stock will be determined by agreement among us and the
49
Table of Contents
representatives of the underwriters and may not be indicative of the price at which shares of our common stock will trade in the public market, if any, after this offering. The market value of our common stock may decrease from the initial public offering price. Furthermore, an inactive market may also impair our ability to raise capital in the future by selling shares of our common stock.
We are an emerging growth company and a smaller reporting company, and we cannot be certain if the reduced reporting requirements applicable to emerging growth companies and smaller reporting companies will make our common stock less attractive to investors.
We are an “emerging growth company,” as defined in the JOBS Act. We will remain an “emerging growth company” until the earliest to occur of:
| • | the last day of the fiscal year during which our total annual revenue equals or exceeds $1.235 billion (subject to adjustment for inflation); |
| • | the last day of the fiscal year following the fifth anniversary of this offering; |
| • | the date on which we have, during the previous three-year period, issued more than $1 billion in non-convertible debt; or |
| • | the date on which we are deemed to be a “large accelerated filer” under the Exchange Act. |
As a result of our “emerging growth company” status, we may take advantage of exemptions from various reporting requirements that would otherwise be applicable to public companies including, but not limited to, reduced disclosure obligations regarding executive compensation in our periodic reports and proxy statements and exemptions from the requirements of holding a nonbinding advisory vote on executive compensation and stockholder approval of any golden parachute payments not previously approved.
We are also a “smaller reporting company,” meaning that the market value of our stock held by non-affiliates plus the proposed aggregate amount of gross proceeds to us as a result of this offering is less than $700.0 million and our annual revenue is less than $100.0 million during the most recently completed fiscal year. We may continue to be a smaller reporting company after this offering if either (i) the market value of our stock held by non-affiliates is less than $250.0 million or (ii) our annual revenue is less than $100.0 million during the most recently completed fiscal year and the market value of our stock held by non-affiliates is less than $700.0 million.
If we are a smaller reporting company at the time we cease to be an emerging growth company, we may continue to rely on exemptions from certain disclosure requirements that are available to smaller reporting companies. Specifically, as a smaller reporting company we may choose to present only the two most recent fiscal years of audited financial statements in our annual report on Form 10-K and, similar to emerging growth companies, smaller reporting companies have reduced disclosure obligations regarding executive compensation. Investors may find our common stock less attractive because we may rely on these exemptions. If some investors find our common stock less attractive as a result, there may be a less active trading market for our common stock and the market price of our common stock may be adversely affected and more volatile.
We will incur increased costs and become subject to additional regulations and requirements as a result of becoming a public company, which could lower our profits or make it more difficult to run our business.
As a public company, we will incur significant legal, accounting, and other expenses that we have not incurred as a private company, including costs associated with public company reporting requirements. We will also incur costs associated with the Sarbanes-Oxley Act and related rules implemented by the SEC and the exchange on which our securities are listed. The expenses generally incurred by public companies for reporting and corporate governance purposes have been increasing. We expect these rules and regulations to increase our legal and financial compliance costs and to make some activities more time-consuming and costly, although we are currently unable to estimate these costs with any degree of certainty. These laws and regulations also could
50
Table of Contents
make it more difficult or costly for us to obtain certain types of insurance, including director and officer liability insurance, and we may be forced to accept reduced policy limits and coverage or incur substantially higher costs to obtain the same or similar coverage.
These laws and regulations could also make it more difficult for us to attract and retain qualified persons to serve on our board of directors, on our board committees or as our executive officers. Furthermore, if we are unable to satisfy our obligations as a public company, we could be subject to delisting of our common stock, fines, sanctions, other regulatory action, and potentially civil litigation.
If we are unable to design, implement, and maintain effective internal control over financial reporting in the future, investors may lose confidence in the accuracy and completeness of our financial reports and the market price of our common stock may decline.
As a public company, we will be required to maintain internal control over financial reporting and to report any material weaknesses in such internal controls. In addition, beginning with our second annual report on Form 10-K, we will be required to furnish a report by management on the effectiveness of our internal control over financial reporting, pursuant to the rules and regulations of the SEC regarding compliance with Section 404 of the Sarbanes-Oxley Act. The process of designing, implementing and testing the internal control over financial reporting required to comply with this obligation is time consuming, costly and complicated. See the risk factor titled “We have identified two material weaknesses in our internal control over financial reporting. These material weaknesses could adversely affect our ability to report our results of operations and financial condition accurately and in a timely manner.” Any failure to maintain internal control over financial reporting could severely inhibit our ability to accurately report our financial condition, results of operations, or cash flows.
Further, if we are unable to comply with the requirements of Section 404 of the Sarbanes-Oxley Act in a timely manner, or if we or, if required, our auditors, are unable to assert that our internal control over financial reporting is effective, investors may lose confidence in the accuracy and completeness of our financial reports and the market price of our common stock could decline, and we could also become subject to investigations by the stock exchange on which our common stock is listed, the SEC or other regulatory authorities, which could require additional financial and management resources. Failure to remedy any material weakness in our internal control over financial reporting, or to implement or maintain other effective control systems required of public companies, could also restrict our future access to the capital markets.
In addition, as we continue to scale and improve our operations, including our internal systems and processes, we are currently evaluating the implementation of a variety of critical systems, such as supply chain management, customer relationship management, billing, human resource information systems and accounting systems. We cannot assure you that new systems, including any increases in scale or related improvements, will be successfully implemented or that appropriate personnel will be available to facilitate and manage these processes. Failure to implement necessary systems and procedures, transition to new systems and processes or hire the necessary personnel could result in higher costs, compromised internal reporting and processes and system errors or failures. For example, we are evaluating an upgrade to or replacement of our existing ERP system that facilitates orderly maintenance of books and records and the preparation of financial statements. ERP system implementations are complex projects that require significant investment of capital and human resources, the reengineering of many business processes and the attention of many employees who would otherwise be focused on other aspects of our business. The implementation and transition to any new critical system, including a new ERP system, may be disruptive to our business if they do not work as planned or if we experience issues related to such implementation or transition, which could have a material adverse effect on our operations.
We do not intend to pay dividends in the foreseeable future. As a result, your ability to achieve a return on your investment will depend on appreciation in the market price of our common stock.
We have never declared or paid cash dividends on our capital stock, and we do not currently intend to pay any cash dividends on our capital stock in the foreseeable future. We currently intend to retain all available funds
51
Table of Contents
and any future earnings to fund the development and expansion of our business. Any future determination related to dividend policy will be made at the discretion of our board of directors, subject to applicable laws, and will depend upon, among other factors, our results of operations, prospects, financial condition, contractual restrictions and capital requirements. In addition, our ability to pay cash dividends on our capital stock may be limited by the terms of any future debt or preferred securities we issue or any future credit facilities we enter into.
Accordingly, investors must for the foreseeable future rely on sales of their common stock after price appreciation, which may never occur, as the only way to realize any future gains on their investments.
If our operating and financial performance in any given period does not meet any guidance that we provide to the public, the market price of our common stock may decline.
We may, but are not obligated to, provide public guidance on our expected operating and financial results for future periods. Any such guidance will be comprised of forward-looking statements subject to the risks and uncertainties described in this prospectus and in our other public filings and public statements. Our actual results may not always be in line with or exceed any guidance we have provided, especially in times of economic uncertainty. If actual circumstances differ from those in our assumptions, our operating and financial results could fall below our publicly announced guidance or the expectations of investors. If, in the future, our operating or financial results for a particular period do not meet any guidance we provide or the expectations of investment analysts or investors generally, or if we reduce our guidance for future periods, the market price of our common stock may decline. Even if we do issue public guidance, there can be no assurance that we will continue to do so in the future.
We will have broad discretion in the use of net proceeds to us from this offering and may not use them effectively.
We will have broad discretion in the application of the net proceeds to us from this offering, including for any of the purposes described in “Use of Proceeds,” and you will not have the opportunity as part of your investment decision to assess whether the net proceeds are being used appropriately. Because of the number and variability of factors that will determine our use of the net proceeds from this offering, their ultimate use may vary substantially from their currently intended use. We intend to use a portion of the net proceeds to fund research and product development activities, including clinical studies and research and development to advance our GI-tract CapsoCam endoscopic capsules and CapsoCloud. If we do not use the net proceeds that we receive in this offering effectively, our business, financial condition, results of operations, and prospects could be harmed, and the market price of our common stock could decline. Pending their use, we may invest the net proceeds from this offering in short-term, investment-grade, interest-bearing securities such as money market accounts, certificates of deposit, commercial paper, and guaranteed obligations of the U.S. government that may not generate a high yield for our stockholders. These investments may not yield a favorable return to our investors.
Investors in this offering will experience immediate and substantial dilution.
The initial public offering price of our common stock is expected to be substantially higher than the pro forma as adjusted net tangible book value per share of our common stock. Therefore, if you purchase shares of our common stock in this offering, you will pay a price per share that substantially exceeds our pro forma as adjusted net tangible book value per share after this offering. Based on the initial public offering price of $ per share, the midpoint of the price range set forth on the cover page of this prospectus, you will experience immediate dilution of $ per share, representing the difference between our pro forma as adjusted net tangible book value per share after giving effect to this offering and the initial public offering price. Furthermore, if the underwriters exercise their option to purchase additional shares, or outstanding options and warrants are exercised, you could experience further dilution. For a further description of the dilution that you will experience immediately after this offering, see the section titled “Dilution.”
52
Table of Contents
We may require additional capital to support business growth, and this capital might not be available on terms favorable to us, or at all, and may dilute existing stockholders’ ownership of our common stock.
We intend to continue to make investments to support our business growth and may require additional funds to respond to business challenges and opportunities, including the need to develop new products, enhance our existing products, enhance our operating infrastructure, potentially expand internationally, and potentially acquire complementary businesses and technologies. In order to achieve these objectives, we may make future commitments of capital resources. Accordingly, we may need to engage in equity or debt financings to secure additional funds. If we raise additional funds through further issuances of equity or convertible debt securities, our existing stockholders could suffer significant dilution, and any new equity securities we issue could have rights, preferences, and privileges superior to those of holders of our common stock. In addition, the incurrence of indebtedness would increase our fixed obligations and include covenants or other restrictions that would impede our ability to manage our operations. Further, if additional financing is needed, we may not be able to obtain additional financing on terms favorable to us or at all. Our inability to obtain adequate financing or financing on terms satisfactory to us, when we require it, could significantly limit our ability to continue supporting our business growth and responding to business challenges and opportunities.
Sales of a substantial number of shares of our common stock in the public market could cause our stock price to fall.
If our existing stockholders sell, or indicate an intention to sell, substantial amounts of our common stock in the public market after the lock-up, market standoff, and other legal restrictions on resale discussed in this prospectus lapse, the trading price of our common stock could decline. Based upon the number of shares outstanding as of , 2025 and assuming (i) the conversion of our outstanding convertible preferred stock as of , 2025 into an aggregate of shares of our common stock immediately prior to the completion of this offering, (ii) no exercise of the underwriters’ option to purchase additional shares of common stock, and (iii) no exercise of outstanding options or warrants subsequent to , 2025, upon the closing of this offering, we will have outstanding a total of shares of common stock. Of these shares, all of the shares of our common stock sold in this offering, plus any shares sold upon exercise of the underwriters’ option to purchase additional shares, will be freely tradable, without restriction, in the public market immediately following this offering, other than shares purchased by our “affiliates” (as such term is defined in Rule 144 under the Securities Act of 1933). See the section titled “Shares Eligible for Future Sale” in this prospectus for restrictions applicable to our affiliates. We anticipate that we, our directors, executive officers, and the holders of the outstanding shares of our common stock (but limited to such holders that hold an aggregate of not more than 95% of our outstanding common shares on an as converted to common basis) have entered or will enter into lock-up agreements with the underwriters prior to the commencement of this offering. The lock-up agreements pertaining to this offering will expire 6 months after the date of closing of this offering (the “Lock-Up Period”). After the expiration of the lock-up agreements and the market standoff restrictions described below, as of , 2025, up to approximately additional shares of common stock will be eligible for sale in the public market, approximately % of which shares are owned by directors, executive officers, and other owners of more than 5% of our outstanding common stock, stock options, warrants and securities convertible into our common stock and will be subject to Rule 144 under the Securities Act of 1933, as amended (the “Securities Act”). The representatives of the underwriters may, however, in their sole discretion, permit our officers, directors, and other stockholders who are subject to these lock-up agreements to sell shares prior to the expiration of the lock-up agreements. Furthermore, our outstanding common stock, stock options, warrants, and other securities convertible into or exercisable or exchangeable for our common stock are subject to market standoff restrictions with us that include restrictions on the sale, transfer, or other disposition of shares during the Lock-Up Period. As a result of the foregoing, substantially all of our outstanding common stock, stock options, warrants, and other securities convertible into or exercisable or exchangeable for our common stock are subject to a lock-up agreement or market standoff provisions during the Lock-Up Period. We have agreed to enforce all such market standoff restrictions on behalf of the underwriters and not to release, amend, or waive any such market standoff provisions during the Lock-Up Period without the prior consent of Benchmark, on behalf of the
53
Table of Contents
underwriters, provided that we may release shares from such restrictions to the extent that it would be permissible to release such shares under the form of lock-up agreement with the underwriters signed by or that will be signed by certain record holders of our securities as described herein. In addition, as of , 2025, shares of common stock that are subject to outstanding options or subject to outstanding warrants will become eligible for sale in the public market to the extent permitted by the provisions of various vesting schedules, lock-up agreements, market standoff restrictions, and Rule 144 and Rule 701 under the Securities Act. If these additional shares of common stock are sold, or if it is perceived that they will be sold, in the public market, the market price of our common stock could decline. After this offering, based upon the number of shares outstanding as of , 2025, the holders of approximately shares of our common stock, or approximately % of our total outstanding common stock, will be entitled to rights with respect to the registration of their shares under the Securities Act, subject to the lock-up agreements and market standoff restrictions described above. Registration of these shares under the Securities Act would result in the shares becoming freely tradable without restriction under the Securities Act, except for shares purchased by affiliates. Any sales of securities by these stockholders could have a material adverse effect on the trading price of our common stock. Record holders of our securities are typically the parties to the lock-up agreements with the underwriters and the market standoff restrictions referred to above, while holders of beneficial interests in our shares who are not also record holders in respect of such shares are not typically subject to any such agreements or other similar restrictions. Accordingly, we believe that holders of beneficial interests who are not record holders and are not bound by market standoff restrictions or lock-up agreements could enter into transactions with respect to those beneficial interests that negatively impact our stock price. In addition, a security holder who is neither subject to market standoff restrictions with us nor a lock-up agreement with the underwriters may be able to sell, short sell, transfer, pledge, or otherwise dispose of or attempt to sell, short sell, transfer, hedge, pledge, or otherwise dispose of their equity interests at any time.
Provisions in our charter documents and under Delaware law could discourage a takeover that stockholders may consider favorable and may lead to entrenchment of management.
Our amended and restated certificate of incorporation and amended and restated bylaws that will be in effect immediately prior to the consummation of this offering will contain provisions that could delay or prevent changes in control or changes in our management without the consent of our board of directors. These provisions will include the following:
| • | A classified board of directors with three-year staggered terms, which may delay the ability of stockholders to change the membership of a majority of our board of directors; |
| • | No cumulative voting in the election of directors, which limits the ability of minority stockholders to elect director candidates; |
| • | The exclusive right of our board of directors to elect a director to fill a vacancy created by the expansion of the board of directors or the resignation, death, or removal of a director, which prevents stockholders from being able to fill vacancies on our board of directors; |
| • | The ability of our board of directors to authorize the issuance of shares of preferred stock and to determine the price and other terms of those shares, including preferences and voting rights, without stockholder approval, which could be used to significantly dilute the ownership of a hostile acquiror; |
| • | The ability of our board of directors to alter our amended and restated bylaws without obtaining stockholder approval; |
| • | The required approval of at least % of the shares entitled to vote at an election of directors to adopt, amend or repeal our amended and restated bylaws or to repeal certain provisions of our amended and restated certificate of incorporation; |
| • | A prohibition on stockholder action by written consent, which forces stockholder action to be taken at an annual or special meeting of our stockholders; |
54
Table of Contents
| • | The requirement that a special meeting of stockholders may be called only by our board of directors, which may delay the ability of our stockholders to force consideration of a proposal or to take action, including the removal of directors; and |
| • | Advance notice procedures that stockholders must comply with in order to nominate candidates to our board of directors or to propose matters to be acted upon at a stockholders’ meeting, which may discourage or deter a potential acquiror from conducting a solicitation of proxies to elect the acquiror’s own slate of directors or otherwise attempting to obtain control of us. |
We are also subject to the anti-takeover provisions contained in Section 203 of the General Corporation Law of the State of Delaware (the “Delaware General Corporation Law”). Under Section 203, a corporation may not, in general, engage in a business combination with any holder of 15% or more of its capital stock unless the holder has held the stock for three years or, among other exceptions, the board of directors has approved the transaction. For a description of our capital stock, see the section titled “Description of Capital Stock.”
Claims for indemnification by our directors, officers, and other employees or agents may reduce our available funds to satisfy successful third-party claims against us and may reduce the amount of money available to us.
Our amended and restated certificate of incorporation and amended and restated bylaws will provide that we will indemnify our directors and officers, in each case to the fullest extent permitted by Delaware law. In addition, as permitted by Section 145 of the Delaware General Corporation Law, our amended and restated bylaws to be effective immediately prior to the completion of this offering and our indemnification agreements that we have entered into with our directors, officers and certain other employees will provide that:
| • | We will indemnify our directors and officers for serving us in those capacities or for serving other business enterprises at our request, to the fullest extent permitted by Delaware law. Delaware law provides that a corporation may indemnify such person if such person acted in good faith and in a manner such person reasonably believed to be in or not opposed to the best interests of the registrant and, with respect to any criminal proceeding, had no reasonable cause to believe such person’s conduct was unlawful. |
| • | We may, in our discretion, indemnify employees and agents in those circumstances where indemnification is permitted by applicable law. |
| • | We are required to advance expenses, as incurred, to our directors and officers in connection with defending a proceeding, except that such directors or officers shall undertake to repay such advances if it is ultimately determined that such person is not entitled to indemnification. |
| • | We will not be obligated pursuant to our amended and restated bylaws to indemnify a person with respect to proceedings initiated by that person against us or our other indemnitees, except with respect to proceedings authorized by our board of directors or brought to enforce a right to indemnification. |
| • | The rights conferred in our amended and restated bylaws are not exclusive, and we are authorized to enter into indemnification agreements with our directors, officers, employees, and agents and to obtain insurance to indemnify such persons. |
| • | We may not retroactively amend our amended and restated bylaws provisions to reduce our indemnification obligations to directors, officers, employees, and agents. |
Our amended and restated certificate of incorporation will provide that the Court of Chancery of the State of Delaware will be the exclusive forum for certain disputes between us and our stockholders, which could limit our stockholders’ ability to obtain a favorable judicial forum for disputes with us or our directors, officers, or employees.
Our amended and restated certificate of incorporation will provide that, unless we consent in writing to the selection of an alternative forum, the Court of Chancery of the State of Delaware (or, in the event that the Court
55
Table of Contents
of Chancery does not have jurisdiction, the federal district court for the District of Delaware or other state courts of the State of Delaware) is the exclusive forum for any derivative action or proceeding brought on our behalf, any action asserting a claim of breach of fiduciary duty owed by any of our directors, officers, or stockholders to us or to our stockholders, any action asserting a claim against us arising pursuant to the Delaware General Corporation Law, our amended and restated certificate of incorporation, or our amended and restated bylaws (as either may be amended from time to time), or any action asserting a claim against us that is governed by the internal affairs doctrine; provided that, the exclusive forum provision will not apply to suits brought to enforce any liability or duty created by the Exchange Act or any other claim for which the federal courts have exclusive jurisdiction; and provided further that, if and only if the Court of Chancery of the State of Delaware dismisses any such action for lack of subject matter jurisdiction, such action may be brought in another state or federal court sitting in the State of Delaware. Our amended and restated certificate of incorporation will also provide that the federal district courts of the United States of America will be the exclusive forum for the resolution of any complaint asserting a cause of action against us or any of our directors, officers, employees, or agents and arising under the Securities Act. Nothing in our amended and restated certificate of incorporation or amended and restated bylaws precludes stockholders that assert claims under the Exchange Act from bringing such claims in state or federal court, subject to applicable law.
If any action the subject matter of which is within the scope described above is filed in a court other than a court located within the State of Delaware (a “Foreign Action”), in the name of any stockholder, such stockholder shall be deemed to have consented to the personal jurisdiction of the state and federal courts located within the State of Delaware in connection with any action brought in any such court to enforce the applicable provisions of our amended and restated certificate of incorporation and having service of process made upon such stockholder in any such action by service upon such stockholder’s counsel in the Foreign Action as agent for such stockholder. Although our amended and restated certificate of incorporation will contain the choice of forum provision described above, it is possible that a court could find that such a provision is inapplicable for a particular claim or action or that such provision is unenforceable.
We believe these provisions may benefit us by providing increased consistency in the application of Delaware law and federal securities laws by chancellors and judges, as applicable, particularly experienced in resolving corporate disputes, efficient administration of cases on a more expedited schedule relative to other forums, and protection against the burdens of multi-forum litigation. However, this choice of forum provision may limit a stockholder’s ability to bring a claim in a judicial forum that it finds favorable for disputes with us or any of our directors, officers, other employees, or stockholders, which may discourage lawsuits with respect to such claims, although our stockholders will not be deemed to have waived our compliance with federal securities laws and the rules and regulations thereunder. Furthermore, the enforceability of similar choice of forum provisions in other companies’ certificates of incorporation has been challenged in legal proceedings, and it is possible that a court could find these types of provisions to be inapplicable or unenforceable.
The market price of our common stock may be volatile, which could cause the value of your investment to decline.
Even if an active trading market develops, the market price of our common stock may be highly volatile and could be subject to wide fluctuations. Securities markets worldwide experience significant price and volume fluctuations. This market volatility, as well as general economic, market, or political conditions, could reduce the market price of our common stock regardless of our operating performance. In addition, our results of operations could be below the expectations of public market analysts and investors due to a number of potential factors, including variations in our quarterly results of operations, additions or departures of key management personnel, failure to meet analysts’ earnings estimates, publication of research reports about our industry, litigation and government investigations, data privacy and security-related events, changes or proposed changes in laws or regulations or differing interpretations or enforcement thereof affecting our business, adverse market reaction to any indebtedness we may incur or securities we may issue in the future, changes in market valuations of similar companies or speculation in the press or investment community, announcements by our competitors, adverse
56
Table of Contents
publicity about the medical device industry, or individual scandals, and, in response, the market price of our common stock could decrease significantly.
You may be unable to resell your shares of common stock at or above the initial public offering price. Stock markets experience extreme price and volume fluctuations. In the past, following periods of volatility in the overall market and the market price of a company’s securities, securities class action litigation has often been instituted against these companies. Such litigation, if instituted against us, could result in substantial costs and a diversion of our management’s attention and resources.
If securities or industry analysts do not publish research or reports about our business, or if they issue an adverse or misleading opinion regarding our stock, our stock price and trading volume could decline.
The trading market for our common stock will be influenced by the research and reports that industry or securities analysts publish about us or our business. We do not currently have and may never obtain research coverage by securities and industry analysts. If no or few securities or industry analysts commence coverage of us, the market price for our stock would be negatively impacted. In the event we obtain securities or industry analyst coverage, if any of the analysts who cover us issue an adverse or misleading opinion regarding us, our business model, our intellectual property or our stock performance, or if our results of operations fail to meet the expectations of analysts, our stock price would likely decline. If one or more of these analysts cease coverage of us or fail to publish reports on us regularly, we could lose visibility in the financial markets, which in turn could cause our stock price or trading volume to decline.
57
Table of Contents
We estimate that we will receive net proceeds from the sale of shares of common stock in this offering of approximately $ million (or approximately $ million if the underwriters’ option to purchase additional shares of common stock from us is exercised in full), based upon an assumed initial public offering price of $ per share, the midpoint of the range set forth on the cover page of this prospectus, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
We intend to use the net proceeds from this offering as follows:
(i) approximately $ to fund research and development and clinical investigation of our CapsoCam’s accuracy in detecting cancerous and precancerous pancreatic neoplasia by visualizing abnormalities of the duodenal papilla;
(ii) approximately $ to fund research and development and clinical investigation of our CapsoCam’s accuracy in screening esophageal varices in cirrhotic patients with portal hypertension; and
(iii) the remainder for general corporate purposes, including working capital, operating expenses and capital expenditures.
The expected use of net proceeds from this offering represents our intentions based upon our present plans and business conditions. We cannot predict with certainty all of the particular uses for the proceeds of this offering or the amounts that we will actually spend on the uses set forth above. Accordingly, our management will have broad discretion in applying the net proceeds from this offering. The timing and amount of our actual expenditures will be based on many factors, including cash flows from operations and the anticipated growth of our business. Pending their use, we intend to invest the net proceeds from this offering in a variety of capital-preservation investments, including government securities and money market funds.
58
Table of Contents
We have never declared or paid any cash or other dividends or distributions on our capital stock. We currently intend to retain earnings, if any, to finance the growth and development of our business. We do not expect to pay any cash dividends on our common stock in the foreseeable future. Payment of future dividends, if any, will be at the discretion of our board of directors and will depend on our financial condition, results of operations, capital requirements, restrictions contained in any financing instruments, provisions of applicable law and other factors the board of directors deems relevant.
59
Table of Contents
The following table sets forth our cash and capitalization as of December 31, 2024:
| • | on an actual basis; |
| • | on a pro forma basis, to reflect the Preferred Stock Conversion; and |
| • | on a pro forma as adjusted basis, after giving effect to the Preferred Stock Conversion set forth above, the sale of shares of our common stock in this offering at an assumed initial public offering price of $ (the midpoint of the range set forth on the cover page of this prospectus), and our receipt of the estimated $ in net proceeds from this offering, after deducting underwriting commissions and estimated offering expenses payable by us. |
You should read this capitalization table together with “Use of Proceeds,” “Summary Financial Data,” “Management’s Discussion and Analysis of Financial Condition and Results of Operations” and our financial statements and the related notes appearing elsewhere in this prospectus. The pro forma as adjusted information below is illustrative only and our capitalization following the completion of this offering will depend on the actual initial public offering price and other terms of this offering determined at pricing.
| As of December 31, 2024 | ||||||||||||
| Actual | Pro Forma | Pro Forma as Adjusted |
||||||||||
| (in thousands, except share and per share amounts) | ||||||||||||
| Cash |
$ | $ | $ | |||||||||
|
|
|
|
|
|
|
|||||||
| Convertible preferred stock, par value $0.001 per share; shares authorized; shares issued and outstanding, on an actual basis; no shares issued and outstanding, on a pro forma basis and on a pro forma as adjusted basis |
$ | $ | $ | |||||||||
| Stockholders’ deficit: |
||||||||||||
| Common stock, par value $0.001 per share; shares authorized; shares issued and outstanding, on an actual basis; shares authorized, shares issued and outstanding, on a pro forma basis; and shares authorized, shares issued and outstanding, on a pro forma as adjusted basis |
||||||||||||
| Additional paid-in capital |
||||||||||||
| Accumulated deficit |
||||||||||||
|
|
|
|
|
|
|
|||||||
| Total stockholders’ deficit |
$ | $ | $ | |||||||||
|
|
|
|
|
|
|
|||||||
| Total capitalization |
$ | $ | $ | |||||||||
|
|
|
|
|
|
|
|||||||
Each $1.00 increase or decrease in the assumed initial public offering price of $ per share of our common stock, which is the midpoint of the estimated price range set forth on the cover page of this prospectus, would increase or decrease, as applicable, each of our pro forma as adjusted cash and cash equivalents, additional paid-in-capital, total stockholders’ equity (deficit), and total capitalization by approximately $ million, assuming the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same, and after deducting estimated underwriting discounts and commissions and estimated offering expenses payable by us. Similarly, each increase or decrease of 1.0 million shares in the number of shares of common stock offered by us would increase or decrease, as applicable, each of our pro forma as adjusted cash and cash equivalents, additional paid-in-capital, total stockholders’ equity (deficit), and total capitalization by approximately
60
Table of Contents
$ million, assuming the assumed initial public offering price of $ per share of our common stock, which is the midpoint of the estimated price range set forth on the cover page of this prospectus, remains the same, and after deducting estimated underwriting discounts and commissions and estimated offering expenses payable by us.
If the underwriters exercise in full their option to purchase up to additional shares of common stock at the assumed initial public offering price of $ per share of our common stock, which is the midpoint of the estimated price range set forth on the cover page of this prospectus, after deducting estimated underwriting discounts and commissions and estimated offering expenses payable by us, pro forma as adjusted cash, additional paid-in capital, total stockholders’ equity (deficit), total capitalization, and shares of common stock outstanding as of December 31, 2024 would be $ , $ , $ , $ , and shares, respectively.
The number of shares of our common stock to be outstanding after this offering is based on shares of our common stock outstanding as of December 31, 2024 and reflects the Preferred Stock Conversion.
The number of shares of our common stock to be outstanding after this offering does not include:
| • | 50,000 shares of our common stock issuable upon the exercise of outstanding warrants as of December 31, 2024 with a weighted-average exercise price of $1.45 per share; |
| • | shares of our common stock issuable upon the exercise of outstanding stock options as of December 31, 2024, with a weighted-average exercise price of $ per share; |
| • | shares of our common stock issuable upon the exercise of outstanding stock options granted subsequent to December 31, 2024, with a weighted-average exercise price of $ per share; and |
| • | shares of our common stock reserved for future issuance under our 2025 Plan, which will become effective as of the date immediately prior to the date our registration statement relating to this offering becomes effective, as well as any future increases in the number of shares of common stock reserved for issuance under the 2025 Plan. |
61
Table of Contents
Our net tangible book value as of December 31, 2024 was approximately $ , or $ per share of common stock based upon shares of common stock outstanding on that date. Net tangible book value per share represents the amount of our total tangible assets reduced by the amount of our total liabilities, divided by the total number of shares of common stock outstanding.
Our as adjusted net tangible book value will be $ or $ per share. As adjusted net tangible book value per share represents as adjusted net tangible book value divided by the total number of shares outstanding after giving effect to the sale of the shares in this offering at the assumed initial public offering price of $ per share, the midpoint of the range set forth on the cover page of this prospectus, after deducting underwriting discounts and commissions and other estimated offering expenses payable by us. This represents an immediate increase in as adjusted net tangible book value of $ per share to existing stockholders and an immediate dilution of $ per share to investors purchasing shares of common stock in this offering at the assumed public offering price.
The following table illustrates this dilution on a per share basis to new investors:
| Assumed initial public offering price per share |
$ | |||||||
| Historical net tangible book value per share as of December 31, 2024 |
$ | |||||||
| Increase in net tangible book value per share attributable to new investors |
$ | |||||||
|
|
|
|
|
|||||
| Pro forma net tangible book value per share after giving effect to this offering |
$ | |||||||
|
|
|
|
|
|||||
| Dilution in net tangible book value per share to new investors |
$ | |||||||
|
|
|
|
|
The following tables set forth, as of December 31, 2024, the number of shares of common stock purchased from us, the total consideration paid to us and the average price per share paid by the existing holders of our common stock and the price to be paid by new investors at the public offering price.
| Shares Purchased | Total Consideration | Average Price Per Share |
||||||||||||||||||
| Number | Percent | Amount | Percent | |||||||||||||||||
| Existing stockholders |
||||||||||||||||||||
| Investors purchasing shares in this offering |
||||||||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
| Total |
100 | % | 100 | % | ||||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||||||
The number of shares of common stock to be outstanding after this offering is based on shares outstanding as of December 31, 2024.
If the underwriters exercise their overallotment option, our as pro forma net tangible book value following the offering will be $ per share, and the dilution to new investors in the offering will be $ per share.
A $1.00 increase or decrease in the assumed public offering price per share would increase or decrease our as adjusted net tangible book value after this offering by approximately $ , and dilution per share to new investors by approximately $ .
62
Table of Contents
MANAGEMENT’S DISCUSSION AND ANALYSIS OF FINANCIAL CONDITION AND RESULTS OF OPERATIONS
The following discussion should be read in conjunction with the section titled “Summary Financial Data” and our financial statements and related notes thereto included elsewhere in this prospectus. In addition to historical information, this discussion contains forward-looking statements that involve risks, uncertainties, and assumptions that could cause actual results to differ materially from management’s expectations. See the section titled “Cautionary Note Regarding Forward-Looking Statements” included elsewhere in this prospectus. Factors that could cause or contribute to such differences include, but are not limited to, those discussed in the section titled “Risk Factors.” Our historical results are not necessarily indicative of the results that may be expected for any period in the future. We are not undertaking any obligation to update any forward-looking statements or other statements we may make in the following discussion or elsewhere in this prospectus even though these statements may be affected by events or circumstances occurring after the forward-looking statements or other statements were made.
Overview
We are a commercial-stage medical technology company that develops advanced imaging and AI technologies, deployed in our capsule endoscopy solutions to identify abnormalities of the GI tract for diagnostic and screening purposes. We were founded in 2005 and are headquartered in Saratoga, California.
We developed our first capsule endoscope system, currently comprising the CapsoCam Plus single-use capsule and the CapsoCloud and CapsoView software, to panoramically visualize the small-bowel mucosa to detect abnormalities such as obscure GI bleeding and Crohn’s disease.
We are (i) in the process of updating CapsoCam Plus to add our self-developed AI assisted reading technology and (ii) targeting related FDA 510(k) clearance of the updated capsule for late 2025, with commercialization shortly thereafter. Our AI assisted reading tools detect and highlight suspected abnormalities for a clinician, reducing their time to review the video and making capsule endoscopy more financially attractive to their practice.
We began sales of our small bowel capsule system to our provider customers (i.e., primarily gastroenterologists practicing in clinics and/or hospitals) both internationally (in 2012) and in the U.S. (in 2017) through our global sales and marketing team. In the U.S., we sell to customers directly. Internationally we sell both directly and through qualified exclusive distributors in specified regions. Our largest international markets (based on our sales) are Germany, Sweden, and the U.K. In 2023, we established a direct sales team in Germany to better serve our customers and strengthen our market presence in this key market. We plan to (i) further grow our existing sales and marketing team to increase small-bowel-related sales and (ii) leverage our existing sales and marketing team to sell future product additions to our GI-tract capsule endoscopy solution.
We currently manufacture and intend to continue manufacturing our CapsoCam capsules included in our GI-tract capsule endoscopy solution (including CapsoCam Colon capsules). To assist us in manufacturing our GI-tract capsule endoscopy products, we rely on component suppliers and assembly manufacturers based in Asia (particularly Taiwan and Japan).
Driven by increased acceptance of CapsoCam Plus and growth in the small-bowel capsule endoscopy market, our revenue has increased in each year since we began U.S. direct sales in 2020. Our revenues for the year ended December 31, 2023 and 2024 totaled approximately $9.8 million and $[ ] million, respectively, representing a year-over-year growth of approximately [ ]% from 2023 to 2024. In 2023 and 2024, international sales accounted for 26% and [ ]% of total revenue. As of December 31, 2024, our small bowel capsule has been used by over 120,000 patients worldwide. For 2024, our customer retention rate was approximately 90% (calculated based on new 2022 and 2023 customers that have adopted our CapsoCam solution and remained customers in 2024).
63
Table of Contents
As part of our effort to expand and grow our revenues beyond small-bowel-related revenues, we are developing our next pipeline capsule endoscope product, CapsoCam Colon. Our CapsoCam Colon capsule (i) leverages CapsoCam Plus’s existing capsule design with its panoramic view and (ii) incorporates both our self-developed AI to automatically detect polyps in the video and our polyp-size measurement tool enabled by a 3D sensor in the capsule (polyp size being highly correlated with a polyp’s risk of becoming cancer). Based on our current regulatory development plan (focused on first obtaining requisite U.S. FDA 510(k) clearance of both our capsule and the incorporated AI technology), we are targeting CapsoCam Colon revenues beginning in mid-2026 after receiving FDA 510(k) clearance of our second generation of CapsoCam Colon system, designed with a larger field of view and better image quality to improve accuracy (measured in terms of polyp-detection sensitivity and specificity). In particular, our ability to grow CapsoCam Colon revenue is dependent on receiving requisite FDA and EU clearance for an initial indicated patient population followed by subsequent regulatory clearances and/or approvals for use of our capsule endoscopy by a broader indicated patient population.
Longer term, we believe our CapsoCam family of products, incorporating our panoramic imaging solution, can be adapted to address new GI medical indications. Potential new medical indications include pancreatic cancer and esophageal medical conditions (such as esophageal varices and Barrett’s esophagus). In 2025, we plan to commence clinical investigation of CapsoCam’s accuracy in (i) detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) and (ii) screening esophageal varices in cirrhotic patients with portal hypertension.
Accumulated and Continued Near-term Losses; Substantial Doubt About Our Ability to Continue as a Going Concern; Planned Continued Investment
We have incurred net losses since inception, and we expect to incur additional losses and negative cashflows in the foreseeable future. In addition, our 2023 audited financial statements include a footnote raising substantial doubt about our ability to continue as a going concern. The 2023 audited financial statements do not include any adjustments to reflect the future effects on the recoverability and classification of assets or the amounts and classification of liabilities if we are unable to continue as a going concern. For the year ended December 31, 2023, we incurred net losses of $11.3 million and expect additional losses and negative cash flows in 2024 and 2025. As of December 31, 2023, we had an accumulated deficit of $110.5 million and, as of December 31, 2024, we had approximately $9 million in cash, including $20.4 million raised in 2023 and $15.0 million raised in 2024, net of issuance costs, through the preferred stock issuances to our existing investors.
Our accumulated deficit, to date funded solely through equity issuances, reflects significant front-end spending and investment related to both completed and ongoing key operational milestones, including: (i) the initial and continued development of CapsoCam Plus and CapsoCloud; (ii) development of our pipeline CapsoCam Colon capsule; (iii) initial development and ongoing improvements to our AI assisted reading tools and technologies; and (iv) funding of completed and ongoing related clinical and other studies.
We plan to grow and expand our near-term revenue by further penetrating the small bowel market and generating initial CapsoCam Colon sales targeting mid-2026. As we execute on our business plan to grow our business and revenues, we will continue to incur development costs and clinical study expenses and will make additional investments. In particular, as described below, we intend to continue to make significant investments in research and development efforts to develop our next generation products and expand patient and medical and patient indications for our GI tract capsule endoscopy solution.
Based on our operating plan, we expect the proceeds from this offering, together with existing cash balances, will be sufficient to fund our operations for at least the next 12 months. Our underlying assumptions could be wrong and our ability to execute against our business plan is subject to a number of risks (including regulatory, competitive and supply chain related risks). As a result, we may need to undertake additional capital-raising activities sufficient to fund our operations and investments to grow our revenues and business as contemplated. See “—Liquidity and Capital Resources—Funding Requirements” below and “Risk Factors—Risks Relating to Our Common Stock and this Offering—We may require additional capital to support business
64
Table of Contents
growth, and this capital might not be available on terms favorable to us, or at all, and may dilute existing stockholders’ ownership of our common stock.” above.
Key Factors Affecting Our Results of Operations and Performance
We believe there are several important factors that we expect will impact our operating performance and results of operations for the foreseeable future. These factors include:
| • | Success in further penetrating the small bowel capsule endoscopy market. Until commercialization of our CapsoCam Colon capsule (targeted for mid-2026), to grow our revenues (and, in turn, reduce our expected losses and negative cash flows), we intend to grow our small-bowel-related revenues by, among other things: (i) retaining and growing our customer base; (ii) cost-effectively increasing the size and effectiveness of our U.S. and international sales teams and our customer-support function; (iii) pursuing the pediatric market (with children comprising a significant portion of the Crohn’s disease patient population); (iv) introducing complementary products such as our (a) capsule delivery device (availability expected in 2025) and (b) patency capsule (for verifying a capsule endoscope can pass through the bowel without retention prior to an exam) (tentative FDA 510(k) submission planned by mid-2025); (v) facilitating increased telemedicine adoption following recent FDA clearance of remote ingestion of our CapsoCam Plus, allowing patients to ingest our capsules in the comfort of their own homes with remote provider supervision; and (vi) following related FDA 510(k) clearance (targeted for late 2025), commercializing our updated CapsoCam Plus which incorporates our AI assisted reading technology. |
| • | Success in our clinical development efforts and managing related costs. Prior FDA, EU and other regulatory clearances are required (as applicable) to, among other things, (i) commercialize our GI-tract capsule endoscopy solution (including newly developed capsules or major hardware or software enhancements and improvements to a previously approved capsule or related product), (ii) target a new or expanded indicated patient population for our solution or (iii) target new medical conditions. These clearances must be supported by satisfactory clinical study data, among other things. By their nature, clinical development and related expenses often require significant upfront investment in terms of time and cost before revenue generation is ensured. Consequently, to generate or achieve and maintain revenue, we must effectively manage our planned clinical development efforts in terms of successful results and time and costs expended. In particular, our clinical study expenses may fluctuate significantly due to numerous factors, including the nature of a clinical study and relatedly, the study’s protocols, size in terms of patients and clinical sites and duration and the ability to timely enroll sufficient qualified patients (which may be impacted by competing clinical trials). We may also be required to conduct additional clinical trials or other testing of any of our products beyond those that are contemplated or if we experience significant delays in enrollment in any clinical trials, we could incur significant additional costs and the clinical development timeline for our products may be delayed. |
| • | Success in our research and development efforts. We plan to continue investing in research and development to build upon and expand our GI-tract capsule endoscopy solution. Our research and development initiatives are focused on introducing new capsule products and enhancements and improvements to existing products, aimed at increasing the value provided by our GI-tract capsule endoscopy solution to patients and providers. We plan to appropriately modify our combined CapsoCam capsule and CapsoCloud and/or CapsoView solution to address new GI indications, for which we have begun and/or expect to soon begin the technical validation process for several additional indications and clinical-use cases beyond the small bowel. |
| • | Success in further developing our AI product features. We plan to continue investing to (i) improve the pathology-detection and classification accuracy and the scope of our AI algorithms and (ii) apply AI, including large language models, to streamline the diagnostic and medical-report-generation processes, which in turn improves the efficiency and effectiveness of our healthcare provider customers. Our CapsoCloud platform in the ordinary course collects patient videos and physician diagnostic reports that, with patient consent, we can utilize as a dataset for training and validating our AI algorithms. This ever-expanding patient pathology dataset is unique among capsule endoscopy companies, and together with |
65
Table of Contents
| our inhouse team of AI experts, provides a competitive advantage. Ongoing investment is required to apply expert labels to the accumulated image data and to modify, train, and test the algorithms on this labeled data to expand applications beyond the current horizon. |
Components of our Results of Operations and Balance Sheet
Revenue
To date, with limited exception for capsule video reading service revenues, all our revenue is generated from the sale of our CapsoCam capsules to our customers and the use of CapsoCloud or CapsoView as the customer selected capsule video delivery option for (i) streaming the capsule videos (via CapsoCloud) for clinician review or (ii) downloading the capsule videos (via CapsoView) for clinician review. Sales of capsules and revenue related to CapsoView (including the associated data access reading device) constitute product sales. CapsoCloud data access related revenue is accounted for as service revenue as a software-as-a-service offering. CapsoCloud related revenues totaled $0.6 million in 2023 and $[ ] million in 2024. Use of CapsoView for viewing capsule videos is prevalent internationally, particularly in Europe due to the EU Cybersecurity Act, officially known as Regulation (EU) 2019/881. In the U.S., approximately 90% of our customers utilize CapsoCloud for viewing small bowel capsule videos.
Fees paid to GPOs that act as procurement agents for their underlying medical practice, clinic, or hospital members are deducted from revenue in the period the related revenue is recognized.
Costs of Revenue
We currently manufacture and intend to continue manufacturing our CapsoCam capsules included in our GI-tract capsule endoscopy solution. To assist us in doing so, we rely on component suppliers and assembly manufacturers based in Asia (particularly Taiwan and Japan).
Costs of revenue include materials, direct labor, and manufacturing overhead costs related to sold products, as well as certain period costs such as non-allocated overhead, scrap, and outbound freight costs, fees paid to physicians for providing reading services, and the costs of operating CapsoCloud as a software-as-a-service offering for video delivery of capsule videos such as shipping costs, processing costs, and data storage costs. All shipping and handling costs directly related to bringing products to their final point of sale are included in costs of revenue. As we expand our product offerings, acquire new customers and existing customers increase their use of our CapsoCam solutions, we expect that our costs of revenue will continue to increase in absolute terms in line with increased revenues.
Gross Profit and Gross Margin
We calculate gross profit as revenue less costs of revenue. Gross margin represents gross profit as a percentage of revenue. We expect gross profit and gross margin to change and be affected by various factors going forward, including selling prices, product costs, customer mix and production volumes and, relatedly, product mix once sale of CapsoCam Colon capsules begin with CapsoCam Colon capsules expected to have a higher sales price and related costs of revenue as compared to CapsoCam Plus, given the additional optics and other features incorporated into the capsule.
Accounts Receivable, Net
Our current payment terms are net 30-days, and only in special cases do we allow more than 30-days for payment. For our international distributors we require pre-payment for the first year and, subject to a favorable credit worthiness determination, we convert them to our standard 30-day term. Our average accounts receivable balance for 2024 was $[ ] million with less than [ ]% of our accounts receivable aged more than 60 days at year-end. We do not have a significant customer credit concentration with no single customer accounting for more than [ ]% of our accounts receivable as of December 31, 2024.
66
Table of Contents
Inventory
Finished goods inventory is stored onsite at our facility in Saratoga, California. We generally average a 3-month supply of finished goods. To date, we have not had a materials or finished goods supply issue that disrupted our ability to supply our customers. Our suppliers have begun to build up raw material supplies for our products and indicated that they will be able to ramp-up production availability according to our internal sales forecasts for both CapsoCam Plus and CapsoCam Colon. During 2025, we will evaluate potentially increasing our on-hand finished goods inventory to a 6-month supply. In 2025, we plan to evaluate all our suppliers to, among other things, (i) ensure they are able to meet anticipated demand for our CapsoCam Colon capsule and (ii) mitigate any supply chain and geopolitical issues.
Operating Expenses
Selling and Marketing
Selling and marketing expenses include costs directly attributable to actively marketing our products and services using both direct employees and outside contractors or vendors. These costs include salaries, bonuses, benefits, and stock-based compensation, sales commissions, travel costs and expense reimbursements, and the costs of sponsoring programs, events, and conferences.
We expect our selling and marketing expenses to increase in the foreseeable future as we continue to increase the size of our in-house sales organization and market penetration in the U.S. and internationally, and expand indications. However, we expect selling and marketing expenses to decrease as a percentage of revenue primarily as, and to the extent, our revenue grows.
Research and Development
Research and development costs are expensed as incurred in accordance with ASC 730. See Note 3 to our financial statements included elsewhere in this prospectus for further details. Our research and development team includes hardware and software engineers with deep expertise in medical technology, optics, data science, AI, and cloud-based data and security architecture and individuals with extensive clinical development expertise.
Our GI-tract capsule endoscopy solution research and development activities include both (i) activities focused on increasing the value of our solution (such as developing new capsule products, including the associated software component, and new enhancements and features) and (ii) related clinical trial development efforts. Related research and development expenses include salaries, bonuses, benefits, and stock-based compensation for our employees focused on research and development or clinical trials, costs of independent contractors and outside vendors, costs of supplies consumed in or product inventory utilized in clinical trials, and the costs of the clinical trials themselves as charged by trial sites or vendors responsible for multiple trial sites.
During the year ended December 31, 2023 and 2024, our research and development expenses included $2.4 million and $[ ] million of expenses related to ongoing clinical studies, primarily related to the first arm of our pivotal study for our CapsoCam Colon capsule involving approximately 1,200 patients at 20 sites throughout the U.S. At our expense, in accordance with the clinical study protocol, each participant first completed an optical colonoscopy followed by a capsule colonoscopy using our CapsoCam Colon.
In the near term, we expect our clinical development expenses to vary as a percent of revenue as, among others, we (i) finalize the first arm of our CapsoCam Colon large-scale pivotal study and begin the second arm of our CapsoCam Colon pivotal study involving the second generation of that capsule (incorporating further advanced features designed to improve the accuracy of CapsoCam Colon) with the second arm expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S and (ii) commence clinical development of our updated small bowel CapsoCam Plus capsule incorporating our AI assisted reading technology.
67
Table of Contents
In addition to clinical trials costs, our research and development costs also includes engineering for our AI technology, hardware development, and regulatory personnel.
General and Administrative
General and administrative expenses consist primarily of personnel expenses, including salaries, bonuses, benefits, and stock-based compensation expense for personnel in executive, administrative, finance, human resources, and other supporting functions. General and administrative expenses also include professional fees for legal services, consulting services, tax matters and audits, as well as information technology expenses, office expenses, rent, insurance, and foreign exchange gains (losses).
We expect that our general and administrative expenses will increase in the foreseeable future as we increase our headcount to support the continued growth of our business. We also anticipate incurring additional expenses associated with operating as a public company, including increased expenses related to audit, legal, regulatory, compliance, director and officer insurance, investor and public relations, and tax-related services associated with maintaining compliance with the rules and regulations of the SEC and standards applicable to companies listed on a national securities exchange. However, we expect general and administrative expenses to decrease as a percentage of revenue primarily as, and to the extent, our revenue grows.
Provision for Income Taxes
We have recorded deferred tax assets for U.S. federal income taxes for which we provide a full valuation allowance. These deferred tax assets primarily include net operating loss carryforwards of $24.0 million, capitalized research and development expenses of $3.3 million, and research and development tax credits of $1.5 million, as of December 31, 2023, which begin expiring in 2027. We expect to maintain this full valuation allowance for the foreseeable future as it is not more likely than not the deferred tax assets will be realized based on our history of losses.
Results of Operations
The following tables set forth our results of operations for the periods presented and as a percentage of our revenue for those periods. The period-to-period comparison of financial results is not necessarily indicative of financial results to be achieved in future periods.
| Year Ended December 31, | ||||||||||||||||
| 2023 | 2024 | $ Change | % Change | |||||||||||||
| (in thousands, except percentages) | ||||||||||||||||
| Revenue |
$ | 9,753 | % | |||||||||||||
| Costs of revenue |
4,262 | % | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Gross profit |
5,491 | % | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Operating expenses: |
||||||||||||||||
| Selling and marketing |
5,533 | % | ||||||||||||||
| Research and development |
9,333 | % | ||||||||||||||
| General and administrative |
1,972 | % | ||||||||||||||
| Total operating expenses |
16,838 | % | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Operating loss |
(11,347 | ) | % | |||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total non-operating income, net |
54 | |||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Loss before provision for income taxes |
(11,293 | ) | % | |||||||||||||
| Provision for income taxes |
11 | % | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Net loss |
$ | (11,304 | ) | $ | $ | % | ||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
68
Table of Contents
| Year Ended December 31, | ||||||||
| 2023 | 2024 | |||||||
| Revenue |
100 | % | % | |||||
| Costs of revenue |
44 | % | % | |||||
|
|
|
|
|
|||||
| Gross profit |
56 | % | % | |||||
|
|
|
|
|
|||||
| Operating expenses: |
||||||||
| Selling and marketing |
57 | % | % | |||||
| Research and development |
96 | % | % | |||||
| General and administrative |
20 | % | % | |||||
|
|
|
|
|
|||||
| Total operating expenses |
173 | % | % | |||||
|
|
|
|
|
|||||
| Operating loss |
(116 | )% | % | |||||
| Total non-operating income, net |
* | % | ||||||
|
|
|
|
|
|||||
| Loss before provision for income taxes |
(116 | )% | % | |||||
| Provision for income taxes |
* | % | ||||||
|
|
|
|
|
|||||
| Net loss |
(116 | )% | % | |||||
* Less than 1%
| Year Ended December 31, | ||||||||
| 2023 | 2024 | |||||||
| (in thousands) | ||||||||
| Revenue by Geography and Type of Customer |
||||||||
| U.S. – sales to medical practices and hospitals: |
||||||||
| Products (capsules and CapsoView) |
$ | 6,612 | ||||||
| Services (CapsoCloud and reading services) |
627 | |||||||
| Outside U.S. – sales to distributors(1): |
||||||||
| Products (capsules and CapsoView) |
2,514 | |||||||
|
|
|
|||||||
| Total revenue |
$ | 9,753 | ||||||
|
|
|
|
|
|||||
| (1) | Includes direct sales in Germany, which began in 2023, of approximately $320,000 in 2023 and $[ ] in 2024. |
Comparison of the Years Ended December 31, 2023 and 2024
Revenue
| Year Ended December 31, |
||||||||||||||||
| 2023 | 2024 | $ Change | % Change | |||||||||||||
| (in thousands, except percentages) | ||||||||||||||||
| Revenue |
$ | 9,753 | $ | $ | % | |||||||||||
Revenue increased $[ ] million, or [ ]%, for the year ended December 31, 2024, compared to the year ended December 31, 2023. The increase was driven by an increase in our CapsoCam Plus sales due to increased acceptance of CapsoCam Plus and growth in the small-bowel capsule endoscopy market.
Costs of Revenue
| Year Ended December 31, |
||||||||||||||||
| 2023 | 2024 | $ Change | % Change | |||||||||||||
| (in thousands, except percentages) | ||||||||||||||||
| Costs of revenue |
$ | 4,262 | $ | $ | % | |||||||||||
69
Table of Contents
As we continued to scale our business, costs of revenue increased $[ ] million, or [ ]%, for the year ended December 31, 2024, compared to the year ended December 31, 2023 reflecting increased unit sales of CapsoCam Plus for the small-bowel and the related software component.
Gross Profit and Gross Margin
| Year Ended December 31, |
||||||||||||||||
| 2023 | 2024 | $ Change | % Change | |||||||||||||
| (in thousands, except percentages) | ||||||||||||||||
| Gross profit |
$ | 5,491 | $ | $ | % | |||||||||||
| Gross margin |
56 | % | % | % | % | |||||||||||
Gross profit increased $[ ] million, or [ ]%, for the year ended December 31, 2024, compared to the year ended December 31, 2023, in line with increased CapsoCam Plus unit sales and the related software component.
Operating Expenses
| Year Ended December 31, |
||||||||||||||||
| 2023 | 2024 | $ Change | % Change | |||||||||||||
| (in thousands, except percentages) | ||||||||||||||||
| Selling and marketing |
$ | 5,533 | $ | $ | % | |||||||||||
| Research and development |
9,333 | % | ||||||||||||||
| General and administrative |
1,972 | % | ||||||||||||||
|
|
|
|
|
|
|
|
|
|||||||||
| Total operating expenses |
$ | 16,838 | $ | $ | % | |||||||||||
Selling and Marketing Expenses
Selling and marketing expenses increased $[ ] million, or [ ]%, for the year ended December 31, 2024, compared to the year ended December 31, 2023. Approximately $[ ] million of the total increase was due to the expansion of our direct sales team in the U.S. and expanded distribution network in Europe.
Research and Development Expenses
Research and development expenses increased $[ ] million, or [ ]%, for the year ended December 31, 2024, compared to the year ended December 31, 2023. Of the total increase, $[ ] million related to an increase in clinical trials costs associated with our CapsoCam Colon pivotal study and less than $[ ] million related to other research and development activities. Costs associated with clinical trials are included in our research and development expenses.
General and Administrative Expenses
General and administrative expenses increased $[ ] million, or [ ]%, for the year ended December 31, 2024 compared to the year ended December 31, 2023. Of the total increase, $[ ] million was due to increased consulting and accounting fees associated with the audit of our financial statements, and $[ ] million was due to headcount related expenses, expanded office space, and increased recruitment expenses.
Liquidity and Capital Resources
Overview
To date, we have financed our operations primarily through the net proceeds we have received from the sales of our convertible preferred stock and common stock as well as cash generated from sales of our CapsoCam
70
Table of Contents
Plus capsule endoscopy solution. We have generated losses from our operations as reflected in our accumulated deficit of $110.5 million as of December 31, 2023, and used $10.8 million in operating activities for the year ended December 31, 2023.
Our losses primarily resulted from the costs incurred in the development and sales and marketing of our products and providing general and administrative support for our operations. We expect to continue to incur losses in the foreseeable future and to expend significant amounts of cash in the foreseeable future as we continue to scale our business, invest in research and development activities, increase sales and marketing expenses to support commercial expansion, and increase general and administrative expenses to support being a publicly-traded company.
Our audited financial statements for the year ended December 31, 2023 included in this prospectus note that there is substantial doubt about our ability to continue as a going concern within one year after the date of issuance of those financial statements; and in the auditors’ report accompanying our audited financial statements included herein, our independent registered public accounting firm included an explanatory paragraph stating that our recurring losses, negative cash outflows from operations and stockholders’ deficit as of December 31, 2023 raise substantial doubt as to our ability to continue as a going concern. This means that our management and our independent registered public accounting firm have expressed substantial doubt about our ability to continue our operations without an additional infusion of capital from external sources. Our audited financial statements have been prepared on a going concern basis and do not include any adjustments to reflect the future effects on the recoverability and classification of assets or the amounts and classification of liabilities that may be necessary should we be unable to continue as a going concern. If we are unable to finance our operations, our business would be in jeopardy and we might not be able to continue operations and might have to liquidate our assets. In that case, investors might receive less than the value at which those assets are carried on our audited financial statements for the year ended December 31, 2023, and it is likely that investors would lose all or a part of their investment. See “Risk Factors—Risks Relating to Our Business and Industry—Our audited financial statements for the year ended December 31, 2023 include a footnote raising substantial doubt about our ability to continue as a “going concern” and, even if this offering is successful, we may still need to raise additional financing to fund our business and revenue growth plans.”
We have lease obligations and other contractual obligations and commitments as part of our ordinary course of business. See Note 7 to our financial statements included elsewhere in this prospectus for information about our lease obligations. In addition, see Note 8 to our financial statements included elsewhere in this prospectus for information about our other commitments and contingencies.
Source of Liquidity
As of December 31, 2024, our principal source of liquidity consisted of $9 million in cash.
Funding Requirements
Based on our current operating plan, we believe that the estimated net proceeds from this offering together with existing cash balances, will be sufficient to fund our operations for at least the next 12 months. We have based this estimate on assumptions that may prove to be wrong, and we could deplete our capital resources sooner than we expect. We may experience lower than expected cash generated from operating activities or greater than expected costs of revenue or operating expenses, and may need to raise additional capital to fund operations, further research, and development activities, or acquire, invest in, or in-license other businesses, assets, or technologies.
Our future capital needs will depend upon our ability to execute our revenue growth plans (as described in greater detail above) and many factors, including:
| • | the cost and pace of developing new products, enhancements to existing products and our research and development activities; |
71
Table of Contents
| • | the market acceptance of our products; |
| • | our ability to develop and commercialize our CapsoCam capsule endoscopy solution for new indications, patient populations and clinical use cases; |
| • | our ability to successfully complete any required clinical or other studies and obtain and maintain any required regulatory approval or clearances; |
| • | insurer and third-party reimbursement of the costs associated with our GI-tract capsule endoscopy solution; |
| • | successful management of our global supply chain including our component suppliers and assembly manufacturers for our CapsoCam solution, many of which are located in Asia (particularly Taiwan and Japan) and some of which are currently single-source suppliers; |
| • | successful growth and leveraging of our global sales team (including, where appropriate, distributors) and marketing team to sell and market our GI-tract capsule endoscopy solution; |
| • | the costs of attaining, defending, and enforcing our intellectual property rights; |
| • | whether we acquire third-party products or technologies; |
| • | the amount and nature of competition from other GI-tract diagnostic products or procedures; |
| • | our ability to raise additional funds to finance our operations; and |
| • | the costs associated with being a public company. |
If these sources of cash are insufficient to satisfy our liquidity requirements, we may need to engage in equity or debt financings to secure additional funds. If we raise additional funds through further issuances of equity or convertible debt securities, our existing stockholders could suffer significant dilution, and any new equity securities we issue could have rights, preferences, and privileges superior to those of holders of our common stock. In addition, the incurrence of indebtedness would increase our fixed obligations and include covenants or other restrictions that could impede our ability to manage our operations. Our ability to raise additional funds may be adversely impacted by deteriorating global economic conditions and the disruptions to and volatility in the credit and financial markets in the U.S. and fluctuations in interest rates, resulting from factors that include but are not limited to, inflation, the conflict between Russia and Ukraine, political tensions between China and Taiwan and other factors, diminished liquidity and credit availability, declines in consumer confidence, declines in economic growth, increases in unemployment rates and interest rates, and uncertainty about economic stability. If the equity and credit markets deteriorate, it may make any necessary debt or equity financing more difficult, more costly, and more dilutive. Further, if additional financing is needed, we may not be able to obtain additional financing on terms favorable to us or at all. Our inability to obtain adequate financing or financing on terms satisfactory to us, when we require it, could significantly limit our ability to continue supporting our business growth and responding to business challenges and opportunities.
Cash Flows
The following table shows a summary of our cash flows for each of the periods presented:
| Year Ended December 31, | ||||||||
| 2023 | 2024 | |||||||
| (in thousands) | ||||||||
| Net cash used in operating activities |
$ | (10,804) | $ | |||||
| Net cash used in investing activities |
$ | (754) | $ | |||||
| Net cash provided by (used in) financing activities |
$ | 20,529 | $ | |||||
| Net increase in cash |
$ | 8,971 | $ | |||||
72
Table of Contents
Net Cash Used in Operating Activities
Net cash used in operating activities during the year ended December 31, 2023, consisted primarily of our net loss of $11.3 million (including $5.5 million in gross profit) and also reflecting (i) a $0.7 million increase in inventory and (ii) a $0.8 million decrease in prepaid expenses primarily related to prepaid 2023 clinical trial related activities.
Net Cash Used in Investing Activities
Net cash used in investing activities during the year ended December 31, 2023 was $0.8 million, and consisted of purchases of property and equipment.
Net Cash Provided by (Used in) Financing Activities
Net cash provided by financing activities during the year ended December 31, 2023, consisted primarily of $20.4 million in proceeds from the sale of our Series H convertible preferred stock and $0.2 million in proceeds from the exercise of options and warrants.
Critical Accounting Estimates
Our management’s discussion and analysis of our financial condition and results of operations are based on our financial statements, which have been prepared in accordance with U.S. GAAP. Preparation thereof requires us to make estimates and judgments that affect the reported amounts of assets, liabilities, revenues, expenses, and certain items included in the footnotes to our financial statements. See Note 3 to our financial statements included elsewhere in this prospectus for information about our significant accounting policies. While we believe the processes used in developing our estimates to be reasonable, actual amounts may differ from estimates. The following reflects the critical accounting estimates used in the preparation of our financial statements. The term “critical accounting estimates” refers to those estimates that involve a significant level of estimation uncertainty that have had, or are reasonably likely to have, a material impact on our financial condition or results of operations.
Research and Development Expenses
Research and development expenses, generally expensed as incurred, include salaries, bonuses, benefits, and stock-based compensation for employees focused on research and development or clinical trials, costs of independent contractors and outside vendors, costs of supplies consumed in or product inventory utilized in clinical trials, costs of the clinical trials themselves as charged by trial sites or vendors responsible for multiple trial sites, and purchased or in-licensed intellectual property used in new product development where there is no alternative future use. We capitalize prepayments for goods or services, including trial device inventory or pre-paid clinical trial amounts, that will be used, consumed, or rendered for future research and development activities and recognize expense as the related goods are delivered or services are performed. We also record expenses and accruals for estimated costs of research and development activities that have not yet been billed to us, including services for clinical trials.
Research and development expenses possess significant estimation uncertainty that have been, or are reasonably likely to be, material and include costs incurred with clinical trial sites and related vendors.
Such estimation uncertainty arises because: (i) the total time periods over which costs are expected to be incurred, and attributed to, may contract or expand based upon difficult-to-predict outcomes such as better than expected progress in, or delays in, clinical trials (including due to study protocol amendments); (ii) the costs’ behavior may be variable, or fixed, requiring judgments regarding proper attribution methodologies; and (iii) visibility of the precision of the exact progress of a particular study subject through the phases of the trial may be very limited, directly implicating necessary assumptions regarding informed consent, enrollment, or completion and speed of progress.
73
Table of Contents
Revenue Recognition
We recognize revenue using a five-step model prescribed by U.S. GAAP resulting in revenue being recognized as performance obligations within a contract are satisfied. Judgment is required to apply the principles-based, five-step model for revenue recognition. Management is required to make certain estimates and assumptions about our contracts with our customers. To date, with limited exception for capsule video reading service revenues, all our revenue is generated from the sale of our CapsoCam capsules to our customers (classified as product revenue) and the use of CapsoCloud or CapsoAccess (together with CapsoView) as the customer selected capsule video delivery option for (i) streaming the capsule videos (via CapsoCloud) for clinician review (classified as service revenue) or (ii) downloading the capsule videos (via CapsoAccess and CapsoView) for clinician review (classified as product revenue).
Those aspects of the five-step revenue recognition model prescribed by U.S. GAAP that give rise to significant levels of estimation uncertainty and have been, or are reasonably likely to be, material to our financial condition or results of operations include (i) the estimation of stand-alone selling prices for performance obligations that are not, or are rarely, sold separately or possess a wide range of observed historical selling prices and (ii) the measurement of applicable time periods to be used for recognition of revenue which is initially deferred at the time of sale and subsequently recognized over time or at a point in time.
The estimation of stand-alone selling prices, where we rely upon a cost-plus-expected margin approach, variability in our expected profit margin assumptions can, in turn, cause variation in estimated stand-alone selling prices and therefore the amounts of revenue allocated to different performance obligations.
For the estimation of applicable time periods impacting recognition of revenue deferrals, estimates are determined using historical customer behavior and deliverable timing data. Usage of historical data requires consideration of trends, patterns, and changes in the underlying data, judgments regarding sampling, and judgments as to whether historical data needs to be adjusted for future expectations. Changes in these underlying inputs have not significantly impacted service revenues or deferrals in the most recent period.
Stock-Based Compensation
We maintain equity incentive plans to provide long-term incentives for employees, consultants and members of our board of directors. These plans allow for the issuance of non-statutory and incentive stock options to employees and non-statutory stock options to consultants and non-employee directors. We are required to determine the fair value of equity incentive awards and recognize compensation expense for all equity incentive awards granted, including employee stock options.
We account for stock-based compensation awards, including stock options to employees and non-employees, based on their estimated grant date fair value. We recognize the fair value of stock options, which vest based on continued service, on a straight-line basis over the requisite service period, which is generally four years. As described in detail in Note 10 to our financial statements included elsewhere in this prospectus, determining the fair value of stock-based compensation to be recorded typically involves the use of an option-pricing model (which, for our Company, is the Black-Scholes option-pricing model).
A significant level of estimation uncertainty arises from certain inputs to the Black-Scholes model. Those inputs include (i) the fair value of the underlying common stock and (ii) the volatility of that fair value.
Recently Issued Accounting Pronouncements
See Note 3 to our financial statements included elsewhere in this prospectus for a description of recent accounting pronouncements applicable to our financial statements.
74
Table of Contents
Internal Control Over Financial Reporting
Prior to our initial public offering, we have been a private company with limited accounting and financial reporting personnel and other resources to address our internal controls and procedures. In connection with the audit of our financial statements as of December 31, 2023, we identified two control deficiencies in our financial reporting process that constitute material weaknesses as of the year ended December 31, 2023. The first material weakness relates to our failure to design or maintain sufficient controls over implementation of information technology general controls or complementary user entity controls for applications (such as our ERP, payroll, and stock option management IT systems) used in the preparation of our financial statements. More specifically, we did not design or maintain sufficient controls related to user access provisioning and monitoring, change management, program development and data management. The second material weakness relates to a lack of segregation of duties in the financial reporting function due to a limited number of staff performing the financial reporting function. More specifically, there is a limited level of multiple reviews among those tasked with preparing our financial records and with respect to our existing ERP system proper segregation of duties was not enforced (i.e., for journal entries, we did not always divide responsibilities with one person responsible for making the journal entry and a different person responsible for approving the journal entry).
As defined in the standards established by the PCAOB, a “material weakness” is a deficiency, or a combination of deficiencies, in internal control over financial reporting such that there is a reasonable possibility that a material misstatement of the company’s annual or interim financial statements will not be prevented or detected on a timely basis.
To respond to these material weaknesses, we have devoted, and plan to continue to devote, significant effort and resources to the remediation and improvement of our internal control over financial reporting. For example, in addition to hiring a Corporate Controller in Q1 2025, we are also evaluating the current and future headcount and other needs of the accounting department to ensure proper segregation of duties. We are also evaluating an upgrade to or replacement of our existing ERP system which would help address many of the control issues contributing to these material weaknesses. We believe these measures will assist us with meeting the Sarbanes-Oxley Act compliance requirements and improving our overall internal controls. However, we cannot assure you that these measures may fully address the material weaknesses in our internal control over financial reporting or that we may conclude that they have been fully remediated.
Assuming we complete remediation efforts in the near term, we may not have sufficient time to test the effectiveness of our internal control over financial reporting. In addition, we cannot assure you that we will be able to successfully remediate these material weaknesses and, even if we do, we cannot assure you that we will not suffer from other material weaknesses in the future. Except for additional personnel costs, costs of consultants, and costs to upgrade or replace our existing ERP system, we do not expect to incur any material costs related to our remediation efforts.
The process of designing and implementing an effective financial reporting system is a continuous effort that requires us to anticipate and react to changes in our business and the economic and regulatory environments and to expend significant resources to maintain a financial reporting system that is adequate to satisfy our reporting obligation. See “Risk Factors—Risks Relating to Our Common Stock and this Offering—We have identified two material weaknesses in our internal control over financial reporting. These material weaknesses could adversely affect our ability to report our results of operations and financial condition accurately and in a timely manner.”
Suppliers and Assembly Manufacturers
To assist us in manufacturing our GI-tract capsule endoscopy products, we rely on component suppliers and assembly manufacturers based in Asia (particularly Taiwan and Japan). Many of our suppliers/manufacturers are single source and located in Asia. For example, the single source suppliers of our lens modules and ASICs are
75
Table of Contents
located in Taiwan and the single source supplier of our CMOS image sensors is located in Japan. Currently, assembled CapsoCam capsules are shipped by our supplier from Taiwan to our U.S. facility where we complete the manufacturing process before distributing the capsules to our distribution network. Any disruption to our supply chain could significantly harm our ability to effectively manufacture and deliver our CapsoCam capsules and, in turn materially harm, our financial results. For additional information regarding risks relating to our suppliers and assembly manufacturers, see “Risk Factors—Risks Relating to Our Business and Industry—We rely on various suppliers to assist us in the assembly and manufacture of our GI-tract capsule endoscopy solution and sourcing of critical and other components and many of these suppliers are single source suppliers located in Asia (particularly Taiwan and Japan); any disruption in our supply chain could adversely affect our ability to meet the demand for our products and fulfil our orders.”
Political and Natural Disaster Risk in Asia
Political instability in Asia (including tensions between China and Taiwan) or other events such as trade and other international disputes, conflicts or war, restrictions on international trade (such as tariffs and other controls on imports or exports of goods, technology, or data) could disrupt or require changes to our supply chain, increase our costs and harm our business. In addition, Taiwan and Japan are also susceptible to natural disasters, such as earthquakes and typhoons that could similarly disrupt or impact our business. For additional information regarding political and other risks in Asia, see “Risk Factors—Risks Relating to Our Business and Industry—We face substantial geopolitical and trade risks (including a possible trade war and imposition of tariffs and supply chain interruptions) and natural disaster risks associated with our business operations in Asia (particularly Taiwan and Japan).”
Inflation
Inflation generally affects us by increasing our cost of labor, manufacturing overhead costs related to sold products and salaries. Our operating costs have increased and may continue to increase because of these pressures and we may not be able to fully offset these cost increases by raising prices for our CapsoCam capsules or the associated software, CapsoCloud and CapsoView.
Emerging Growth Company and Smaller Reporting Company Status
We are an emerging growth company, as defined in the JOBS Act. The JOBS Act permits an “emerging growth company” such as us to take advantage of an extended transition period for complying with new or revised accounting standards. This provision allows an emerging growth company to delay the adoption of some accounting standards until those standards would otherwise apply to private companies. We have elected to use the extended transition period for any other new or revised accounting standards during the period in which we remain an emerging growth company; however, we may adopt certain new or revised accounting standards early. As a result, we will not be subject to the same new or revised accounting standards as other public companies that are not emerging growth companies and our financial statements may not be comparable to other public companies that comply with new or revised accounting pronouncements as of public company effective dates. The JOBS Act also exempts us from having to provide an attestation and report from our independent registered public accounting firm on the assessment of our internal control over financial reporting pursuant to the Sarbanes-Oxley Act.
We will remain an emerging growth company until the earliest of: (i) the last day of the fiscal year following the fifth anniversary of the consummation of this offering; (ii) the last day of the fiscal year in which we have total annual gross revenue of at least $1.235 billion; (iii) the last day of the fiscal year in which we are deemed to be a “large accelerated filer” as defined in Rule 12b-2 under the Exchange Act, which would occur if the market value of our common stock held by non-affiliates exceeded $700.0 million as of the last business day of the second fiscal quarter of such year; or (iv) the date on which we have issued more than $1.0 billion in non-convertible debt securities during the prior three-year period.
76
Table of Contents
We are also a “smaller reporting company” as defined in the Exchange Act. We may continue to be a smaller reporting company even after we are no longer an emerging growth company. We may take advantage of certain of the scaled disclosures available to smaller reporting companies and will be able to take advantage of these scaled disclosures for so long as our voting and non-voting common stock held by non-affiliates is less than $250.0 million measured on the last business day of our second fiscal quarter, or our annual revenue is less than $100.0 million during the most recently completed fiscal year and our voting and non-voting common stock held by non-affiliates is less than $700.0 million measured on the last business day of our second fiscal quarter. We cannot predict if investors will find our shares of common stock less attractive because we may rely on these exemptions. If some investors find our shares of common stock less attractive as a result, there may be a less active trading market for shares of our common stock and our share price may be more volatile.
77
Table of Contents
Overview
We are a commercial-stage medical technology company focused on creating diagnostic and screening products to identify abnormalities of the GI tract. We develop advanced imaging and AI technologies in creating such products while maximizing the flexibility, convenience, profitability, and safety of patient care. We are a Delaware company, incorporated in 2005. Our corporate headquarters is located in Saratoga, California.
Currently, our GI-tract capsule endoscopy solution comprises our single-use CapsoCam capsule and the associated software, CapsoCloud and CapsoView. The CapsoCam capsule, with its panoramic view, acquires and stores video images in onboard memory while moving through the GI tract and the software component allows healthcare providers to view the video retrieved from the capsule by either streaming it from the cloud, where it is securely stored, anywhere at their convenience using our CapsoCloud software or downloading data from the capsule themselves and reviewing it in our CapsoView software. Our first U.S. Food and Drug Administration (“FDA”) cleared capsule endoscopy is our small bowel capsule (the current generation of which we refer to as CapsoCam Plus), for which we began commercial sales in Europe in 2012, subsequent to which we received 510(k) clearance in 2016 and began commercial sales in the U.S. in 2017. CapsoCam Plus is used to visualize the small bowel mucosa to detect abnormalities of the small bowel in adults and children aged 2 years and above. As of December 31, 2024, our CapsoCam Plus has been used for over 120,000 patients. For the year ended December 31, 2023 and 2024, we generated approximately $9.8 million and $ million, respectively, in revenue from sales of CapsoCam Plus, an increase of approximately % over the prior year. Driven by increased acceptance of CapsoCam Plus and growth in the small-bowel capsule endoscopy market, our revenue has increased in each year since we began U.S. direct sales in 2020. We are (i) in the process of updating CapsoCam Plus to add our self-developed AI assisted reading technology and (ii) targeting related FDA 510(k) clearance of the updated capsule for late 2025, with commercialization shortly thereafter. Our AI assisted reading tools detect and highlight suspected abnormalities for a clinician, reducing their time to review a capsule video and making capsule endoscopy more financially attractive to their practice.
Building upon the commercial success and the existing design of our CapsoCam Plus capsule, we developed our next pipeline capsule endoscope, CapsoCam Colon. Our CapsoCam Colon capsule (i) leverages CapsoCam Plus’s existing capsule design with its panoramic view and (ii) incorporates both our self-developed AI to automatically detect polyps in the video and our polyp-size measurement tool enabled by a 3D sensor in the capsule (polyp size being highly correlated with a polyp’s risk of becoming cancer). We are currently (i) concluding a pivotal study to support 510(k) clearance of CapsoCam Colon (including the use of our AI technology) for use by currently indicated patients who are a subset of the colorectal cancer screening and surveillance populations, (ii) planning to submit our 510(k) application, with two existing capsule endoscopies as predicate devices, to the FDA in the first half of 2025 with a goal of obtaining 510(k) clearance from the FDA in late 2025, (iii) planning to extend our pivotal study to include a second arm for our second generation CapsoCam Colon capsule, which will include improvements such as a new lens and illumination optics with an increased field of view and improved image quality, and (iv) planning to use the clinical results of the second arm to support submission to the FDA of a 510(k) application in late 2025 with a goal of obtaining 510(k) clearance in the first half of 2026. As a part of this second 510(k) application and to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation. We plan to commercialize our CapsoCam Colon capsule shortly thereafter.
We currently sell CapsoCam Plus capsules in the U.S. to our customers, which primarily include gastroenterologists practicing in clinics and/or hospitals, primarily through our in-house sales team. Outside the U.S., we sell CapsoCam Plus through a combination of our in-house sales team and qualified distributors. In 2023 and 2024, international sales accounted for 26% and % of total revenue.
78
Table of Contents
We believe that our GI-tract capsule endoscopy solution is positioned to benefit from (i) our existing sales and marketing structure (with our in-house sales team and marketing team targeting sales of our various approved GI tract diagnostic products to the same target customer base), (ii) technological advancements (including improvements to our proprietary AI and other technologies and third-party supplier improvements in optics and storage capacity) and (iii) increased telemedicine adoption (following FDA clearance in December 2024 of remote ingestion of our CapsoCam Plus). We also believe our solution can be adapted to address new GI indications. Potential new medical indications include pancreatic cancer and esophageal medical conditions (such as esophageal varices and Barrett’s esophagus). In 2025, we plan to commence clinical investigation of CapsoCam’s accuracy in (i) detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) and (ii) screening esophageal varices in cirrhotic patients with portal hypertension.
CapsoCam Plus
Diseases of the small bowel include obscure GI bleeding, chronic iron-deficiency anemia, Crohn’s disease, tumors, and polyposis. Capsule endoscopy is the first-line modality for imaging the mucosa of the small bowel including the pathologies characterizing these diseases. Various methods of enteroscopy for reaching the entirety of the small bowel, which is approximately 20 feet long, are invasive, time consuming, and require a high level of endoscopy skill. Enteroscopy is still required for biopsy or to provide certain therapies, but for diagnostic visualization, capsule endoscopy is preferred for its simplicity, non-invasiveness, and relatively low cost. Competitor capsule endoscopy systems that are currently available in the market consist of capsules with end-view systems, which provide only limited “tunnel” or partial “wall” views of the small bowel, and wired data recorders worn on the body, which incur an upfront capital expense and clinical workflow complications for providers and discomfort and multiple clinical visits for patients.
We believe our CapsoCam Plus is a superior capsule endoscopy system, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. Our CapsoCam captures a full 360° panoramic video, providing a complete view of the GI mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature, ultimately resulting in superior diagnostic yield. This was demonstrated by a large, single center retrospective study comparing the clinical performance of the CapsoCam system to competitor systems between 2012 and 2018. The same study also noted CapsoCam’s operational benefits, which included higher exam completion rates and less lost data. In particular, our CapsoCam is a zero-capex “wire-free” data collection solution for providers as it stores the entire video in onboard memory. Following retrieval, our cloud-based platform, CapsoCloud, gives providers in the U.S. the ability to remotely access data from the cloud and stream in vivo videos anywhere at their convenience. Outside of the U.S., providers review procedure videos using CapsoView software (primarily due to foreign data privacy and access regulations).
In the first half of 2025, we plan to file for FDA and EU clearance for the use of AI in our CapsoCam Plus capsule. Also, we are currently developing a capsule delivery device (availability expected in 2025) and a patency capsule (for verifying a capsule endoscope can pass through the bowel without retention prior to an exam) (tentative FDA 510(k) submission planned by mid-2025). The recent indication for children aged 2 years and above and the delivery device should enable faster penetration of the pediatric market. Of the patients who are unable to swallow a capsule, many are children. Patency capsules are used primarily with Crohn’s disease patients to verify that a capsule endoscope can pass through the GI tract without retention at a stricture, a narrowing of the small bowel which can result from inflammation and scarring associated with Crohn’s disease. The patency capsule is the same diameter as a capsule endoscope such that if it passes without delay, the CapsoCam is also likely to pass without retention (but will dissolve if retained). The U.S. capsule endoscopy market is estimated at approximately $[ ] in [ ], and is forecasted to reach approximately $[ ] in [ ]. The small bowel market is currently the dominant sector within the broader capsule endoscopy market in the U.S.
79
Table of Contents
CapsoCam Colon
A colon polyp is a clump of cells that forms on the lining of the colon. Most colon polyps are harmless, but, over time, some colon polyps develop into CRC. The size of a polyp is highly correlated with its risk of becoming cancerous. Currently, optical colonoscopy, accompanied by polypectomy and biopsy, is considered the gold-standard for the detection of colorectal polyps and cancers. Colon capsule endoscopy provides non-invasive visualization of the entire colon from the cecum to the rectum, and it has demonstrated good sensitivity and specificity for the detection of colon polyps. It is intended to be used for (a) patients after an incomplete optical colonoscopy with adequate preparation and a complete evaluation of the colon was not technically possible and (b) patients with evidence of GI bleeding of lower GI origin with major risks for colonoscopy or moderate sedation, but who can tolerate colonoscopy and moderate sedation in the event a clinically significant colon abnormality is identified on capsule endoscopy. Currently, only two competitive products are available in the market for these indications–only one of which, Medtronic’s PillCam COLON 2, is available in the U.S. These competitor capsules have end-view systems, which provide only limited “tunnel” views of the colon, and wired data recorders worn on the body, which incur an upfront capital expense and clinical workflow complications for providers and discomfort and multiple clinical visits for patients.
We believe our CapsoCam Colon, once FDA cleared, will be a superior capsule endoscopy system, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. Our CapsoCam Colon utilizes our self-developed AI for automated polyp detection. AI improves diagnostic yield and provides more consistent accuracy with reduced dependency on the experience level of the physician video reader and their level of fatigue and distraction. Our CapsoCam Colon also incorporates our proprietary 3D-sensing technology to more accurately measure the size of polyps in the GI tract, enabling physicians to more confidently decide that patients with small (e.g., less than 6mm) polyps may forgo a follow-on colonoscopy, increasing the utility of the procedure for healthcare providers and patients alike. Similar to our CapsoCam Plus, our CapsoCam Colon also captures a full 360° panoramic video, is a zero-capex “wire-free” data collection solution for providers and utilizes our cloud-based platform, CapsoCloud, for providers in the U.S. and CapsoView software for providers outside of the U.S. Further, CapsoCam Colon is a panenteric capsule which, when programmed with slightly different operating parameters, can be used to visualize both the small bowel and the colon in one procedure for the evaluation of Crohn’s disease, ulcerative colitis, irritable bowel syndrome and obscure GI bleeding.
Current indications for colon capsule endoscopy are limited to patients with evidence of lower-GI bleeding (such as a positive stool test) for whom the risk of colonoscopy or moderate sedation is significant and for patients who have had an incomplete colonoscopy, with adequate preparation. The addressable population in the U.S. includes approximately [ ] million patients. The estimated annual U.S. total addressable market opportunity is approximately $[ ].
We have established a competitive advantage through multiple strategic initiatives, including investing substantial resources to create our intellectual property portfolio. As of December 31, 2024, we had over 140 issued patents covering multiple aspects of our capsules and technology.
We invest in research and development initiatives that are focused on introducing enhancements and improvements aimed at increasing the value provided by our GI-tract capsule endoscopy solution. Our research and development team includes hardware and software engineers with deep expertise in medical technology, optics, data science, AI, and cloud-based data and security architecture and individuals with extensive clinical development expertise.
80
Table of Contents
Our Program
We developed our first commercial product, the CapsoCam capsule endoscopy solution (the current generation of which we refer to as CapsoCam Plus), for visualization of the small bowel mucosa. The following table summarizes key information about our clinical program for the small bowel capsule:
| Indication |
Clinical Trials |
Estimated Timeline | ||
| Visualize small bowel and detect pathologies | Completed | Received FDA 510(k) clearance in 20161 | ||
| Data Collection (AI) Study | FDA 510(k) submission expected in 1st half of 2025 |
| 1 | Received FDA 510(k) clearance in December 2024 for pediatric (children aged 2 years and above) use and telemedicine supervision (i.e., remote ingestion) indications. |
We have developed our next pipeline product, the CapsoCam Colon capsule endoscopy system, for visualization of the colon and detection and measurement of polyps. The following table summarizes key information about our clinical program for CapsoCam Colon:
| Indication |
Clinical Trials |
Estimated Timeline | ||
| Visualize colon and detect/measure polyps | Feasibility Study | Completed | ||
| Pilot Study | Completed | |||
| Pivotal Study (first arm)—1st generation capsule | FDA 510(k) submission expected in 1st half of 2025 | |||
| Pivotal Study (second arm)—improved 2nd generation capsule | FDA 510(k) submission expected in late 2025 |
Our Strengths
We believe the continued growth of our Company will be driven by the following factors:
| • | Sole capsule endoscope with a 360° panoramic view available in the market. Only our CapsoCam Plus boasts a 360° panoramic lateral view. It houses four high-resolution cameras around its circumference, and the images from each are stitched into a single panoramic image. Compared to competitive end-view systems, a 360° panoramic lateral view provides a complete view of the GI mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature, resulting in demonstrated superior diagnostic yield. |
| • | Telemedicine-enabled and zero-capex “wire-free” data collection and remote data analysis. Our CapsoCam is a zero-capex “wire-free” data collection solution for providers, as it stores the entire video in onboard memory. In December 2024, we received FDA 510(k) clearance for telemedicine supervision (i.e., remote ingestion) of our CapsoCam Plus, allowing patients to ingest our capsule in the comfort of their own homes, under the remote supervision of providers. The CapsoCam solution frees up exam-room schedules for providers and provides flexibility to administer capsules any day at any time. A provider’s practice can easily scale to multiple capsules per day with no added cost, and there is no equipment to recover from patients. Our CapsoCam Plus solution includes our cloud-based platform, CapsoCloud, which provides a flexible, trackable, streamlined, and capital-equipment-free workflow for providers in the U.S. It also allows clinicians to track procedures and stream in vivo videos anywhere at their convenience, generate reports, store and manage patient data, and transfer data to third-party reading services. We believe at-home procedures and remote analysis via CapsoCloud will be attractive to providers and patients alike, particularly for future screening indications. |
| • | Automated pathology detection due to usage of AI. CapsoCam Colon (subject to FDA clearance) incorporates deep learning AI for automated pathology detection of polyps, a capability that competitive |
81
Table of Contents
| systems lack. In the first half of 2025, we also plan to file for FDA and EU clearance for the use of AI in our CapsoCam Plus capsule. Our AI assisted reading tools detect and highlight suspected abnormalities for a clinician, reducing their time to review the video and making capsule endoscopy more financially attractive to their practice. CapsoCloud continuously acquires consenting patients’ clinical data, enabling our in-house AI experts to develop ever-improving automated lesion detection and classification. |
| • | 3D-sensing technology informs follow-on care decisions. Our CapsoCam Colon (subject to FDA clearance) incorporates our proprietary 3D-sensing technology to more accurately measure polyp sizes. Polyp size is highly correlated with its risk of becoming cancer. No other capsule endoscope currently in the market has 3D-enabled measurement capability. With automated pathology detection and the ability to manually review video frames adjacent to an identified polyp, physicians can more confidently decide that patients with small (e.g., less than 6mm) polyps may forgo a follow-on colonoscopy, increasing the utility of the procedure for healthcare providers and patients alike. |
| • | Experienced leadership team. Our senior management team consists of industry professionals with deep industry expertise across various disciplines, including medical technology, engineering, optics, sales and marketing, finance, operations, data science, AI, and clinical operations and research. |
Our Growth Strategies
Our long-term, lifesaving vision is an ingestible capsule that, in a single convenient non-invasive procedure, screens for multiple cancers—esophageal, gastric, pancreatic, small-bowel, and colorectal—at early and precancerous stages, utilizing AI to analyze thousands of images captured in the GI tract. We are building towards this goal with a planned succession of FDA-cleared indications, targeting existing and nascent markets. Until then, key elements of our nearer-term growth strategy include:
| • | Obtain 510(k) clearance of CapsoCam Colon. We have developed our next pipeline product, CapsoCam Colon, for visualization of the colon and detection and measurement of polyps. In addition to having a 360° panoramic lateral view, it incorporates deep learning AI for automated pathology detection of polyps and 3D-sensing technology to more accurately measure polyp sizes. We are currently concluding a pivotal study to support 510(k) clearance of CapsoCam Colon for use by currently indicated patients who are a subset of the colorectal cancer screening and surveillance populations. The pivotal study involved approximately 1,200 patients enrolled at 20 sites throughout the U.S. Based on the clinical results from the pivotal study, we are planning to submit our 510(k) application, with two existing capsule endoscopies as predicate devices (one for the capsule and one for the incorporated AI), to the FDA in the first half of 2025 with a goal of obtaining 510(k) clearance in late 2025. To enhance the sensitivity and specificity of CapsoCam Colon for detecting and measuring polyps, we are developing our second generation CapsoCam Colon capsule, which will include improvements such as a new lens and illumination optics with an increased field of view and improved image quality, and will extend our pivotal study to include a second arm, which is expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S. We plan to use the clinical results of the second arm to support submission to the FDA of a 510(k) application in late 2025 with a goal of obtaining 510(k) clearance in the first half of 2026. Current indications for colon capsule endoscopy are limited to patients with evidence of lower-GI bleeding (such as a positive stool test) for whom the risk of colonoscopy or moderate sedation is significant and for patients who have had an incomplete colonoscopy, with adequate preparation. As a part of this second 510(k) application, to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation. The addressable population in the U.S. includes approximately [ ] million patients. The estimated annual U.S. total addressable market opportunity is approximately $[ ]. |
| • | Expand clinical use cases and accessories for CapsoCam Plus. In December 2024, we received 510(k) clearance of our CapsoCam Plus for pediatric use in children aged 2 years and above and telemedicine supervision (i.e., remote ingestion). Also, we are currently developing a capsule delivery device (availability expected in 2025) for patients who are unable to swallow the capsule, many of whom are |
82
Table of Contents
| children. The capsule delivery device should enable faster penetration of the newly indicated pediatric market. We are also currently developing a patency capsule (tentative FDA 510(k) submission planned by mid-2025), which is used primarily with Crohn’s disease patients to verify that a capsule endoscope can pass through the GI tract without retention at a stricture, a narrowing of the small bowel which can result from inflammation and scarring associated with Crohn’s disease. |
| • | Continue to improve and innovate our GI-tract capsule endoscopy solution. Our research and development initiatives are focused on introducing enhancements and improvements aimed at increasing the value provided by our GI-tract capsule endoscopy solution. In particular, we are working on improvements to our CapsoCam, including a new lens and illumination optics with an increased field of view, improved image quality and higher peak frame rate. For our AI assisted reading technology, we plan to continue investing to (i) improve the pathology-detection and classification accuracy and the scope of our AI algorithms and (ii) apply AI, including large language models, to streamline the diagnostic and medical-report-generation processes, which in turn improves the efficiency and effectiveness of our healthcare provider customers. We also plan to continue making improvements to our CapsoCloud and CapsoView software. |
| • | Expand into new indications and clinical use cases beyond small bowel and colon. In 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) by visualizing abnormalities of the duodenal papilla. A dilated, or unnaturally opened, duodenal papilla is correlated with GI tract content reflux into the pancreas duct and pancreatic neoplasia and may indicate the presence of, or elevated risk of developing, serious abnormalities like pancreatitis or a tumor of the pancreas. A dilated papilla is frequently not observable by endoscopy due to physical perturbation/disruption in connection with the procedure and heavy patient sedation. Our CapsoCam capsule, with its panoramic view, can observe the papilla in its natural state. Prior studies have demonstrated the CapsoCam’s superior detection of the normal papilla relative to non-panoramic systems. According to the ACS, pancreatic cancer is estimated to be diagnosed in over 67,000 patients in the U.S. in 2025, and approximately 52,000 patients are estimated to die from it. The overall 5-year survival rate is approximately 13%, which is the lowest survival rate of all cancer types. On the other hand, the 5-year survival rate is around 44% if the pancreatic cancer is still in the early stages at the time of diagnosis. There is currently no effective screening for pancreatic cancer. Also, in 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in screening esophageal varices in cirrhotic patients with portal hypertension. |
Market Overview
Overview and Challenges of Visualizing Small Bowel and Detecting Small Bowel Pathologies
Diseases of the small bowel include obscure GI bleeding, chronic iron-deficiency anemia, Crohn’s disease, tumors, and polyposis.
Obscure GI bleeding is recurrent or persistent GI bleeding of uncertain origin. 5% of all GI bleeding is obscure GI bleeding. In approximately 80% of obscure GI bleeding cases, the origin is localized to the small bowel. Chronic iron-deficiency anemia is a condition in which blood lacks adequate healthy red blood cells. Red blood cells carry oxygen to the body’s tissues. Chronic iron-deficiency is the most common nutrient deficiency in the world and a significant common cause of anemia worldwide. Crohn’s disease is a type of inflammatory bowel disease that causes swelling and irritation of the tissues, called inflammation, in the digestive tract. This can lead to belly pain, severe diarrhea, fatigue, weight loss and malnutrition. Inflammation caused by Crohn’s disease can affect different areas of the digestive tract in different people. Crohn’s disease most commonly affects the end of the small intestine and the beginning of the large intestine. The inflammation often spreads into the deeper layers of the bowel. Crohn’s disease often begins in a person’s teens or twenties, though some patients experience symptoms even earlier.
83
Table of Contents
Capsule endoscopy is the first-line modality for imaging the mucosa of the small bowel including the pathologies characterizing these diseases. Various methods of enteroscopy for reaching the entirety of the small bowel, which is approximately 20 feet long, are invasive, time consuming, and require a high level of endoscopy skill. Enteroscopy is still required for biopsy or to provide certain therapies. For diagnostic visualization, however, capsule endoscopy is preferred given its simplicity, non-invasiveness, and relatively low cost.
After obtaining negative esophagogastroduodenoscopy and colonoscopy procedure results, capsule endoscopy of the small bowel is the first-line investigational procedure for obscure GI bleeding. The reported diagnostic yield of capsule endoscopy ranges from 30% to 70%, exceeding that of alternative diagnostic methods, such as push enteroscopy (which has a reported diagnostic yield estimated at approximately 31%), double-balloon enteroscopy (which has a reported diagnostic yield estimated at approximately 23%), and small-bowel series (which has a reported diagnostic yield estimated at approximately 5%). Capsule endoscopy has the additional advantage of non-invasiveness. Due to the often-intermittent nature of GI bleeding, performing capsule endoscopy within 14 days of a bleeding episode maximizes the diagnostic yield.
The sources of bleeding found by capsule endoscopy include ulcers, vascular anomalies, tumors, polyps, and non-specific enteritis. Sometimes blood is detected without identifying a source lesion. Over one half of patients regularly taking non-steroidal anti-inflammatory drugs have small-bowel lesions with the potential to bleed. Capsule endoscopy identifies the type, location, and extent of lesions, and helps determine therapeutic interventions and monitor their effectiveness. The localization of lesions determines the route (upper or lower) of follow-on device-assisted enteroscopy.
Capsule endoscopy is indicated for patients with chronic iron-deficiency anemia when obscure GI bleeding is merely suspected, after obtaining negative esophagogastroduodenoscopy and colonoscopy procedure results. In particular, capsule endoscopy can be considered for patients with severe anemia—requiring blood transfusions, hemoglobin level <100 g/L, or persistent or recurrent iron-deficiency anemia despite iron replacement therapy.
Capsule endoscopy is also indicated for patients presenting with clinical features consistent with Crohn’s disease, after negative ileocolonoscopy and imaging studies. Capsule endoscopy has shown good sensitivity (91%-100%) and specificity (91%-92%) when using ileocolonoscopy as the reference test. A prospective study reported that capsule endoscopy aided Crohn’s disease diagnosis in 83% of cases, influenced decision making in 72%, and changed management in 78%. Capsule endoscopy can be used to provide additional information influencing disease management in patients with known Crohn’s disease, when ileocolonoscopy or imaging results do not fully explain the clinical presentation. It further can be used to assess mucosal healing or when Crohn’s disease is recurrent after colectomy.
Capsule endoscopy is recommended for ongoing surveillance of polyps in patients with intestinal polyposis syndromes, especially Peutz-Jeghers Syndrome. Hereditary polyposis syndromes (including Peutz-Jeghers Syndrome) present a significant risk of complications including bleeding and intussusception. Capsule endoscopy has better diagnostic yield than endoscopy for polyps beyond the duodenum. Capsule endoscopy also has demonstrated detection of more polyps and smaller polyps compared to other non-invasive imaging modalities, such as radiography or magnetic resonance enterography, and similar detection compared to device-assisted enteroscopy.
Our Addressable Market Opportunity in Visualizing Small Bowel and Detecting Small Bowel Pathologies
The U.S. capsule endoscopy market is estimated at approximately $[ ] in [ ] and is forecasted to reach approximately $[ ] in [ ]. The small bowel market is currently the dominant sector within the broader capsule endoscopy market in the U.S.
84
Table of Contents
Overview and Challenges of Detecting Colon Polyps
A colon polyp is a clump of cells that forms on the lining of the colon. Most colon polyps are harmless, but, over time, some colon polyps develop into CRC. The size of a polyp is highly correlated with its risk of becoming cancerous.
Currently, optical colonoscopy, accompanied by polypectomy and biopsy, is considered the gold-standard for the detection of colorectal polyps and cancers. Each year in the U.S., there are approximately 153,000 new cases of CRC and approximately 53,000 deaths. It is widely accepted that CRC is among the most preventable, yet least-prevented cancers. CRC can take up to 10-15 years to progress from a pre-cancerous lesion to metastatic cancer and death. Patients who are diagnosed early in the progression of the disease—with pre-cancerous lesions or early-stage cancer—are more likely to have a complete recovery and to be treated less expensively.
Colon capsule endoscopy provides non-invasive visualization of the entire colon from the cecum to the rectum, and it has demonstrated good sensitivity and specificity for the detection of colon polyps. It is intended to be used for (a) patients after an incomplete optical colonoscopy with adequate preparation and a complete evaluation of the colon was not technically possible and (b) patients with evidence of GI bleeding of lower GI origin with major risks for colonoscopy or moderate sedation, but who can tolerate colonoscopy and moderate sedation in the event a clinically significant colon abnormality is identified on capsule endoscopy. As a part of our 510(k) application for our second generation CapsoCam Colon and to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation.
Incomplete colonoscopies are reported at rates from 4% to 25%. Of these, approximately half result from inadequate bowel preparation and otherwise due to the tortuosity of the colon, adhesions from previous surgeries, angulation, fixation of bowel loops, diverticulosis, or ineffective sedation. A large multicenter study found that 50% of screening patients had a significant dysplastic lesion in the proximal colon. The risk of proximal cancer is 2 times greater when colonoscopy is incomplete. A complete colonoscopy can often be achieved in a second attempt, especially by highly skilled endoscopists employing specialized equipment and techniques, taking extra time, and choosing an appropriate sedation. However, often a non-invasive procedure to visualize the proximal colon is preferred. Computed tomography colonography (“CTC”) and colon capsule endoscopy are both non-invasive options. CTC, also referred to as virtual colonoscopy, is an imaging examination of the entire colon and rectum. CTC uses computed tomography to acquire images and advanced two-dimensional and three-dimensional image display techniques for interpretation. Following incomplete colonoscopy, compared to CTC, colon capsule endoscopy has demonstrated a 61.5% higher diagnostic yield for polyps greater than 5mm and 3 times greater for all polyps. In a multicenter prospective study, among 286 analyzable subjects of average risk in a screening population, the sensitivity for polyps greater than or equal to 6mm for colon capsule endoscopy was 78.2% compared to only 26.8% for CTC.
Many patients have an elevated risk for colonoscopy or the moderate sedation that typically accompanies it. For them, the risk of a screening or surveillance colonoscopy may not be justified without prior evidence that a significant lesion requiring polypectomy is present. Colon capsule endoscopy is a non-invasive low-risk procedure that does not require sedation. It may be used as a preliminary test to determine if a significant polyp is present. However, colon capsule endoscopy is not yet widely used for these patients. A major reason may be that the labeled indications stipulate that these patients have evidence of bleeding from a lower GI origin, such as a positive stool test, prior to colon capsule endoscopy. In most cases, after a positive stool test, providers in the U.S. are likely to refer patients to colonoscopy directly, with colon capsule endoscopy reserved for those cases where the patient refuses colonoscopy or the risk associated with colonoscopy or sedation is especially high.
Our Addressable Market Opportunity in Colon Capsule Endoscopy
We estimate our total addressable market opportunity at over $[ ] annually in the U.S., which represents the potential opportunity from the sale of colon capsule endoscopies to healthcare providers and hospitals.
85
Table of Contents
Other Potential Opportunities
In 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) by visualizing abnormalities of the duodenal papilla. A dilated, or unnaturally opened, duodenal papilla is correlated with GI tract content reflux into the pancreas duct and pancreatic neoplasia and may indicate the presence of, or elevated risk of developing, serious abnormalities like pancreatitis or a tumor of the pancreas. A dilated papilla is frequently not observable by endoscopy due to physical perturbation/disruption in connection with the procedure and heavy patient sedation. Our CapsoCam capsule, with its panoramic view, can observe the papilla in its natural state. Prior studies have demonstrated the CapsoCam’s superior detection of the normal papilla relative to non-panoramic systems. According to the ACS, pancreatic cancer is estimated to be diagnosed in over 67,000 patients in the U.S. in 2025, and approximately 52,000 patients are estimated to die from it. Due to the asymptomatic early stages, in most cases this disease is detected too late, making pancreatic cancer one of the most lethal malignant neoplasms. The overall 5-year survival rate is approximately 13%, which is the lowest survival rate of all cancer types. On the other hand, the 5-year survival rate is around 44% if the pancreatic cancer is still in the early stages at the time of diagnosis. A definitive diagnosis is currently made through a series of investigations, including imaging scans, blood tests and biopsy, which are usually only performed in symptomatic patients. There is currently no effective screening for pancreatic cancer.
Further, in 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in screening esophageal varices in cirrhotic patients with portal hypertension. Patients with cirrhosis who develop portal hypertension are at risk for complications, including bleeding from esophageal varices. Portal hypertension is the result of resistance to portal blood flow, which most often occurs in the liver and with increases in portal blood flow. When esophageal varices rupture, bleeding may be severe and life-threatening.
Despite its advantages, colon capsule endoscopy has not yet been granted approval for a CRC screening indication in the U.S., and is not widely used for screening in other markets. Although many studies suggest that colon capsule endoscopy has polyp-detection accuracy similar to that of a colonoscopy, the quantity of high-quality evidence establishing its effectiveness in a screening population is currently considered insufficient. Colon capsule endoscopy adoption is also hindered by a relatively high percentage of procedures that do not afford complete visualization of the colon, due to inadequate bowel preparation or a battery life shorter than the GI transit time. In many cases, significant polyps detected by colon capsule endoscopy are not detected during subsequent colonoscopy. In part, this reflects the imperfect sensitivity of colonoscopy.
Colon capsule endoscopy has advantages relative to other non-invasive tests that are used for CRC screening. Compared to CTC, colon capsule endoscopy has demonstrated superior polyp detection accuracy and diagnostic yield. Stool tests and blood tests have demonstrated good sensitivity for cancer, but sensitivity for advanced precancerous lesions is under 50%. Relatively high false-positive rates for multi-targeted stool DNA tests, such as Cologuard, lead to unnecessary colonoscopies. In a study with over 10,000 average-risk screening patients, advanced neoplasia was not detected by colonoscopy in 75% of Cologuard-positive cases.
We believe the large, underserved population of unscreened and inadequately screened patients represents a significant opportunity for our CapsoCam Colon capsule endoscopy solution. CRC is the second leading cause of cancer deaths in the U.S. and the leading cause of cancer deaths in the U.S. among non-smokers. Each year in the U.S., there are approximately 153,000 new cases of CRC and approximately 53,000 deaths. It is widely accepted that CRC is among the most preventable, yet least-prevented cancers. CRC can take up to 10-15 years to progress from a pre-cancerous lesion to metastatic cancer and death. Patients who are diagnosed early in the progression of the disease—with pre-cancerous lesions or early-stage cancer—are more likely to have a complete recovery and to be treated less expensively. The ACS recommends that men and women over the age of 45 receive an optical colonoscopy every 10 years or other structural test, such as sigmoidoscopy or CTC, every 5 years or alternatively, a multi-targeted stool DNA test, such as Cologuard, should be performed every 3 years, or other stool-based test, such as FIT or FOBT, should be performed every year. The addressable annual U.S. patient population is at least [ ].
86
Table of Contents
In the future, we intend to conduct clinical studies to support FDA approval of a premarket approval application (“PMA”) for use of CapsoCam Colon as a method for non-invasive detection of cancer and pre-cancerous lesions in the colon. In other words, we would like to seek FDA approval of CapsoCam Colon as a screening test for both the potential prevention of and the detection of cancer.
Our Solutions
CapsoCam Plus
Our CapsoCam Plus capsule endoscopy system is intended for visualization of the small bowel mucosa, to detect abnormalities of the small bowel in adults and children aged 2 years and above. Capsule endoscopy is the first-line modality for imaging the mucosa of the small bowel. After first launching our small bowel capsule in Europe in 2012, we launched in the U.S. in 2017, and as of December 31, 2024, our CapsoCam Plus has been used for over 120,000 patients.
Key Benefits of CapsoCam Plus
We believe that our CapsoCam Plus is a superior capsule endoscopy system compared to competitor systems, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. The benefits delivered by the CapsoCam Plus include the following:
| • | Sole capsule endoscope with a 360° panoramic view available in the market. Our CapsoCam is the only capsule endoscope currently available in the market that captures a full 360° panoramic video, providing a complete view of the GI mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature. Our CapsoCam houses four high-resolution cameras around the capsule circumference, and the images from each are stitched into a single panoramic image. Our competitors’ end-view systems have a camera at one end of the capsule, which affords a limited “tunnel view” when the capsule is in a straight section of the small bowel or a partial “wall view” of the outer curvature when it is in a curved section of the small bowel. |
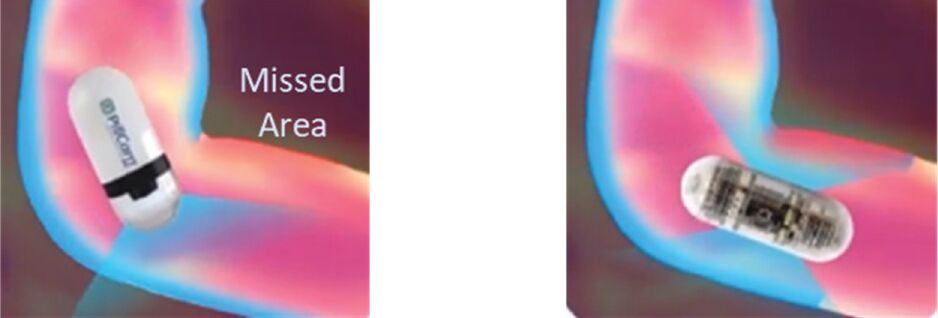
| Competitor’s “tunnel view” can miss inner curvature |
CapsoCam’s direct mucosal view sees inner curvature |
87
Table of Contents
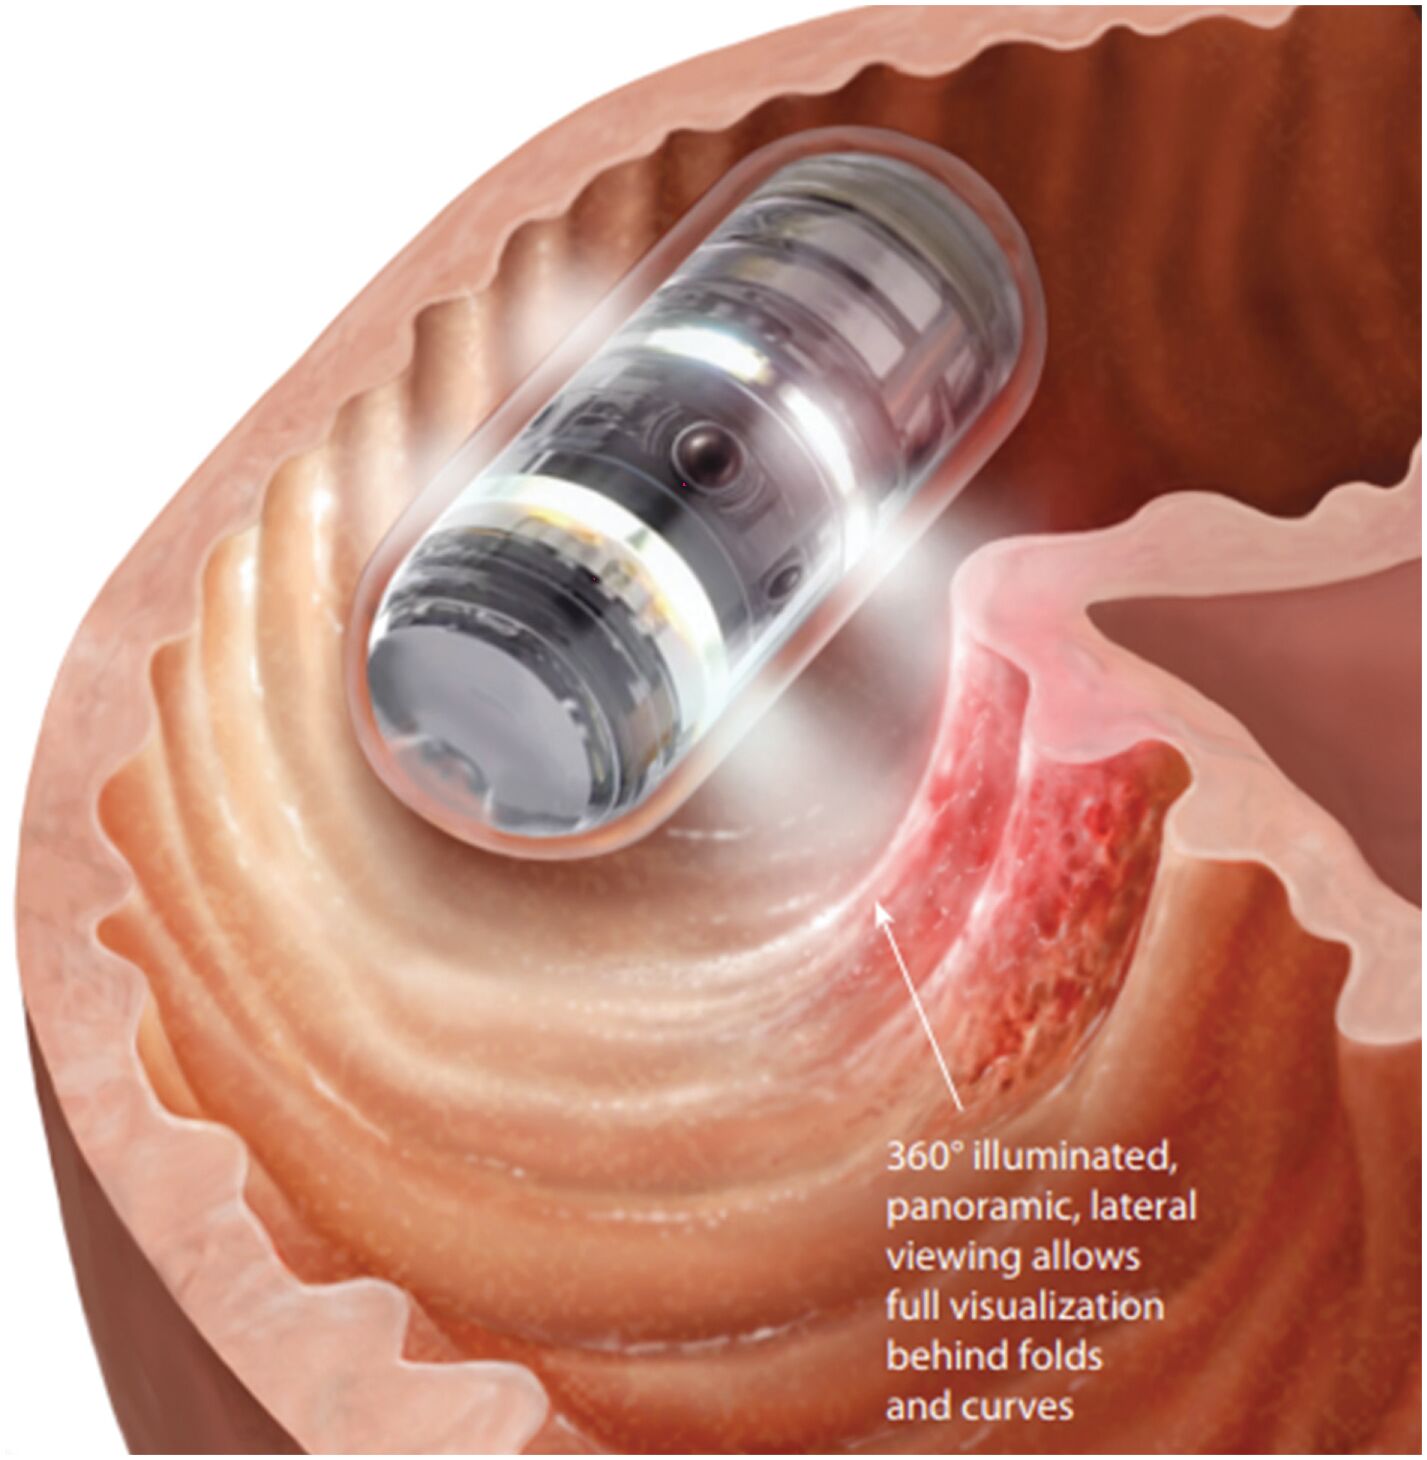
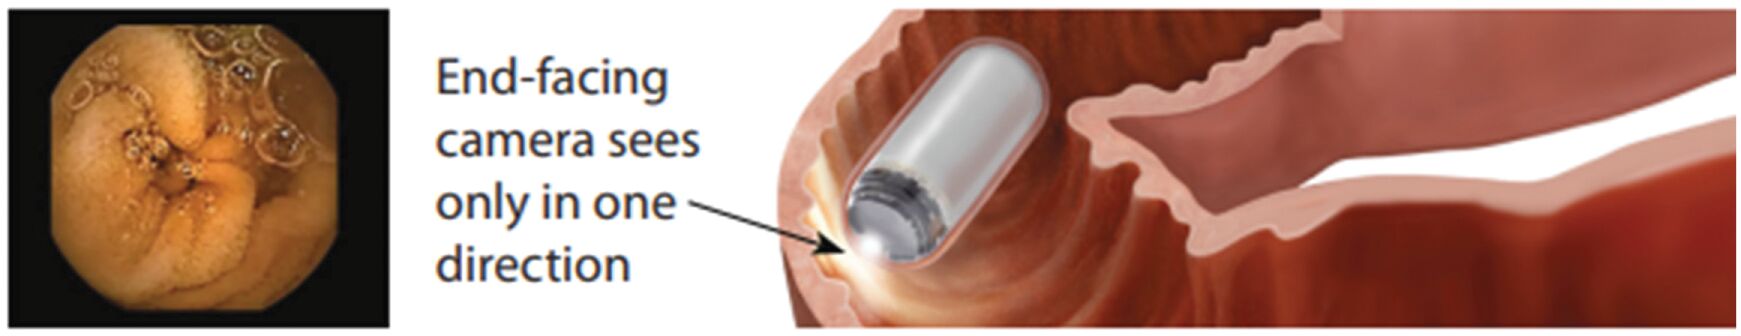
88
Table of Contents
CapsoCam captures a complete 360° view of the GI mucosa:
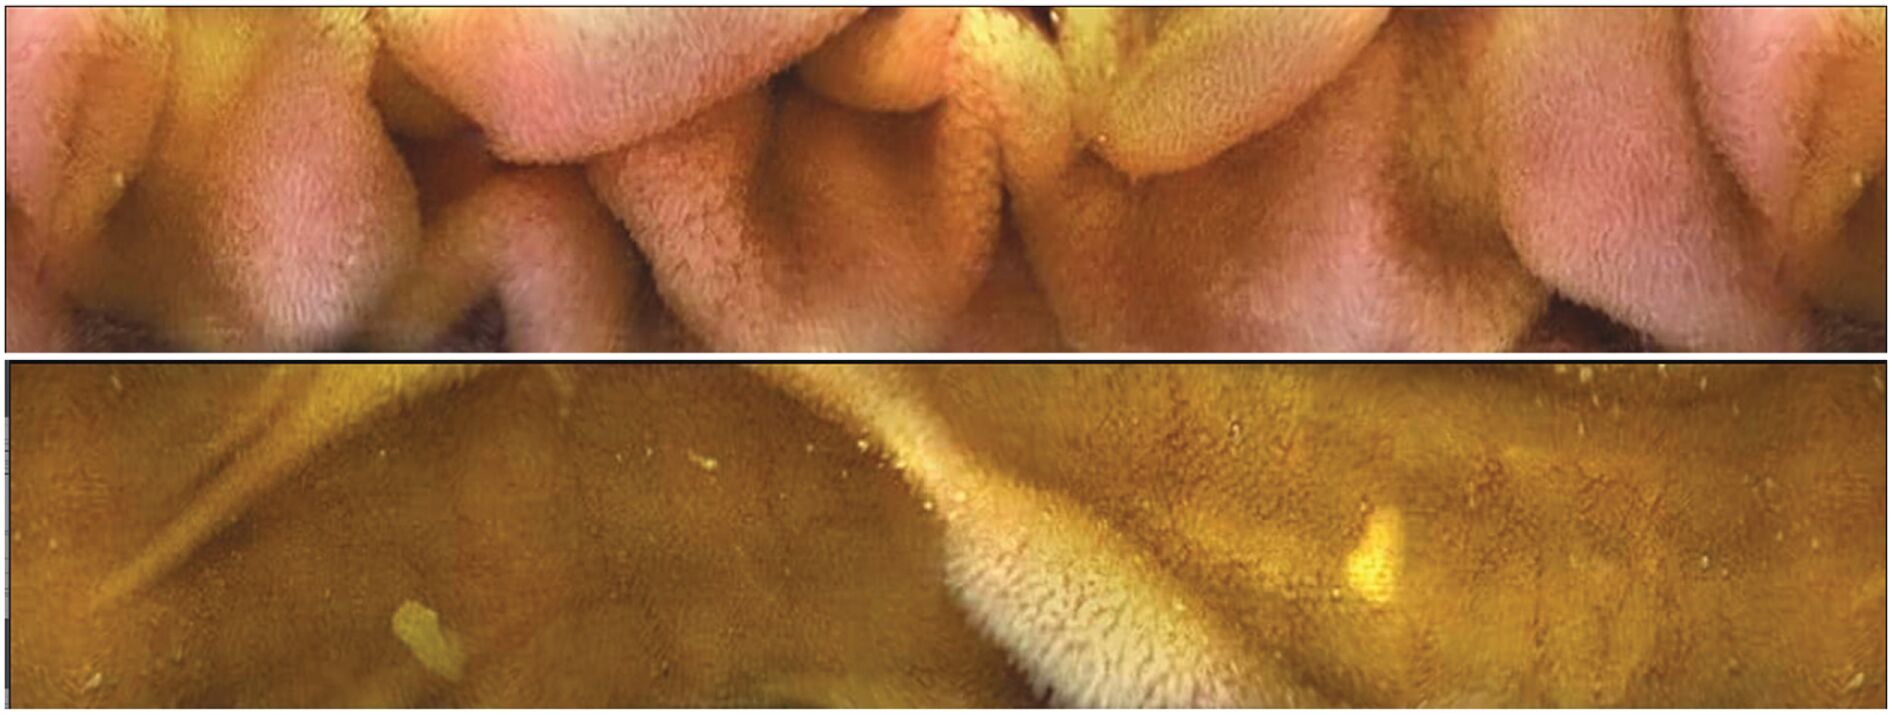
Competitive capsules have a “tunnel view” in straight sections of the small bowel:
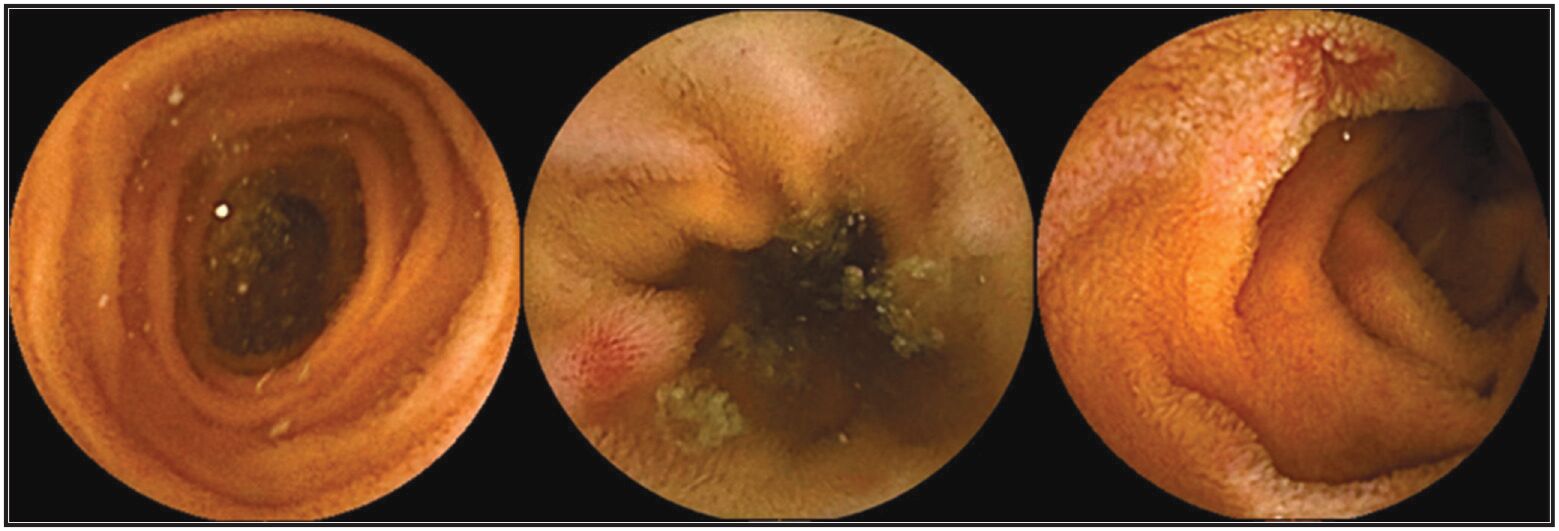
Competitive capsules have a partial wall view in curved sections of the small bowel:
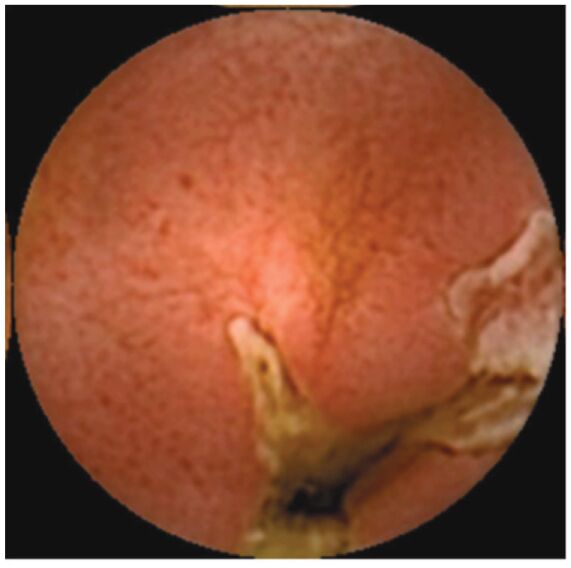
| • | Telemedicine-enabled and zero-capex “wire-free” data collection. Our CapsoCam is a zero-capex solution for providers as it stores the entire video in onboard memory. In addition, patients are able to ingest our capsule in the comfort of their own homes, under the remote supervision of providers. The CapsoCam solution frees up exam-room schedules for providers and provides flexibility to administer capsules any day at any time. A provider’s practice can easily scale to multiple capsules per day with no added cost. In contrast, our competitors’ data recorder comprises a belt that must be worn close to |
89
Table of Contents
| the skin around the patient’s abdomen, which is connected by wires to a recorder module. We believe this system is an uncomfortable and inconvenient encumbrance for patients. In addition, this system is a burden for providers as it is a piece of capital equipment with upfront cost. The equipment must be recovered from the patient, cleaned and batteries recharged, before it is ready to be fitted for the next patient. The data must also be downloaded from the recorder to a computer, which takes additional time. |
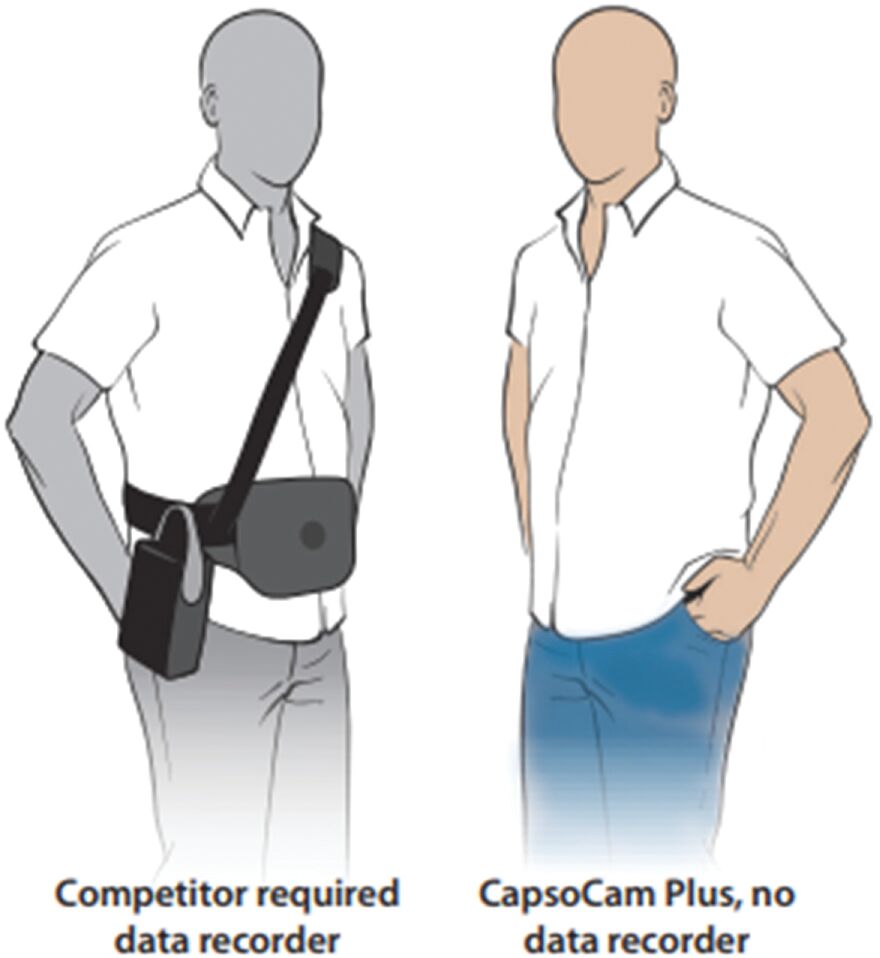
| • | Zero-capex remote data analysis via cloud-based platform. Our cloud-based platform, CapsoCloud, provides a flexible, trackable, streamlined, and capital-equipment-free workflow for providers in the U.S. By giving providers the ability following capsule retrieval to remotely access data obtained from the capsule endoscopy, CapsoCloud allows clinicians to track procedures and stream in vivo videos anywhere at their convenience, generate reports, store and manage patient data, and transfer data to third-party reading services. In contrast, our competitors’ data recorder equipment must be returned to the provider after the procedure. |
Key Components of CapsoCam Plus
The principal component of the CapsoCam Plus capsule endoscopy system is a single-use ingestible capsule that acquires and stores video images in onboard memory while moving through the GI tract, propelled by natural peristalsis. The patient or an authorized caregiver retrieves the capsule after it is egested using the retrieval kit and may return it directly to the clinic, or, in the U.S., mail it in a pre-paid envelop to a CapsoVision-affiliated download center. Thereupon, the capsule is cleaned and disinfected and the procedure data is downloaded using the CapsoAccess system.
90
Table of Contents
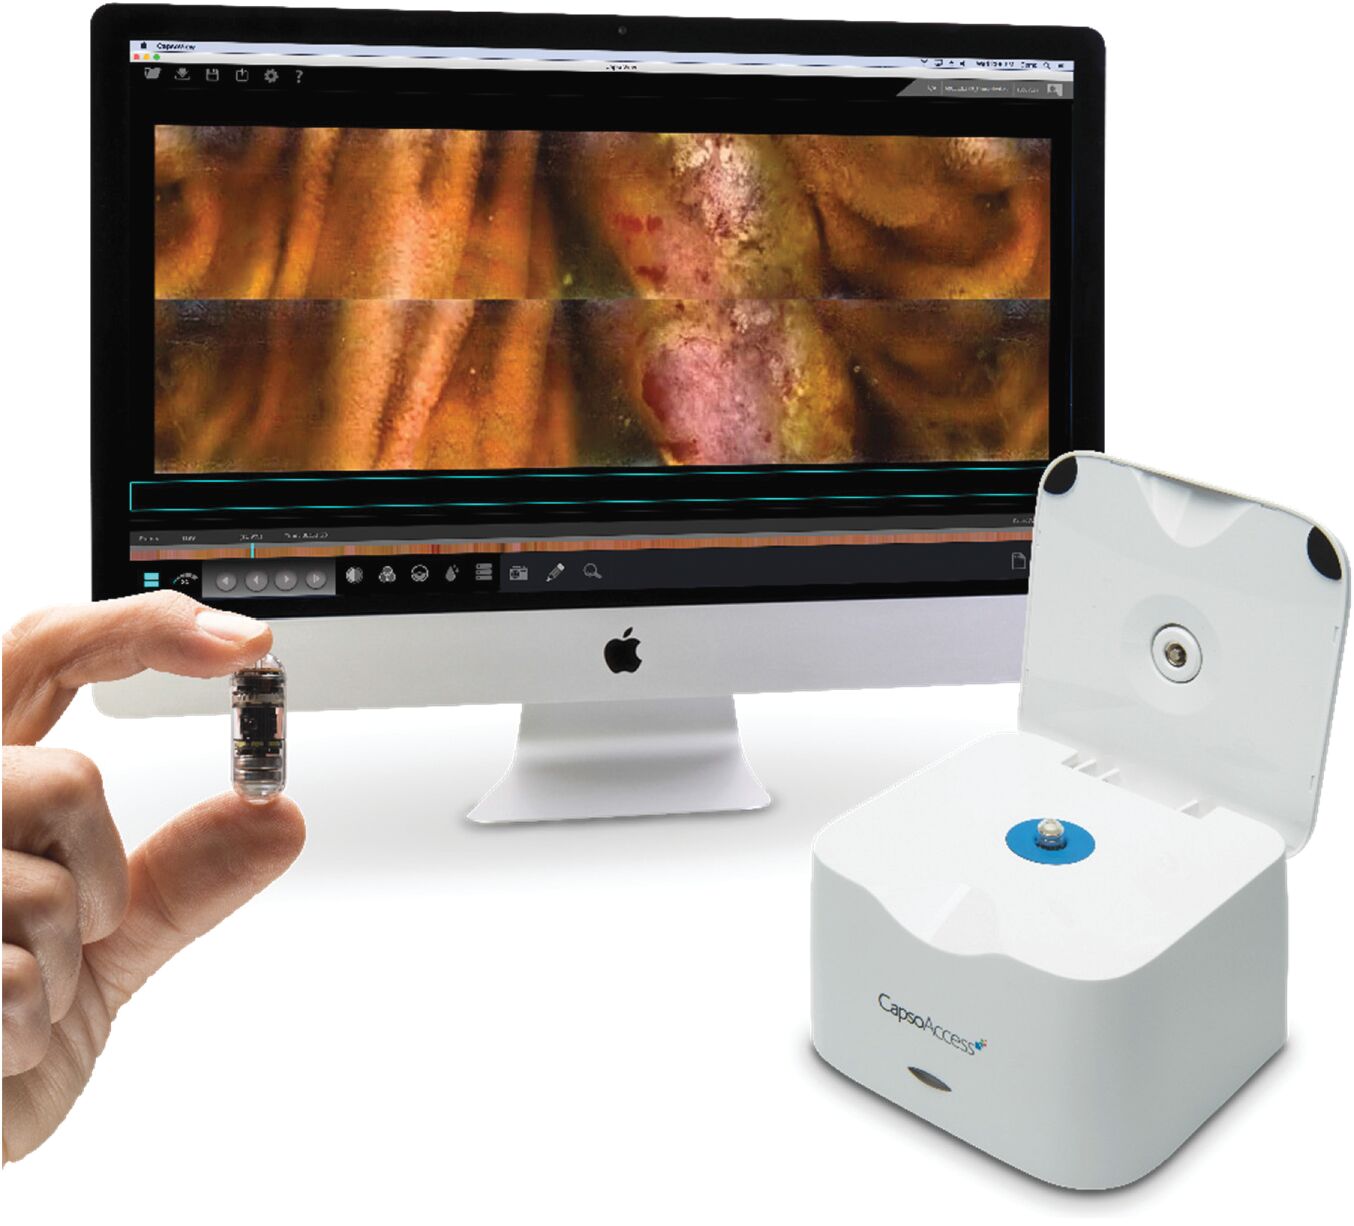
The components of a CapsoCam Plus capsule include the capsule housing, a lens module, the image sensor, the multichip package, an illumination system, a battery pack, a switch, the optical transceiver and a printed circuit board assembly.
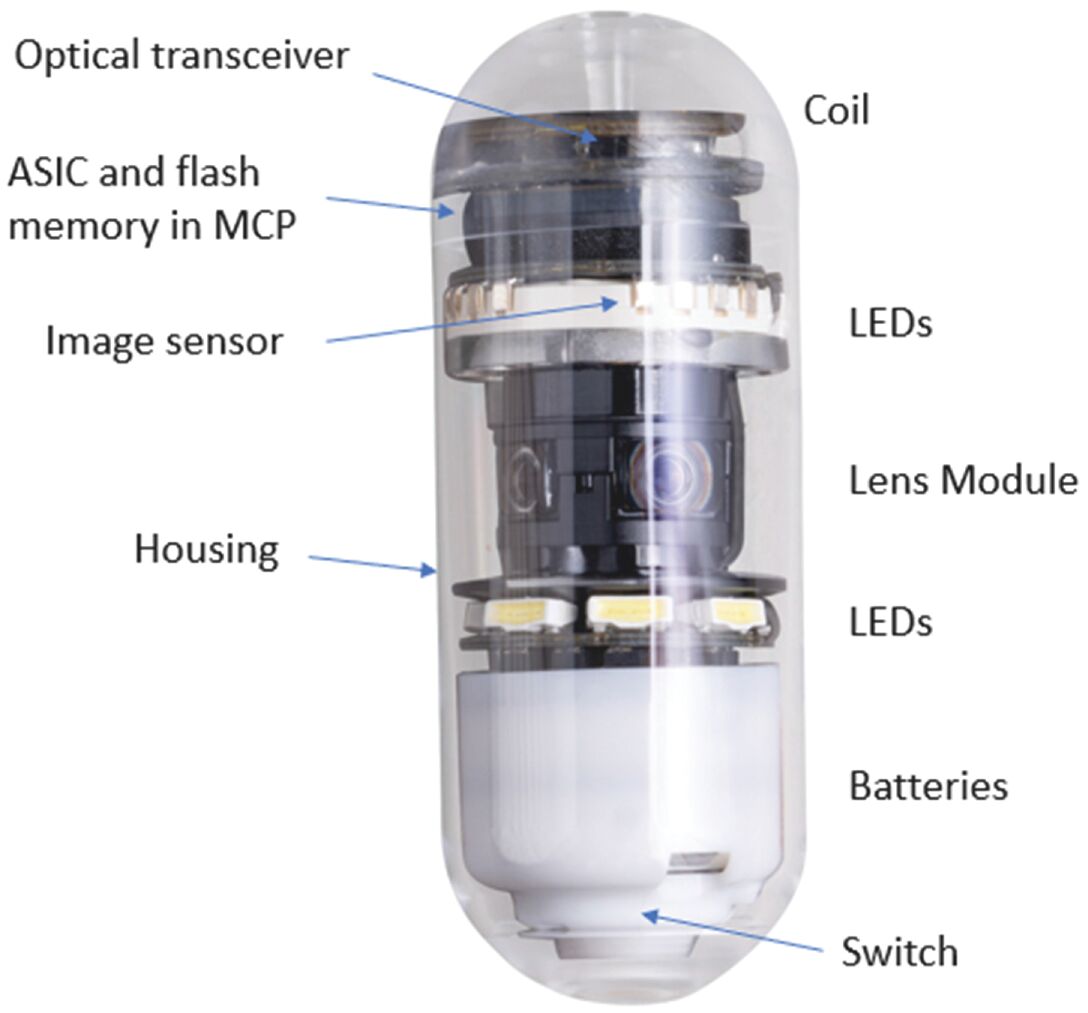
91
Table of Contents
In particular, our CapsoCam Plus capsule consists of a diagnostic imaging system in a capsule housing. The four cameras within the image sensor share a single custom CMOS image sensor, and, as the capsule passes through the GI tract, they together capture a full 360° panorama of the GI tract and the high-resolution color images and other data are stored in the onboard flash memory. The multichip package contains 1.5 GB memory to store the procedure data and an application-specific integrated circuit for controlling the capsule and performing image processing functions, including image compression.
Specifications:
| Length | 30.5 mm | |
| Diameter | 11.3 mm | |
| Mass | 3.7 grams | |
| Camera combined field of view | 74° x 360° | |
| Depth of field | 0 to 18 mm | |
| Video Format | JPEG color images | |
| Video resolution | 1152 x 212 pixels | |
| Frame rate | Up to 20 fps, 3-5 fps per camera | |
| Operating time (typical) | 15-20 hours |
The CapsoCam Plus procedure comprises the following steps:
| i. | Preparation. To ensure that the upper GI tract is free of food, the patient abstains from eating the morning of the procedure and may begin fasting at some point the prior day, depending on the physician’s instructions. A light dose of osmotic laxative is recommended to further clean the small bowel. |
| ii. | Ingestion. The patient ingests the capsule with a glass of water under the clinician’s supervision. Unlike competing capsule endoscopy systems, the patient need not be fitted with a data recorder and the in-office procedure consists only of the capsule ingestion and instructing the patient on the retrieval process. In the U.S., the patient may swallow the capsule at home or another remote location with telemedicine supervision by the clinician. Removing the capsule from its package activates it. Once the patient has successfully swallowed the capsule, they are free to go about their day, and after an additional four hours of fasting, they may eat a light meal. |
| iii. | Capsule Recovery. Patients place a rubber pan on the toilet to collect the capsule after excretion. When the patient next has a bowel movement, typically the next morning, they use the retrieval kit, CapsoRetrieve, to recover the capsule using a magnetic wand and package it for return to the clinic or to a data download center. The capsule is typically excreted within 3 to 30 hours after swallowing. |
| iv. | Data Access. The Capsule Data Access system, CapsoAccess, is used to recover the procedure video from the capsule after it has been cleaned and disinfected, which may be done at the clinic if the patient returns the capsule there. Alternatively, the patient may use a pre-paid envelope to mail the capsule to an authorized download center where the data is accessed and transmitted via the Internet to the clinician. |
| v. | Video Review. The physician reviews the procedure video and generates a report, using either CapsoView software or by streaming the video from CapsoCloud. CapsoCloud, a cloud-based software application, provides the physician the flexibility to review the procedure on any computer or Apple iPad connected to the Internet, at their convenience. CapsoCloud also stores the videos and reports, allows management and tracking of the procedure, and facilitates video review by third-party services or multiple physician colleagues. |
92
Table of Contents
CapsoRetrieve major elements—retrieval pan, magnetic wand, and capsule container:
CapsoAccess accesses procedure data from the CapsoCam Plus capsule after its retrieval:
CapsoCam Colon
We have developed our next pipeline product, CapsoCam Colon, for visualization of the colon and detection and measurement of polyps. We are currently concluding a pivotal study to support 510(k) clearance of CapsoCam Colon for use by currently indicated patients who are a subset of the colorectal cancer screening and surveillance populations. The pivotal study involved approximately 1,200 patients enrolled at 20 sites throughout the U.S. Based on the clinical results from the pivotal study, we are planning to submit our 510(k) application, with two existing capsule endoscopies as predicate devices, to the FDA in the first half of 2025 with a goal of obtaining 510(k) clearance in late 2025. To enhance the sensitivity and specificity of CapsoCam Colon for detecting and measuring polyps, we are developing our second generation CapsoCam Colon capsule and will extend our pivotal study to include a second arm. Our second generation CapsoCam Colon capsule will include improvements such as a new lens and illumination optics with an increased field of view and improved image quality. The second arm of the pivotal study is expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S. We plan to use the clinical results of the second arm to support submission to the FDA of a 510(k) application in late 2025 with a goal of obtaining 510(k) clearance in the first half of 2026. As a part of this second 510(k) application and to increase the population of indicated patients, we intend to seek FDA clearance of expanded indications that remove the requirement for evidence of GI bleeding of lower GI origin for patients with major risks for colonoscopy or moderate sedation.
93
Table of Contents
Key Benefits of CapsoCam Colon
We believe that our CapsoCam Colon, once FDA cleared, will be a superior capsule endoscopy system compared to competitor systems, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. The benefits delivered by the CapsoCam Colon include the following:
| • | Sole capsule endoscope with a 360° panoramic view. Our CapsoCam is the only colon capsule endoscope that captures a full 360° panoramic video, providing a complete view of the colon mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature. Our CapsoCam houses four high-resolution cameras around the capsule circumference, and the images from each are stitched into a single panoramic image. Our competitors’ end-view systems have a camera at each end of the capsule, which affords a limited “tunnel view” and polyps may be hidden by pockets (haustra) between folds. Our CapsoCam, by contrast, has a panoramic view of the colon and can visualize polyps on folds or within the pockets (haustra) between folds. In addition, the inflatable balloon which extends the CapsoCam Colon capsule helps to stabilize its movement through the intestines, resulting in a smoother video. |
CapsoCam captures a complete 360° view of the colon mucosa:
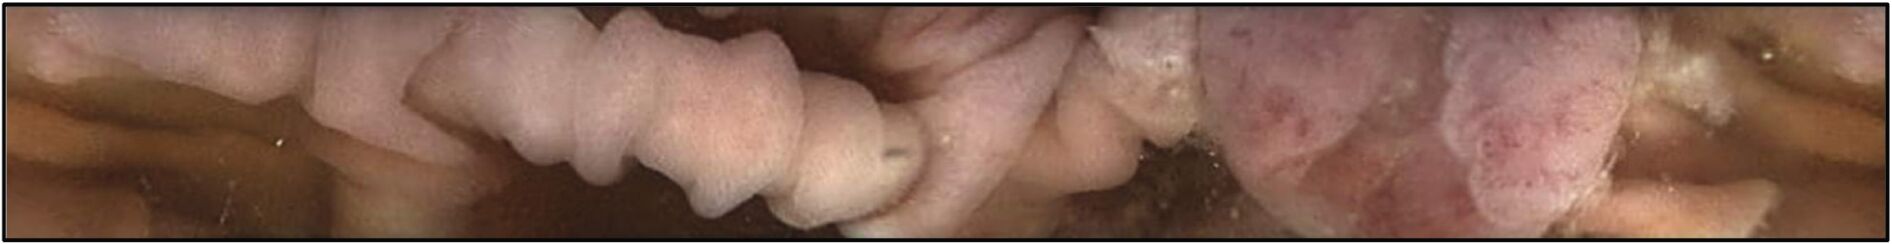
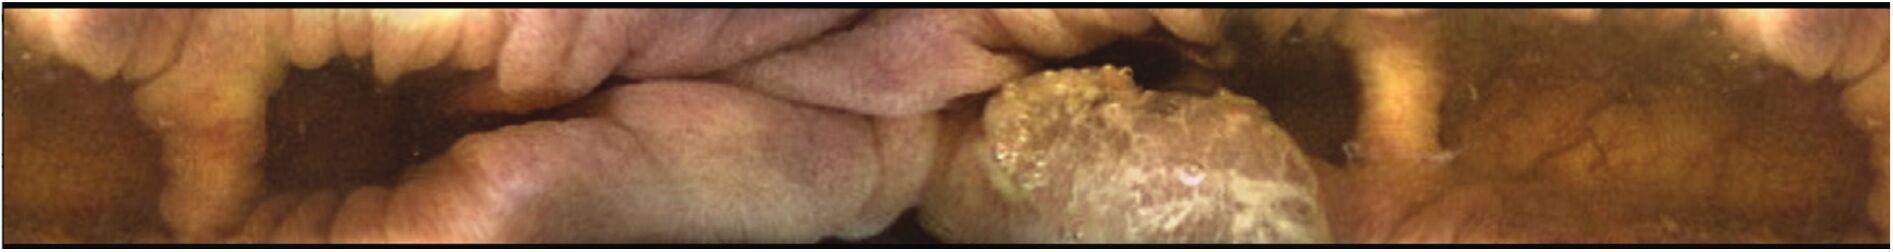
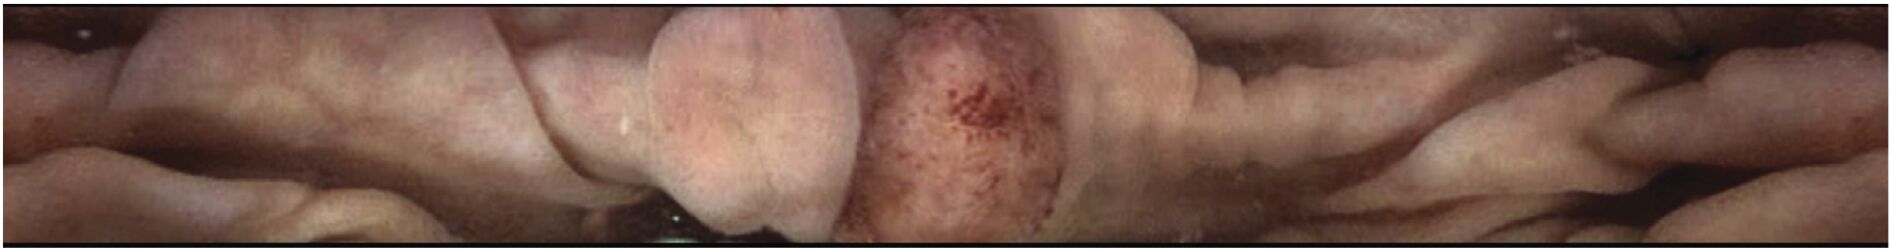
94
Table of Contents
Competitive capsules have a “tunnel view” of the colon:
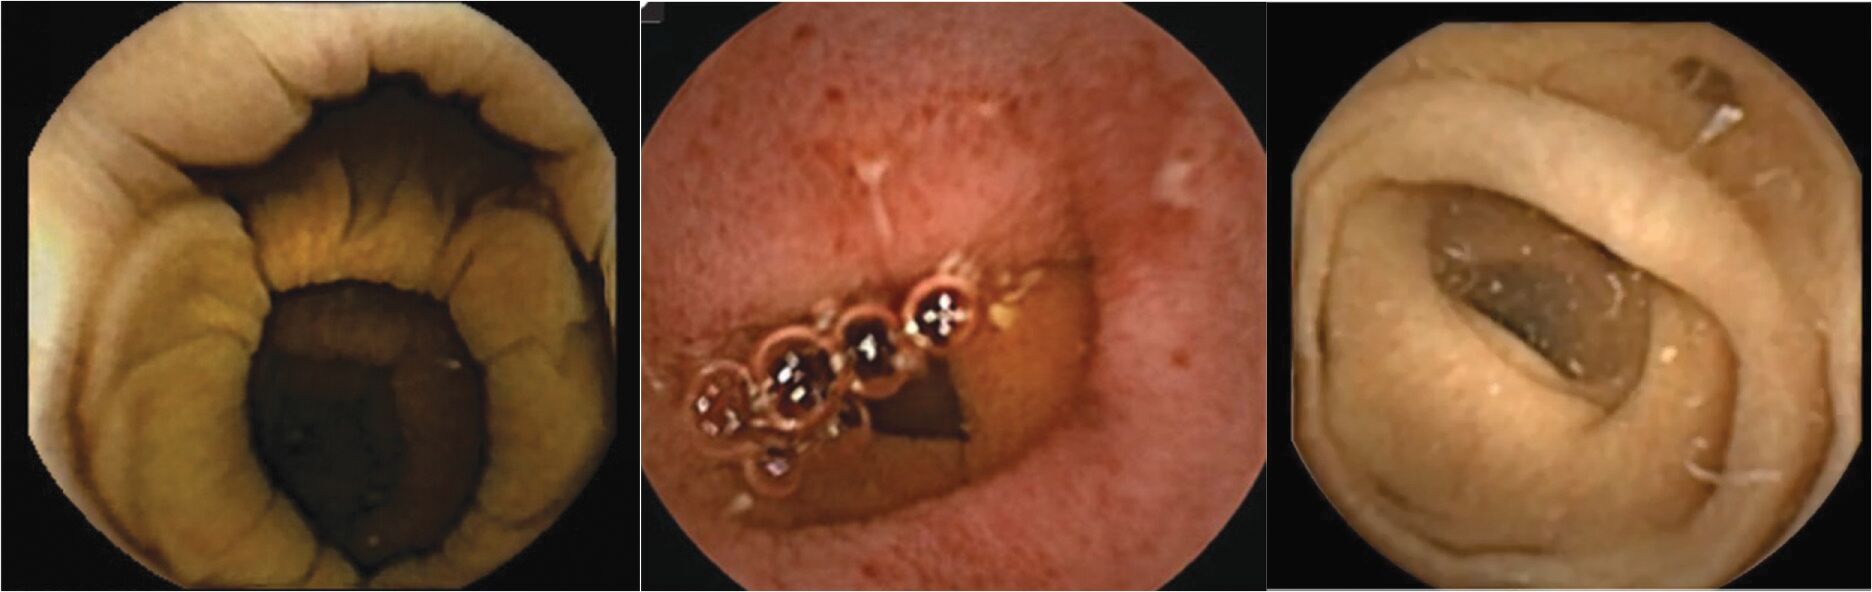
| • | Automated pathology detection due to usage of AI. Our CapsoCam Colon is the only colon capsule endoscopy system (subject to FDA clearance) that utilizes AI for automated polyp detection. Instead of reviewing the entire procedure video, clinicians can focus on those frames with suspected lesions, highlighted with a bonding box, reducing the time taken to review the video and making colon capsule endoscopy financially attractive to their practice. AI improves diagnostic yield and provides more consistent accuracy with reduced dependency on the experience level of the physician video reader and their level of fatigue and distraction. Without AI, physicians can overlook polyps captured in the video. In addition, CapsoCloud automatically accumulates patient data critical to develop and train improved AI-based lesion-detection and classification capabilities. We have an in-house AI development team of experts who will leverage archived patient data on CapsoCloud as it grows during commercial use of CapsoCam Colon to make ongoing improvements to the AI algorithm. In contrast, with competitor capsule endoscopy systems, the clinic downloads patient data from the data recorder and the manufacturer does not have ready access to that data. |
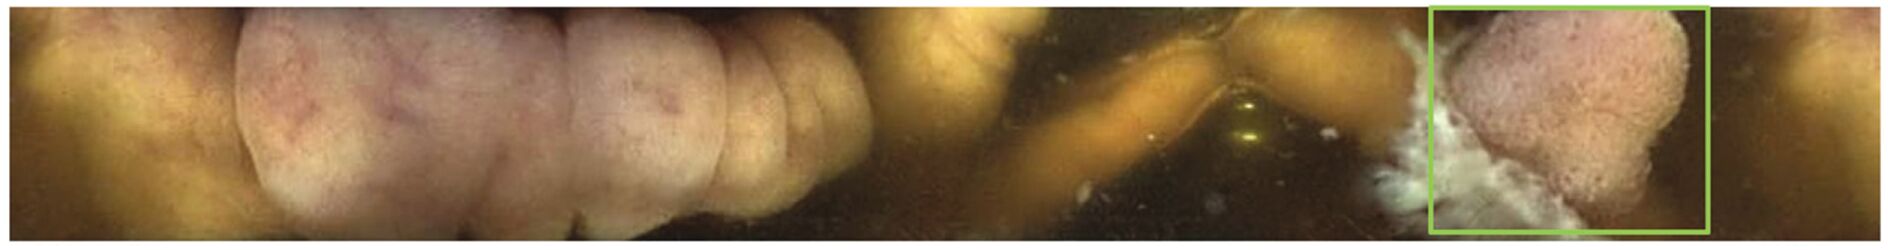
| • | 3D-sensing technology informs follow-on care decisions. Our CapsoCam Colon (subject to FDA clearance) also incorporates our proprietary 3D-sensing technology to more accurately measure polyp sizes. Polyp size is highly correlated with its risk of becoming cancer. No other capsule endoscope currently in the market has 3D-enabled measurement capability. With automated pathology detection and the ability to manually review video frames adjacent to an identified polyp, physicians can more confidently decide that patients with small (e.g., less than 6mm) polyps may forgo a follow-on colonoscopy, increasing the utility of the procedure for healthcare providers and patients alike. |
| • | Telemedicine-enabled and zero-capex “wire-free” data collection. Our CapsoCam is a zero-capex solution for providers as it stores the entire video in onboard memory. In contrast, our competitors’ data recorder comprises a belt that must be worn close to the skin around the patient’s abdomen, which is connected by wires to a recorder module. We believe this system is an uncomfortable and inconvenient encumbrance for patients. In addition, this system is a burden for providers as it is a piece of capital equipment with upfront cost. The equipment must be recovered from the patient, cleaned and batteries recharged, before it is ready to be fitted for the next patient. The data must also be downloaded from the recorder to a computer, which takes additional time. With no data recorder, our CapsoCam frees up exam-room schedules for providers and provides flexibility to administer capsules any day at any time. A provider’s practice can easily scale to multiple capsules per day with no added cost. |
95
Table of Contents
| • | Zero-capex remote data analysis via cloud-based platform. Our cloud-based platform, CapsoCloud, provides a flexible, trackable, streamlined, and capital-equipment-free workflow for providers in the U.S. In contrast, our competitors’ data recorder equipment must be returned to the provider after the procedure. Following the commercial launch of our CapsoCam Colon solution, providers will initially utilize CapsoCloud to download in vivo videos for remote review. Within one year of commercial launch, we plan to introduce user-friendly streaming functions to facilitate via CapsoCloud remote in vivo video review, procedure report generation and image annotation. |
| • | Panenteric capsule. CapsoCam Colon is a panenteric capsule which, when programmed with slightly different operating parameters, can be used to visualize both the small bowel and the colon in one procedure for the evaluation of Crohn’s disease, ulcerative colitis, irritable bowel syndrome and obscure GI bleeding. |
Key Components of CapsoCam Colon
The principal component of the CapsoCam Colon capsule endoscopy system is a single-use ingestible capsule that acquires and stores video images in onboard memory while moving through the GI tract, propelled by natural peristalsis. The patient or an authorized caregiver retrieves the capsule after it is egested using the retrieval kit and may return it directly to the clinic, or, in the U.S., mail it in a pre-paid envelop to a CapsoVision-affiliated download center. Thereupon, the capsule is cleaned and disinfected and the procedure data is downloaded using the CapsoAccess system.
The components of a CapsoCam Colon capsule include the capsule housing, a balloon, a shell, a lens module, the image sensor, the multichip package, an illumination system, a structured-light projector, a battery pack, a switch, the optical transceiver and a printed circuit board assembly.
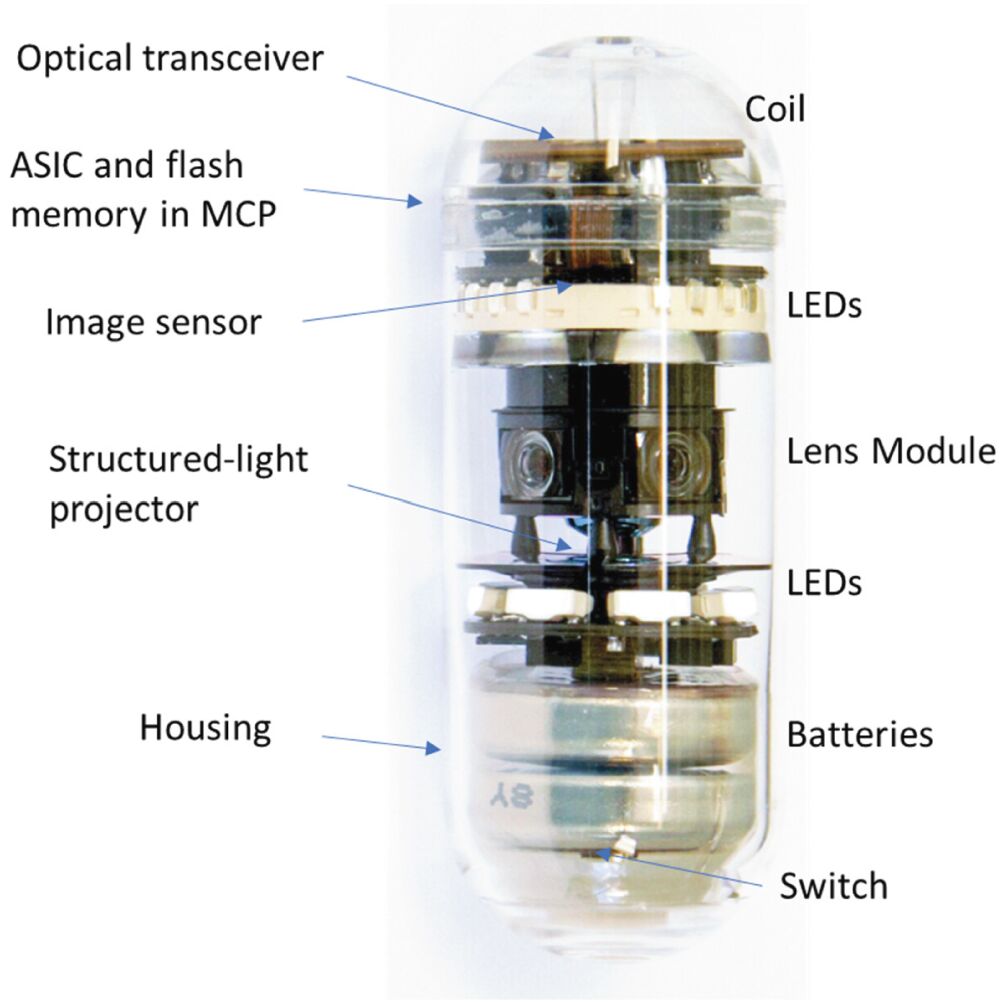
|
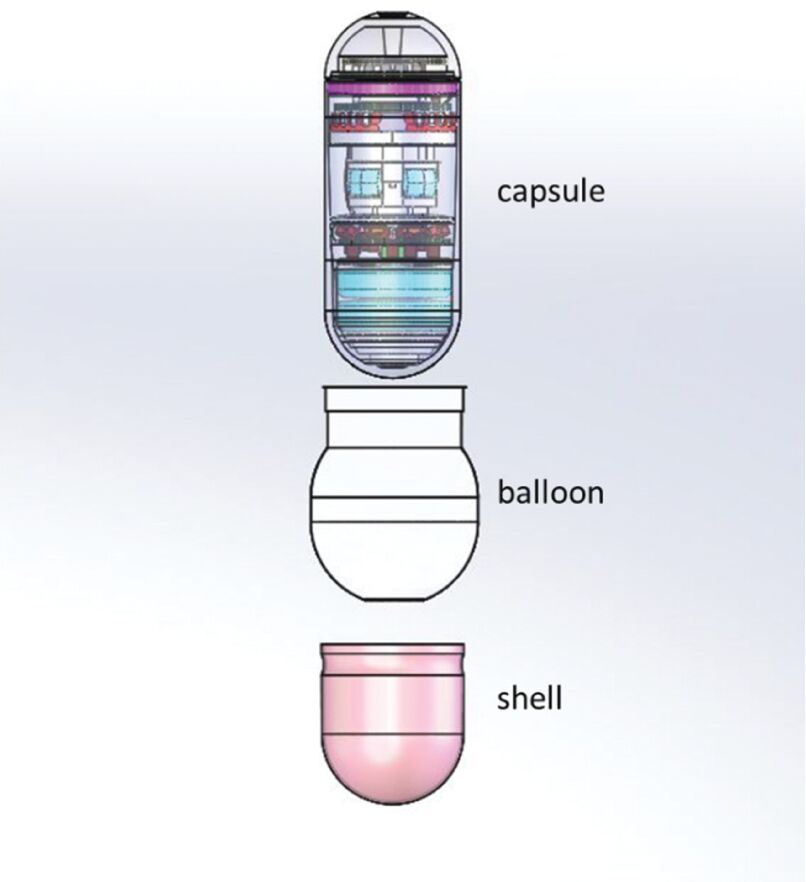
|
In particular, our CapsoCam Colon capsule consists of a diagnostic imaging system in a capsule housing with an attached inflatable balloon, where the inflatable balloon is protected by an outer dissolvable shell covering only the balloon end of the camera capsule. Once inside a liquid environment, the shell softens and dissolves. When the balloon is fully inflated, typically after about 2-3 hours, the capsule is slightly buoyant, enabling it to move through the intestines more readily with flowing liquid, increasing the likelihood of a complete colon visualization.
96
Table of Contents
Specifications:
| Length (at ingestion / at max inflation) | 32.6 mm / 38 mm | |
| Diameter (at ingestion / at max inflation) | 11.8 mm / 12.4 mm | |
| Mass | 3.8 grams | |
| Camera combined field of view | 74° x 360° | |
| Depth of field | 0 to 18 mm | |
| Video Format | JPEG color images | |
| Video resolution | 1152 x 144 pixels | |
| Frame rate | Up to 48 fps, 12 fps per camera | |
| Operating time (typical) | 24 hours |
The CapsoCam Colon procedure comprises the following steps:
| i. | Preparation. The colon is cleansed with a preparation similar to that needed for a colonoscopy. The exact protocol is determined by the patient’s physician, but the recommended process comprises a low-residue diet for several days, clear liquids the day before and on the day of ingestion, and 4 liters of PEG osmotic laxative solution split between the night before and the morning of ingestion. Shortly before ingesting the capsule, the patient may be administered a prokinetic to speed the capsule’s transit through the stomach and small bowel. |
| ii. | Ingestion. The patient ingests the capsule with a glass of water under the clinician’s supervision. Unlike competing capsule endoscopy systems, the patient need not be fitted with a data recorder and the in-office procedure consists only of the capsule ingestion and instructing the patient on the retrieval process. Where authorized, the patient may swallow the capsule at home or another remote location with telemedicine supervision by the clinician. Removing the capsule from its package activates it. The patient continues a clear-liquid diet until the capsule is egested or the following morning, whichever comes first. The patient drinks low-volume osmotic laxative boosters every few hours for a total of 2-4 doses, stopping if the capsule is egested. If the capsule has not yet been egested, a bisacodyl suppository in the evening and upon arising the next day is recommended. |
| iii. | Capsule Recovery. Patients place a rubber pan on the toilet to collect the capsule after excretion. When the patient next has a bowel movement, they use the retrieval kit, CapsoRetrieve, to recover the capsule using a magnetic wand and package it for return to the clinic or to a data download center. The capsule is typically excreted within 1 to 24 hours after swallowing. |
| iv. | Data Access. The Capsule Data Access system, CapsoAccess, is used to recover the procedure video from the capsule after it has been cleaned and disinfected, which may be done at the clinic if the patient returns the capsule there. Alternatively, the patient may use a pre-paid envelope to mail the capsule to an authorized download center where the data is accessed and transmitted via the Internet to the clinician. |
| v. | Video Review. The physician reviews the procedure video and generates a report, using either CapsoView software or, when and where available, by streaming the video from CapsoCloud. Following the commercial launch of our CapsoCam Colon solution, providers will initially utilize CapsoCloud to download in vivo videos for remote review. Within one year of commercial launch, we plan to introduce user-friendly streaming functions to facilitate via CapsoCloud remote in vivo video review, procedure report generation and image annotation. |
Clinical Development
CapsoCam Colon
As of the date of this prospectus, we are actively performing data cleanup for the first arm of our pivotal study for CapsoCam Colon, in preparation for our 510(k) submission. To enhance the sensitivity and specificity
97
Table of Contents
of CapsoCam Colon for detecting and measuring polyps, we are developing our second generation CapsoCam Colon capsule and will extend our pivotal study to include a second arm. Our second generation CapsoCam Colon capsule will include improvements such as a new lens and illumination optics with an increased field of view and improved image quality. The second arm of the pivotal study is expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S. Previously, we completed our pilot study in 2021.
As of the date of this prospectus, the FDA has only authorized one colon capsule imaging system, the Medtronic PillCam COLON 2, introduced in 2014, which does not include any AI-driven analytics; and one AI-driven capsule endoscopy analysis software, the AnX Robotics NaviCam ProScan, introduced in 2023, which exclusively supports AnX Robotics’ small bowel capsule system. These devices will respectively serve as predicate devices for our CapsoCam Colon 510(k) submission. In particular, to receive initial FDA clearance for our first generation CapsoCam Colon, the clinical results must demonstrate that:
| • | CapsoCam Colon, considered without the integration of AI, demonstrates a polyp-detection accuracy that is, at a minimum, comparable to the performance of the predicate device (i.e., Medtronic PillCam COLON 2); and |
| • | our AI technology can (i) reliably and accurately identify and analyze images and video of the colon to detect abnormalities as quantified by diagnostic accuracy measures such as sensitivity and specificity, and, in doing so, (ii) aid qualified physicians in achieving improved diagnostic performance relative to not using AI. |
Pivotal Study
First Arm. As of the date of this prospectus, we are actively performing data cleanup for the first arm of our pivotal study in preparation for our 510(k) submission seeking FDA clearance of our initial, first generation CapsoCam Colon capsule endoscope.
This study is a prospective, open label, pivotal study of the accuracy of CapsoCam Colon in detecting colonic polyps, using colonoscopy as the reference. Enrollment began in 2021 and was completed in October 2024. Approximately 1,200 patients were enrolled at 20 sites throughout the U.S. The patient population were males and females between 45-75 years of age referred for colonoscopy.
The purpose of this pivotal study is to evaluate the safety and effectiveness of our CapsoCam Colon capsule endoscope system in the visualization of the colon and in the detection and size measurement of colonic polyps and to show that AI-based computer assisted detection improves the polyp-detection accuracy and efficiency of capsule video readers.
The coprimary endpoints are (a) sensitivity of the CapsoCam Colon for detecting the presence in a patient of the largest polyp detected by optical colonoscopy if that polyp is ≥ 6mm, where a match is considered to have occurred if a polyp detected by the CapsoCam Colon is assessed as having a size within plus or minus 50% of the size of the polyp detected by optical colonoscopy and as having a location within the same or an adjacent colon segment and (b) specificity of the CapsoCam Colon for not detecting any polyp ≥ 6mm in a patient for whom optical colonoscopy did not detect any polyp ≥ 6mm.
The secondary endpoints include, among others, the sensitivity and specificity of non-computer assisted detection reading versus computer assisted detection reading for polyps ≥ 6mm and for polyps with histopathology-defined advanced neoplasia of the same size and ≥ 10mm.
Second Arm. The second arm of the pivotal study (modeled after the first arm of the pivotal study) will evaluate the safety and effectiveness of our second-generation CapsoCam Colon capsule in a similar patient population expected to involve approximately 200 patients enrolled at up to 10 sites in the U.S.
98
Table of Contents
Pilot Study
Previously, we completed our pilot study for our CapsoCam Colon in 2021. That study was a prospective, open label, pilot study of the CapsoCam Colon capsule endoscope compared to optical colonoscopy. Enrollment began in January 2020 and was completed in February 2021. A total of 112 patients ingested the capsule at 5 sites in the U.S. and results from 105 patients were included in the analysis. The patient population were males and females between 50-75 years of age who had been referred for colonoscopy and had an increased risk for having colon polyps.
This was a non-significant risk pilot study designed to evaluate the safety and performance of the CapsoCam Colon capsule endoscope in patients who met the eligibility criteria and were scheduled for colonoscopy.
Primary Efficacy Assessments: Sensitivity/Specificity
The coprimary endpoints were (a) sensitivity of the CapsoCam Colon for detecting the presence in a patient of the largest polyp detected by optical colonoscopy if that polyp was ≥ 6mm, where a match was considered to have occurred if a polyp detected by the CapsoCam Colon was assessed as having a size within plus or minus 50% of the size of the polyp detected by optical colonoscopy and as having a location within the same or an adjacent colon segment and (b) specificity of the CapsoCam Colon for not detecting any polyp ≥ 6mm in a patient for whom optical colonoscopy did not detect any polyp ≥ 6mm.
The overall sensitivity/specificity for polyps ≥ 6mm for 105 subjects were 84.8% and 92.7% respectively with a prevalence of 21.9%. Overall sensitivity/specificity for polyps ≥ 10mm were 87.5% and 99.0% respectively with a prevalence of 7.6%. The 95% confidence interval for sensitivity for polyps ≥ 6mm was 71.1-98.5%.
Safety Analysis
Safety of the CapsoCam Colon was confirmed with no reports of device-related adverse events, serious adverse events or unanticipated adverse device effects. All events and observations reported in the pilot study were not related to the study or related to medications for bowel preparation or to the prokinetic boosters. There were no serious adverse events reported for bowel preparation or booster consumption.
CapsoCam Plus
In the first half of 2025, we plan to seek FDA 510(k) clearance for the use of AI in our small bowel capsule, CapsoCam Plus. The incorporation of AI technology into our GI-tract capsule endoscopy solution (including the associated software, CapsoCloud and CapsoView) requires appropriate FDA regulatory authorization, supported by requisite clinical and other studies, prior to its commercialization in the U.S. Under the FDA’s current regulatory framework, our capsule endoscopy solution and the AI technology are assessed separately, with each component falling under a different regulatory classification and subject to distinct regulatory requirements. This bifurcated approach, set by precedent, ensures that each component meets the necessary safety, effectiveness, and performance expectations.
In seeking 510(k) clearance for our AI technology as incorporated into CapsoCam Plus we are (i) utilizing the NaviCam ProScan as the predicate device (confirmed with the FDA) and (ii) currently conducting a retrospective clinical study of the CapsoCam Plus solution with the AI technology incorporated to analyze in vivo videos from completed, real-world clinical cases, to assess the performance of the AI technology for small bowel. Similar to the first arm of our CapsoCam Colon pivotal study, the study seeks to demonstrate that the AI technology can (i) reliably and accurately identify and analyze images and video of the small bowel to detect abnormalities as quantified by diagnostic accuracy measures such as sensitivity and specificity, and, in doing so, (ii) aid qualified physicians in achieving improved diagnostic performance relative to not using AI.
99
Table of Contents
Sales, Marketing and Distribution
We primarily generate revenues from the sales of our CapsoCam Plus capsule to our customers, including gastroenterologists practicing in clinics and/or hospitals, which typically place their orders on a monthly or bimonthly basis, as well as our distributors. We engage our customers by using data mining platforms to identify potential customers, initiating their comprehensive evaluation of our CapsoCam capsule and solutions, followed by our strategic sales efforts that ensure a high retention rate. Our customer retention rate stands at approximately 90% for 2024, showcasing our commitment to customer satisfaction and loyalty.
We operate with two sales groups and conduct our marketing strategies differently: one for the U.S. market and one for the international market, each tailored to effectively serve different geographical markets. Our U.S. market sales group, primarily using a direct sales model, focuses on the U.S. domestic market. Our international market sales group is comprised primarily of qualified distributors responsible for selling to customers outside the U.S. Using distributors internationally allows us to extend our global reach and effectively penetrate diverse markets. We are confident that our skilled sales force can be effectively leveraged to promote and distribute our CapsoCam Plus capsules and future CapsoCam capsules (including, once FDA cleared, our CapsoCam Colon capsules).
Our revenues in the year ended December 31, 2023 and 2024 totaled approximately $9.8 million and $ million, respectively, representing a year-over-year growth of approximately % from 2023 to 2024. In 2023 and 2024, international sales accounted for 26% and % of total revenue. Our average accounts receivable balance for 2024 was $ million with less than 5% of our accounts receivable aged more than 60 days at year-end. We do not have a significant customer credit concentration with no single customer accounting for more than 10% of our accounts receivable as of December 31, 2024.
U.S. Market
We currently market our CapsoCam Plus capsule endoscopy solution in the U.S. through a combination of our in-house sales team, integrated GPOs, and to a limited extent, select independent sales representatives. We intend to expand our direct sales force by increasing the number of sales employees and independent sales representatives and contractors. This strategic growth should enhance our market reach and drive higher sales performance. As of December 31, 2024, we had 26 full-time sales employees, including 4 sales directors and 20 territory managers, and 3 independent sales representatives covering all of the states in the U.S. We offer comprehensive training programs to our sales employees, sales representatives and contractors, aimed at enhancing sales performance and ensuring strict compliance with FDA regulations and any other applicable regulations.
Since the launch of our CapsoCam Plus capsule in 2017, we have endeavored to market our CapsoCam Plus capsule endoscopy solution effectively by leveraging the network outreach of independent sales representatives, who managed to bring our products to the attention of GI physicians across the U.S. Meanwhile, we also invested in building our in-house sales team by establishing the role of territory managers for overseeing and coordinating our sales effort in a specific state or states. We believe our hybrid marketing and sales team will enable our CapsoCam capsules to further penetrate into the healthcare market.
In-house Sales Team
We have built a dedicated team of sales directors and territory managers to promote our CapsoCam capsules. Based on our current sales output and strategic planning, we have put in place four regional sales directors, each overseeing a distinct region within the U.S. Our regional sales directors are responsible for supervising the work of territory managers within their region, providing them with technical support and training, as well as setting overall sales strategies for the region. Our regional sales directors are industry veterans who possess rich experience in medical device sales and distribution matters.
100
Table of Contents
Our territory managers are our front-line sales persons who interact with physicians, clinics, hospitals and other healthcare providers on a daily basis. Utilizing the data-mining platforms we have procured, each of our territory managers generates a list of potential customers, and reaches out to introduce them to our CapsoCam capsule. After this introduction, subject to their protocols and practices, the potential customers will generally make an initial purchase of 5 to 10 capsules for evaluation, before placing subsequent orders. We provide constant assistance and monitoring throughout the customer evaluation process to answer questions, identify potential issues and obtain relevant feedback to a potential customer’s purchase decision. Following the evaluation process, we seek to onboard the customer, with customers typically ordering capsules, subject to their needs, on a monthly or bi-monthly basis and we strive to meet their expected timeline of delivery.
Our territory managers provide continued service and support to our customers, including periodic education of physicians, nurses and other healthcare providers about our capsule, feedback collection to understand and attend to any issues encountered in the course of using our capsule and CapsoCloud, and tracking any adverse events. As a result, we were able to secure a high customer retention rate of approximately 90% for 2024.
Independent Sales Representatives
We engage independent sales representatives to complement our in-house sales team. We pay our independent sales representatives commissions and bonuses, if eligible, based on their sales results. We discourage our independent sales representatives from concurrently representing our competitors in promoting similar products.
Marketing Methods
We also use various sales and marketing methods to generate awareness and demand for our CapsoCam Plus capsule, such as traditional marketing through direct sales, channel partners and trade shows as well as digital marketing through websites, social media, videos and organic and paid searches.
Sales Arrangement with GPOs
We have entered into agreements with several GPOs, which are organizations that leverage the collective purchasing power of their members, such as hospitals, clinics, and other healthcare providers, to negotiate favorable pricing and terms for medical products and services. Our arrangement with GPOs can significantly enhance our ability to meet the stringent requirements set by hospitals, clinics, and other healthcare providers. By leveraging the collective purchasing power and compliance expertise of GPOs, we can ensure adherence to industry standards and regulations more effectively. Under these agreements, the GPOs’ members may elect to purchase our CapsoCam capsules from us on a direct sale basis. During the term of these agreements, which typically ranges from two to three years and are subject to renewal, we are generally prohibited from entering into new contracts outside of these agreements with the GPOs’ members for our CapsoCam capsules. The agreements also contain firm pricing provisions, which may limit our ability to adjust our prices in response to market conditions or competitive pressures. However, we may offer bulk purchase options or special promotions to the GPOs’ members, subject to certain conditions and approvals. We also agree to provide the GPOs’ members with pricing, terms, and conditions that are equal or better than those offered to our other customers. In addition, we agree to negotiate in good faith to equitably adjust the pricing for any current products under the agreements if it is affected by any new technology. We pay administrative fees to the GPOs based on a percentage of sales for products their members purchase from us.
International Market
We initially began commercial sales in Europe in 2012 for our GI-tract capsule endoscopy solution (i.e., our small bowel capsule). Since then, we have expanded into other non-U.S. countries or regions, and our
101
Table of Contents
international markets now include Germany, Sweden and the U.K. Our distributors typically have exclusivity in their respective country or territory. As of December 31, 2024, we had approximately 50 exclusive distributors covering multiple non-U.S. countries or regions, including Europe and Africa. We also have one exclusive distributor for sales of our CapsoCam capsule for the veterinary market worldwide. Although we have historically utilized exclusive distributors to access various countries or regions, we may choose to implement a direct sales model. For example, we (i) in 2023, we established a direct sales team in Germany to better serve our customers and strengthen our market presence in this key market and (ii) are transitioning to a direct sales model in some pivotal countries, including the G7 countries, to enhance market penetration and drive growth.
For both our direct sales and distributor sales, we are responsible for obtaining and maintaining the regulatory permits for our current CapsoCam Plus capsule, future CapsoCam capsules and associated software, CapsoCloud and CapsoView, in the relevant countries and regions.
Exclusive Distribution Agreements
We typically enter into exclusive distribution agreements with our international distributors, which typically specify the purchase price, payment terms, minimum purchase obligations, territory, term, renewal, termination, and other rights and obligations of the parties. The purchase price is usually agreed in the distribution agreement and may be revised through mutual agreement, taking into account the prevailing market prices. Such revision usually occurs no more than once per year. The payment terms vary depending on the distributor’s creditworthiness and performance, but generally range from 30 to 60 days from the date of invoice. The minimum purchase obligations are based on the projected sales volume and growth potential of the territory. The territory is defined as the country or region where the distributor has the exclusive right to sell and market our CapsoCam capsule endoscopy solution. The term of the distribution agreement is initially one to five years, and may be renewed for additional one to three years upon agreement or automatically, unless terminated earlier by either party for cause or convenience. Our exclusive distributors usually have the right of first offer for any new products that we develop.
We are responsible for the manufacturing, supply, packaging, and labeling of our CapsoCam capsules and for obtaining and maintaining the regulatory permit to manufacture and market our capsule endoscopy solution in the U.S. and other countries or regions where we export our products. We are also required to report any death, serious injury, or malfunction incidents involving our CapsoCam product to the relevant authorities and to conduct any necessary product recalls or corrective actions. Our distributors agree to supply us with any information and assistance related to such incidents and to comply with our instructions and policies regarding product safety and quality. Our distributors also agree to forward any customer and/or regulatory complaints or correspondence regarding our CapsoCam capsule endoscopy solution to us promptly and to cooperate with us in resolving any such issues.
We are prohibited from selling our CapsoCam capsules or engaging another distributor to sell our CapsoCam capsules in the territory of our exclusive distributors, unless we terminate the distribution agreement with the existing distributor. Our distributors are prohibited from advertising, marketing, or selling our CapsoCam capsules outside their territory or soliciting orders from persons or entities located outside their territory.
We may terminate our distribution agreements with our exclusive distributors for cause, such as breach of contract, failure to meet minimum purchase obligations, insolvency, or misconduct, or for convenience, subject to a prior notice period and a break-up fee based on the anticipated margin, if applicable. We also agree to honor our obligations to the distributor’s key customers (including governmental bodies) for a period of at least one year following the termination of the distribution agreement, unless we assign such obligations to a successor distributor.
102
Table of Contents
Coverage and Reimbursement for CapsoCam
In the U.S., we derive substantially all of our revenue from gastroenterologists practicing in clinics and/or hospitals that use our CapsoCam Plus capsule endoscopy solution. These providers, in turn, bill third-party payers, including private insurers, Medicare, and Medicaid, for the services and items they provide to patients. The CapsoCam Plus enables our customers to operate under the existing reimbursement structure for optical capsule endoscopy of the small bowel, which has well-established reimbursement levels via CPT codes that varies by state. Subject to patient and provider compliance with guidelines around recommended procedures, government and commercial payers generally provide coverage for optical capsule endoscopy of the small bowel under this framework.
Similarly, our CapsoCam Colon, once FDA cleared, is expected to be covered under the existing reimbursement structure for colon capsule endoscopy for our initially indicated patient group, which has reimbursement levels via CPT codes.
Research and Development
We invest in research and development initiatives that are focused on introducing enhancements and improvements aimed at increasing the value provided by our GI-tract capsule endoscopy solution. Our solution is currently comprised of our CapsoCam Plus capsule for the small bowel, our planned CapsoCam Colon (subject to FDA clearance), CapsoCloud, our cloud-based platform, and our CapsoView software. In doing so, we hope to realize our vision of creating an ingestible capsule that, in a single convenient non-invasive procedure, cost-effectively screens and/or identifies multiple pathologies in a broad patient population. In particular, for CRC, we hope to screen for both cancerous and precancerous polyps and lesions, recognizing that early detection can assist in the prevention and treatment of CRC. Our research and development team includes hardware and software engineers with deep expertise in medical technology, optics, data science, AI, and cloud-based data and security architecture and individuals with extensive clinical development expertise.
Initial and recent research and development efforts focused on developing our first capsule endoscopy, the small bowel capsule (510(k) clearance received in 2016) and improved versions thereof (the current generation of which we refer to as CapsoCam Plus), development of our AI assisted reading technology and improving on CapsoCam Plus to develop CapsoCam Colon (with 510(k) clearance targeted for late 2025). CapsoCam Colon incorporates our self-developed AI technology and other multiple self-developed proprietary technologies designed to assist in effectively visualizing the colon and detecting and measuring polyps. In addition to having AI for automated pathology detection of polyps, it incorporates a 360° panoramic lateral view and 3D-sensing technology to more accurately measure polyp sizes. We are researching and developing improvements to our CapsoCam Plus capsule to incorporate our AI assisted reading technology. We are working on improvements to our CapsoCam, including a new lens and illumination optics with an increased field of view, improved image quality and higher peak frame rate. For our AI assisted reading technology, we plan to incorporate improvements designed to deliver increased sensitivity for identifying polyps and pre-cancerous lesions, reduced viewing times, enhanced usability of the polyp size measurement tool, improved camera dynamic range for a clearer view of dark areas and improvements to increase completion rates. We also plan to continue making improvements to our CapsoCloud and CapsoView software.
Certain of our research and development efforts may also include conducting and overseeing clinical studies and obtaining FDA clearance, which may or may not be granted.
We incurred approximately $9.3 million in research and development expenses for the year ended December 31, 2023. For additional information, see “Management’s Discussion and Analysis of Financial Condition and Results of Operations—Components of our Results of Operations and Balance Sheet—Operating Expenses.”
103
Table of Contents
Manufacturing and Supply
We manage all aspects of manufacturing, supply chain and distribution of our GI-tract capsule endoscopy products from our headquarters in Saratoga, California. We manufacture our products using component suppliers and assembly manufacturers based in Asia. The critical components of our products, such as lens modules, CMOS image sensors, and ASICs are provided by single-source suppliers. In particular, our lens modules are sourced from Largan, a single supplier based in Taiwan who we have an over 16-year relationship with, our CMOS image sensors are sourced from Toshiba, a single supplier based in Japan who we have an over 10-year relationship with, and our ASICs are sourced from Moai Electronics Corporation, a single supplier based in Taiwan who we have an over 10-year relationship with. As a result of our long-standing relationships with these suppliers, we are able to source those component parts on favorable terms and within reasonable lead-times.
In particular, we have entered into a memorandum of understanding with Largan Precision Company, Ltd. (“Largan”), which is based in Taiwan, in May 2008, for the supply of lens modules for our first generation capsules, pursuant to which we make purchases on a purchase order basis. The terms of the memorandum of understanding were subsequently amended in January 2010 (as amended, the “First Largan MOU”). In addition, we entered into another memorandum of understanding with Largan in October 2022, for the supply of lens modules for our second generation capsules (together with the First Largan MOU, collectively, the “Largan MOUs”). Each Largan MOU was effective upon signing and will continue until terminated in accordance thereunder. Either party may terminate either Largan MOU for certain breaches of the agreement by the other party and that party fails to cure the breach within 45 days after notice of such breach from the terminating party.
We have also entered into a development agreement with Toshiba Corporation (“Toshiba”), which is based in Japan, in August 2013 (the “Toshiba Agreement”), for the supply of our CMOS image sensors, pursuant to which we make purchases on a purchase order basis. Pursuant to the Toshiba Agreement, Toshiba may discontinue the supply of CMOS image sensors to us if we do not place purchase orders totaling more than a specified number of units in any consecutive 12 months. The Toshiba Agreement was effective upon signing and will continue until terminated in accordance thereunder.
We have also entered into a development and manufacturing agreement with Moai Electronics Corporation (“Moai”), which is based in Taiwan, in June 2014, for the supply of our ASICs, pursuant to which we make purchases on a purchase order basis. The terms of the development and manufacturing agreement were subsequently amended in March 2015 (as amended, the “Moai Agreement”). The Moai Agreement was effective upon signing and will continue until terminated in accordance thereunder. We may terminate the Moai Agreement for certain breaches of the agreement by Moai and Moai fails to cure the breach within 30 days after notice of such breach. The Moai Agreement prohibits Moai from entering into or continuing any discussions to develop, make or otherwise commercialize any competing products during the term thereof and for a period ending on the first anniversary of the termination thereof.
Currently, assembled CapsoCam capsules are shipped from Taiwan to our U.S. facility where we complete the manufacturing process before distributing the capsules to our distribution network. Once the assembled products in test form are delivered to us, we are responsible for the testing, cleaning, packaging and labeling of our products at our headquarters in Saratoga, California.
We are also in the process of planning or implementing various mitigation measures to address supply chain risks (including qualifying a backup supplier for certain critical components and looking to build reserve supplies of capsules and critical components to address unanticipated delays).
Competition
We operate in a highly competitive industry. The medical device market, particularly in the area of diagnostic imaging and endoscopic procedures, is characterized by rapid technological advancements and the
104
Table of Contents
continuous introduction of new products. The competition that we face for our current and future GI-tract capsule endoscopy solutions will vary based on numerous factors including: the nature of the pathology(ies) or medical condition(s); the ease of use of our products; the relevant patient population and related indicated patients; available indicated means to diagnose or treat the pathology/condition and related accuracy, reliability and safety; patient considerations (such as adherence, comfort, safety and convenience); provider considerations (such as clinical workflow and convenience); and economic and time considerations (such as cost, availability of third-party reimbursement and provider time required).
CapsoCam Plus (small bowel). The competition that we face for our small bowel CapsoCam Plus is primarily from traditional enteroscopy procedures performed by trained physicians in hospital or clinical settings and other capsule-based imaging solutions manufactured by companies such as Medtronic, IntroMedic, JinShan and Ankon. Those competitors include well-established companies with significant resources and brand recognition such as Medtronic, that are constantly developing and marketing innovative products that may offer superior features or lower costs. With respect to Medtronic, it also enjoys other competitive advantages including (i) a “first mover advantage” as the first manufacturer of a small bowel capsule endoscopy and a colon capsule endoscopy (described below); (ii) exclusive supply arrangements (sometimes up to three years) with some of the larger GI practices and hospitals (particularly in the Northeast region of the U.S.) which our sales team also targets; and (iii) greater brand recognition and financial resources.
The competitor capsule endoscopy systems currently available in the market (i) consist of capsules with end-view systems, providing only limited “tunnel” or partial “wall” views of the small bowel and (ii) utilize wired data recorders worn on the patient’s. As a result, providers are forced to incur upfront capital expenses and clinical workflow complications and patients experience discomfort and multiple clinical visits. We believe our CapsoCam Plus is a superior capsule endoscopy system, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. Our CapsoCam captures a full 360° panoramic video, providing a complete view of the GI mucosa—unobstructed by folds and with complete coverage of the bending intestine’s inner curvature, ultimately resulting in superior diagnostic yield. Also, our CapsoCam is a zero-capex “wire-free” data collection solution for providers as it stores the entire video in onboard memory. Following retrieval, our cloud-based platform, CapsoCloud, gives providers in the U.S. the ability to remotely access data from the cloud and stream in vivo videos anywhere at their convenience. Outside of the U.S., providers review procedure videos using CapsoView software (primarily due to foreign data privacy and access regulations).
CapsoCam Colon. The competition that we face for CapsoCam Colon is primarily from (i) procedure-based detection technologies such as optical colonoscopy, flexible sigmoidoscopy and CTC (or “virtual” colonoscopy); (ii) stool-based DNA tests such as Cologuard (initial FDA clearance in 2014); and (iii) other capsule-based imaging solutions like PillCam COLON 2 (initial FDA clearance in 2014). Other sources of competition include (a) other common CRC screening tests, such as the FOBT and the FIT, and (b) other screening technologies including liquid biopsy tests, such as Epi proColon (510(k) clearance received in 2016) and C-Scan (CE Mark obtained in 2019). Those competitors include well-established companies with significant resources such as Medtronic, and are constantly developing and marketing innovative products that may offer superior features or lower costs.
We believe that our CapsoCam Colon, once FDA cleared, will compare favorably to other available products and services. All other colorectal cancer detection methods in use today are constrained by some combination of poor sensitivity, poor adherence, or high cost. For example, colonoscopy requires advanced dietary restrictions and bowel cleansing, potential time away from work, someone to drive the patient home from the procedure and can also be uncomfortable, time-consuming, hazardous, and expensive. Fecal blood testing, including FIT testing, suffers from poor sensitivity, with only a 74% detection rate for cancer and 24% detection rate for advanced precancerous lesions. Stool tests and blood tests have demonstrated good sensitivity for cancer, but sensitivity for advanced precancerous lesions is under 50%. Relatively high false-positive rates for multi-targeted stool DNA tests, such as Cologuard, lead to unnecessary colonoscopies. In a study with over 10,000 average-risk screening patients, advanced neoplasia was not detected by colonoscopy in 75% of Cologuard-positive cases.
105
Table of Contents
Colon capsule endoscopy provides non-invasive visualization of the entire colon from the cecum to the rectum, and it has demonstrated good sensitivity and specificity for the detection of colon polyps. Currently, the only two competitor products that are available in the market for a limited subset of indicated patients have end-view systems and require wired data recorders to be worn on the body. We believe our CapsoCam Colon, once FDA cleared, will be a superior capsule endoscopy system, both in how it captures and presents images of the GI tract and the clinical workflow and patient experience that it enables. In addition to the competitive advantages described above for our CapsoCam Plus, which also apply to our CapsoCam Colon, CapsoCam Colon utilizes our self-developed AI for automated polyp detection and incorporates our proprietary 3D-sensing technology to more accurately measure the size of polyps in the GI tract.
CapsoCloud. Our cloud-based platform, CapsoCloud, provides a flexible, trackable, streamlined, and capital-equipment-free workflow for providers in the U.S. By giving providers the ability following capsule retrieval to remotely access data obtained from the capsule endoscopy, CapsoCloud (in connection with the CapsoCam Plus) allows clinicians to track procedures and stream in vivo videos anywhere at their convenience, generate reports, store and manage patient data, and transfer data to third-party reading services. Following the commercial launch of our CapsoCam Colon solution, providers will initially utilize CapsoCloud to download in vivo videos for remote review. Within one year of commercial launch, we plan to introduce user-friendly streaming functions to facilitate via CapsoCloud remote in vivo video review, procedure report generation and image annotation.
CapsoView. Outside of the U.S., providers review procedure videos using CapsoView software (primarily due to foreign data privacy and access regulations).
We might not be able to compete successfully in our market, particularly as we seek to obtain the required FDA clearance and seek to commercialize CapsoCam Colon. If our competitors introduce new diagnostic tests that compete with or surpass the accuracy, price or ease of use of our products, we may be unable to satisfy existing customers or attract new customers at the prices and levels that would allow us to generate attractive rates of return on our investment. Increased competition could result in price reductions and revenue shortfalls, loss of customers and loss of market share, which could harm our business, prospects, financial condition and operating results.
Intellectual Property
Intellectual property rights are important to us. We seek to protect our intellectual property and proprietary technologies through combined means, including by pursuing patent applications that cover our technologies and product candidates, as well as any other relevant inventions and improvements that are considered commercially important to the development of our business. We have developed, and are continuing to develop, a comprehensive intellectual property portfolio related to our capsule camera products and their portable usage, including structural and compartmental design, imaging mechanics and algorithms for optimizing the screening and detection performance.
Our success depends in part on our ability to: (a) obtain, maintain, protect and enforce intellectual property and other proprietary rights for our current and future technology, inventions, improvements, and know-how we consider important to our business, (b) preserve the confidentiality of our trade secrets, (c) defend and enforce our intellectual property rights, (d) prevent others from infringing, misappropriating, or violating our intellectual property and other proprietary rights, and (e) operate without infringing, misappropriating, or violating the intellectual property and other proprietary rights of others. Our objective is to seek to protect our proprietary position by, among other methods, pursuing and obtaining patent protection in the U.S. and in jurisdictions outside of the U.S. related to our proprietary technology, inventions, and improvements that are important to the development and implementation of our business. Our patent portfolio is intended to cover components of our products, as well as any other inventions that are important to our business. We also rely on trademarks, trade secrets, and know-how to develop and maintain our proprietary position.
106
Table of Contents
As set forth in the table below, our patent portfolio, as of December 31, 2024, contains over 140 total issued patents worldwide with anticipated expiration dates as indicated.
| Jurisdiction |
Title |
Patent No. |
Status |
Date of Expiration |
||||||||
| US |
In vivo autonomous camera with on-board data storage or digital wireless transmission in regulatory approved band | 8073223 | Issued | 9/19/2026 | ||||||||
| US |
Capture control for in vivo camera | 7940973 | Issued | 9/19/2026 | ||||||||
| US |
Image capture control for in vivo autonomous camera | 7792344 | Issued | 9/19/2026 | ||||||||
| US |
Onboard data storage and method | 7495993 | Issued | 10/25/2026 | ||||||||
| US |
System and method for capsule camera with on-board storage | 8472795 | Issued | 11/22/2026 | ||||||||
| US |
Methods to compensate Manufacturing Variations and Design Imperfections in a Capsule Camera | 9307233 | Issued | 1/9/2027 | ||||||||
| US |
Methods to compensate manufacturing variations and design imperfections in a display device | 11019318 | Issued | 1/9/2027 | ||||||||
| US |
Methods to compensate manufacturing variations and design imperfections in a display device | 10499029 | Issued | 1/20/2027 | ||||||||
| US |
Systems and methods for capsule camera control | 8213698 | Issued | 6/23/2027 | ||||||||
| US |
Panoramic imaging system | 7817354 | Issued | 3/7/2028 | ||||||||
| US |
Lighting control for in vivo capsule camera | 7796870 | Issued | 6/5/2028 | ||||||||
| US |
Detection of when a capsule camera enters into or goes out of a human body and associated operations | 9025017 | Issued | 1/2/2029 | ||||||||
| US |
In vivo camera with multiple sources to illuminate tissue at different distances | 8956281 | Issued | 5/29/2029 | ||||||||
| US |
Retrieval pan | D737,959 | Issued | 9/1/2029 | ||||||||
| US |
In vivo autonomous camera with on-board data storage or digital wireless transmission in regulatory approved band | 7983458 | Issued | 11/24/2029 | ||||||||
| US |
Detection of when a capsule camera enters into or goes out of a human body and associated operations | 8187174 | Issued | 4/10/2030 | ||||||||
| US |
Capture control for in vivo camera | 7974454 | Issued | 5/10/2030 | ||||||||
| US |
Multiple capsule camera apparatus and methods for using the same | 9041785 | Issued | 6/1/2030 | ||||||||
| US |
Methods to compensate manufacturing variations and design imperfections in a capsule camera | 8405711 | Issued | 6/14/2030 | ||||||||
| US |
System and method for multiple viewing-window display of capsule images | 8150124 | Issued | 6/20/2030 | ||||||||
| US |
System and method for image enhancement of dark areas of capsule images | 8150123 | Issued | 9/3/2030 | ||||||||
| US |
Capsule imaging system having a folded optical axis | 9001187 | Issued | 9/25/2030 | ||||||||
| US |
System and method for display of panoramic capsule images | 8724868 | Issued | 10/3/2030 | ||||||||
| US |
Methods to compensate manufacturing variations and design imperfections in a capsule camera | 9007478 | Issued | 10/29/2030 | ||||||||
| US |
Data communication between capsulated camera and its external environments | 9285670 | Issued | 12/7/2030 | ||||||||
| US |
Multi-stream image decoding apparatus and method | 8803961 | Issued | 1/12/2031 | ||||||||
| US |
In vivo image capturing system including capsule enclosing a camera | 8773500 | Issued | 1/18/2031 | ||||||||
| US |
System and method for capsule camera with capture control and motion-compensated video compression | 8165374 | Issued | 6/9/2031 | ||||||||
| US |
Imaging system having a folded optical axis | 8717413 | Issued | 6/10/2031 | ||||||||
| US |
In vivo camera with multiple sources to illuminate tissue at different distances | 8636653 | Issued | 8/3/2031 | ||||||||
107
Table of Contents
| Jurisdiction |
Title |
Patent No. |
Status |
Date of Expiration |
||||||||
| US |
Multiple capsule camera apparatus and methods for using the same | 8300091 | Issued | 8/31/2031 | ||||||||
| US |
Camera system with multiple pixel arrays on a chip | 9118850 | Issued | 11/3/2031 | ||||||||
| US |
In vivo camera with multiple sources to illuminate tissue at different distances | 10244929 | Issued | 2/5/2032 | ||||||||
| US |
Camera System with Mutiple Pixel Arrays On A Chip | 9621825 | Issued | 4/12/2032 | ||||||||
| US |
In vivo camera with multiple sources to illuminate tissue at different distances | 11103129 | Issued | 2/2/2033 | ||||||||
| US |
System and Method for Displaying Annotated Capsule Images | 9626477 | Issued | 4/16/2033 | ||||||||
| US |
Capsule orientation detection for capsule docking system with inductive power drive circuit | 10159400 | Issued | 5/2/2033 | ||||||||
| US |
Optical docking system with inductive powering for capsule camera | 10602913 | Issued | 5/2/2033 | ||||||||
| US |
Docking system with inductive powering for capsule camera | 10869594 | Issued | 5/2/2033 | ||||||||
| US |
Docking system with inductive powering for capsule camera | 10881282 | Issued | 5/2/2033 | ||||||||
| US |
System and method for displaying bookmarked capsule images | 10154226 | Issued | 9/30/2033 | ||||||||
| US |
System and Method for Displaying Annotated Capsule Images | 9304669 | Issued | 3/1/2034 | ||||||||
| US |
Method of overlap-dependent image stitching for images captured using a capsule camera | 9324172 | Issued | 5/19/2034 | ||||||||
| US |
Power source control for medical capsules | 11129516 | Issued | 7/10/2034 | ||||||||
| US |
Image sensor with integrated power conservation control | 9357150 | Issued | 8/12/2034 | ||||||||
| US |
In vivo capsule device with electrodes | 10531786 | Issued | 8/20/2034 | ||||||||
| US |
Reconstruction of images from an in vivo multi-camera capsule | 10068334 | Issued | 5/28/2035 | ||||||||
| US |
Power source control for medical capsules | 10206557 | Issued | 9/13/2035 | ||||||||
| US |
Method and apparatus for endoscope with distance measuring for object scaling | 10402992 | Issued | 10/16/2035 | ||||||||
| US |
Single image sensor for capturing mixed structured-light images and regular images | 10447950 | Issued | 10/16/2035 | ||||||||
| US |
Endoscope employing structured light providing physiological feature size measurement | 10531074 | Issued | 10/16/2035 | ||||||||
| US |
Endoscope with images optimized based on depth map derived from structured light images | 10624533 | Issued | 10/16/2035 | ||||||||
| US |
Single image sensor for capturing mixed structured-light images and regular images | 10742909 | Issued | 10/16/2035 | ||||||||
| US |
Single image sensor for capturing mixed structured-light images and regular images | 10785428 | Issued | 10/16/2035 | ||||||||
| US |
Method and apparatus of sharpening of gastrointestinal images based on depth information | 10943333 | Issued | 10/16/2035 | ||||||||
| US |
Single image sensor for capturing mixed structured-light images and regular images | 11102428 | Issued | 10/16/2035 | ||||||||
| US |
Endoscope employing structured light providing physiological feature size measurement | 11019327 | Issued | 10/16/2035 | ||||||||
| US |
Reconstruction with Object Detection for Images Captured from a Capsule Camera | 9672620 | Issued | 12/18/2035 | ||||||||
| US |
Capsule orientation detection for capsule docking system with inductive power drive circuit | 10010241 | Issued | 12/28/2035 | ||||||||
| US |
Single image sensor control for capturing mixed mode images | 10201266 | Issued | 1/28/2036 | ||||||||
| US |
Reconstruction of images from an in vivo multi-camera capsule with two-stage confidence matching | 11116390 | Issued | 7/12/2037 | ||||||||
108
Table of Contents
| Jurisdiction |
Title |
Patent No. |
Status |
Date of Expiration |
||||||||
| US |
Single Image Sensor for Capturing Mixed Structured-light Images and Regular Images | 9936151 | Issued | 3/9/2036 | ||||||||
| US |
Reconstruction of images from an in vivo multi-camera capsule with two-stage confidence matching | 11120547 | Issued | 3/28/2036 | ||||||||
| US |
Method and apparatus of sharpening of gastrointestinal images based on depth information | 11354783 | Issued | 4/15/2036 | ||||||||
| US |
Single image sensor for capturing mixed structured-light images and regular images | 10484629 | Issued | 7/31/2036 | ||||||||
| US |
Method and apparatus for image stitching of images captured using a capsule camera | 10943342 | Issued | 2/4/2037 | ||||||||
| US |
Capsule device having variable specific gravity | 10098526 | Issued | 2/10/2037 | ||||||||
| US |
Capsule device having variable specific gravity | RE48181 | Issued | 2/10/2037 | ||||||||
| US |
De-ghosting of images captured using a capsule camera | 10015372 | Issued | 3/18/2037 | ||||||||
| US |
Method of image processing and display for images captured by a capsule camera | 11074672 | Issued | 4/19/2037 | ||||||||
| US |
Method and apparatus of lens alignment for capsule | 10638920 | Issued | 6/30/2037 | ||||||||
| US |
Method and apparatus for estimating area or volume of object of interest from gastrointestinal images | 10580157 | Issued | 8/4/2037 | ||||||||
| US |
Method and apparatus for estimating area or volume of object of interest from gastrointestinal images | 10736559 | Issued | 8/4/2037 | ||||||||
| US |
Method and apparatus for area or volume of object of interest from gastrointestinal images | 10346978 | Issued | 8/18/2037 | ||||||||
| US |
Capsule enteric coating for controlling balloon expansion start time | 10674899 | Issued | 7/9/2038 | ||||||||
| US |
Method and apparatus for capturing images and associated 3D model based on a single image sensor and structured-light patterns in the visible spectrum | 10593055 | Issued | 9/12/2038 | ||||||||
| US |
Method and apparatus for travelled distance measuring by a capsule camera in the gastrointestinal tract | 10506921 | Issued | 10/11/2038 | ||||||||
| US |
Method and apparatus for travelled distance measuring by a capsule camera in the gastrointestinal tract | 10835113 | Issued | 10/11/2038 | ||||||||
| US |
Method and apparatus for detecting missed areas during endoscopy | 11219358 | Issued | 3/2/2040 | ||||||||
| US |
Method and apparatus for leveraging residue energy of capsule endoscope | 11612303 | Issued | 9/29/2041 | ||||||||
| US |
Method and apparatus for objective assessment of gastrointestinal conditions based on images captured in the GI tract | 11948303 | Issued | 4/7/2042 | ||||||||
| Jurisdiction |
Title |
Patent No. | Status | |||||||
| Japan |
In Vivo Sensor with Panoramic Camera | JP5,695,142B | Granted | |||||||
| Japan |
In Vivo Sensor with Panoramic Camera | JP5,523,713B | Granted | |||||||
| Japan |
Multi-Stream Image Decoding Apparatus And Method | JP5,368,469B | Granted | |||||||
| Japan |
Method to Compensate Manufactoring Variation and Design Imperfections In A Capsule Camera | JP5,926,345B | Granted | |||||||
| Japan |
Multi-Stream Image Decoding Apparatus And Method | JP5,592,358 | Granted | |||||||
| Japan |
Optical Wireless Docking System for Capsule Camera | JP6,177,315 | Granted | |||||||
| Japan |
Power Source Control For Medical Capsulese | JP6,280,876 | Granted | |||||||
| Japan |
Reconstruction of Images from an in Vivo Multi-Cameras Capsule | JP6,501,800 | Granted | |||||||
109
Table of Contents
| Jurisdiction |
Title |
Patent No. | Status | |||||||
| Japan |
System and Method for Capsule Device with Multiple Phases of Density | JP6,510,591 | Granted | |||||||
| Japan |
Methods to Compensate Manufacturing Variations and Design Imperfections In A Display Device | JP6,737,937 | Granted | |||||||
| Japan |
Methods to Compensate Manufacturing Variations and Design Imperfections In A Display Device | JP6,563,870 | Granted | |||||||
| Japan |
Single Image Sensor for Capturing Mixed Structured-light Images and Regular Images | JP6,803,908 | Granted | |||||||
| Japan |
Single Image Sensor for Capturing Mixed Structured-light Images and Regular Images | JP7,114,666 | Granted | |||||||
| Japan |
Single Image Sensor for Capturing Mixed Structured-light Images and Regular Images | JP7,114,667 | Granted | |||||||
| Japan |
Camera System With Multiple Pixel Arrays On A Chip | JP5,368,469 | Granted | |||||||
| Japan |
Endoscope Employing Structured Light Providing Physiological Feature Size Measurement | JP6,891,345 | Granted | |||||||
| Europe |
Data communication between capsulated camera and its external environments | EP2198342 | Granted | |||||||
| Europe |
In vivo sensor with panoramic camera | EP1974240 | Granted | |||||||
| Europe |
Camera system with multiple pixel arrays on a chip | EP2225877 | Granted | |||||||
| Europe |
Power source control for medical capsules | EP 2816946 | Granted | |||||||
| Europe |
Capsule endoscopic docking system | EP 2858549 | Granted | |||||||
| Europe |
Multi-stream image decoding apparatus and method | EP2291119 | Granted | |||||||
| Europe |
Optical wireless docking system for capsule camera | EP 2849627 | Granted | |||||||
| Europe |
Power source control for medical capsules | EP 3272270 | Granted | |||||||
| Europe |
In vivo camera with multiple sources to illuminate tissue at different distances | EP 2299895 | Granted | |||||||
| Europe |
Capsule device having variable specific gravity | EP 3270761 | Granted | |||||||
| Europe |
Reconstruction of images from an in vivo multi-camera capsule with confidence matching | EP 3148399 | Granted | |||||||
| Europe |
Capsule coating for image capture control | EP 3226745 | Granted | |||||||
| Europe |
Single image sensor for capturing structured-light images and regular images and its method of operation | EP3362989 | Granted | |||||||
| Taiwan |
Reconstruction of images from an in vivo multi-camera capsule | TWI532460 | Granted | |||||||
| Taiwan |
Method and Apparatus for Endoscope with Distance Measuring for Object Scaling | TW-I596931 | Granted | |||||||
| Taiwan |
Capsule Orientation Detection for Capsule Docking System with Inductive Power Drive Circuit | TW-I592129 | Granted | |||||||
| China |
Image Capture Control For In Vivo Camera | ZL201180011653.4 | Granted | |||||||
| China |
Multi-Stream Image Decoding Apparatus And Method | ZL200980129312.X | Granted | |||||||
| China |
In Vivo Camera with Multiple Sources to Illuminate Tissue At different Distances | ZL200980120587.7 | Granted | |||||||
| China |
Data Communication Between Capsulated Camera and Its External Environments | ZL200820106599.X | Granted | |||||||
| China |
In Vivo Autonomous Camera with On-Board Data Storage or Digital Wireless Transmission in Regulatory Approved Band | ZL200680040026.2 | Granted | |||||||
| China |
Camera System With Multiple Pixel Arrays On A Chip | ZL200880125638.0 | Granted | |||||||
| China |
Method to Compensate Manufactoring Variation and Design Imperfections In A Capsule Camera | ZL200780052067.8 | Granted | |||||||
| China |
Method to Compensate Manufactoring Variation and Design Imperfections In A Capsule Camera | ZL201210082410.6 | Granted | |||||||
| China |
Power Source Control for Medical Capsules | ZL201380004147.1 | Granted | |||||||
110
Table of Contents
| Jurisdiction |
Title |
Patent No. | Status | |||||||
| China |
Optical Wireless Docking System for Capsule Camera | ZL201380026186.1 | Granted | |||||||
| China |
In Vivo Camera With Multiple Sources To Illuminate Tissue At Different Distance | ZL201410534382.6 | Granted | |||||||
| China |
In Vivo Capsule Device with Electrodes | ZL201310346793.8 | Granted | |||||||
| China |
Capsule Camera Device With Multi-Spectral Light Sources | ZL201380081896.4 | Granted | |||||||
| China |
In Vivo Sensor with Panoramic Camera | ZL200680050987.1 | Granted | |||||||
| China |
Reconstruction of Images From An In Vivo Multi-Camera Capsule | ZL201480030874.X | Granted | |||||||
| China |
In Vivo Camera With Multiple Sources To Illuminate Tissue At Different Distances | ZL201410532920.8 | Granted | |||||||
| China |
In Vivo Camera With Multiple Sources To Illuminate Tissue At Different Distances | ZL201811307576.7 | Granted | |||||||
| China |
Method of Overlap-Dependent Image Stitching for Images Captured Using a Capsule Camera | ZL201680020175.6 | Granted | |||||||
| China |
Capsule Device Having Variable Specific Gravity | ZL201680016549.7 | Granted | |||||||
| China |
Reconstruction With Object Detection For Images Captured From A Capsule Camera | ZL201680047653.2 | Granted | |||||||
| China |
Single Image Sensor for Capturing Mixed Structured-light Images and Regular Images | ZL201680059418.7 | Granted | |||||||
| China |
Method and Apparatus for Gastric Examination Using a Capsule Camera | ZL201780097479.7 | Granted | |||||||
| China |
Image Sensor with Integrated Power Conservation Control | ZL201580079351.9 | Granted | |||||||
| China |
Method and Apparatus of Lens Alignment for Capsule Camera | ZL201780036954.X | Granted | |||||||
| China |
Method and Apparatus for Area or Volume of Object of Interest From Gastrointestinal Images | ZL201710768987.5 | Granted | |||||||
| China |
De-ghosting of Images Captured Using a Capsule Camera | ZL201711016971.5 | Granted | |||||||
| China |
Method and Apparatus for Image Stitching of Images Captured Using a Capsule Camera | ZL201680091275.8 | Granted | |||||||
| China |
Method and Apparatus for Travelled Distance Measuring by a Capsule Camera in the Gastrointestinal Tract | ZL201910119473.6 | Granted | |||||||
| China |
Endoscope Employing Structured Light Providing Physiological Feature Size Measurement | ZL201880045690.9 | Granted | |||||||
| China |
Method of Image Processing and Display for Images Captured by a Capsule Camera | ZL201780089753.6 | Granted | |||||||
| China |
Method and Apparatus for Capturing Images and Associated 3D Model Based on a Single Image Sensor and Structured-Light Patterns in the Visible Spectrum | ZL201910228496.0 | Granted | |||||||
In addition to patents, we also rely upon trademarks, trade secrets, know-how, and continuing technological innovation to develop and maintain our competitive position. We maintain and are seeking registered trademarks. We have certain know-how and trade secrets relating to our capsule camera and other technologies. We rely on trade secrets to protect certain aspects of our technology. See the section titled “Risk factors—Risks Relating to Our Intellectual Property” for a more comprehensive description of risks related to our intellectual property.
Employees and Human Capital Resources
As of December 31, 2024, we had approximately 90 full time employees. None of our employees are represented by a labor union or party to a collective bargaining agreement. We consider our relationship with our employees to be good.
Our human capital resources objectives include, as applicable, identifying, recruiting, retaining, incentivizing, and integrating our existing and additional employees.
111
Table of Contents
Facilities
Our corporate headquarters is in Saratoga, California, where we lease an approximately 2,000 square foot office facility pursuant to a lease agreement which initially commenced on March 14, 2017 and was modified by follow-on addendums, and will expire on December 31, 2027. We lease two office facilities in Taiwan, where a small group of our employees are stationed, pursuant to two lease agreements which commenced on February 15, 2024 and May 16, 2024, respectively, and will expire on February 14, 2025 and May 15, 2025, respectively.
Our existing facilities will continue to support our research and development, finance, marketing, and administrative teams. We believe that our facilities are adequate to support our expansion through the end of the facilities’ lease periods. We believe that suitable additional or alternative space would be available in the future as required on commercially reasonable terms.
Legal Proceedings
From time to time, we may be involved in various legal proceedings arising from the normal course of business activities. We are not presently a party to any litigation the outcome of which, we believe, if determined adversely to us, would individually or taken together, materially and adversely affect our business, financial condition, or results of operations. Future litigation may be necessary to defend ourselves, our partners, and our customers by determining the scope, enforceability, and validity of third-party proprietary rights, to establish our proprietary rights or for other matters. Involvement in such proceedings is costly and can impose a significant burden on management and employees. The results of any current or future litigation cannot be predicted with certainty, and regardless of the outcome, litigation can have an adverse impact on us because of legal expenses and settlement costs, diversion of management attention, and resources and other factors.
Government Regulation
Our products and operations are subject to extensive regulation by the FDA and other federal and state authorities in the U.S., as well as comparable authorities in foreign jurisdictions. Our product candidates are subject to regulation as medical devices in the U.S. under the Federal Food, Drug, and Cosmetic Act (the “FDCA”), as implemented and enforced by the FDA.
United States Regulation of Medical Devices
The FDA regulates the development, design, non-clinical and clinical research, manufacturing, safety, efficacy, labeling, packaging, storage, installation, servicing, recordkeeping, premarket clearance or approval, adverse event reporting, advertising, promotion, marketing and distribution, and import and export of medical devices to ensure that medical devices distributed domestically are safe and effective for their intended uses and otherwise meet the requirements of the FDCA.
FDA premarket clearance and approval requirements
Unless an exemption applies, each medical device commercially distributed in the U.S. requires either FDA clearance of a 510(k) premarket notification, or approval of a PMA. Under the FDCA, medical devices are classified into one of three classes—Class I, Class II or Class III—depending on the degree of risk associated with each medical device and the extent of manufacturer and regulatory control needed to ensure its safety and effectiveness. Class I includes devices with the lowest risk to the patient for which safety and effectiveness can be assured by adherence to the FDA’s General Controls for medical devices, which include compliance with the applicable portions of the QSR, facility registration and product listing, reporting of adverse medical events, and truthful and non-misleading labeling, advertising, and promotional materials. Class II devices are subject to the FDA’s General Controls, as well as any special controls deemed necessary by the FDA to ensure the safety and effectiveness of the device. These special controls can include performance standards, post-market surveillance, patient registries and FDA guidance documents.
112
Table of Contents
While most Class I devices are exempt from the 510(k) premarket notification requirement, manufacturers of most Class II devices are required to submit to the FDA a premarket notification under Section 510(k) of the FDCA, requesting permission to commercially distribute the device. The FDA’s permission to commercially distribute a device subject to a 510(k) premarket notification is generally known as 510(k) clearance. Devices deemed by the FDA to pose the greatest risks, such as life-sustaining, life-supporting and some implantable devices, devices that have a new intended use, or devices that use advanced technology that is not substantially equivalent to that of a legally marketed device, are placed in Class III, requiring approval of a PMA. Some pre-amendment devices are unclassified, but are subject to FDA’s premarket notification and clearance process in order to be commercially distributed. The product that we currently market, the CapsoCam Plus, is classified as a Class II device and has received FDA marketing authorization through the 510(k) clearance process.
510(k) Clearance marketing pathway
To obtain 510(k) clearance, a manufacturer must submit to the FDA a premarket notification demonstrating that the proposed device is “substantially equivalent” to a predicate device already on the market. A predicate device is a legally marketed device that is not subject to premarket approval, i.e., a device that was legally marketed prior to May 28, 1976 (pre-amendments) and for which a PMA is not required, a device that has been reclassified from Class III to Class II or I, or a device that was found substantially equivalent through the 510(k) process. The FDA’s 510(k) clearance process usually takes from three to twelve months, but may take longer. The FDA may require additional information, including clinical data, to make a determination regarding substantial equivalence. FDA collects user fees for certain medical device submissions and annual fees and for medical device establishments.
If the FDA agrees that the device is substantially equivalent to a predicate device currently on the market, it will grant 510(k) clearance to commercially market the device. If the FDA determines that the device is “not substantially equivalent” to a previously-cleared device, the device is automatically designated as a Class III device. The device sponsor must then fulfill more rigorous PMA requirements. The PMA process requires that the manufacturer demonstrate that the device is safe and effective for its intended uses, which generally requires the submission of extensive data, including results from pre-clinical studies and human clinical trials. A PMA must also contain a full description of the device and its components, the methods, facilities, and controls used for manufacturing, and proposed labeling. The PMA process is burdensome, and in practice, the FDA’s review of a PMA application may take up to several years following initial submission. Alternatively, a manufacturer can request a risk-based classification determination for a novel device in accordance with the “de novo” process, described below. We currently do not market any medical devices pursuant to a PMA.
After a device receives 510(k) clearance or de novo classification, any modification that could significantly affect its safety or effectiveness, or that would constitute a major change or modification in its intended use, will require a new 510(k) clearance or, depending on the modification, PMA approval or de novo classification. The FDA requires each manufacturer to determine whether the proposed change requires submission of a 510(k), de novo request or a PMA in the first instance, but the FDA can review any such decision and disagree with a manufacturer’s determination. If the FDA disagrees with a manufacturer’s determination, the FDA can require the manufacturer to cease marketing and/or request the recall of the modified device until 510(k) marketing clearance or PMA approval is obtained or a de novo request is granted. In these circumstances, the manufacturer may be subject to significant regulatory fines or penalties.
De novo classification process
Medical device types that the FDA has not previously classified as Class I, II, or III are automatically classified into Class III regardless of the level of risk they pose. The Food and Drug Administration Modernization Act of 1997 established a route to market for low-to-moderate risk medical devices that are automatically placed into Class III due to the absence of a predicate device, called the “Request for Evaluation of Automatic Class III Designation,” or the de novo classification procedure. This procedure allows a manufacturer
113
Table of Contents
whose novel device is automatically classified into Class III to request down-classification of its medical device into Class I or Class II on the basis that the device presents low or moderate risk, rather than requiring the submission and approval of a PMA application. Pursuant to the Food and Drug Administration Safety and Innovation Act (the “FDASIA”) manufacturers may request de novo classification directly without first submitting a 510(k) pre-market notification to the FDA and receiving a not-substantially-equivalent determination. De novo classification requests are subject to the payment of user fees.
Under FDASIA, FDA is required to classify the device within 120 days following receipt of the de novo request, although the process may take significantly longer. If the manufacturer seeks reclassification into Class II, the manufacturer must include a draft proposal for special controls that are necessary to provide a reasonable assurance of the safety and effectiveness of the medical device. If FDA grants the de novo request, the device may be legally marketed in the U.S. However, the FDA may reject the request if the FDA identifies a legally marketed predicate device that would be appropriate for a 510(k) notification, determines that the device is not low-to-moderate risk, or determines that General Controls would be inadequate to control the risks and/or special controls cannot be developed. After a device receives de novo classification, any modification that could significantly affect its safety or efficacy, or that would constitute a major change or modification in its intended use, will require a new 510(k) clearance or, depending on the modification, another de novo request or even PMA approval.
Medical device clinical trials
Clinical trials are sometimes required to support 510(k) or de novo submissions. All clinical investigations of devices to determine safety and effectiveness must be conducted in accordance with the FDA’s IDE, regulations which govern investigational device labeling, prohibit promotion of the investigational device, and specify an array of recordkeeping, reporting and monitoring responsibilities of study sponsors and study investigators. If the device presents a “significant risk” to human health, as defined by the FDA, the FDA requires the device sponsor to submit an IDE application to the FDA, which must become effective prior to commencing human clinical trials. If the device under evaluation does not present a significant risk to human health, then the device sponsor is not required to submit an IDE application to the FDA before initiating human clinical trials, but must still comply with abbreviated IDE requirements when conducting such trials. A significant risk device is one that presents a potential for serious risk to the health, safety or welfare of a patient and either is implanted, used in supporting or sustaining human life, substantially important in diagnosing, curing, mitigating or treating disease or otherwise preventing impairment of human health, or presents a potential for serious risk to a patient in some other way. An IDE application must be supported by appropriate data, such as animal and laboratory test results, showing that it is safe to test the device in humans and that the testing protocol is scientifically sound. The IDE will automatically become effective 30 days after receipt by the FDA unless the FDA notifies the company that the investigation may not begin. If the FDA determines that there are deficiencies or other concerns with an IDE for which it requires modification, the FDA may permit a clinical trial to proceed under a conditional approval.
Regardless of the degree of risk presented by the medical device, clinical studies must be approved by, and conducted under the oversight of, an Institutional Review Board (the “IRB”), for each clinical site. The IRB is responsible for the initial and continuing review of the clinical study, and may pose additional requirements for the conduct of the study. If an IDE application is approved by the FDA and one or more IRBs, human clinical trials may begin at a specific number of investigational sites with a specific number of patients, as approved by the FDA. If the device presents a non-significant risk to the patient, a sponsor may begin the clinical trial after obtaining approval for the trial by one or more IRBs without separate approval from the FDA, but must still follow abbreviated IDE requirements, such as monitoring the investigation, ensuring that the investigators obtain informed consent, and labeling and record-keeping requirements. Acceptance of an IDE application for review does not guarantee that the FDA will allow the IDE to become effective and, if it does become effective, the FDA may or may not determine that the data derived from the trials support the safety and effectiveness of the device or warrant the continuation of clinical trials.
114
Table of Contents
During a study, the sponsor is required to comply with the applicable FDA requirements, including, for example, trial monitoring, selecting clinical investigators and providing them with the investigational plan, ensuring IRB review, adverse event reporting, record keeping and prohibitions on the promotion of investigational devices or on making safety or effectiveness claims for them. The clinical investigators in the clinical study are also subject to FDA’s regulations and must obtain patient informed consent, rigorously follow the investigational plan and study protocol, control the disposition of the investigational device, and comply with all reporting and recordkeeping requirements. Additionally, after a trial begins, we, the FDA or the IRB could suspend or terminate a clinical trial at any time for various reasons, such as strategic business decisions or a belief that the risks to study subjects may outweigh the anticipated benefits.
Expedited development and review programs
Following passage of the 21st Century Cures Act, the FDA implemented the Breakthrough Devices Program, which is a voluntary program offered to manufacturers of certain medical devices and device-led combination products that may provide for more effective treatment or diagnosis of life-threatening or irreversibly debilitating diseases or conditions. The goal of the program is to provide patients and health care providers with more timely access to qualifying devices by expediting their development, assessment and review, while preserving the statutory standards for PMA approval, 510(k) clearance and de novo classification. The program is available for medical devices that meet certain eligibility criteria, including that the device provides more effective treatment or diagnosis of life-threatening or irreversibly debilitating diseases or conditions, and that: (i) the device represents a breakthrough technology, (ii) no approved or cleared alternatives exist, (iii) the device offers significant advantages over existing approved or cleared alternatives, or (iv) the availability of the device is in the best interest of patients. Breakthrough Device Designation provides certain benefits to device developers, including more interactive and timely communications with FDA staff; use of post-market data collection, when scientifically appropriate, to facilitate expedited and efficient development and review of the device; opportunities for more efficient and flexible clinical study design; and prioritized review of premarket submissions. When reviewing Breakthrough Device Designation requests, the FDA may require a combination of literature or preliminary bench, animal or clinical data to demonstrate a reasonable likelihood of clinical and technological success. Receiving a Breakthrough Device Designation from the FDA does not guarantee that the FDA will grant marketing authorization for the device. In 2025, we plan to commence clinical investigation of our CapsoCam’s accuracy in detecting abnormalities indicative of cancerous and precancerous pancreatic neoplasia (abnormal cell growth) by visualizing abnormalities of the duodenal papilla. Thereafter, we hope to secure Breakthrough Device Designation.
Post-market regulation
After a device is cleared or approved for marketing, numerous and pervasive regulatory requirements continue to apply. These include:
| • | establishment registration and device listing with the FDA; |
| • | QSR requirements, which require manufacturers, including third-party manufacturers, to follow stringent design, testing, control, documentation and other quality assurance procedures during all aspects of the design and manufacturing process; |
| • | labeling regulations and FDA prohibitions against the promotion of investigational products, or the promotion of “off-label” uses of cleared or approved products; |
| • | requirements related to promotional activities; |
| • | clearance or approval of product modifications to cleared devices or devices authorized through the de novo classification process that could significantly affect safety or effectiveness, or that would constitute a major change in intended use of such devices, or approval of certain modifications to PMA-approved devices; |
115
Table of Contents
| • | medical device reporting regulations, which require that a manufacturer report to the FDA if a device it markets may have caused or contributed to a death or serious injury, or has malfunctioned and the device or a similar device that it markets would be likely to cause or contribute to a death or serious injury, if the malfunction were to recur; |
| • | correction, removal and recall reporting regulations, which require that manufacturers report to the FDA field corrections and product recalls or removals if undertaken to reduce a risk to health posed by the device or to remedy a violation of the FDCA that may present a risk to health; |
| • | the FDA’s recall authority, whereby the agency can order device manufacturers to recall from the market a product that is in violation of governing laws and regulations; and |
| • | post-market surveillance activities and regulations, which apply when deemed by the FDA to be necessary to protect the public health or to provide additional safety and effectiveness data for the device. |
Manufacturing processes for medical devices are required to comply with the applicable portions of the QSR, which cover the methods and the facilities and controls for the design, manufacture, testing, production, processes, controls, quality assurance, labeling, packaging, distribution, installation and servicing of finished devices intended for human use. The QSR also requires, among other things, maintenance of a device master file, device history file, and complaint files. As a manufacturer, we are subject to periodic scheduled or unscheduled inspections by the FDA. Failure to maintain compliance with the QSR requirements could result in the shutdown of, or restrictions on, manufacturing operations and the recall or seizure of marketed products. The discovery of previously unknown problems with marketed medical devices, including unanticipated adverse events or adverse events of increasing severity or frequency, whether resulting from the use of the device within the scope of its clearance or off-label by a physician in the practice of medicine, could result in restrictions on the device, including the removal of the product from the market or voluntary or mandatory device recalls.
The FDA has broad regulatory compliance and enforcement powers. If the FDA determines that a manufacturer has failed to comply with applicable regulatory requirements, it can take a variety of compliance or enforcement actions, which may result in any of the following sanctions, among others:
| • | warning letters, untitled letters, it has come to our attention letters, fines, injunctions, consent decrees, and civil penalties; |
| • | recalls, withdrawals, or administrative detention or product seizures; |
| • | operating restrictions or partial suspension or total shutdown of production; |
| • | refusing or delaying requests for 510(k) marketing clearance or PMA approvals of new products or modified products; |
| • | withdrawing 510(k) clearances or PMA approvals that have already been granted; |
| • | refusal to grant export approvals for devices being shipped to foreign markets; or |
| • | criminal prosecution. |
We are also subject to regulation by the California Department of Public Health Food and Drug Branch (“FDB”) through the Medical Device Safety Program. We must maintain a California Medical Device Manufacturing license. Our facilities may be subjected to scheduled or unscheduled inspections by the FDB.
Healthcare Fraud and Abuse Laws
In the U.S., we are subject to a number of federal and state healthcare regulatory laws that restrict business practices in the healthcare industry. These laws include, but are not limited to, federal and state anti-kickback, false claims, and other healthcare fraud and abuse laws.
116
Table of Contents
The federal Anti-Kickback Statute prohibits, among other things, any person or entity from knowingly and willfully offering, paying, soliciting, receiving or providing any remuneration, directly or indirectly, overtly or covertly, to induce or in return for purchasing, leasing, ordering, or arranging for or recommending the purchase, lease, or order of any good, facility, item, or service reimbursable, in whole or in part, under Medicare, Medicaid or other federal healthcare programs. A person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation.
The federal false claims laws, including the civil False Claims Act, prohibit, among other things, any person or entity from knowingly presenting, or causing to be presented, a false, fictitious or fraudulent claim for payment to, or approval by, the federal government, knowingly making, using, or causing to be made or used a false record or statement material to a false or fraudulent claim to the federal government, or knowingly making a false statement to avoid, decrease or conceal an obligation to pay money to the federal government. A claim includes “any request or demand” for money or property presented to the government. Actions under the civil False Claims Act may be brought by the Attorney General or as a qui tam action by a private individual in the name of the government. Moreover, a claim including items or services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the federal civil False Claims Act.
In addition, the civil monetary penalties statute, subject to certain exceptions, prohibits, among other things, the offer or transfer of remuneration, including waivers of copayments and deductible amounts (or any part thereof), to a Medicare or state healthcare program beneficiary if the person knows or should know it is likely to influence the beneficiary’s selection of a particular provider, practitioner, or supplier of services reimbursable by Medicare or a state healthcare program.
The federal Health Insurance Portability and Accountability Act of 1996 (“HIPAA”) created additional federal criminal statutes that prohibit, among other actions, knowingly and willfully executing, or attempting to execute, a scheme to defraud any healthcare benefit program, including private third party payers, knowingly and willfully embezzling or stealing from a healthcare benefit program, willfully obstructing a criminal investigation of a healthcare offense, and knowingly and willfully falsifying, concealing or covering up a material fact or making any materially false, fictitious, or fraudulent statement in connection with the delivery of or payment for healthcare benefits, items, or services. Similar to the U.S. federal Anti-Kickback Statute, a person or entity does not need to have actual knowledge of the statute or specific intent to violate it in order to have committed a violation.
HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act of 2009 (“HITECH”), and their respective implementing regulations, impose requirements relating to the privacy, security and transmission of individually identifiable health information on certain covered healthcare providers, health plans, and healthcare clearinghouses, as well as business associates, independent contractors or agents of covered entities that create, receive, maintain or transmit protected health information in connection with providing a service for or on behalf of a covered entity. HITECH also created new tiers of civil monetary penalties, amended HIPAA to make civil and criminal penalties directly applicable to business associates, and gave state attorneys general new authority to file civil actions for damages or injunctions in federal courts to enforce the federal HIPAA laws and seek attorneys’ fees and costs associated with pursuing federal civil actions.
The federal Physician Payments Sunshine Act requires certain manufacturers of drugs, devices, biologics and medical supplies for which payment is available under Medicare, Medicaid or the Children’s Health Insurance Program, with specific exceptions, to report annually to the CMS, information related to payments or other transfers of value made to physicians (defined to include doctors, dentists, optometrists, podiatrists, and chiropractors), certain other healthcare professionals such as physician assistants and nurse practitioners, and teaching hospitals, and applicable manufacturers and applicable group purchasing organizations to report annually to CMS ownership and investment interests held by physicians and their immediate family members.
Several states have also adopted fraud and abuse laws similar to those described above. The scope of these laws and the interpretations of them vary from state to state and are enforced by state courts and regulatory
117
Table of Contents
authorities, each with broad discretion. Some state fraud and abuse laws apply to items or services reimbursed by any payer, including patients and commercial insurers, not just those reimbursed by a federally funded healthcare program.
Violations of fraud and abuse laws, including federal and state anti-kickback and false claims laws, may be punishable by criminal and civil sanctions, including fines and civil monetary penalties, the possibility of exclusion from federal healthcare programs (including Medicare and Medicaid), disgorgement, and corporate integrity agreements, which impose, among other things, rigorous operational and monitoring requirements on companies. Similar sanctions and penalties, as well as imprisonment, also can be imposed upon executive officers and employees of such companies.
Coverage and Reimbursement Regulation
In the U.S., our commercial success depends in part on the extent to which governmental authorities, private health insurers and other third-party payers provide coverage for and establish adequate reimbursement levels for our products and related services. Use of the CapsoCam Plus capsule endoscopy system is reimbursed under existing physician and hospital codes. We do not bill any third-party payers for the CapsoCam Plus. Instead, we invoice healthcare providers and the cost is bundled into the reimbursement received by healthcare providers when the CapsoCam Plus is used. Failure by physicians, hospitals, and other users of our products to obtain adequate reimbursement from third-party payers for services performed with our products, or adverse changes in government and private third-party payers’ coverage and reimbursement policies, could adversely impact demand for our products.
Coverage and reimbursement for use of the CapsoCam Plus capsule endoscopy system can differ significantly from payer to payer. Third-party payers are increasingly auditing and challenging the prices charged for medical products and services with concern for upcoding, miscoding, using inappropriate modifiers, or billing for inappropriate care settings. Some third-party payers must approve coverage for new or innovative devices before they will reimburse healthcare providers who use the products or therapies. Even though a new product may have been cleared for commercial distribution by the FDA, we may find limited demand for the product unless and until reimbursement approval has been obtained from governmental and private third-party payers.
In addition to uncertainties surrounding coverage policies, there are periodic changes to reimbursement levels. Third-party payers regularly update reimbursement amounts and also from time to time revise the methodologies used to determine reimbursement amounts. This includes routine updates to payments to hospitals under the IPPS. These updates could directly impact the demand for our products.
We believe the overall escalating cost of medical products and services being paid for by the government and private health insurance has led to, and will continue to lead to, increased pressures on the healthcare and medical device industries to reduce the costs of products and services. Third-party payers are developing increasingly sophisticated methods of controlling healthcare costs through prospective reimbursement and capitation programs, group purchasing, redesign of benefits, and exploration of more cost-effective methods of delivering healthcare. In the U.S., some insured individuals enroll in managed care programs, which monitor and often require pre-approval of the services that a member will receive. Some managed care programs pay their providers on a per capita (patient) basis, which puts the providers at financial risk for the services provided to their patients by paying these providers a predetermined payment per member per month and, consequently, may limit the willingness of these providers to use our products.
As we also sell into international markets, we note that reimbursement and healthcare payment systems vary significantly by country, and many countries have instituted price ceilings on specific product lines and procedures. In particular, in Europe, reimbursement is entirely regulated at member state level, varies significantly between countries, and member states are facing increased pressure to limit public healthcare spending. There can be no assurance that our products will be considered cost-effective by third party payers, that
118
Table of Contents
an adequate level of reimbursement will be available or that the third-party payers’ reimbursement policies will not adversely affect our ability to sell our products profitably. More and more, local, product specific reimbursement law is applied as an overlay to medical device regulation, which has provided an additional layer of clearance requirements.
Healthcare Reform
The U.S. and some foreign jurisdictions are considering or have enacted a number of legislative and regulatory proposals to change the healthcare system in ways that could affect our ability to sell our products profitably. Among policy makers and payers in the U.S. and elsewhere, there is significant interest in promoting changes in healthcare systems with the stated goals of containing healthcare costs, improving quality or expanding access. Current and future legislative proposals to further reform healthcare or reduce healthcare costs may limit coverage of or lower reimbursement for the procedures associated with the use of our products. The cost containment measures that payers and providers are instituting and the effect of any healthcare reform initiative implemented in the future could impact our revenue from the sale of our products.
The implementation of the Affordable Care Act (the “ACA”) in the U.S., for example, has substantially changed healthcare financing and delivery by both governmental and private insurers, and significantly affected medical device manufacturers. The ACA, among other things, provided incentives to programs that increase the federal government’s comparative effectiveness research, and implemented payment system reforms including a national pilot program on payment bundling to encourage hospitals, physicians, and other providers to improve the coordination, quality and efficiency of certain healthcare services through bundled payment models. Additionally, the ACA expanded eligibility criteria for Medicaid programs and created a new Patient-Centered Outcomes Research Institute to oversee, identify priorities in, and conduct comparative clinical effectiveness research, along with funding for such research. Since its enactment, there have been judicial, executive and political challenges to certain aspects of the ACA. On June 17, 2021, the U.S. Supreme Court dismissed a judicial challenge to the ACA brought by several states without specifically ruling on its constitutionality.
Other legislative changes have been proposed and adopted since the ACA was enacted. For example, the Budget Control Act of 2011, among other things, reduced Medicare payments to providers, effective on April 1, 2013 and, due to subsequent legislative amendments to the statute, will remain in effect through 2032, with the exception of a temporary suspension from May 1, 2020 through March 31, 2022, unless additional Congressional action is taken. Additionally, the American Taxpayer Relief Act of 2012, among other things, further reduced Medicare payments to several providers, including hospitals, and increased the statute of limitations period for the government to recover overpayments to providers from three to five years.
We expect additional state and federal healthcare reform measures to be adopted in the future, any of which could limit the amounts that federal and state governments will pay for healthcare products and services, which could result in reduced demand for our products or additional pricing pressure.
State Corporate Practice of Medicine and Fee-Splitting Laws
Our arrangements with contracted telemedicine providers who provide reading services to certain customers are subject to various state laws, including those commonly referred to as corporate practice of medicine and fee-splitting laws, which are intended to prevent unlicensed persons from interfering with or influencing the physician’s professional judgment, and prohibiting the sharing of professional service fees with non-professional or business interests. These laws vary from state to state and are subject to broad interpretation and enforcement by state regulators and other bodies. A determination that we and/or our contracted providers are not compliant with such laws could lead to adverse judicial or administrative action, civil or criminal penalties, receipt of cease and desist orders from state regulators, loss of customers, and/or restructuring of these contractual arrangements.
119
Table of Contents
Data Privacy and Security Laws
Numerous state, federal, and foreign laws, regulations, and standards govern the collection, use, disclosure, access to, confidentiality, and security of health-related and other personal information, and could apply now or in the future to our operations or the operations of our collaborators, third-party providers, and others upon whom we commercially rely upon. In the U.S., numerous federal and state laws and regulations, including data breach notification laws, health information privacy and security laws, and consumer protection laws and regulations govern the collection, use, disclosure and protection of health-related and other personal information. In addition, certain foreign laws govern the privacy and security of personal data, including health-related data. Privacy and security laws, regulations, and other obligations are constantly evolving, may conflict with each other to complicate compliance efforts, and can result in investigations, proceedings or actions that lead to significant civil and/or criminal penalties and restrictions on data processing.
Regulation of Medical Devices Outside the United States
Outside of the U.S., the regulation of medical devices is also complex. In Europe, for instance, products are subject to extensive regulatory requirements. In 2021, a new regulatory scheme for medical devices, the Medical Devices Regulation (“MDR”), became effective in EU member states. The MDR requires that medical devices may only be placed on the market if they do not compromise safety and health when properly installed, maintained, and used in accordance with their intended purpose. The MDR has significant requirements for many medical devices, including requirements for clinical evidence and documentation, device identification and traceability, registration of economic operators throughout the distribution chain and post-market surveillance. In some regions, the level of government regulation of medical devices is increasing, which can lengthen time to market and increase registration and approval costs. In many countries, the national health or social security organizations require products to be qualified before they can be marketed and considered eligible for reimbursement.
In many instances, global regulatory agencies have come together in an attempt to harmonize medical device regulatory requirements. In 2011, the regulatory agencies of the U.S., Canada, Brazil, Australia and Japan came together and established the International Medical Device Regulators Forum (the “IMDRF”). The IMDRF continues to grow and now has a management committee of regulatory agency representatives from 11 countries and affiliate members representing 7 countries. One example of the IMDRF harmonizing medical device regulatory requirements is the Medical Device Single Audit Program (the “MDSAP”), whereby a medical device manufacturer can have a single Quality Management System audit of their facility which covers the regulatory requirements of Australia, Brazil, Canada, Japan and the U.S. Instead of having periodic quality inspections from regulators of each of these countries, a single comprehensive inspection is performed. We are audited in compliance with the MDSAP.
Other regional groups working to harmonize regulatory requirements are the Asia-Pacific Economic Cooperation group, Global Harmonization Working Party and African Medical Devices Forum. While regulatory requirements are constantly evolving, regulatory agencies recognize the impact and are attempting to harmonize their efforts.
While the list of regulated countries continues to grow, many of the regulated countries leverage device approvals from the U.S. or Europe, meaning that the testing and clinical studies required to satisfy device safety and efficacy requirements of the U.S. and Europe, often carry over to other geographies.
120
Table of Contents
Directors and Executive Officers
The following table sets forth the names and ages of all of our directors and executive officers as of the date of this prospectus:
| Name |
Age | Position | ||||
| Executive Officers |
||||||
| Kang-Huai (Johnny) Wang |
68 | President, Chief Executive Officer and Director | ||||
| Kevin Lundquist |
57 | Chief Financial Officer | ||||
| Rebecca Petersen |
61 | Director of Clinical Affairs | ||||
| Douglas Atkinson |
56 | Head of Global Sales | ||||
| Non-Employee Directors |
||||||
| Eliyahou Harari |
79 | Director | ||||
| Chen Lung Tsai |
72 | Director | ||||
Set forth below is biographical information about each of the individuals named in the tables above:
Executive Officers
Kang-Huai (Johnny) Wang is a co-founder of our Company and a member of our board of directors, and has served as our President and Chief Executive Officer since November 2024, and before then our President and Chief Technology Officer from October 2006 to October 2024. Prior to joining the Company, Mr. Wang was Vice President of Engineering for ESS Technology, a technology company from September 2003 to November 2004; Chief Executive Officer and co-founder of Divio, Inc, a video processing chip design company, from March 1996 to August 2003; and as Design Manager for C-Cube Mircrosystem, an IC design house from March 1992 to August 1995. Prior to 1992, Mr. Wang held various engineering roles in the semiconductor industry. Mr. Wang holds a B.S. in electrical engineering from National Chiao-Tung University in Taiwan, and a M.S. in electrical engineering from Texas A&M University. We believe that Mr. Wang is qualified to serve on our board of directors due to the valuable expertise and perspective he brings in his capacity as a co-founder and our chief executive officer and because of his extensive experience and knowledge of our industry.
Kevin Lundquist joined the Company in November 2024 and currently serves as our Chief Financial Officer. Prior to joining the Company, Mr. Lundquist served as Chief Financial Officer of Abzena Biologics, a biotechnology company, from January 2022 to December 2023; as Vice President of finance of Revance, Inc, a biotechnology company from August 2020 to January 2022; as Senior Director of Global Manufacturing Finance of Roche, a pharmaceutical company from August 2014 to August 2020; and as Chief Financial Officer for Caterpillar Japan, a heavy machinery company from May 2010 to January 2014; as Director of Finance Operations for Caterpillar India, a heavy machinery company from August 2006 to May 2010; as Director of International Business Development for Abbott Laboratories, a pharmaceutical company from May 2003 to August 2006. Mr. Lundquist holds an M.B.A. in International Finance from Utah State University and a B.S. in accounting and finance from the University of Utah.
Rebecca Petersen joined the Company and has served as our Director of Clinical Affairs since March 2022, Associate Director from January 2022 to March 2022, and our Senior Clinical Project Manager from June 2020 to January 2022. Ms. Petersen has worked in both device and pharmaceutical clinical operations during her 19 years of career in clinical research. Ms. Peterson worked as Senior Clinical Trials Manager at Gilead Sciences, a pharmaceutical company from March 2017 to July 2018 and before then in various clinical research roles from October 2011 to March 2017; and as a Senior Clinical Research Associate at Abbott Vascular, a medical device company from December 2006 to October 2011. Ms. Petersen also has a nursing background working in direct patient care. Ms. Peterson received a B.S. in computer science from California State University Hayward.
121
Table of Contents
Douglas Atkinson joined the Company and has served as our Head of Global Sales since October 2016. Prior to joining the Company, Mr. Atkinson was Director of Sales for MedTech Micro-Fixation, a distributor of Zimmer-Biomet, a medical device company, from May 2015 to October 2016; as Regional Sales Director at Medtronic PLC, a gastrointestinal medical device company, from January 2005 to April 2015. Before serving at Medtronic, Mr. Atkinson also served as Regional Director of Sales and National Accounts at Stryker Corporation from May 1995 to December 2004. At Stryker, Mr. Atkinson was recognized for multiple sales leadership and revenue growth awards. Mr. Atkinson received a B.A. in economics and political science from Duke University.
Non-Employee Directors
Eliyahou Harari has served as a member of our board of directors since 2021. Dr. Harari has also served as the Chairman and CEO of SunRise Memory since 2016, a memory technology company; and as the founder, Chairman and CEO of SanDisk Corp., from 1988 to 2010. In 1976, Dr. Harari invented the first practical Electrically Erasable Programmable Read Only Memory which laid the foundation for today’s Flash Memory industry. Dr. Harari received a Ph.D. in Solid State Physics from Princeton University in 1973. Dr. Harari is a member of the US National Academy of Engineering. In 2012, Dr. Harari was the recipient of the U.S. National Medal of Technology and Innovation. Dr. Harari was also awarded the IEEE Robert Noyce Medal by the Institute of Electrical and Electronics Engineers in 2009 and the Dr. Morris Chang Exemplary Leadership Award by the Global Semiconductor Alliance in 2008. We believe Dr. Harari is qualified to serve on our board of directors because of his extensive experience and leadership in the memory technology industry, his scientific background and training, and his significant contributions to the advancement of flash memory.
Chen Lung Tsai has served as a member of our board of directors since 2014. Mr. Tsai also served as the Chairman of the Board for One Test Systems, a technology innovation company, from 2018 to 2023; as USA Representative for TeraPower, a nuclear power plant, from 2016 to 2018; as Chief Operations Officer and board director for Lucis-Tech, an innovative technology manufacturer, from 2015 to 2016; as President of Operations and Senior Vice President (“SVP”) of Manufacturing Operations for Greenliant Systems, a solid state storage company, from 2010 to 2015; as SVP of Worldwide Backend Operations and a board director for Silicon Storage Technology, a data storage technology company from 1996 to 2010, as well as various other engineering roles in the semiconductor manufacturing industry. Mr. Tsai holds a B.S. from Show Chu University, and a M.S. in Physics and Electrical Engineering from the Florida Institute of Technology. We believe Mr. Tsai is qualified to serve on our board of directors because of his leadership in global semiconductor manufacturing, innovation in secure storage technologies, and significant contributions to the advancement of semiconductor testing and operations.
Director Independence
In accordance with our amended and restated certificate of incorporation, which will be effective immediately prior to the completion of this offering, our board of directors will consist of directors. We are currently working to appoint additional individuals to our board of directors. The rules of the Nasdaq Stock Market, or the Nasdaq Rules, require a majority of a listed company’s board of directors to be composed of independent directors within one year of listing. In addition, the Nasdaq Rules require that, subject to specified exceptions, each member of a listed company’s audit, compensation and nominating and governance committees be independent. Under the Nasdaq Rules, a director will only qualify as an independent director if, in the opinion of our board of directors, that person does not have a relationship that would interfere with the exercise of independent judgment in carrying out the responsibilities of a director. The Nasdaq Rules also require that audit committee members satisfy independence criteria set forth in Rule 10A-3 under the Exchange Act. In order to be considered independent for purposes of Rule 10A-3, a member of an audit committee of a listed company may not, other than in his or her capacity as a member of the audit committee, the board of directors, or any other board committee, accept, directly or indirectly, any consulting, advisory, or other compensatory fee from the listed company or any of its subsidiaries or otherwise be an affiliated person of the listed company or any of its
122
Table of Contents
subsidiaries. In considering the independence of compensation committee members, the Nasdaq Rules require that our board of directors must consider additional factors relevant to the duties of a compensation committee member, including the source of any compensation we pay to the director and any affiliations with the Company.
Classified Board of Directors
In accordance with our amended and restated certificate of incorporation, which will be effective immediately prior to the completion of this offering, our board of directors will be divided into three classes with staggered three-year terms. At each annual general meeting of stockholders, the successors to directors whose terms then expire will be elected to serve from the time of election and qualification until the third annual meeting following election. Our directors will be divided among the three classes as follows:
| • | The Class I directors will be , , and , and their terms will expire at the annual meeting of stockholders to be held in 2026; |
| • | The Class II directors will be , , and , and their terms will expire at the annual meeting of stockholders to be held in 2027; and |
| • | The Class III directors will be , , and , and their terms will expire at the annual meeting of stockholders to be held in 2028. |
We expect that any additional directorships resulting from an increase in the number of directors will be distributed among the three classes so that, as nearly as possible, each class will consist of one-third of the directors. The division of our board of directors into three classes with staggered three-year terms may delay or prevent a change of our management or a change in control.
Leadership Structure of the Board
Our amended and restated bylaws and corporate governance guidelines to be in place immediately prior to the consummation of this offering will provide our board of directors with flexibility to combine or separate the positions of Chair of the board of directors and Chief Executive Officer and to implement a lead director in accordance with its determination regarding which structure would be in the best interests of our Company.
Our board of directors has concluded that our current leadership structure is appropriate at this time. However, our board of directors will continue to periodically review our leadership structure and may make such changes in the future as it deems appropriate.
Board Committees
Our board of directors will establish an audit committee, a compensation committee, and a nominating and corporate governance committee, immediately prior to the completion of this offering. Our board of directors may establish other committees to facilitate the management of our business. The composition and functions of each committee are described below. Members will serve on these committees until their resignation or until otherwise determined by our board of directors. Each committee will adopt a written charter that satisfies the applicable rules and regulations of the SEC and Nasdaq Rules, which we will post on our website at capsovision.com upon the completion of this offering.
Audit Committee
Our audit committee will oversee our corporate accounting and financial reporting process. Among other matters, the audit committee will:
| • | appoint our independent registered public accounting firm; |
| • | evaluate the independent registered public accounting firm’s qualifications, independence, and performance; |
123
Table of Contents
| • | determine the engagement of the independent registered public accounting firm; |
| • | review and approve the scope of the annual audit and pre-approves the audit and non-audit fees and services; |
| • | review and approve all related-party transactions on an ongoing basis; |
| • | establish procedures for the receipt, retention, and treatment of any complaints received by us regarding accounting, internal accounting controls, or auditing matters; |
| • | discuss with management and our independent registered public accounting firm the results of the annual audit and the review of our quarterly financial statements; |
| • | approve the retention of the independent registered public accounting firm to perform any proposed permissible non-audit services; |
| • | discuss on a periodic basis, or as appropriate, with management our policies and procedures with respect to risk assessment and risk management, including information security, financial, and regulatory compliance related risks; |
| • | be responsible for reviewing our financial statements and our management’s discussion and analysis of financial condition and results of operations to be included in our annual and quarterly reports to be filed with the SEC; |
| • | be responsible for our controls and procedures for mitigating cybersecurity and other information technology risks, including our plans to respond to data breaches; |
| • | review and investigate complaints regarding accounting, internal control over financial reporting, and auditing matters received through our compliance helpline (and other means) pursuant to our whistleblower policy; and |
| • | review the audit committee charter and the audit committee’s performance on an annual basis. |
Our audit committee will consists of , , and . Our board of directors has determined that and are independent under the Listing Rules and Rule 10A-3(b)(1) of the Exchange Act. The chair of our audit committee will be . Our board of directors has determined that is an “audit committee financial expert” as such term is currently defined in Item 407(d)(5) of Regulation S-K. Our board of directors has also determined that each member of our audit committee can read and understand fundamental financial statements, in accordance with applicable requirements.
Compensation Committee
Our compensation committee will oversee policies relating to compensation and benefits of our officers and employees. Among other matters, the compensation committee will:
| • | review and approve or recommends corporate goals and objectives relevant to compensation of our Chief Executive Officer; |
| • | evaluate the performance of the Chief Executive Officer in light of those goals and objectives and approves, or make recommendations to the board of directors regarding, the compensation of the Chief Executive Officer based on such evaluations; |
| • | review and approve or makes recommendations to our board of directors regarding the issuance of stock options and other awards under our stock plans to our Chief Executive Officer, other executive officers, employees, and other service providers; |
| • | oversee the evaluation of our executive officers (other than our Chief Executive Officer) and, after considering such evaluation, review and approve, or make recommendations to the board of directors regarding, the compensation of such executive officers; and |
124
Table of Contents
| • | review the compensation committee charter and the compensation committee’s performance on annual basis. |
Our compensation committee will consist of , , and . Our board of directors has determined that and are independent under the Listing Rules and are non-employee directors, as defined by Rule 16b-3 promulgated under the Exchange Act. The chair of our compensation committee is .
Nominating and Corporate Governance Committee
Our nominating and corporate governance committee will oversee policies relating to our corporate governance. Among other matters, the nominating and corporate governance committee will:
| • | identify and recommend candidates for membership on our board of directors, including the consideration of nominees submitted by stockholders, and on each of the board’s committees; |
| • | review and recommend our corporate governance guidelines and policies; |
| • | oversee the process of evaluating the performance of our board of directors; |
| • | oversee management succession planning; |
| • | assist our board of directors on corporate governance matters, including strategy and risks related to environmental, social, and governance issues; and |
| • | review the nominating and corporate governance committee charter on an annual basis and the nominating and corporate governance committee’s performance periodically. |
Our nominating and corporate governance committee will consist of , and . Our board of directors has determined that and are independent under the Listing Rules. The chair of our nominating and corporate governance committee is .
Compensation Committee Interlocks and Insider Participation
None of the members of our compensation committee is currently, or has been at any time, one of our executive officers or employees. None of our executive officers currently serves, or has served during the last year, as a member of the board of directors or compensation committee of any entity that has one or more executive officers serving as a member of our board of directors or on our compensation committee.
Code of Business Conduct and Ethics
In connection with this offering, our board of directors intends to adopt a written code of business conduct and ethics that applies to all of our directors, officers, and employees, including our principal executive officer, principal financial officer, and principal accounting officer or controller, or persons performing similar functions, and agents and representatives. The full text of our code of business conduct and ethics will be posted on our website at capsovision.com upon the completion of this offering. The audit committee of our board of directors will be responsible for overseeing our code of business conduct and ethics and any waivers applicable to any director, executive officer, or employee. We intend to disclose any future amendments to certain provisions of our code of business conduct and ethics, or waivers of such provisions applicable to our directors, officers, and employees, including our principal executive officer, principal financial officer, principal accounting officer or controller, or persons performing similar functions, and agents and representatives, on our website identified above or in public filings.
125
Table of Contents
Policies on Clawback and Recovery of Compensation
In connection with this offering, we will adopt, prior to the listing of our shares on Nasdaq, a clawback policy to address the recovery of erroneously-awarded incentive compensation in compliance with the requirements of the Dodd-Frank Act, final SEC rules and applicable listing standards.
Limitations on Liability and Indemnification Matters
Our amended and restated certificate of incorporation and our amended and restated bylaws, both of which will become effective immediately prior to the completion of this offering, limit our directors’ and officers’ liability and provide that we shall indemnify our directors and officers to the fullest extent permitted under the General Corporation Law of the State of Delaware (the “DGCL”). The DGCL provides that directors and officers of a corporation will not be personally liable for monetary damages for breach of their fiduciary duties as directors or officers, except for liability for any:
| • | transaction from which the director or officer derives an improper personal benefit; |
| • | act or omission not in good faith or that involves intentional misconduct or a knowing violation of law; |
| • | unlawful payment of dividends or redemption of shares; or |
| • | breach of a director’s or officer’s duty of loyalty to the corporation or its stockholders. |
These limitations of liability do not apply to liabilities arising under federal securities laws and do not affect the availability of equitable remedies such as injunctive relief or recession.
The DGCL, our amended and restated certificate of incorporation, and our amended and restated bylaws provide that we will, in certain situations, indemnify our directors and officers and may indemnify other employees and other agents, to the fullest extent permitted by law. Any indemnified person is also entitled, subject to certain limitations, to advancement, direct payment, or reimbursement of reasonable expenses (including attorneys’ fees and disbursements) in advance of the final disposition of the proceeding.
In addition, we have entered, and intend to continue to enter, into separate indemnification agreements with our directors and officers. These indemnification agreements, among other things, require us to indemnify our directors and officers for certain expenses, including attorneys’ fees, judgments, fines, and settlement amounts incurred by a director or officer in any action or proceeding arising out of their services as a director or officer, or any other company or enterprise to which the person provides services at our request.
In connection with this offering, we will put in place a directors’ and officers’ insurance policy pursuant to which our directors and officers will be insured against liability for actions taken in their capacities as directors and officers. We believe that these provisions in our amended and restated certificate of incorporation and amended and restated bylaws and these indemnification agreements are necessary to attract and retain qualified persons as directors and officers.
Insofar as indemnification for liabilities arising under the Securities Act may be permitted to directors, officers, or control persons, in the opinion of the SEC, such indemnification is against public policy, as expressed in the Securities Act and is therefore unenforceable.
126
Table of Contents
EXECUTIVE AND DIRECTOR COMPENSATION
This section describes the material components of the executive compensation program for our executive officers and directors. This discussion may contain forward-looking statements that are based on our current plans, considerations, expectations and determinations regarding future compensation programs. Actual compensation programs that we adopt following the completion of this offering may differ materially from the existing and currently planned programs summarized or referred to in this discussion.
Executive Compensation Program
In connection with this offering, we intend to develop a compensation program that is designed to align executives’ compensation with our business objectives and the creation of stockholder value, while helping us to continue to attract, motivate and retain individuals who contribute to the long-term success of the company. We anticipate that compensation for our executive officers will have three primary components: base salary, an annual cash incentive bonus opportunity, and long-term equity-based incentive compensation.
Decisions on the design and implementation of the executive compensation program will be made by the compensation committee, as established in connection with this offering. The executive compensation program actually adopted will depend on the judgment of the members of the compensation committee.
The table below sets forth the compensation for the year ended December 31, 2024 awarded to or earned by our chief executive officer and the other executive officers employed by us on that date (the “NEOs”).
Summary Compensation Table — Fiscal Year 2024
| Year | Salary ($) |
Bonus ($)(1) |
Stock Awards ($)(2) |
Option Awards ($)(2) |
Non-Equity Incentive Plan Compensation ($)(1) |
All Other Compensation ($) |
Total ($) |
|||||||||||||||||||||||||
| Kang-Huai (Johnny) Wang |
||||||||||||||||||||||||||||||||
| President and Chief Executive Officer |
2024 | 204,091 | — | — | [ ] | — | — | [ ] | ||||||||||||||||||||||||
| Kevin Lundquist(3) |
||||||||||||||||||||||||||||||||
| Chief Financial Officer |
2024 | 54,167 | — | — | [ ] | — | — | [ ] | ||||||||||||||||||||||||
| Rebecca Petersen |
||||||||||||||||||||||||||||||||
| Director of Clinical Affairs |
2024 | 196,100 | — | — | — | — | — | 196,100 | ||||||||||||||||||||||||
| Douglas Atkinson |
||||||||||||||||||||||||||||||||
| Head of Global Sales |
2024 | 283,150 | 140,550 | — | — | — | — | 423,700 | ||||||||||||||||||||||||
| (1) | The amount reported in this column for Mr. Atkinson represents sales commissions earned during fiscal 2024. The NEOs did not receive any bonuses for services performed during 2024. |
| (2) | The amounts reported in these columns reflect the grant date fair value of stock awards and option awards granted to the NEOs during 2024 under the CapsoVision, Inc. Amended and Restated 2005 Stock Plan and are accounted for in accordance with FASB ASC Topic 718. Please see Note 10 to the Financial Statements included elsewhere in this prospectus for a discussion of the relevant assumptions used in calculating these amounts. |
| (3) | Mr. Lundquist commenced employment with us effective November 1, 2024. |
Outstanding Equity Awards as of December 31, 2024
The following table provides information regarding outstanding options to purchase shares of our common stock held by each of the NEOs as of December 31, 2024, including the vesting dates for the portions of these awards that had not vested as of that date. Each of the options held by the NEOs reported in the table below, to
127
Table of Contents
the extent then outstanding and unvested, will vest in full upon a change in control of the Company. The NEOs did not hold any other outstanding equity awards as of that date.
| Option Awards |
||||||||||||||||||||
| Name |
Number of Securities Underlying Unexercised Options (#) Exercisable |
Number of Securities Underlying Unexercised Options (#) Unexercisable |
Equity Incentive Plan Awards: Number of Securities Underlying Unexercised Unearned Options (#) |
Option Exercise Price ($) |
Option Expiration Date |
|||||||||||||||
| Kang-Huai (Johnny) Wang |
50,000 | — | — | 0.09 | 7/2/2027 | |||||||||||||||
| 50,000 | — | — | 0.09 | 10/3/2028 | ||||||||||||||||
| 68,333 | 11,667 | (1) | — | 0.11 | 6/14/2031 | |||||||||||||||
| 33,333 | 16,667 | (2) | — | 0.11 | 3/22/2032 | |||||||||||||||
| — | 887,500 | (3) | — | 0.17 | 10/23/2034 | |||||||||||||||
| Kevin Lundquist |
— | 1,387,500 | (3) | — | 0.17 | 11/12//2034 | ||||||||||||||
| Rebecca Petersen |
25,000 | — | — | 0.11 | 10/27/2031 | |||||||||||||||
| 46,666 | 23,334 | (2) | — | 0.11 | 3/22/2032 | |||||||||||||||
| Douglas Atkinson |
120,000 | — | — | 0.086 | 11/9/2026 | |||||||||||||||
| 85,416 | 14,583 | (1) | — | 0.11 | 6/14/2031 | |||||||||||||||
| 75,000 | 125,000 | (4) | — | |||||||||||||||||
| (1) | These options vest as to 25% of the shares subject to the option on July 1, 2022 and as to the remaining 75% of the shares in 36 equal monthly installments thereafter through July 1, 2025. |
| (2) | These options vest as to 25% of the shares subject to the option on April 1, 2023 and as to the remaining 75% of the shares in 36 equal monthly installments thereafter through April 1, 2026. |
| (3) | These options vest as to 25% of the shares subject to the option on November 1, 2025 and as to the remaining 75% of the shares in 36 equal monthly installments thereafter through November 1, 2028. |
| (4) | This option vests as to 25% of the shares subject to the option on June 16, 2024 and as to the remaining 75% of the shares in 36 equal monthly installments thereafter through June 16, 2027. |
Employment Agreements
We have entered into offer letters with each of the NEOs (other than Mr. Wang). Each letter provides for the NEO to receive a base salary and initial option grant. In addition, Mr. Lundquist’s letter provides for him to receive an annual bonus of $40,000, and Mr. Atkinson’s letter provides for him to have the opportunity to receive an annual bonus based on achievement of certain sales targets. His target bonus for 2024 was $148,100. The NEOs’ offer letters do not include any severance provisions.
Equity Incentive Plans
As of , 2025, our employees, consultants and directors held outstanding stock options to purchase up to shares of our common stock. These awards were granted under our Amended and Restated 2005 Stock Plan (the “2005 Plan”). As of , 2025, the outstanding options were vested with respect to shares and were unvested with respect to shares. The exercise prices of those options ranged from $ per share to $ per share, and each of the options had a maximum term of 10 years from the applicable date of grant.
The following sections provide more detailed information concerning our benefit plans and, with respect to our equity compensation plans, the shares that are available for future awards under these plans. Each summary below is qualified in its entirety by the full text of the relevant plan document, which has been filed with the SEC as an exhibit to the registration statement of which this prospectus is a part and is available through the SEC’s internet site at http://www.sec.gov.
128
Table of Contents
2005 Stock Plan
Under the 2005 Plan, we are generally authorized to grant options and other equity-based awards to our employees, directors, officers and consultants and those of our subsidiaries. Options under the 2005 Plan are either incentive stock options, within the meaning of Section 422 of the Internal Revenue Code, or nonqualified stock options. All options granted under the plan expire no later than ten years from their date of grant. As of , 2025, we had reserved shares of our common stock for issuance under the 2005 Plan and shares remained available for future grant. No new awards will be granted under the 2005 Plan after the consummation of this offering.
Our board of directors, or a committee appointed by the board, administers the 2005 Plan. As is customary in incentive plans of this nature, the number of shares subject to outstanding awards under the 2005 Plan and the exercise prices of those awards, are subject to adjustment in the event of changes in our capital structure, reorganizations and other extraordinary events. If a merger or similar transaction involving the Company occurs, or there is a sale of substantially all shares or assets of the Company occurs, the board of directors may provide for outstanding options to either be assumed, continued or substituted by the acquirer or successor entity or, alternatively, to terminate upon the transaction. The plan administrator may also provide for the accelerated vesting of awards granted under the 2005 Plan.
Our board of directors may amend or terminate the 2005 Plan at any time. The 2005 Plan requires that certain amendments specified in the plan be submitted to stockholders for their approval.
2025 Equity Incentive Plan
We expect our board of directors to adopt a 2025 Equity Incentive Plan, or the 2025 Plan, prior to the consummation of this offering to provide an additional means through the grant of awards to attract, motivate, retain and reward selected employees and other eligible persons. We also intend to obtain approval of this plan from our stockholders prior to the consummation of this offering. The below summary of the 2025 Plan is what we expect the terms of the plan will be. Employees, officers, directors and consultants that provide services to us or one of our subsidiaries may be selected to receive awards under the 2025 Plan.
Our compensation committee will administer the 2025 Plan. The compensation committee has broad authority to:
| • | select participants and determine the types of awards that they are to receive; |
| • | determine the number of shares that are to be subject to awards and the terms and conditions of awards, including the price (if any) to be paid for the shares or the award and establish the vesting conditions (if applicable) of such shares or awards; |
| • | cancel, modify or waive our rights with respect to, or modify, discontinue, suspend or terminate any or all outstanding awards, subject to any required consents; |
| • | construe and interpret the terms of the 2025 Plan and any agreements relating to the plan; |
| • | accelerate or extend the vesting or exercisability or extend the term of any or all outstanding awards subject to any required consent; |
| • | subject to the other provisions of the 2025 Plan, make certain adjustments to an outstanding award and authorize the termination, conversion, substitution or succession of an award; and |
| • | allow the purchase price of an award or shares of our common stock to be paid in the form of cash, check or electronic funds transfer, by the delivery of previously-owned shares of our common stock or by a reduction of the number of shares deliverable pursuant to the award, by services rendered by the recipient of the award, by notice and third party payment or cashless exercise on such terms as the administrator may authorize or any other form permitted by law. |
129
Table of Contents
A total of shares of our common stock will initially be authorized for issuance with respect to awards granted under the 2025 Plan. Shares subject to outstanding awards granted under the 2005 Plan that are not paid, delivered or exercised before they expire or are canceled or terminated will be available for award grants under the 2025 Plan. In addition, the share limit will automatically increase on the first trading day in January of each year (commencing with 2026) by an amount equal to lesser of (1) % of the total number of shares of our common stock that are issued and outstanding on the last trading day in December in the prior year, or (2) such lesser number as determined by our board of directors. Any shares subject to awards that are not paid, delivered or exercised before they expire or are canceled or terminated, fail to vest, as well as shares used to pay the purchase or exercise price of awards or related tax withholding obligations, will become available for other award grants under the 2025 Plan. As of the date of this prospectus, no awards have been granted under the 2025 Plan, and the full number of shares currently authorized under the 2025 Plan is available for award purposes.
Awards under the 2025 Plan may be in the form of incentive or nonqualified stock options, stock appreciation rights, stock bonuses, restricted stock, stock units and other forms of awards including cash awards. Awards under the plan generally will not be transferable other than by will or the laws of descent and distribution, except that the plan administrator may authorize certain transfers.
Nonqualified and incentive stock options may not be granted at prices below the fair market value of the shares of our common stock on the date of grant. Incentive stock options must have an exercise price that is at least equal to the fair market value of our common stock, or 110% of fair market value of our common stock for incentive stock option grants to any 10% owner of our common stock, on the date of grant. The maximum term of options and stock appreciation rights granted under the plan is 10 years. These and other awards may also be issued solely or in part for services. Awards are generally paid in cash or shares of our common stock. The plan administrator may provide for the deferred payment of awards and may determine the terms applicable to deferrals.
As is customary in incentive plans of this nature, the number and type of shares available under the 2025 Plan and any outstanding awards, as well as the exercise or purchase prices of awards, will be subject to adjustment in the event of certain reorganizations, mergers, combinations, recapitalizations, stock splits, stock dividends or other similar events that change the number or kind of shares outstanding, and extraordinary dividends or distributions of property to the stockholders. In no case (except due to an adjustment referred to above or any repricing that may be approved by our stockholders) will any adjustment be made to a stock option or stock appreciation right award under the 2025 Plan (by amendment, cancellation and regrant, exchange or other means) that would constitute a repricing of the per-share exercise or base price of the award.
Generally, and subject to limited exceptions set forth in the 2025 Plan, if we dissolve or undergo certain corporate transactions such as a merger, business combination or other reorganization, or a sale of all or substantially all of its assets, all awards then-outstanding under the 2025 Plan will become fully vested or paid, as applicable, and will terminate or be terminated in such circumstances, unless the plan administrator provides for the assumption, substitution or other continuation of the award. The plan administrator also has the discretion to establish other change in control provisions with respect to awards granted under the 2025 Plan. For example, the administrator could provide for the acceleration of vesting or payment of an award in connection with a corporate event that is not described above and provide that any such acceleration shall be automatic upon the occurrence of any such event.
Our board of directors may amend or terminate the 2025 Plan at any time, but no such action will affect any outstanding award in any manner materially adverse to a participant without the consent of the participant. Plan amendments will be submitted to stockholders for their approval as required by applicable law or any applicable listing agency. The 2025 Plan is not exclusive—our board of directors and compensation committee may grant stock and performance incentives or other compensation, in stock or cash, under other plans or authority.
The plan will terminate on , 2035. However, the plan administrator will retain its authority until all outstanding awards are exercised or terminated.
130
Table of Contents
Defined Contribution Plans
As part of its overall compensation program, we provide all full-time employees, including each of the NEOs, with the opportunity to participate in a defined contribution 401(k) plan. The plan is intended to qualify under Section 401 of the Internal Revenue Code so that employee contributions and income earned on such contributions are not taxable to employees until withdrawn. Employees may elect to defer a percentage of their eligible compensation (not to exceed the statutorily prescribed annual limit) in the form of elective deferral contributions to the plan. The 401(k) plan also has a “catch-up contribution” feature for employees aged 50 or older (including those who qualify as “highly compensated” employees) who can defer amounts over the statutory limit that applies to all other employees. The Company does not currently make any matching or other contributions to participants’ accounts under the 401(k) plan.
DIRECTOR COMPENSATION
Two of our current non-employee directors (Eliyahou Harari and Chen Lung Tsai) will continue to serve on our board of directors following this offering. We are currently evaluating the compensation to be provided to our non-employee directors following this offering and have not yet determined the terms of our director compensation policy. Mr. Wang, who is employed by us, will also serve on our board following the closing.
Director Compensation Table — Fiscal Year 2024
The following table sets forth certain information concerning compensation paid to members of our board of directors who are not employed by us or any of our subsidiaries (“non-employee directors”) and who served on our board during the year ended December 31, 2024. The compensation for the year ended December 31, 2024 for Mr. Wang is reported in the “Summary Compensation Table—Fiscal Year 2024” above. Mr. Wang did not receive any compensation for his service on our board of directors during 2024.
| Name |
Fees Earned or Paid in Cash ($) |
Stock Awards ($)(1)(2)(3) |
Option Awards ($)(1)(2)(3) |
All Other Compensation ($) |
Total ($) |
|||||||||||||||
| Min-Liang Chen(6) |
— | — | — | — | — | |||||||||||||||
| Eliyahou Harari |
— | — | — | — | — | |||||||||||||||
| Hui Ying Kuo(6) |
— | — | — | — | — | |||||||||||||||
| Howard Lee(6) |
— | — | — | — | — | |||||||||||||||
| Heidi Chung Sutardja(6) |
— | — | — | — | — | |||||||||||||||
| Chen Lung Tsai |
— | — | — | 13,108 | (4) | 13,108 | ||||||||||||||
| Wen-Hung Tsai(5)(6) |
— | — | [ | ] | — | [ | ] | |||||||||||||
| Pen-Jung Tseng(6) |
— | — | — | — | — | |||||||||||||||
| (1) | In November 2024, the Company granted Mr. Wen-Hung Tsai an option to purchase 100,000 shares of our common stock at an exercise price of $0.17 per share. The option vests over four years, subject to Mr. Tsai’s continued services to the Company, and would vest in full upon a change in control of the Company. |
| (2) | The amounts reported in these columns reflect the grant date fair value of the stock and option awards granted to the non-employee directors during 2024 under the 2005 Plan described in note (1) above and are accounted for in accordance with FASB ASC Topic 718. Please see Note 10 to the Financial Statements included elsewhere in this prospectus for a discussion of the relevant assumptions used in calculating these amounts. |
131
Table of Contents
| (3) | As of December 31, 2024, the non-employee directors listed in the table above held outstanding Stock Awards and Option Awards with respect to the number of shares of our common stock set forth below: |
| Name |
Stock Awards (#) |
Option Awards (#) |
||||||
| Min-Liang Chen |
— | — | ||||||
| Eliyahou Harari |
— | — | ||||||
| Hui Ying Kuo |
— | — | ||||||
| Howard Lee |
— | — | ||||||
| Heidi Chung Sutardja |
— | — | ||||||
| Chen Lung Tsai |
— | — | ||||||
| Wen-Hung Tsai |
— | 100,000 | ||||||
| Pen-Jung Tseng |
— | — | ||||||
| (4) | The amount reported in this column for Mr. Chen Lung Tsai represents reimbursement of expenses for health insurance. |
| (5) | Mr. Wen-Hung Tsai resigned as a member of our board on February 26, 2024 and continues to serve as an advisor to the Company. The option grant described in note (1) above represents his sole compensation for these services. |
| (6) | It is currently anticipated that none of these directors will remain on our board upon the completion of this offering. |
132
Table of Contents
CERTAIN RELATIONSHIPS AND RELATED PARTY TRANSACTIONS
The following includes a summary of transactions since January 1, 2022 to which we have been a party in which (1) the amount involved exceeded or will exceed $120,000, and (2) any of our directors, executive officers, or holders of more than 5% of our capital stock, or any affiliate or member of the immediate family of the foregoing persons, had or will have a direct or indirect material interest, other than compensation and other arrangements that are described under the section titled “Executive and Director Compensation.”
Preferred Stock Financings
Series H Preferred Stock Financing. Since January 1, 2022, we issued and sold to investors in private placements an aggregate of 33,120,656 shares of our Series H preferred stock at a purchase price of $1.45 per share, for aggregate consideration of approximately $48.02 million.
The following table sets forth the aggregate number of shares of our capital stock acquired by beneficial owners of more than 5% of our capital stock in the financing transaction described above. Each share of our preferred stock identified in the following table will convert into shares of common stock immediately upon the closing of this offering.
|
Name(1) |
Series H Preferred Stock (#) |
Aggregate Cash Purchase Price ($) |
||||||
| Eliyahou Harari |
8,086,655 | $ | 11,725,649.18 | |||||
| Ching-Hang Shen |
12,997,319 | $ | 18,846,112.43 | |||||
| (1) | For additional information regarding these shareholders and their equity holdings, see “Security Ownership of Certain Beneficial Owners and Management.” |
Investors’ Rights Agreement
We entered into an amended and restated investor rights agreement on November 21, 2019 with the holders of our preferred stock, including holders of more than 5% of our capital stock and [entities with which certain of our directors are affiliated]. The agreement provides for certain rights relating to the registration of such holders’ common stock, including shares issuable upon conversion of preferred stock, and a right of first refusal to purchase future securities sold by us. See “Description of Capital Stock—Registration Rights” for additional information.
Voting Agreement
We entered into an amended and restated voting agreement on November 21, 2019 with certain holders of our common stock and preferred stock, including certain of our directors and executive officers, holders of more than 5% of our capital stock, and [entities with which certain of our directors are affiliated]. Upon the consummation of this offering, the amended and restated voting agreement will terminate.
Right of First Refusal and Co-Sale Agreement
We entered into an amended and restated right of first refusal and co-sale agreement on November 21, 2019 with certain holders of our common stock and preferred stock, including certain of our directors and executive officers, holders of more than 5% of our capital stock, and [entities with which certain of our directors are affiliated]. This agreement provides for rights of first refusal and co-sale relating to the shares of our common stock held by the parties to the agreement. Upon the consummation of this offering, the amended and restated right of first refusal and co-sale agreement will terminate.
133
Table of Contents
Executive Officer and Director Compensation
Please see the section titled “Executive and Director Compensation” for information regarding the compensation of our directors and executive officers.
Employment Agreements
We have entered into employment agreements with our named executive officers. For more information regarding the agreements with our named executive officers, see the section titled “Executive and Director Compensation—Executive Compensation Program.”
Indemnification Agreements
We intend to enter into indemnification agreements with each of our directors and executive officers. These agreements, among other things, require us or will require us to indemnify each director (and in certain cases their related venture capital funds) and executive officer to the fullest extent permitted by Delaware law, including indemnification of expenses such as attorneys’ fees, judgments, fines and settlement amounts incurred by the director or executive officer in any action or proceeding, including any action or proceeding by or in right of us, arising out of the person’s services as a director or executive officer. For further information, see the section titled “Management—Limitations on Liability and Indemnification Matters.”
Stock Option Grants to Executive Officers and Directors
We have granted stock options to our executive officers and certain of our directors as more fully described in the section entitled “Executive and Director Compensation.”
Other Transactions with Related Parties
Transactions with VeriSilicon
We entered into the ASIC Design, Manufacturing and Product Sales Agreement (“MSA”) with VerSilicon, Inc. (“VeriSilicon”) in May 2022. Pursuant to the MSA, we paid VeriSilicon for certain R&D and manufacturing services. VeriSilicon holds an insignificant percentage of our capital stock. The R&D expenses incurred in 2023 and 2024 were $962,836 and $[ ], respectively. As of December 31, 2023 and 2024, we have no payables due to VeriSilicon, respectively.
Policies and Procedures for Related Party Transactions
Our audit committee charter will provide that our audit committee will be responsible for reviewing and approving in advance any related party transaction. This will cover, with certain exceptions set forth in Item 404 of Regulation S-K under the Securities Act, any transaction, arrangement or relationship, or any series of similar transactions, arrangements or relationships in which we were or are to be a participant, where the amount involved exceeds $120,000 and a related person had or will have a direct or indirect material interest, including, without limitation, purchases of goods or services by or from the related person or entities in which the related person has a material interest, indebtedness, guarantees of indebtedness and employment by us of a related person. All of the transactions described in this section occurred prior to the creation of our audit committee and the adoption of this policy, and, as such, they were not conducted on an arms’ length basis.
134
Table of Contents
SECURITY OWNERSHIP OF CERTAIN BENEFICIAL OWNERS AND MANAGEMENT
The following table sets forth information, as of December 31, 2024, regarding beneficial ownership of our common stock by:
| • | each of our directors; |
| • | each of our executive officers; |
| • | all directors and executive officers as a group; and |
| • | each person, or group of affiliated persons, known by us to beneficially own more than five percent of our shares of common stock. |
The percentage ownership information under the column titled “Beneficial Ownership Prior to this Offering” is based on 135,719,248 shares of our common stock outstanding as of December 31, 2024, including 128,756,397 shares of our common stock resulting from the Preferred Stock Conversion, as if this conversion had occurred as of December 31, 2024. The percentage ownership information under the column titled “Beneficial Ownership After this Offering” assumes the foregoing and the issuance of shares of common stock in this offering and assumes no exercise of the underwriters’ option to purchase additional shares.
Beneficial ownership is determined according to the rules of the SEC, and generally means that person has beneficial ownership of a security if he or she possesses sole or shared voting or investment power of that security, and includes options that are currently exercisable or exercisable within 60 days. Each director or officer, as the case may be, has furnished us with information with respect to beneficial ownership. Except as otherwise indicated, we believe that the beneficial owners of common stock listed below, based on the information each of them has given to us, have sole investment and voting power with respect to their shares, except where community property laws may apply. Except as otherwise noted below, the address for each person or entity listed in the table is c/o CapsoVision, Inc, 18805 Cox Ave Suite 250 Saratoga, CA 95070.
| Shares Beneficially Owned Prior to This Offering |
Shares Beneficially Owned After This Offering |
|||||||||||||||||||||||
| Number of Outstanding Shares Beneficially Owned |
Number of Shares Exercisable Within 60 Days |
Number of Shares Beneficially Owned |
% of Beneficial Ownership |
Number of Shares Beneficially Owned |
% of Beneficial Ownership |
|||||||||||||||||||
| 5% and Greater Stockholders: |
||||||||||||||||||||||||
| Eliyahou Harari(1) |
14,340,669 | — | 14,340,669 | 10.6 | % | |||||||||||||||||||
| Ching-Hang Shen(2) |
12,997,319 | — | 12,997,319 | 9.6 | % | |||||||||||||||||||
| KIOXIA Corporation(3) |
11,478,261 | — | 11,478,261 | 8.5 | % | |||||||||||||||||||
| CID Greater China Venture Capital Fund III, L.P.(4) |
10,408,694 | — | 10,408,694 | 7.7 | % | |||||||||||||||||||
| Shih-Chang Stephen Wu(5) |
7,858,948 | — | 7,858,948 | 5.8 | % | |||||||||||||||||||
| Named Executive Officers and Directors: |
||||||||||||||||||||||||
| Kang-Huai (Johnny) Wang(6) |
2,188,172 | 209,792 | 2,397,964 | 1.8 | % | |||||||||||||||||||
| Kevin Lundquist |
— | — | — | — | ||||||||||||||||||||
| Rebecca Ann Petersen(7) |
— | 76,042 | 76,042 | * | ||||||||||||||||||||
| Douglas Patrick Atkinson(8) |
— | 295,000 | 295,000 | * | ||||||||||||||||||||
| Eliyahou Harari(1) |
14,340,669 | — | 14,340,669 | 10.6 | % | |||||||||||||||||||
| Chen Lung Tsai(9) |
247,490 | — | 247,490 | * | ||||||||||||||||||||
| Directors and executive officers as a group |
||||||||||||||||||||||||
135
Table of Contents
Notes:
| * | Represents less than 1% of our total outstanding shares on an as converted basis as of the date of this prospectus. |
| (1) | Consists of (i) 280,000 shares of common stock directly held by Eliyahou Harari; (ii) 222,222 shares of Series D-1 preferred stock, 934,588 shares of Series D-2 preferred stock, 434,782 shares of Series E preferred stock, 434,783 shares of Series F-1 preferred stock, 2,000,000 shares of Series F-2 preferred stock, 740,741 shares of Series G preferred stock, 1,206,898 shares of Series G-1 preferred stock and 4,638,377 shares of Series H preferred stock directly held by Harari Family Trust; (iii) 1,724,139 shares of Series H preferred stock directly held by Harari 2010 Children Remainder Trust – DAH; and (iv) 1,724,139 shares of Series H preferred stock directly held by Harari 2010 Children Remainder Trust – MHG (together with Harari Family Trust, Harari 2010 Children Remainder Trust – DAH, the “Harari Family Trusts”). Eliyahou Harari and Britt Harari, Eliyahou Harari’s wife, are trustees of Harari Family Trusts and have the voting power and investment power attached to and may be deemed as the beneficial owners of the shares of preferred stock held by the Harari Family Trusts. All shares of the preferred stock held by Harari Family Trusts will be converted into shares of common stock immediately prior to the completion of this offering. The business address of Mr. Eliyahou Harari is 225 Charcot, San Jose, California, U.S. |
| (2) | Consists of 12,997,319 shares of Series H preferred stock directly held by Ching-Hang Shen. All shares of the preferred stock held by Ching-Hang Shen will be converted into shares of common stock immediately prior to the completion of this offering. Mr. Ching-Hang Shen’s business address is No.112, Ln. 189, Sec. 2, Zhongshan N. Rd. Tamsui Dist. New Taipei City 251, Taiwan, R.O.C. |
| (3) | Consists of 11,478,261 shares of Series E preferred stock directly held by KIOXIA Corporation. KIOXIA Corporation is wholly-owned by KIOXIA Holdings Corporation, a company listed on the Tokyo Stock Exchange Prime Market, which may be deemed to have the beneficial ownership of the shares of the preferred stock directly held by KIOXIA Corporation. All shares of the preferred stock held by KIOXIA Corporation will be converted into shares of common stock immediately prior to the completion of this offering. The business address of KIOXIA Corporation is 1-21, Shibaura 3-Chome, Minato-ku, Tokyo 108-0023 Japan. |
| (4) | Consists of (i) 6,999,999 shares of Series D preferred stock; (ii) 869,565 shares of Series E preferred stock; (iii) 1,739,130 shares of Series F-1 preferred stock; and (iv) 800,000 shares of Series F-2 preferred stock directly held by CID Greater China Venture Capital Fund III, L.P., the general partner of which is CID Venture Capital General Partner III, Limited, which is wholly-owned by Altenza International Limited, which is controlled by Ching Yi Chang and Chih Cheng Chang. Each of these entities and individuals may be deemed to have the beneficial ownership of the shares of the preferred stock directly held CID Greater China Venture Capital Fund III, L.P. All shares of the preferred stock held by China Venture Capital Fund III, L.P. will be converted into shares of common stock immediately prior to the completion of this offering. The business address of CID Greater China Venture Capital Fund III, L.P. is Walker House, 87 Mary Street, George Town, Grand Cayman, KY1-9005 Cayman Islands. |
| (5) | Consists of (i) 150,000 shares of common stock; (ii) 2,222,138 shares of Series A preferred stock; (iii) 4,285,682 shares of Series B preferred stock; (iv) 1,034,462 shares of Series C preferred stock; and (v) 166,666 shares of Series D preferred stock directly held by Mr. Shih-Chang Stephen Wu. All shares of the preferred stock held by Mr. Shih-Chang Stephen Wu will be converted into shares of common stock immediately prior to the completion of this offering. The business address of Mr. Shih-Chang Stephen Wu is No. 498, Sec. 2, Bentian Road, Tainan, Taiwan, R.O.C. |
| (6) | Consists of (i) 700,000 shares of common stock; (ii) 1,407,407 shares of Series A preferred stock; (iii) 33,707 shares of Series C-1 preferred stock; (iv) 11,111 shares of Series D preferred stock; (v) 35,947 shares of Series H preferred stock directly held by Kang-Huai (Johnny) Wang; and (vi) 209,792 shares of common stock subject to options exercisable within 60 days of December 31, 2024. All shares of the preferred stock held by Kang-Huai (Johnny) Wang will be converted into shares of common stock immediately prior to the completion of this offering. |
| (7) | Consists of 76,042 shares of common stock subject to options exercisable within 60 days of December 31, 2024. |
136
Table of Contents
| (8) | Consists of 295,000 shares of common stock subject to options exercisable within 60 days of December 31, 2024. |
| (9) | Consists of (i) 70,000 shares of common stock, 86,956 shares of Series E preferred stock, 21,568 shares of Series H preferred stock directly held by Chen Lung Tsai; and (ii) 68,966 shares of Series H preferred stock directly held by The Tsai Family Trust. Chen Lung Tsai and Mei Man Tsai, Chen Lung Tsai’s wife, are trustees of The Tsai Family Trust and have the voting power and investment power attached to and may be deemed as the beneficial owners of the shares directly held by The Tsai Family Trust. All shares of the preferred stock held by Chen Lung Tsai and The Tsai Family Trust will be converted into shares of common stock immediately prior to the completion of this offering. |
137
Table of Contents
The following description of our capital stock and provisions of our amended and restated certificate of incorporation and amended and restated bylaws are summaries. You should also refer to the amended and restated certificate of incorporation, the amended and restated bylaws, and the amended and restated investors’ rights agreement, which are filed as exhibits to the registration statement of which this prospectus is a part.
General
Upon the completion of this offering and the filing of our amended and restated certificate of incorporation, our authorized capital stock will consist of shares of common stock, par value $0.001 per share, and shares of preferred stock, par value $0.001 per share.
Common Stock
Outstanding Shares
As of , we had shares of common stock outstanding, held of record by stockholders, after giving effect to the Preferred Stock Conversion.
Voting Rights
Each holder of common stock is entitled to one vote for each share on all matters submitted to a vote of the stockholders, including the election of directors. Our stockholders do not have cumulative voting rights in the election of directors. Accordingly, holders of a majority of the voting shares are able to elect all of the directors.
Dividends
Subject to preferences that may be applicable to any then outstanding preferred stock, holders of our common stock are entitled to receive ratably any dividends that our board of directors may declare out of funds legally available.
Liquidation
In the event of our liquidation, dissolution, or winding up, holders of our common stock will be entitled to share ratably in the net assets legally available for distribution to stockholders after the payment of all of our debts and other liabilities, subject to the satisfaction of any liquidation preference granted to the holders of any then outstanding shares of preferred stock.
Rights and Preferences
Holders of our common stock have no preemptive, conversion, or subscription rights, and there are no redemption or sinking-fund provisions applicable to our common stock. The rights, preferences, and privileges of the holders of our common stock are subject to, and may be adversely affected by, the rights of the holders of shares of any series of our preferred stock that we may designate and issue in the future.
Preferred Stock
Upon the completion of this offering, all of our currently outstanding shares of preferred stock will convert into common stock, and we will not have any preferred shares outstanding. Immediately prior to the completion of this offering, our certificate of incorporation will be amended and restated to delete all references to such shares of preferred stock. Under the amended and restated certificate of incorporation, our board of directors will
138
Table of Contents
have the authority, without further action by the stockholders, to issue up to shares of preferred stock in one or more series, to establish from time to time the number of shares to be included in each such series, to fix the rights, preferences, and privileges of the shares of each wholly unissued series and any qualifications, limitations, or restrictions thereon, and to increase or decrease the number of shares of any such series, but not below the number of shares of such series then outstanding.
Our board of directors may authorize the issuance of preferred stock with voting or conversion rights that could adversely affect the voting power or other rights of the holders of common stock. The issuance of preferred stock, while providing flexibility in connection with possible acquisitions and other corporate purposes, could, among other things, have the effect of delaying, deferring, or preventing a change in control of our Company that may otherwise benefit holders of our common stock and may adversely affect the market price of the common stock and the voting and other rights of the holders of common stock. We have no current plans to issue any shares of preferred stock.
Stock Options
As of , 2025, shares of common stock were issuable upon the exercise of outstanding stock options, at a weighted-average exercise price of $ per share. For additional information regarding terms of our equity incentive plans, see the section titled “Executive and Director Compensation—Equity Incentive Plans.”
Warrants
The following table sets forth information about outstanding warrants to purchase shares of our stock as of December 31, 2024.
| Class of Stock Underlying | Issue Date | Number of Shares of Common Stock Underlying Warrants |
Exercise Price Per Share |
Expiration Date | ||||||||||||
| Common Stock |
1/12/2021 | 50,000 | $ | 1.45 | January 12, 2026 | |||||||||||
|
|
|
|
|
|||||||||||||
Registration Rights
Upon the completion of this offering and subject to the lock-up agreements entered into in connection with this offering and federal securities laws, certain holders of shares of our common stock, including those shares of our common stock that will be issued upon the conversion of our preferred stock in connection with this offering, will initially be entitled to certain rights with respect to registration of such shares under the Securities Act. These shares are referred to as registrable securities. The holders of these registrable securities possess registration rights pursuant to the terms of our amended and restated investors’ rights agreement, as amended, and are described in additional detail below. The registration of shares of our common stock pursuant to the exercise of the registration rights described below would enable the holders to trade these shares without restriction under the Securities Act when the applicable registration statement is declared effective. We will pay the registration expenses, other than underwriting discounts, selling commissions, stock transfer taxes, fees and disbursements of more than one special counsel for the holders, and the compensation of regular employees of the company, of the shares registered pursuant to the demand, piggyback, and Form S-3 registrations described below.
Generally, in an underwritten offering, the managing underwriter, if any, has the right, subject to specified conditions and limitations, to limit the number of shares the holders may include. The demand, piggyback, and Form S-3 registration rights described below will terminate upon the earliest to occur of (1) the date five years after the consummation of this offering or (2) with respect to each stockholder, such time after the completion of this offering at which Rule 144 of the Securities Act (“Rule 144”) or another similar exemption under the Securities Act is available for the sale of all of such stockholder’s shares without limitation, during a three-month period without registration.
139
Table of Contents
Demand Registration Rights
Upon the completion of this offering, holders of up to approximately million shares of our common stock issuable upon conversion of our outstanding preferred stock will be entitled to certain demand registration rights. Beginning on the earlier of (i) November 21, 2024 and (ii) six months following the effectiveness of the registration statement of which this prospectus is a part, investors holding, collectively, not less than 50% of registrable securities may, on not more than two occasions, request that we register all or a portion of their shares, subject to certain specified exceptions. Such request for registration must cover securities the anticipated aggregate offering price of which is at least $5.0 million. If such holders exercise their demand registration rights, then holders of approximately million shares of our common stock issuable upon conversion of our outstanding preferred stock will be entitled to register their shares, subject to specified conditions and limitations in the corresponding offering.
Piggyback Registration Rights
In the event that we propose to register any of our securities under the Securities Act in another offering, either for our own account or for the account of other security holders, the holders of registrable securities will be entitled to certain “piggyback” registration rights allowing them to include their shares in such registration, subject to specified conditions and limitations.
S-3 Registration Rights
Upon the completion of this offering, the holders of up to approximately million shares of our common stock issuable upon conversion of our outstanding preferred stock will initially be entitled to certain Form S-3 registration rights. Any holder or holders of registrable securities may, with respect to not more than two such registrations within any 12-month period, request that we register all or a portion of their shares on Form S-3 if we are qualified to file a registration statement on Form S-3, subject to specified exceptions. Such request for registration on Form S-3 must cover securities with aggregate proceeds, net of underwriting discounts and expenses related to the issuance, which equal or exceed $1.0 million. The right to have such shares registered on Form S-3 is further subject to other specified conditions and limitations.
Anti-Takeover Provisions of Delaware Law and Our Amended and Restated Certificate of Incorporation and Amended and Restated Bylaws
Section 203 of the Delaware General Corporation Law
We are subject to Section 203 of the DGCL, which prohibits a Delaware corporation from engaging in any business combination with any interested stockholder for a period of three years after the date that such stockholder became an interested stockholder, with the following exceptions:
| • | before such date, the board of directors of the corporation approved either the business combination or the transaction that resulted in the stockholder becoming an interested stockholder; |
| • | upon completion of the transaction that resulted in the stockholder becoming an interested stockholder, the interested stockholder owned at least 85% of the voting stock of the corporation outstanding at the time the transaction began, excluding for purposes of determining the voting stock outstanding (but not the outstanding voting stock owned by the interested stockholder) those shares owned (i) by persons who are directors and also officers and (ii) employee stock plans in which employee participants do not have the right to determine confidentially whether shares held subject to the plan will be tendered in a tender or exchange offer; or |
| • | on or after such date, the business combination is approved by the board of directors and authorized at an annual or special meeting of the stockholders, and not by written consent, by the affirmative vote of at least % of the outstanding voting stock that is not owned by the interested stockholder. |
140
Table of Contents
In general, Section 203 defines a “business combination” to include the following:
| • | any merger or consolidation involving the corporation and the interested stockholder; |
| • | any sale, transfer, pledge, or other disposition of 10% or more of the assets of the corporation involving the interested stockholder; |
| • | subject to certain exceptions, any transaction that results in the issuance or transfer by the corporation of any stock of the corporation to the interested stockholder; |
| • | any transaction involving the corporation that has the effect of increasing the proportionate share of the stock or any class or series of the corporation beneficially owned by the interested stockholder; and |
| • | the receipt by the interested stockholder of the benefit of any loans, advances, guarantees, pledges, or other financial benefits by or through the corporation. |
In general, Section 203 defines an “interested stockholder” as an entity or person who, together with the person’s affiliates and associates, beneficially owns, or within three years prior to the time of determination of interested stockholder status did own, 15% or more of the outstanding voting stock of the corporation.
The statute could prohibit or delay mergers or other takeover or change in control attempts and, accordingly, may discourage attempts to acquire us even though such a transaction may offer our stockholders the opportunity to sell their stock at a price above the prevailing market price.
Amended and Restated Certificate of Incorporation and Amended and Restated Bylaws
Among other things, our amended and restated certificate of incorporation and amended and restated bylaws will:
| • | permit our board of directors to issue up to shares of preferred stock, with any rights, preferences, and privileges as they may designate; |
| • | provide that the authorized number of directors may be changed only by resolution of our board of directors; |
| • | provide that our board of directors will be classified into three classes of directors, divided as nearly as equal in number as possible; |
| • | provide that, subject to the rights of any series of preferred stock to elect directors, directors may only be removed for cause, which removal may be effected, subject to any limitation imposed by law, by the holders of at least % of the voting power of all of our then-outstanding shares of the capital stock entitled to vote generally at an election of directors; |
| • | provide that all vacancies, including newly created directorships, may, except as otherwise required by law, be filled by the affirmative vote of a majority of directors then in office, even if less than a quorum; |
| • | require that any action to be taken by our stockholders must be effected at a duly called annual or special meeting of stockholders and not be taken by written consent or electronic transmission; |
| • | provide that stockholders seeking to present proposals before a meeting of stockholders or to nominate candidates for election as directors at a meeting of stockholders must provide advance notice in writing and also specify requirements as to the form and content of a stockholder’s notice; |
| • | provide that special meetings of our stockholders may be called only by our board of directors pursuant to a resolution adopted by a majority of the total number of directors constituting the board, and not by our stockholders; and |
| • | not provide for cumulative voting rights, therefore allowing the holders of a majority of the shares of common stock entitled to vote in any election of directors to elect all of the directors standing for election, if they should so choose. |
141
Table of Contents
The amendment of any of these provisions would require approval by the holders of at least % of the voting power of all of our then-outstanding common stock.
The combination of these provisions will make it more difficult for our existing stockholders to replace our board of directors as well as for another party to obtain control of us by replacing our board of directors. Because our board of directors has the power to retain and discharge our officers, these provisions could also make it more difficult for existing stockholders or another party to effect a change in management. In addition, the authorization of undesignated preferred stock makes it possible for our board of directors to issue preferred stock with voting or other rights or preferences that could impede the success of any attempt to effect a change in control of our Company.
These provisions are intended to enhance the likelihood of continued stability in the composition of our board of directors and its policies and to discourage coercive takeover practices and inadequate takeover bids. These provisions are also designed to reduce our vulnerability to hostile takeovers and to discourage certain tactics that may be used in proxy fights. However, such provisions could have the effect of discouraging others from making tender offers for our shares and may have the effect of delaying changes in control or management of our Company. As a consequence, these provisions may also inhibit fluctuations in the market price of our stock that could result from actual or rumored takeover attempts. We believe that the benefits of these provisions, including increased protection of our potential ability to negotiate with the proponent of an unfriendly or unsolicited proposal to acquire or restructure our Company, outweigh the disadvantages of discouraging takeover proposals because negotiation of takeover proposals could result in an improvement of their terms.
Choice of Forum
Our amended and restated certificate of incorporation will provide that, unless we consent in writing to the selection of an alternative forum, the Court of Chancery of the State of Delaware (or, in the event that the Court of Chancery does not have jurisdiction, the federal district court for the District of Delaware or other state courts of the State of Delaware) is the exclusive forum for any derivative action or proceeding brought on our behalf; any action asserting a claim of breach of fiduciary duty; any action asserting a claim against us arising pursuant to the DGCL, our amended and restated certificate of incorporation, or our amended and restated bylaws (as either may be amended from time to time); or any action asserting a claim against us that is governed by the internal affairs doctrine; provided that, the exclusive forum provision will not apply to suits brought to enforce any liability or duty created by the Exchange Act or any other claim for which the federal courts have exclusive jurisdiction; and provided further that, if and only if the Court of Chancery of the State of Delaware dismisses any such action for lack of subject matter jurisdiction, such action may be brought in another state or federal court sitting in the State of Delaware. Our amended and restated certificate of incorporation will also provide that the federal district courts of the United States of America will be the exclusive forum for the resolution of any complaint asserting a cause of action against us or any of our directors, officers, employees, or agents and arising under the Securities Act. Nothing in our amended and restated certificate of incorporation or amended and restated bylaws precludes stockholders that assert claims under the Exchange Act from bringing such claims in state or federal court, subject to applicable law.
If any Foreign Action is filed in a court other than a court located within the State of Delaware, in the name of any stockholder, such stockholder shall be deemed to have consented to the personal jurisdiction of the state and federal courts located within the State of Delaware in connection with any action brought in any such court to enforce the applicable provisions of our amended and restated certificate of incorporation and having service of process made upon such stockholder in any such action by service upon such stockholder’s counsel in the Foreign Action as agent for such stockholder. Although our amended and restated certificate of incorporation will contain the choice of forum provision described above, it is possible that a court could find that such a provision is inapplicable for a particular claim or action or that such provision is unenforceable.
This choice of forum provision may limit a stockholder’s ability to bring a claim in a judicial forum that it finds favorable for disputes with us or any of our directors, officers, other employees, or stockholders, which
142
Table of Contents
may discourage lawsuits with respect to such claims, although our stockholders will not be deemed to have waived our compliance with federal securities laws and the rules and regulations thereunder. Furthermore, the enforceability of similar choice of forum provisions in other companies’ certificates of incorporation has been challenged in legal proceedings, and it is possible that a court could find these types of provisions to be inapplicable or unenforceable.
Limitations on Liability and Indemnification
For a discussion of liability and indemnification, see the section titled “Management—Limitations on Liability and Indemnification Matters.”
Listing
In connection with this offering, we intend to apply to list our common stock on Nasdaq under the trading symbol “CV.”
Transfer Agent and Registrar
Upon completion of this offering, the transfer agent and registrar for our common stock will be . The transfer agent and registrar’s address is .
143
Table of Contents
SHARES ELIGIBLE FOR FUTURE SALE
Future sales of substantial amounts of common stock in the public market after this offering could adversely affect market prices prevailing from time to time and could impair our ability to raise capital through the sale of our equity securities. We are unable to estimate the number of shares of common stock that may be sold in the future.
Upon the closing of this offering, we will have:
| • | shares of common stock outstanding; and |
| • | shares of common stock issuable upon exercise of warrants to be issued to the representative of the underwriters in connection with this offering. |
All of the shares sold in this offering will be freely tradable without restriction under the Securities Act unless purchased by one of our affiliates as that term is defined in Rule 144 under the Securities Act, which generally includes directors, officers or 10% stockholders. None of the holders of shares of our common stock or securities exercisable for or convertible into shares of our common stock have any registration rights.
Lock-Up
Our directors, officers and holders (but limited to such holders that hold an aggregate of not more than 95%) of our outstanding shares of common stock have agreed not to offer, sell, dispose of or hedge any shares of our common stock, subject to specified limited exceptions, during the period continuing through the date that is 6 months after the date of closing of this offering.
Rule 144
Shares of common stock held by any of our affiliates, as that term is defined in Rule 144 of the Securities Act, as well as shares held by our current stockholders, may be resold only pursuant to further registration under the Securities Act or in transactions that are exempt from registration under the Securities Act. In general, under Rule 144 as currently in effect, any person who is or has been our affiliate during the 90 days immediately preceding the sale and who has beneficially owned shares for at least six months is entitled to sell, within any three-month period commencing 90 days after the date of this prospectus, a number of shares that does not exceed the greater of: (i) 1% of the number of shares of common stock then outstanding, or (ii) the average weekly trading volume of the common stock during the four calendar weeks preceding the filing of a Form 144 with respect to the sale.
Sales under Rule 144 by our affiliates will also be subject to manner of sale provisions and notice requirements and to the availability of current public information about us.
Regulation S
Regulation S under the Securities Act provides that securities owned by any person may be sold without registration in the United States, provided that the sale is effected in an “offshore transaction” and no “directed selling efforts” are made in the United States (as these terms are defined in Regulation S) and subject to certain other conditions. In general, this means that our shares of common stock may be sold in some manner outside the United States without requiring registration in the United States.
Rule 701
In general, under Rule 701 as in effect on the date of this prospectus, any of our employees, directors, officers, consultants, or advisors who purchased shares from us in reliance on Rule 701 in connection with a
144
Table of Contents
compensatory stock or option plan or other written agreement before the effective date of this offering, or who purchase shares from us after that date upon the exercise of options granted before that date, are eligible to resell such shares 90 days after the effective date of this offering in reliance upon Rule 144. If such person is not an affiliate, such sale may be made subject only to current public information provisions of Rule 144.
If such a person is an affiliate, such sale may be made under Rule 144 without compliance with the holding period requirement, but subject to the other Rule 144 restrictions described above.
145
Table of Contents
MATERIAL U.S. FEDERAL INCOME TAX CONSEQUENCES FOR NON-U.S. HOLDERS
The following is a summary of material U.S. federal income tax consequences to non-U.S. holders (as defined below) of the purchase, ownership and disposition of our common stock issued pursuant to this offering. This discussion is not a complete analysis of all potential U.S. federal income tax consequences relating thereto, does not address the special tax accounting rules under Section 451(b) of the Internal Revenue Code of 1986, as amended (the “Code”), or any U.S. federal non-income tax consequences, such as estate or gift tax consequences, or any tax consequences arising under any state, local or foreign tax laws. This discussion is based on the Code, Treasury Regulations promulgated thereunder, judicial decisions, and published rulings and administrative pronouncements of the Internal Revenue Service (“IRS”) all as in effect on the date hereof. These authorities are subject to differing interpretations and may change, possibly retroactively, resulting in U.S. federal income tax consequences different from those discussed below. We have not requested a ruling from the IRS with respect to the statements made and the conclusions reached in the following summary, and there can be no assurance that the IRS or a court will agree with such statements and conclusions.
This discussion is limited to non-U.S. holders who purchase our common stock pursuant to this offering and who hold our common stock as a “capital asset” within the meaning of Section 1221 of the Code (generally, property held for investment). This discussion does not address all of the U.S. federal income tax consequences that may be relevant to a non-U.S. holder in light of such non-U.S. holder’s particular circumstances. This discussion also does not consider any specific facts or circumstances that may be relevant to non-U.S. holders subject to special rules under the U.S. federal income tax laws, including:
| • | certain former citizens or long-term residents of the United States; |
| • | partnerships or other entities or arrangements treated as pass-through or disregarded entities for U.S. federal income tax purposes (and investors therein); |
| • | “controlled foreign corporations” as defined in Section 957 of the Code; |
| • | “passive foreign investment companies” as defined in Section 1297 of the Code; |
| • | corporations that accumulate earnings to avoid U.S. federal income tax; |
| • | banks, financial institutions, investment funds, insurance companies, brokers, dealers or traders in securities; |
| • | tax-exempt organizations and governmental organizations; |
| • | tax-qualified retirement plans; |
| • | persons who acquire our common stock through the exercise of an option or otherwise as compensation; |
| • | “qualified foreign pension funds” as defined in Section 897(l)(2) of the Code and entities all of the interests of which are held by qualified foreign pension funds; |
| • | persons that own or have owned, actually or constructively, more than 5% of our common stock; |
| • | persons who have elected to mark securities to market; and |
| • | persons holding our common stock as part of a hedging or conversion transaction or straddle, or synthetic security or a constructive sale, or other risk reduction strategy or integrated investment. |
If an entity or arrangement that is classified as a partnership for U.S. federal income tax purposes holds our common stock, the U.S. federal income tax treatment of a partner in the partnership will generally depend on the status of the partner, the activities of the partnership and certain determinations made at the partner level. Partnerships holding our common stock and the partners in such partnerships are urged to consult their tax advisors about the particular U.S. federal income tax consequences to them of holding and disposing of our common stock.
146
Table of Contents
THIS DISCUSSION IS FOR INFORMATIONAL PURPOSES ONLY AND IS NOT TAX ADVICE. PROSPECTIVE INVESTORS SHOULD CONSULT THEIR TAX ADVISORS REGARDING THE PARTICULAR U.S. FEDERAL INCOME TAX CONSEQUENCES TO THEM OF ACQUIRING, OWNING AND DISPOSING OF OUR COMMON STOCK, AS WELL AS ANY TAX CONSEQUENCES ARISING UNDER ANY STATE, LOCAL OR FOREIGN TAX LAWS AND ANY OTHER U.S. FEDERAL TAX LAWS OR UNDER ANY APPLICABLE INCOME TAX TREATY.
Definition of Non-U.S. Holder
For purposes of this discussion, a non-U.S. holder is any beneficial owner of our common stock that is not a “U.S. holder” or a partnership (including any entity or arrangement treated as a partnership) for U.S. federal income tax purposes. A U.S. holder is any person that, for U.S. federal income tax purposes, is or is treated as any of the following:
| • | an individual who is a citizen or resident of the United States; |
| • | a corporation created or organized under the laws of the United States, any state thereof or the District of Columbia; |
| • | an estate, the income of which is subject to U.S. federal income tax regardless of its source; or |
| • | a trust (1) whose administration is subject to the primary supervision of a U.S. court and which has one or more U.S. persons who have the authority to control all substantial decisions of the trust or (2) that has a valid election in effect under applicable Treasury Regulations to be treated as a U.S. person. |
Distributions on Our Common Stock
If we distribute cash or other property to holders of shares of our common stock, such distributions will constitute dividends for U.S. federal income tax purposes to the extent such distribution is made from our current or accumulated earnings and profits, as determined under U.S. federal income tax principles. Amounts not treated as dividends for U.S. federal income tax purposes will constitute a return of capital and will first be applied against and reduce a holder’s tax basis in our common stock, but not below zero. Any excess will be treated as gain realized on the sale or other disposition of our common stock and will be treated as described under “ Gain on Disposition of Our Common Stock” below.
Subject to the discussion below regarding effectively connected income, backup withholding and FATCA (as defined below), dividends paid to a non-U.S. holder of shares of our common stock generally will be subject to U.S. federal withholding tax at a rate of 30% of the gross amount of the dividends or such lower rate specified by an applicable income tax treaty. To receive the benefit of a reduced treaty rate, a non-U.S. holder must furnish us or our withholding agent with a valid IRS Form W-8BEN (in the case of individuals) or IRS Form W-8BEN-E (in the case of entities), or other appropriate form, certifying such holder’s qualification for the reduced rate. This certification must be provided to us or our withholding agent before the payment of the dividends and must be updated periodically. In the case of a non-U.S. Holder that is an entity, Treasury Regulations and the relevant tax treaty provide rules to determine whether, for purposes of determining the applicability of the tax treaty, dividends will be treated as paid to the entity or to those holding an interest in the entity. If the non-U.S. holder holds our common stock through a financial institution or other agent acting on the non-U.S. holder’s behalf, the non-U.S. holder will be required to provide appropriate documentation to the agent, which then will be required to provide certification to us or our withholding agent, either directly or through other intermediaries.
If a non-U.S. holder holds shares of our common stock in connection with the conduct of a trade or business in the United States, and dividends paid on our common stock are effectively connected with such holder’s U.S. trade or business (and are attributable to such holder’s permanent establishment or fixed base in the United States if required by an applicable tax treaty), the non-U.S. holder will be exempt from U.S. federal withholding tax. To claim the exemption, the non-U.S. holder must generally furnish a valid IRS Form W-8ECI (or applicable successor form) to the applicable withholding agent.
147
Table of Contents
However, any such effectively connected dividends generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a foreign corporation also may be subject to an additional branch profits tax equal to 30% (or such lower rate specified by an applicable income tax treaty) of its effectively connected dividends, as adjusted for certain items.
Non-U.S. holders that do not provide the required certification on a timely basis, but that qualify for a reduced treaty rate, may obtain a refund of any excess amounts withheld by timely filing an appropriate claim for refund with the IRS. Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
Gain on Disposition of Our Common Stock
Subject to the discussion below regarding backup withholding and FATCA, a non-U.S. holder generally will not be subject to U.S. federal income tax on any gain realized on the sale or other disposition of our common stock, unless:
| • | the gain is effectively connected with the non-U.S. holder’s conduct of a trade or business in the United States and, if required by an applicable income tax treaty, is attributable to a permanent establishment or fixed base maintained by the non-U.S. holder in the United States; |
| • | the non-U.S. holder is a nonresident alien individual present in the United States for 183 days or more during the taxable year of the disposition, and certain other requirements are met; or |
| • | our common stock constitutes a “United States real property interest” by reason of our status as a United States real property holding corporation (“USRPHC”) for U.S. federal income tax purposes at any time within the shorter of the five-year period preceding the disposition or the non-U.S. holder’s holding period for our common stock, and our common stock is not “regularly traded” on an established securities market during the calendar year in which the sale or other disposition occurs. |
Gain described in the first bullet point above generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a foreign corporation also may be subject to an additional branch profits tax equal to 30% (or such lower rate specified by an applicable income tax treaty) of its effectively connected earnings and profits for the taxable year, as adjusted for certain items.
Gain described in the second bullet point above will be subject to U.S. federal income tax at a flat 30% rate (or such lower rate specified by an applicable income tax treaty), but may be offset by certain U.S.- source capital losses (even though the individual is not considered a resident of the United States), provided that the non-U.S. holder has timely filed U.S. federal income tax returns with respect to such losses.
Determining whether we are a USRPHC depends on the fair market value of our U.S. real property interests relative to the fair market value of our other trade or business assets and our foreign real property interests. We believe that we are not currently, and we do not anticipate becoming, a USRPHC for U.S. federal income tax purposes, although there can be no assurance we will not in the future become a USRPHC. Even if we are treated as a USRPHC, gain realized by a non-U.S. Holder on a disposition of our common stock will not be subject to U.S. federal income tax so long as (1) the non-U.S. Holder owned, directly, indirectly and constructively, no more than 5% of our common stock at all times within the shorter of (a) the five-year period preceding the disposition or (b) the holder’s holding period and (2) our common stock is regularly traded on an established securities market within the meaning of applicable U.S. Treasury regulations. There can be no assurance that our common stock qualifies as regularly traded on an established securities market. If any gain on a non-U.S. holder’s disposition of our common stock is taxable because we are a USRPHC and such holder’s ownership of our common stock exceeds 5%, such holder will be taxed on such disposition generally in the manner applicable to U.S. persons and in addition, a purchaser of such holder’s common stock may be required to withhold tax with respect to that obligation.
148
Table of Contents
Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
Information Reporting and Backup Withholding
Annual reports are required to be filed with the IRS and provided to each non-U.S. holder indicating the amount of distributions on our common stock paid to such holder and the amount of any tax withheld with respect to those distributions. These information reporting requirements apply even if no withholding was required because the distributions were effectively connected with the holder’s conduct of a U.S. trade or business, or withholding was reduced or eliminated by an applicable income tax treaty. This information also may be made available under a specific treaty or agreement with the tax authorities in the country in which the non-U.S. holder resides or is established. Backup withholding, currently at a 24% rate, generally will not apply to payments to a non-U.S. holder of dividends on or the gross proceeds of a disposition of our common stock provided the non-U.S. holder furnishes the required certification for its non-U.S. status, such as by providing a valid IRS Form W-8BEN, IRS Form W-8BEN-E or IRS Form W-8ECI, or otherwise establishes an exemption, and if the payor does not have actual knowledge, or reason to know, that the holder is a U.S. person who is not an exempt recipient.
Backup withholding is not an additional tax. If any amount is withheld under the backup withholding rules, the non-U.S. holder should consult with a U.S. tax advisor regarding the possibility of and procedure for obtaining a refund or a credit against the non-U.S. holder’s U.S. federal income tax liability, if any.
Withholding on Foreign Entities
Sections 1471 through 1474 of the Code, which are commonly referred to as FATCA, impose a U.S. federal withholding tax of 30% on certain payments made to a “foreign financial institution” (as specially defined under these rules) unless such institution enters into an agreement with the U.S. government to withhold on certain payments and to collect and provide to the U.S. tax authorities substantial information regarding certain U.S. account holders of such institution (which includes certain equity and debt holders of such institution, as well as certain account holders that are foreign entities with U.S. owners) or an exemption applies. FATCA also generally imposes a U.S. federal withholding tax of 30% on certain payments made to a “non-financial foreign entity” (as specially defined under these rules) unless such entity provides the withholding agent a certification that it does not have any “substantial United States owners” or provides information identifying certain direct and indirect U.S. owners of the entity or an exemption applies. An intergovernmental agreement between the United States and an applicable foreign country may modify these requirements. Under certain circumstances, a non-U.S. holder might be eligible for refunds or credits of such taxes. FATCA currently applies to dividends paid on our common stock and would have applied also to payments of gross proceeds from the sale or other disposition of our common stock. However, the U.S. Treasury Department has released proposed regulations under FATCA providing for the elimination of the federal withholding tax of 30% applicable to gross proceeds of a sale or other disposition of from property of a type that can produce U.S. source dividends or interest. Under these proposed Treasury Regulations (which may be relied upon by taxpayers prior to finalization), FATCA will not apply to gross proceeds from sales or other dispositions of our common stock.
Prospective investors are encouraged to consult with their tax advisors regarding the possible implications of FATCA on their investment in our common stock.
EACH PROSPECTIVE INVESTOR SHOULD CONSULT ITS TAX ADVISOR REGARDING THE TAX CONSEQUENCES OF PURCHASING, HOLDING AND DISPOSING OF OUR COMMON STOCK, INCLUDING THE CONSEQUENCES OF ANY RECENT AND PROPOSED CHANGE IN APPLICABLE LAW, AS WELL AS TAX CONSEQUENCES ARISING UNDER ANY STATE, LOCAL, NON-U.S. OR U.S. FEDERAL NON-INCOME TAX LAWS.
149
Table of Contents
The Benchmark Company, LLC, is acting as the representative of the underwriters (“Benchmark” or the “Representative”) and the book-running manager, with respect to the shares sold in this offering. We will enter into an underwriting agreement with the Representative. Subject to the terms and conditions of the underwriting agreement, we have agreed to sell to the underwriters named below, and the underwriters have agreed, severally and not jointly, to purchase from us the number of shares of common stock set forth opposite the underwriter’s name in the following table at the initial public offering price less the underwriting discounts set forth in the cover page of this prospectus:
| Underwriter |
Number of Shares |
|||
| The Benchmark Company, LLC |
||||
|
|
|
|||
| TOTAL |
||||
|
|
|
|||
The underwriters have committed to purchase all of the shares offered by us other than those shares covered by the over-allotment option described below, if they purchase any shares. The obligations of the underwriters may be terminated upon the occurrence of certain events specified in the underwriting agreement. Furthermore, pursuant to the underwriting agreement, the underwriters’ obligations are subject to customary conditions, representations and warranties contained in the underwriting agreement, such as receipt by the underwriters of officers’ certificates and legal opinions.
We have agreed to indemnify the underwriters against specified liabilities, including liabilities under the Securities Act, or to contribute to payments the underwriters may be required to make in respect of those liabilities.
The underwriters are offering the shares, subject to prior sale, when, as and if issued to and accepted by them, subject to approval of legal matters by their counsel and other conditions contained in the underwriting agreement. The underwriters reserve the right to withdraw, cancel or modify offers to the public and to reject orders in whole or in part.
Over-Allotment Option
We have granted the underwriters an option to purchase from us, at the initial public offering price less the underwriting discounts and commissions, up to an additional shares of our common stock, solely to cover over-allotments, if any. The underwriters may exercise this option, in whole or in part, for our common stock, any time during the 30-day period from the date of the closing of this offering. If this option is exercised in full, the total price to the public will be $ and the total net proceeds before expenses to us will be $ .
Underwriting Discount, Commissions and Expenses
The following table shows the per share and total underwriting discounts and commissions to be paid to the underwriters. These amounts are shown assuming both no exercise and full exercise of the underwriters’ option to purchase additional shares of our common stock, assuming an initial public offering price of $ per share, the midpoint of the estimated price range set forth on the cover page of this prospectus.
| Total | ||||||||||||
| Per Share | No Exercise | Full Exercise | ||||||||||
| Initial public offering price(1) |
$ | $ | $ | |||||||||
| Underwriting discounts and commissions(2) |
$ | $ | $ | |||||||||
| Proceeds, before expenses, to us |
$ | $ | $ | |||||||||
150
Table of Contents
| (1) | Assuming an initial public offering price is $ , the midpoint of the range set forth on the cover page of this prospectus. |
| (2) | Represents underwriting discounts equal to (i) 7% per share, which is the underwriting discounts we have agreed to pay on investors in this offering introduced by the underwriters; and (ii) 5% per share, which is the underwriting discounts we have agreed to pay on investors in this offering introduced by us. For purpose of the calculation only, we assume 100% investors in this offering are introduced by the underwriters. |
The Representative has advised us that the underwriters propose initially to offer the shares to the public at the initial public offering price set forth on the cover page of this prospectus and to dealers at that price less a concession not in excess of $ per share. After the initial public offering, the public offering price, concession and discount may be changed.
We have agreed to pay all of the reasonable, documented, necessary and accountable out-of-pocket expenses relating to the offering, including the Representative’s out-of-pocket and accountable expenses up to a maximum aggregate allowance of $182,500 (including, but not limited to, the fees and expenses of underwriters’ legal counsel up to $150,000, book-building costs, “road show” expenses and background checks on our senior management in an amount not to exceed $7,500).
We have paid a $25,000 expense advance to the Representative, which shall be applied against actual out-of-pocket-accountable expenses, which will be returned to us to the extent such out-of-pocket accountable expenses are not actually incurred in accordance with FINRA Rule 5110(g)(4)(A).
We estimate that the total expenses of the offering payable by us, excluding the total underwriting discount, will be approximately $ .
Discretionary Accounts
The underwriters do not intend to confirm sales of the shares offered hereby to any accounts over which they have discretionary authority.
Electronic Distribution
A prospectus in electronic format may be made available on the websites maintained by one or more of the underwriters or selling group members, if any, participating in the offering. Benchmark may allocate a number of shares to the underwriters and selling group members, if any, for sale to their online brokerage account holders. Any such allocations for online distributions will be made by Benchmark on the same basis as other allocations.
Representative’s Warrants
We have agreed to issue to the Representative or its designees at the closing of this offering warrants to purchase the number of common stock equal to 3% of the aggregate number of shares sold in this offering. The warrants will be exercisable at any time and from time to time, in whole or in part, commencing six months after the closing of this offering and may be exercised on a cashless basis. The warrants will be exercisable at a per share price equal to 125% of the initial public offering price per share in the offering. The warrants will terminate five years from the commencement of sales of this offering.
The warrants have been deemed compensation by FINRA and are therefore subject to a 180-day lock-up pursuant to FINRA Rule 5110(e)(1). The Representative warrants will not be sold, transferred, assigned, pledged, or hypothecated or the subject of any hedging, short sale, derivative, put, or call transaction that would result in the effective economic disposition of the warrants or the underlying securities for a period of 180 days from the commencement of sales of this offering except as permitted pursuant to FINRA Rule 5110(e)(2). The shares of common stock underlying the warrants are being registered as a part of the registration statement of which this prospectus forms a part and will be freely tradable upon exercise after the expiration of the FINRA lock-up.
151
Table of Contents
The exercise price and number of shares of common stock issuable upon exercise of the warrants may be adjusted in certain circumstances including in the event of a stock dividend, extraordinary cash dividend or recapitalization, reorganization, merger or consolidation.
Lock-Up Agreements
Our officers and directors, and holders (but limited to such holders that hold an aggregate of not more than 95%) of our outstanding shares of common stock have agreed not to, without the prior written consent of the underwriters, directly or indirectly, offer to sell, sell or otherwise transfer or dispose of any shares of our common stock (or enter into any transaction or device that is designed to, or could be expected to, result in the transfer or disposition by any person at any time in the future of shares of our common stock), enter into any swap or other derivatives transaction that transfers to another, in whole or in part, any of the economic benefits or risks of ownership of shares of common stock, make any demand for or exercise any right or cause to be filed a registration statement, including any amendments thereto, with respect to the registration of any of the shares of common stock or securities convertible into or exercisable or exchangeable for shares of common stock or any other of our securities or publicly disclose the intention to do any of the foregoing, subject to customary exceptions, for a period of 6 months from the date of closing of this offering.
Securities Issuance Standstill
Unless otherwise agreed by the underwriters of this offering, we have agreed that for a period of six (6) months from the closing of this offering, each of us and our successors will not (a) offer, sell, or otherwise transfer or dispose of, directly or indirectly, any shares of our capital stock or any securities convertible into or exercisable or exchangeable for shares of our capital stock; or (b) file or caused to be filed any registration statement with the SEC relating to the offering of any shares of our capital stock or any securities convertible into or exercisable or exchangeable for shares of our capital stock.
Right of First Refusal
We have granted the Representative the right to act as lead or joint-lead investment banker, lead or joint book-runner and/or lead or joint placement agent, for any of our future public and private equity and debt offerings, including all equity linked financings, during the twelve (12) month period following the completion of this initial public offering in compliance with FINRA Rule 5110(g)(6); provided, however, if the underwriting agreement is terminated for cause by the Company, the right of first refusal shall be terminated as provided in FINRA Rule 5110(g)(5)(B).
Determination of Offering Price
The public offering price was negotiated between Benchmark and us. In determining the public offering price of our common stock, Benchmark considered:
| • | the history and prospects for the industry in which we compete; |
| • | our financial information; |
| • | the ability of our management and our business potential and earning prospects; |
| • | the prevailing securities markets at the time of this offering; and |
| • | the recent market prices of, and the demand for, publicly traded shares of generally comparable companies, as well as the recent market price of our Company’s common stock. |
152
Table of Contents
Stabilization
In connection with this offering, the underwriter may engage in stabilizing transactions, over-allotment transactions, syndicate-covering transactions, penalty bids, and purchases to cover positions created by short sales.
| • | Stabilizing transactions permit bids to purchase securities so long as the stabilizing bids do not exceed a specified maximum and are engaged in for the purpose of preventing or retarding a decline in the market price of the securities while the offering is in progress. |
| • | Over-allotment transactions involve sales by the underwriter of securities in excess of the number of securities the underwriter is obligated to purchase. This creates a syndicate short position which may be either a covered short position or a naked short position. In a covered short position, the number of securities over-allotted by the underwriter is not greater than the number of securities that they may purchase in the over-allotment option. In a naked short position, the number of securities involved is greater than the number of securities in the over-allotment option. The underwriter may close out any short position by exercising their over-allotment option and/or purchasing securities in the open market. |
| • | Syndicate covering transactions involves purchases of securities in the open market after the distribution has been completed in order to cover syndicate short positions. In determining the source of securities to close out the short position, the underwriter will consider, among other things, the price of securities available for purchase in the open market as compared with the price at which they may purchase securities through exercise of the over-allotment option. If the underwriter sells more securities than could be covered by exercise of the over-allotment option and, therefore, have a naked short position, the position can be closed out only by buying securities in the open market. A naked short position is more likely to be created if the underwriter is concerned that after pricing there could be downward pressure on the price of the securities in the open market that could adversely affect investors who purchase in the offering. |
| • | Penalty bids permit the underwriter to reclaim a selling concession from a syndicate member when the securities originally sold by that syndicate member are purchased in stabilizing or syndicate covering transactions to cover syndicate short positions. |
These stabilizing transactions, over-allotment transactions, syndicate covering transactions, and penalty bids may have the effect of raising or maintaining the market price of our securities or preventing or retarding a decline in the market price of our securities. As a result, the price of our securities in the open market may be higher than it would otherwise be in the absence of these transactions. Neither we nor the underwriter make any representation or prediction as to the effect that the transactions described above may have on the price of our securities. These transactions may be affected on the Nasdaq Stock Market, in the over-the-counter market or otherwise and, if commenced, may be discontinued at any time.
Passive Market Making
In connection with this offering, underwriter, and selling group members may engage in passive market making transactions in our securities on the Nasdaq Stock Market in accordance with Rule 103 of Regulation M under the Exchange Act, during a period before the commencement of offers or sales of the shares and extending through the completion of the distribution. A passive market maker must display its bid at a price not in excess of the highest independent bid of that security. However, if all independent bids are lowered below the passive market maker’s bid, then that bid must then be lowered when specified purchase limits are exceeded.
Other Relationships
The underwriters and their respective affiliates are full service financial institutions engaged in various activities, which may include sales and trading, commercial and investment banking, advisory, investment
153
Table of Contents
management, investment research, principal investment, hedging, market making, brokerage and other financial and non-financial activities and services. Certain of the underwriters and their respective affiliates have provided, and may in the future provide, a variety of these services to us and to persons and entities with relationships with us, for which they received or will receive customary fees and expenses.
154
Table of Contents
SELLING RESTRICTIONS
Other than in the United States, no action has been taken by us or the underwriter that would permit a public offering of the securities offered by this prospectus in any jurisdiction where action for that purpose is required. The securities offered by this prospectus may not be offered or sold, directly or indirectly, nor may this prospectus or any other offering material or advertisements in connection with the offer and sale of any such securities be distributed or published in any jurisdiction, except under circumstances that will result in compliance with the applicable rules and regulations of that jurisdiction. Persons who come into possession of this prospectus are advised to inform themselves about and to observe any restrictions relating to the offering and the distribution of this prospectus. This prospectus does not constitute an offer to sell or a solicitation of an offer to buy any securities offered by this prospectus in any jurisdiction in which such an offer or a solicitation is unlawful.
Australia
This prospectus:
| • | does not constitute a disclosure document or a prospectus under Chapter 6D.2 of the Corporations Act 2001 (Cth), or the Corporations Act; |
| • | has not been, and will not be, lodged with the Australian Securities and Investments Commission, or ASIC, as a disclosure document for the purposes of the Corporations Act and does not purport to include the information required of a disclosure document for the purposes of the Corporations Act; and |
| • | may only be provided in Australia to select investors who are able to demonstrate that they fall within one or more of the categories of investors, available under section 708 of the Corporations Act, or Exempt Investors. |
The securities may not be directly or indirectly offered for subscription or purchased or sold, and no invitations to subscribe for or buy the securities may be issued, and no draft or definitive offering memorandum, advertisement or other offering material relating to any securities may be distributed in Australia, except where disclosure to investors is not required under Chapter 6D of the Corporations Act or is otherwise in compliance with all applicable Australian laws and regulations. By submitting an application for the securities, you represent and warrant to us that you are an Exempt Investor.
As any offer of securities under this prospectus will be made without disclosure in Australia under Chapter 6D.2 of the Corporations Act, the offer of those securities for resale in Australia within 12 months may, under section 707 of the Corporations Act, require disclosure to investors under Chapter 6D.2 if none of the exemptions in section 708 applies to that resale. By applying for the securities you undertake to us that you will not, for a period of 12 months from the date of issue of the securities, offer, transfer, assign or otherwise alienate those securities to investors in Australia except in circumstances where disclosure to investors is not required under Chapter 6D.2 of the Corporations Act or where a compliant disclosure document is prepared and lodged with ASIC.
Canada
The securities may be sold only to purchasers purchasing, or deemed to be purchasing, as principal that are accredited investors, as defined in National Instrument 45-106 Prospectus Exemptions or subsection 73.3(1) of the Securities Act (Ontario), and are permitted clients, as defined in National Instrument 31-103 Registration Requirements, Exemptions and Ongoing Registrant Obligations. Any resale of the securities must be made in accordance with an exemption from, or in a transaction not subject to, the prospectus requirements of applicable securities laws.
Securities legislation in certain provinces or territories of Canada may provide a purchaser with remedies for rescission or damages if this prospectus (including any amendment thereto) contains a misrepresentation,
155
Table of Contents
provided that the remedies for rescission or damages are exercised by the purchaser within the time limit prescribed by the securities legislation of the purchaser’s province or territory. The purchaser should refer to any applicable provisions of the securities legislation of the purchaser’s province or territory for particulars of these rights or consult with a legal advisor.
Pursuant to section 3A.3 of National Instrument 33-105 Underwriting Conflicts (NI 33-105), the underwriters are not required to comply with the disclosure requirements of NI 33-105 regarding underwriter conflicts of interest in connection with this offering.
Cayman Islands
This prospectus does not constitute an invitation or offer to the public in the Cayman Islands of the securities, whether by way of sale or subscription. The underwriters have not offered or sold, and will not offer or sell, directly or indirectly, any securities in the Cayman Islands.
Dubai International Finance Center, or DIFC
This prospectus relates to an Exempt Offer in accordance with the Markets Rules 2012 of the Dubai Financial Services Authority, or DFSA. This prospectus is intended for distribution only to persons of a type specified in the Markets Rules 2012 of the DFSA. It must not be delivered to, or relied on by, any other person. The DFSA has no responsibility for reviewing or verifying any documents in connection with Exempt Offers. The DFSA has not approved this prospectus supplement nor taken steps to verify the information set forth herein and has no responsibility for this prospectus. The securities to which this prospectus relates may be illiquid and/or subject to restrictions on their resale. Prospective purchasers of the securities offered should conduct their own due diligence on the securities. If you do not understand the contents of this prospectus you should consult an authorized financial advisor.
In relation to its use in the DIFC, this prospectus is strictly private and confidential and is being distributed to a limited number of investors and must not be provided to any person other than the original recipient, and may not be reproduced or used for any other purpose. The interests in the securities may not be offered or sold directly or indirectly to the public in the DIFC.
European Economic Area
In relation to each Member State of the European Economic Area and the United Kingdom, or each a Relevant State, no securities have been offered or will be offered pursuant to this offering to the public in that Relevant State prior to the publication of a prospectus in relation to the securities which has been approved by the competent authority in that Relevant State or, where appropriate, approved in another Relevant State and notified to the competent authority in that Relevant State, all in accordance with the Prospectus Regulation, except that offers of securities may be made to the public in that Relevant State at any time under the following exemptions under the Prospectus Regulation:
(a) to any legal entity which is a qualified investor as defined under the Prospectus Regulation;
(b) to fewer than 150 natural or legal persons (other than qualified investors as defined under the Prospectus Regulation), subject to obtaining the prior consent of the underwriters; or
(c) in any other circumstances falling within Article 1(4) of the Prospectus Regulation,
provided that no such offer of securities shall require us or any underwriter to publish a prospectus pursuant to Article 3 of the Prospectus Regulation or supplement a prospectus pursuant to Article 23 of the Prospectus Regulation and each person who initially acquires any securities or to whom any offer is made will be deemed to
156
Table of Contents
have represented, acknowledged and agreed to and with each of the underwriters and us that it is a “qualified investor” within the meaning of Article 2(e) of the Prospectus Regulation. In the case of any securities being offered to a financial intermediary as that term is used in the Prospectus Regulation, each such financial intermediary will be deemed to have represented, acknowledged and agreed that the securities acquired by it in the offer have not been acquired on a non-discretionary basis on behalf of, nor have they been acquired with a view to their offer or resale to, persons in circumstances which may give rise to an offer of any securities to the public other than their offer or resale in a Relevant State to qualified investors as so defined or in circumstances in which the prior consent of the representative of the underwriters has been obtained to each such proposed offer or resale.
For the purposes of this provision, the expression an “offer to the public” in relation to securities in any Relevant State means the communication in any form and by any means of sufficient information on the terms of the offer and any securities to be offered so as to enable an investor to decide to purchase or subscribe for any securities, and the expression “Prospectus Regulation” means Regulation (EU) 2017/1129.
Hong Kong
The securities have not been offered or sold and will not be offered or sold in Hong Kong, by means of any document, other than (a) to “professional investors” as defined in the Securities and Futures Ordinance (Cap. 571 of the Laws of Hong Kong), or the SFO, of Hong Kong and any rules made thereunder; or (b) in other circumstances which do not result in the document being a “prospectus” as defined in the Companies (Winding Up and Miscellaneous Provisions) Ordinance (Cap. 32) of Hong Kong), or the CO, or which do not constitute an offer to the public within the meaning of the CO. No advertisement, invitation or document relating to the securities has been or may be issued or has been or may be in the possession of any person for the purposes of issue, whether in Hong Kong or elsewhere, which is directed at, or the contents of which are likely to be accessed or read by, the public of Hong Kong (except if permitted to do so under the securities laws of Hong Kong) other than with respect to securities which are or are intended to be disposed of only to persons outside Hong Kong or only to “professional investors” as defined in the SFO and any rules made thereunder.
Israel
This prospectus does not constitute a prospectus under the Israeli Securities Law, 5728-1968, and has not been filed with or approved by the Israel Securities Authority. In Israel, this prospectus may be distributed only to, and is directed only at, investors listed in the first addendum, or the Addendum, to the Israeli Securities Law, consisting primarily of joint investment in trust funds; provident funds; insurance companies; banks, portfolio managers, investment advisors, members of the Tel Aviv Stock Exchange Ltd., underwriters, each purchasing for their own account; venture capital funds; entities with equity in excess of NIS 50 million and “qualified individuals,” each as defined in the Addendum (as it may be amended from time to time), collectively referred to as qualified investors. Qualified investors shall be required to submit written confirmation that they fall within the scope of the Addendum.
Japan
The securities have not been and will not be registered pursuant to Article 4, Paragraph 1 of the Financial Instruments and Exchange Act. Accordingly, none of the securities nor any interest therein may be offered or sold, directly or indirectly, in Japan or to, or for the benefit of, any “resident” of Japan (which term as used herein means any person resident in Japan, including any corporation or other entity organized under the laws of Japan), or to others for re-offering or resale, directly or indirectly, in Japan or to or for the benefit of a resident of Japan, except pursuant to an exemption from the registration requirements of, and otherwise in compliance with, the Financial Instruments and Exchange Act and any other applicable laws, regulations and ministerial guidelines of Japan in effect at the relevant time.
157
Table of Contents
Kingdom of Saudi Arabia
This prospectus may not be distributed in the Kingdom of Saudi Arabia except to such persons as are permitted under the Offers of Securities Regulations as issued by the board of the Saudi Arabian Capital Market Authority, or CMA, pursuant to resolution number 2-11-2004 dated 4 October 2004 as amended by resolution number 1-28-2008, as amended, or the CMA Regulations. The CMA does not make any representation as to the accuracy or completeness of this prospectus and expressly disclaims any liability whatsoever for any loss arising from, or incurred in reliance upon, any part of this prospectus. Prospective purchasers of the securities offered hereby should conduct their own due diligence on the accuracy of the information relating to the securities. If you do not understand the contents of this prospectus, you should consult an authorized financial adviser.
Korea
The securities have not been and will not be registered under the Financial Investments Services and Capital Markets Act of Korea and the decrees and regulations thereunder, or the FSCMA, and the securities have been and will be offered in Korea as a private placement under the FSCMA. None of the securities may be offered, sold or delivered directly or indirectly, or offered or sold to any person for re-offering or resale, directly or indirectly, in Korea or to any resident of Korea except pursuant to the applicable laws and regulations of Korea, including the FSCMA and the Foreign Exchange Transaction Law of Korea and the decrees and regulations thereunder, or the FETL. The securities have not been listed on any of securities exchanges in the world including, without limitation, the Korea Exchange in Korea. Furthermore, the purchaser of the securities shall comply with all applicable regulatory requirements (including but not limited to requirements under the FETL) in connection with the purchase of the securities. By the purchase of the securities, the relevant holder thereof will be deemed to represent and warrant that if it is in Korea or is a resident of Korea, it purchased the securities pursuant to the applicable laws and regulations of Korea.
Kuwait
Unless all necessary approvals from the Kuwait Capital Markets Authority pursuant to Law No. 7 of 2010 Concerning the Establishment of the Capital Markets Authority and Regulating of Securities Activities and the implementing regulations thereto (as amended), and the various resolutions, regulations, instructions and announcements issued from time to time pursuant thereto, or in connection therewith, have been given in relation to the marketing of, and sale of, the securities, the securities may not be marketed, offered for sale, nor sold in the State of Kuwait. Neither this prospectus (including any related document), nor any of the information contained therein is intended to lead to the conclusion of any contract of whatsoever nature within Kuwait.
Malaysia
No prospectus or other offering material or document in connection with the offer and sale of the securities has been or will be registered with the Securities Commission of Malaysia, or Commission, for the Commission’s approval pursuant to the Capital Markets and Services Act 2007. Accordingly, this prospectus and any other document or material in connection with the offer or sale, or invitation for subscription or purchase, of the securities may not be circulated or distributed, nor may the securities be offered or sold, or be made the subject of an invitation for subscription or purchase, whether directly or indirectly, to persons in Malaysia other than (i) a closed end fund approved by the Commission; (ii) a holder of a Capital Markets Services Licence; (iii) a person who acquires the securities, as principal, if the offer is on terms that the securities may only be acquired at a consideration of not less than RM250,000 (or its equivalent in foreign currencies) for each transaction; (iv) an individual whose total net personal assets or total net joint assets with his or her spouse exceeds RM3 million (or its equivalent in foreign currencies), excluding the value of the primary residence of the individual; (v) an individual who has a gross annual income exceeding RM300,000 (or its equivalent in foreign currencies) per annum in the preceding twelve months; (vi) an individual who, jointly with his or her spouse, has a gross annual income of RM400,000 (or its equivalent in foreign currencies), per annum in the preceding twelve
158
Table of Contents
months; (vii) a corporation with total net assets exceeding RM10 million (or its equivalent in a foreign currencies) based on the last audited accounts; (viii) a partnership with total net assets exceeding RM10 million (or its equivalent in foreign currencies); (ix) a bank licensee or insurance licensee as defined in the Labuan Financial Services and Securities Act 2010; (x) an Islamic bank licensee or takaful licensee as defined in the Labuan Financial Services and Securities Act 2010; and (xi) any other person as may be specified by the Commission; provided that, in the each of the preceding categories (i) to (xi), the distribution of the securities is made by a holder of a Capital Markets Services Licence who carries on the business of dealing in securities. The distribution in Malaysia of this prospectus is subject to Malaysian laws. This prospectus does not constitute and may not be used for the purpose of public offering or an issue, offer for subscription or purchase, invitation to subscribe for or purchase any securities requiring the registration of a prospectus with the Commission under the Capital Markets and Services Act 2007.
Mexico
None of the securities or the ordinary shares have been or will be registered with the National Securities Registry (Registro Nacional de Valores) maintained by the Mexican National Banking and Securities Commission (Comision Nacional Bancaria y de Valores), or CNBV, of Mexico and, as a result, may not be offered or sold publicly in Mexico. The securities and the ordinary shares may only be sold to Mexican institutional and qualified investors, pursuant to the private placement exemption set forth in the Mexican Securities Market Law (Ley del Mercado de Valores). As required under the Mexican Securities Market Law, the company will give notice to the CNBV of the offering of the securities under the terms set forth herein. Such notice will be submitted to the CNBV to comply with the Mexican Securities Market Law, and for informational purposes only. The delivery to, and receipt by, the CNBV of such notice does not certify the solvency of the company, the investment quality of the securities, or that the information contained in this prospectus or in any prospectus supplement. The company has prepared this prospectus and is solely responsible for its content, and the CNBV has not reviewed or authorized such content.
People’s Republic of China
This prospectus will not be circulated or distributed in the PRC and the securities will not be offered or sold, and will not be offered or sold to any person for re-offering or resale directly or indirectly to any residents of the PRC except pursuant to any applicable laws and regulations of the PRC. Neither this prospectus nor any advertisement or other offering material may be distributed or published in the PRC, except under circumstances that will result in compliance with applicable laws and regulations.
Singapore
This prospectus has not been registered as a prospectus with the Monetary Authority of Singapore. Accordingly, the underwriters have not offered or sold any securities or caused the securities to be made the subject of an invitation for subscription or purchase and will not offer or sell any securities or cause the securities to be made the subject of an invitation for subscription or purchase, and has not circulated or distributed, nor will it circulate or distribute, this prospectus or any other document or material in connection with the offer or sale, or invitation for subscription or purchase, of the securities, whether directly or indirectly, to any person in Singapore other than:
(a) to an institutional investor (as defined in Section 4A of the Securities and Futures Act (Chapter 289) of Singapore, as modified or amended from time to time, or the SFA) pursuant to Section 274 of the SFA;
(b) to a relevant person (as defined in Section 275(2) of the SFA) pursuant to Section 275(1) of the SFA, or any person pursuant to Section 275(1A) of the SFA, and in accordance with the conditions specified in Section 275 of the SFA; or
159
Table of Contents
(c) otherwise pursuant to, and in accordance with the conditions of, any other applicable provision of the SFA.
Where the securities are subscribed or purchased under Section 275 of the SFA by a relevant person which is:
(a) a corporation (which is not an accredited investor (as defined in Section 4A of the SFA)) the sole business of which is to hold investments and the entire share capital of which is owned by one or more individuals, each of whom is an accredited investor; or
(b) a trust (where the trustee is not an accredited investor) whose sole purpose is to hold investments and each beneficiary of the trust is an individual who is an accredited investor,
securities or securities-based derivatives contracts (each term as defined in Section 2(1) of the SFA) of that corporation or the beneficiaries’ rights and interest (howsoever described) in that trust shall not be transferred within six months after that corporation or that trust has acquired the securities pursuant to an offer made under Section 275 of the SFA except:
(i) to an institutional investor or to a relevant person, or to any person arising from an offer referred to in Section 275(1A) or Section 276(4)(i)(B) of the SFA;
(ii) where no consideration is or will be given for the transfer;
(iii) where the transfer is by operation of law;
(iv) as specified in Section 276(7) of the SFA; or
(v) as specified in Regulation 37A of the Securities and Futures (Offers of Investments) (Securities and Securities-based Derivatives Contracts) Regulations 2018.
State of Qatar
The securities described in this prospectus have not been, and will not be, offered, sold or delivered, at any time, directly or indirectly in the State of Qatar in a manner that would constitute a public offering. This prospectus has not been, and will not be, registered with or approved by the Qatar Financial Markets Authority or Qatar Central Bank and may not be publicly distributed. This prospectus is intended for the original recipient only and must not be provided to any other person. It is not for general circulation in the State of Qatar and may not be reproduced or used for any other purpose.
Switzerland
The securities may not be publicly offered in Switzerland and will not be listed on the SIX Swiss Exchange, or SIX, or on any other stock exchange or regulated trading facility in Switzerland. This prospectus does not constitute a prospectus within the meaning of, and has been prepared without regard to the disclosure standards for issuance prospectuses under art. 652a or art. 1156 of the Swiss Code of Obligations or the disclosure standards for listing prospectuses under art. 27 ff. of the SIX Listing Rules or the listing rules of any other stock exchange or regulated trading facility in Switzerland. Neither this prospectus nor any other offering or marketing material relating to the securities or the offering may be publicly distributed or otherwise made publicly available in Switzerland.
Neither this prospectus nor any other offering or marketing material relating to the offering, the Company, the securities have been or will be filed with or approved by any Swiss regulatory authority. In particular, this prospectus will not be filed with, and the offer of securities will not be supervised by, the Swiss Financial Market Supervisory Authority FINMA (FINMA), and the offer of securities has not been and will not be authorized
160
Table of Contents
under the Swiss Federal Act on Collective Investment Schemes, or CISA. The investor protection afforded to acquirers of interests in collective investment schemes under the CISA does not extend to acquirers of securities.
Taiwan
The securities have not been and will not be registered with the Financial Supervisory Commission of Taiwan pursuant to relevant securities laws and regulations and may not be sold, issued or offered within Taiwan through a public offering or in circumstances which constitutes an offer within the meaning of the Securities and Exchange Act of Taiwan that requires a registration or approval of the Financial Supervisory Commission of Taiwan. No person or entity in Taiwan has been authorized to offer, sell, give advice regarding or otherwise intermediate the offering and sale of the securities in Taiwan.
United Arab Emirates
The securities have not been, and are not being, publicly offered, sold, promoted or advertised in the United Arab Emirates (including the Dubai International Financial Centre) other than in compliance with the laws of the United Arab Emirates (and the Dubai International Financial Centre) governing the issue, offering and sale of securities. Further, this prospectus does not constitute a public offer of securities in the United Arab Emirates (including the Dubai International Financial Centre) and is not intended to be a public offer. This prospectus has not been approved by or filed with the Central Bank of the United Arab Emirates, the Securities and Commodities Authority or the Dubai Financial Services Authority.
United Kingdom
The securities may not be made in the United Kingdom, except that an offer to the public of any securities may be made in the United Kingdom at any time:
(a) to any legal entity which is a qualified investor as defined under Article 2 of the UK Prospectus Regulation;
(b) to fewer than 150 natural or legal persons (other than qualified investors as defined under Article 2 of the UK Prospectus Regulation), subject to obtaining the prior consent of the representative for any such offer; or
(c) in any other circumstances falling within Section 86 of the Financial Services and Markets Act 2000 (as amended, the “FSMA”);
provided that no such offer of securities shall result in the requirement for the publication by us of a prospectus pursuant to Section 85 of the FSMA or supplement a prospectus pursuant to Article 23 of the UK Prospectus Regulation.
For the purposes of this provision, the expression an “offer to the public” in relation to the any securities in the United Kingdom means the communication in any form and by any means of sufficient information on the terms of the offer and the securities to be offered so as to enable an investor to decide to purchase or subscribe for the securities, and the expression “UK Prospectus Regulation” means Regulation (EU) 2017/1129 as it forms part of domestic law by virtue of the European Union (Withdrawal) Act 2018.
This prospectus is only being distributed to and is only directed at: (1) persons who are outside the United Kingdom; (2) investment professionals falling within Article 19(5) of the Financial Services and Markets Act 2000 (Financial Promotion) Order 2005 (the “Order”); or (3) high net worth companies, and other persons to whom it may lawfully be communicated, falling within Article 49(2)(a) to (ed) of the Order (all such persons falling within (1)-(3) together being referred to as “relevant persons”). The securities are only available to, and any invitation, offer or agreement to subscribe, purchase or otherwise acquire the securities will be engaged in only with, relevant persons. Any person who is not a relevant person should not act or rely on this prospectus or any of its contents.
161
Table of Contents
The validity of the securities offered hereby will be passed upon for us by O’Melveny & Myers LLP with respect to certain legal matters as to United States federal securities and New York State law. The underwriters are being represented by Ellenoff Grossman & Schole LLP with respect to certain legal matters as to United States federal securities and New York State law.
162
Table of Contents
The financial statements of CapsoVision, Inc. as of and for the year ended December 31, 2023 appearing in this prospectus, have been audited by Baker Tilly US, LLP, an independent registered public accounting firm, as set forth in their report appearing in this prospectus, in reliance upon such report and upon the authority of said firm as experts in accounting and auditing.
163
Table of Contents
WHERE YOU CAN FIND MORE INFORMATION
We have filed with the SEC a registration statement on Form S-1 under the Securities Act for the shares of common stock being offered by this prospectus. This prospectus, which is part of the registration statement, does not contain all of the information included in the registration statement and the exhibits. For further information about us and the common stock offered by this prospectus, you should refer to the registration statement and its exhibits. References in this prospectus to any of our contracts or other documents are not necessarily complete, and you should refer to the exhibits attached to the registration statement for copies of the actual contract or document. You may read and copy any document that we file at the SEC’s public reference room located at 100 F Street, NE, Washington, D.C. 20549. Please call the SEC at 1-800-SEC-0330 for further information on the public reference rooms. SEC filings are also available to the public at the SEC’s website at www.sec.gov.
We will be subject to the reporting and information requirements of the Exchange Act and, as a result, will file periodic and current reports, proxy statements and other information with the SEC. We expect to make our periodic reports and other information filed with or furnished to the SEC, available, free of charge, through our website as soon as reasonably practicable after those reports and other information are filed with or furnished to the SEC. Additionally, these periodic reports, proxy statements and other information will be available for inspection and copying at the public reference room and website of the SEC referred to above.
164
Table of Contents
| Page | ||||
| F-2 | ||||
| F-3 | ||||
| F-5 | ||||
| Statement of Convertible Preferred Stock and Stockholders’ Deficit |
F-6 | |||
| F-7 | ||||
| F-8 | ||||
Table of Contents
Report of Independent Registered Public Accounting Firm
To the Shareholders and the Board of Directors of CapsoVision, Inc.
Opinion on the Financial Statements
We have audited the accompanying balance sheet of CapsoVision, Inc. (the Company) as of December 31, 2023, the related statements of operations, convertible preferred stock and stockholders’ deficit and cash flows for the year ended December 31, 2023, and the related notes (collectively referred to as the “financial statements”). In our opinion, the financial statements present fairly, in all material respects, the financial position of the Company as of December 31, 2023, and the results of its operations and its cash flows for the year ended December 31, 2023, in conformity with accounting principles generally accepted in the United States of America.
Going Concern Uncertainty
The accompanying financial statements have been prepared assuming that the Company will continue as a going concern. As discussed in Note 2 to the financial statements, the Company has suffered recurring losses, negative cash outflows from operations, and had a stockholders’ deficit as of December 31, 2023, that raise substantial doubt about its ability to continue as a going concern. Management’s plans in regard to these matters are also described in Note 2. The financial statements do not include any adjustments that might result from the outcome of this uncertainty.
Basis for Opinion
These financial statements are the responsibility of the Company’s management. Our responsibility is to express an opinion on the Company’s financial statements based on our audit. We are a public accounting firm registered with the Public Company Accounting Oversight Board (United States) (PCAOB) and are required to be independent with respect to the Company in accordance with the U.S. federal securities laws and the applicable rules and regulations of the Securities and Exchange Commission and the PCAOB.
We conducted our audit in accordance with the standards of the PCAOB and in accordance with auditing standards generally accepted in the United States of America. Those standards require that we plan and perform the audit to obtain reasonable assurance about whether the financial statements are free of material misstatement, whether due to error or fraud. Our audit included performing procedures to assess the risks of material misstatement of the financial statements, whether due to error or fraud, and performing procedures that respond to those risks. Such procedures included examining, on a test basis, evidence regarding the amounts and disclosures in the financial statements. Our audit also included evaluating the accounting principles used and significant estimates made by management, as well as evaluating the overall presentation of the financial statements. We believe that our audit provides a reasonable basis for our opinion.
We have served as the Company’s auditor since 2024.
/s/ Baker Tilly US, LLP
San Jose, California
December 16, 2024
F-2
Table of Contents
CAPSOVISION, INC.
December 31, 2023
| December 31, 2023 | ||||
| ASSETS |
||||
| Current assets |
||||
| Cash |
$ | 14,559,374 | ||
| Accounts receivable, net of allowance for credit losses of $10,089 |
1,865,601 | |||
| Inventory |
2,194,504 | |||
| Prepaid expenses and other current assets |
582,363 | |||
|
|
|
|||
| Total current assets |
19,201,842 | |||
| Property and equipment, net |
772,708 | |||
| Operating lease right-of-use assets |
1,509,706 | |||
| Other long-term assets |
42,616 | |||
|
|
|
|||
| TOTAL ASSETS |
$ | 21,526,872 | ||
|
|
|
|||
| LIABILITIES, CONVERTIBLE PREFERRED STOCK, AND STOCKHOLDERS’ DEFICIT |
||||
| Current liabilities |
||||
| Accounts payable |
$ | 360,642 | ||
| Accrued expenses and other current liabilities |
774,495 | |||
| Deferred revenue |
96,129 | |||
| Operating lease liabilities—current |
276,333 | |||
|
|
|
|||
| Total current liabilities |
1,507,599 | |||
| Operating lease liabilities—long-term |
1,237,504 | |||
|
|
|
|||
| Total liabilities |
2,745,103 | |||
| Commitments and contingencies—Note 8 |
||||
| Convertible preferred stock (each Series: $0.001 par value) |
||||
| Series A: 17,962,675 shares authorized; 17,962,675 shares issued and outstanding; liquidation preference of $4,849,922 as of December 31, 2023 |
4,849,922 | |||
| Series B: 6,000,000 shares authorized; 6,000,000 shares issued and outstanding; liquidation preference of $4,318,938 as of December 31, 2023 |
4,318,938 | |||
| Series C: 5,747,127 shares authorized; 2,931,022 shares issued and outstanding; liquidation preference of $2,549,989 as of December 31, 2023 |
2,549,989 | |||
| Series C-1: 3,876,405 shares authorized; 2,825,835 shares issued and outstanding; liquidation preference of $2,514,993 as of December 31, 2023 |
2,514,993 | |||
| Series D: 2,222,222 shares authorized; 1,733,329 shares issued and outstanding; liquidation preference of $1,559,996 as of December 31, 2023 |
1,559,996 | |||
| Series D-1: 6,766,666 shares authorized; 555,553 shares issued and outstanding; liquidation preference of $499,998 as of December 31, 2023 |
499,998 | |||
| Series D-2: 11,083,333 shares authorized; 9,725,761 shares issued and outstanding; liquidation preference of $17,506,370 as of December 31, 2023 |
8,753,185 | |||
| Series E: 14,000,000 shares authorized; 13,826,084 shares issued and outstanding; liquidation preference of $15,899,997 as of December 31, 2023 |
15,899,997 | |||
| Series F-1: 13,043,479 shares authorized; 4,000,005 shares issued and outstanding; liquidation preference of $4,600,006 as of December 31, 2023 |
4,600,006 | |||
| Series F-2: 12,000,000 shares authorized; 8,320,000 shares issued and outstanding; liquidation preference of $10,400,000 as of December 31, 2023 |
10,400,000 | |||
The accompanying notes are an integral part of these financial statements.
F-3
Table of Contents
CAPSOVISION, INC.
December 31, 2023
| December 31, 2023 | ||||
| Convertible preferred stock (each Series: $0.001 par value), continued |
||||
| Series G: 5,926,000 shares authorized; 5,925,931 shares issued and outstanding; liquidation preference of $8,000,007 as of December 31, 2023 |
$ | 8,000,007 | ||
| Series G-1: 6,896,552 shares authorized; 6,792,389 shares issued and outstanding; liquidation preference of $9,848,964 as of December 31, 2023 |
9,848,964 | |||
| Series H: 37,812,994 shares authorized; 37,812,974 shares issued and outstanding; liquidation preference of $54,828,812 as of December 31, 2023 |
54,828,812 | |||
|
|
|
|||
| Total convertible preferred stock |
128,624,807 | |||
| Stockholders’ deficit |
||||
| Common stock, $0.001 par value: 170,000,000 shares authorized; 6,117,333 shares issued and outstanding as of December 31, 2023 |
6,117 | |||
| Additional paid-in capital |
602,697 | |||
| Accumulated deficit |
(110,451,852 | ) | ||
|
|
|
|||
| Total stockholders’ deficit |
(109,843,038 | ) | ||
|
|
|
|||
| TOTAL LIABILITIES, CONVERTIBLE PREFERRED STOCK, AND STOCKHOLDERS’ DEFICIT |
$ | 21,526,872 | ||
|
|
|
|||
The accompanying notes are an integral part of these financial statements.
F-4
Table of Contents
CAPSOVISION, INC.
Year Ended December 31, 2023
| 2023 | ||||
| REVENUE |
||||
| Revenue |
$ | 9,752,850 | ||
| COSTS OF REVENUE |
||||
| Costs of revenue |
4,262,373 | |||
|
|
|
|||
| Gross profit |
5,490,477 | |||
| OPERATING EXPENSES |
||||
| Selling and marketing |
5,532,852 | |||
| Research and development |
9,332,574 | |||
| General and administrative |
1,972,274 | |||
|
|
|
|||
| Total operating expenses |
16,837,700 | |||
| Operating loss |
(11,347,223 | ) | ||
| NON-OPERATING INCOME |
||||
| Interest income |
49,440 | |||
| Other non-operating income, net |
4,371 | |||
|
|
|
|||
| Total non-operating income, net |
53,811 | |||
| Loss before provision for income taxes |
(11,293,412 | ) | ||
| Provision for income taxes |
11,433 | |||
|
|
|
|||
| Net loss |
$ | (11,304,845 | ) | |
|
|
|
|||
| Net loss per share—basic and diluted |
$ | (1.93 | ) | |
|
|
|
|||
| Weighted average shares—basic and diluted |
5,862,935 | |||
|
|
|
|||
The accompanying notes are an integral part of these financial statements.
F-5
Table of Contents
CAPSOVISION, INC.
STATEMENT OF CONVERTIBLE PREFERRED STOCK AND STOCKHOLDERS’ DEFICIT
Year Ended December 31, 2023
| Convertible Preferred Stock | Common Stock | |||||||||||||||||||||||||||
| Shares | Amount | Shares | Amount | Additional Paid-in- Capital |
Accumulated Deficit | Total Stockholders’ Deficit |
||||||||||||||||||||||
| December 31, 2022 |
104,201,238 | $ | 108,269,831 | 5,651,647 | $ | 5,652 | $ | 386,185 | $ | (99,147,007 | ) | $ | (98,755,170 | ) | ||||||||||||||
| Issuance of Series H convertible preferred stock, net of issuance costs |
14,210,319 | 20,354,976 | — | — | — | — | — | |||||||||||||||||||||
| Issuance of common stock upon exercises of stock options and warrants |
— | — | 465,686 | 465 | 173,877 | — | 174,342 | |||||||||||||||||||||
| Stock-based compensation |
— | — | — | — | 42,635 | — | 42,635 | |||||||||||||||||||||
| Net loss |
— | — | — | — | — | (11,304,845 | ) | (11,304,845 | ) | |||||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||
| December 31, 2023 |
118,411,557 | $ | 128,624,807 | 6,117,333 | $ | 6,117 | $ | 602,697 | $ | (110,451,852) | $ | (109,843,038) | ||||||||||||||||
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|||||||||||||||
The accompanying notes are an integral part of these financial statements.
F-6
Table of Contents
CAPSOVISION, INC.
Year Ended December 31, 2023
| 2023 | ||||
| CASH FLOWS FROM OPERATING ACTIVITIES |
||||
| Net loss |
$ | (11,304,845 | ) | |
| Adjustments to reconcile net loss to net cash used in operating activities: |
||||
| Depreciation and amortization |
141,355 | |||
| Amortization of operating lease right-of-use assets |
246,777 | |||
| Unrealized foreign exchange gains |
(93,152 | ) | ||
| Stock-based compensation |
42,635 | |||
| Bad debt expense |
3,858 | |||
| Changes in operating assets and liabilities: |
||||
| Accounts receivable |
6,962 | |||
| Inventory |
(672,980 | ) | ||
| Prepaid expenses and other current assets |
782,793 | |||
| Other long-term assets |
(5,344 | ) | ||
| Accounts payable |
201,937 | |||
| Accrued expenses and other current liabilities |
101,822 | |||
| Deferred revenue |
21,652 | |||
| Operating lease liabilities |
(277,303 | ) | ||
|
|
|
|||
| Net cash used in operating activities |
$ | (10,803,833 | ) | |
|
|
|
|||
| CASH FLOWS FROM INVESTING ACTIVITIES |
||||
| Purchases of property and equipment |
(754,180 | ) | ||
|
|
|
|||
| Net cash used in investing activities |
$ | (754,180 | ) | |
|
|
|
|||
| CASH FLOWS FROM FINANCING ACTIVITIES |
||||
| Proceeds from issuance of Series H convertible preferred stock, net of issuance costs |
20,354,976 | |||
| Proceeds from exercises of options and warrants on common stock |
174,342 | |||
|
|
|
|||
| Net cash provided by financing activities |
$ | 20,529,318 | ||
|
|
|
|||
| Effect of exchange rate changes on cash |
37,241 | |||
| Net increase in cash |
8,971,305 | |||
| Cash at beginning of year |
5,550,828 | |||
|
|
|
|||
| Cash at end of year |
$ | 14,559,374 | ||
|
|
|
|||
| SUPPLEMENTAL CASH FLOW DISCLOSURES |
||||
| Cash paid for income taxes |
$ | 916 | ||
| Non-cash investing and financing activities |
||||
| Additional right-of-use asset obtained and associated lease liability incurred upon lease modification, net of effect of adjusting existing right-of-use asset and lease liability |
$ | 168,647 | ||
| Purchases of property and equipment in accounts payable and accruals |
$ | 7,100 | ||
The accompanying notes are an integral part of these financial statements.
F-7
Table of Contents
CAPSOVISION, INC.
DECEMBER 31, 2023
| 1. | DESCRIPTION OF BUSINESS AND ORGANIZATIONAL STRUCTURE |
CapsoVision, Inc. (“CapsoVision” or the “Company”) is a commercial stage medical device enterprise whose portfolio of endoscopic medical devices reflects its core mission—empowering through innovation focused on internal imaging of the human gastrointestinal (“GI”) tract. High-quality internal GI imaging facilitates earlier detection of colorectal cancer and other diseases, enabling more timely and effective treatment regimens for patients. The Company is based in Saratoga, California and sells products and services to customers that include, in the U.S., gastrointestinal medical practices, clinics, or hospitals and groups thereof and, in multiple foreign countries, distributor customers who resell products.
The Company’s core technology platform is an orally ingestable endoscopic capsule with key features including multiple cameras facilitating 360 degree imaging, light emitters for measurement, and onboard memory storage and battery life permitting recording and retention of high-resolution video without requiring external data transmission devices or a patient to be present in a physician’s facility throughout the entire digestive cycle. The Company’s product portfolio consists of (i) the in-market CapsoCam Plus® device directed at the small intestines and (ii) the in-development CapsoColon 3D® device directed at the large intestines/colon. The Company’s endocapsule devices co-operate, once retrieved, with a data access device and its enabling software; these permit on-site, on-demand video download and viewing by a physician. The Company’s services include CapsoCloud®, a service offering allowing customers to ship endocapsules off-premise for centralized video downloading; patient video content is made available in a cloud-based software-as-a-service platform for customer viewing, download, and report generation. The Company’s services also include video reading services offered to its customers for video interpretation and related reports.
Since its 2005 inception as a Delaware corporation, the Company initially focused on preclinical and clinical development of products, expanding its technical and product development talent, and building a sales team (the Company’s sales force sells directly to customers in the U.S., and oversees outside-the-U.S. distributor customers). The Company’s organizational structure comprises a single Delaware corporation with a branch office in Taiwan, Republic of China, the location of certain Company suppliers and certain Company personnel.
The Company’s first endocapsule, CapsoCam Plus®, was cleared for human use by the U.S. Food and Drug Administration in 2016 and has since been approved in various other countries. The Company’s second major product, CapsoColon 3D®, has underway a pivotal human clinical study with planned submission of the product for regulatory clearance by the U.S. FDA (and subsequently, regulators in other countries). The Company is subject to risks and uncertainties common to companies in the life sciences sector with significant development programs including, but not limited to, risks associated with successful research, development and manufacturing, competition from other companies, protection of intellectual property, compliance with government regulations, and the ability to secure additional capital to fund operations and research & development. Current and future programs, including for products focused on other areas of the gastrointestinal tract, will require significant research and development efforts, including extensive clinical testing and regulatory approval prior to commercialization.
| 2. | GOING CONCERN |
The Company historically has funded its operations from the issuance of convertible preferred stock and common stock. Since the 2016 launch of the CapsoCam Plus® product, the Company has been able to fund an increasing portion of its operating expenses from sales thereof. However, the Company has incurred operating losses, net losses, and negative operating cash flows since inception and had a stockholders’ deficit as of December 31, 2023. The Company expects to continue to experience similar circumstances until existing
F-8
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
marketed products’ sales and future products’ sales yield further growth in revenues resulting in improved operating income, net income, and positive operating cash flows. As of December 31, 2023, or for the year then ended, the Company had cash of $14,559,374, an operating loss of $11,347,223, a net loss of $11,304,845, cash used in operating activities of $10,803,833, and a stockholders’ deficit of $109,843,038. During 2023, the Company raised $20,354,976 from the issuance of Series H convertible preferred stock, and subsequent to December 31, 2023 through the date of issuance of these financial statements raised approximately $10 million from the issuance of Series H convertible preferred stock. The Company, having evaluated the extant facts and circumstances relating to its ability to continue as a going concern, concluded that the facts and circumstances raise substantial doubt about the Company’s ability to continue as a going concern within one year after the date of issuance of these financial statements. The Company’s existing approved plans do not alleviate this substantial doubt. While the Company intends to consummate additional capital raising activities sufficient to fund the operating and capital expenditures necessary to advance sales growth and product innovation objectives, there is no absolute certainty as to the occurrence, magnitude, or timing of capital raising activities in amounts sufficient to alleviate such substantial doubt.
| 3. | SUMMARY OF SIGNIFICANT ACCOUNTING POLICIES |
Basis of Presentation
The accompanying financial statements are prepared in conformity with accounting principles generally accepted in the United States of America (“U.S. GAAP”). Any reference in these notes as it concerns U.S. GAAP is meant to refer to authoritative pronouncements as found in the Accounting Standards Codification (“ASC”) and Accounting Standards Updates (“ASU”) issued by the Financial Accounting Standards Board (“FASB”).
The accompanying financial statements have been prepared on a going concern basis, which contemplates the realization of assets and the satisfaction of liabilities and commitments in the ordinary course of business.
Reporting Entity
The Company’s organizational structure reflects a single incorporated entity, CapsoVision, Inc.; there are no legal entity subsidiaries or affiliated entities requiring consolidation under the economic interest (variable interest entity) model or voting interest model. This single entity has a foreign branch and physical presence in Taiwan (the Republic of China) which is an extension of the U.S. corporation.
Foreign Currency
The reporting currency for the financial statements of the Company is the United States Dollar. The functional currency of the U.S. corporation, and its offshore branch which is an extension of the U.S. corporation, is the U.S. Dollar. The assets, liabilities, and expenses of the Company’s foreign branch recorded in local currency are remeasured into the U.S. Dollar each period, with associated gains and losses included in operating expenses (as a component of general and administrative expenses) in the statement of operations. Monetary assets and liabilities denominated in currencies other than the functional currency are translated at exchange rates in effect at the balance sheet date with associated gains and losses included in operating expenses (as a component of general and administrative expenses) in the statement of operations. For the year ended December 31, 2023 the Company recorded $126,390 of net foreign exchange gains as a component of general and administrative expenses.
F-9
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Use of Estimates
The preparation of financial statements in conformity with U.S. GAAP requires the making and usage of estimates and assumptions that affect the reported amounts of assets, liabilities, revenue, expenses and the related disclosure of contingent assets and liabilities, if any. The Company bases its estimates on historical experience when available and on other assumptions that management believes are reasonable under the circumstances. The Company seeks to moderate the influence of subjectivity and estimation uncertainty through reliance on useful information from market participants and peers wherever possible. Estimates and assumptions affect various financial statement amounts and their related disclosures, including, but not limited to, the recognition of revenue, stock based compensation, research and development expenses, income tax-related amounts, and allowances for credit losses. Actual results could materially differ from estimates.
Segments
The Company is organized as, and managed as, a single operating (and reportable) segment. Its chief operating decision maker is its President.
Fair Value Measurements
Where required, certain assets and liabilities are carried at fair value. Fair value is defined as the exchange price that would be received for an asset or paid to transfer a liability (an exit price) in the principal or most advantageous market for the asset or liability in an orderly transaction between market participants on the measurement date. Valuation techniques used to measure fair value must maximize the use of observable inputs and minimize the use of unobservable inputs. A framework is used for measuring fair value utilizing a three-tier hierarchy that prioritizes the inputs to valuation techniques used to measure fair value. Financial assets and liabilities carried at fair value are to be classified and disclosed in one of the following three levels of the fair value hierarchy:
| • | Level 1—defined as observable inputs, such as quoted prices unadjusted in active markets for identical assets or liabilities |
| • | Level 2—defined as inputs other than quoted prices included in Level 1 that are either directly or indirectly observable |
| • | Level 3—defined as significant unobservable inputs for which little or no market data exists, therefore necessitating entity-specific assumptions |
To the extent that the valuation is based on models or inputs that are less observable or unobservable in the market, the determination of fair value requires more judgment. An asset or liability’s level within the fair value hierarchy is based on the lowest level of any input that is significant to the fair value.
The carrying amounts of the Company’s financial assets (which include cash and accounts receivable) and liabilities (which include accounts payable) approximate fair value due to their insensitivity to interest rates and/or short time periods to maturity and qualify as Level 1 measurements.
Concentration Risk
Financial instruments which potentially subject the Company to concentrations of credit risk consist principally of demand deposits at well-known financial institutions and accounts receivable. At times, the Company’s cash deposits may be in excess of U.S. Federal deposit insurance limits.
F-10
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
The Company’s sales to customers are typically originated with trade credit terms and receivables are uncollateralized, with limited exceptions for prepayments required for certain customers or time-of-sale payments accepted via credit cards. The Company historically has experienced insignificant levels of bad debt.
The Company’s concentration risk related to revenues relates primarily to its product revenues being derived from a single product. With respect to customer concentration risk, for the year ended, and as of December 31, 2023, a single customer represented approximately 12% of revenue and 28% of accounts receivable, respectively.
The Company’s supply concentration risk relates primarily to its materials procurement and contract manufacturing being substantially concentrated with vendors located in Asia.
Comprehensive Income (Loss)
Comprehensive income (loss) comprises net income (loss) and other comprehensive income (loss), which is defined as all changes in stockholders’ equity (deficit) other than net income (loss) and those resulting from investments by and distributions to stockholders. The Company historically has not had, and during the year ended December 31, 2023 did not have, any transactions requiring presentation in other comprehensive income (loss); as such, net loss equals comprehensive loss.
Revenue Recognition
The Company previously adopted FASB ASC 606, Revenue from Contracts with Customers (“ASC 606”) using the modified retrospective method with effect from January 1, 2020. The guidance provides a single model to determine when and how revenue is recognized. The core principle is that an entity should recognize revenue to depict the transfer of control of promised goods or services to customers in an amount that reflects the consideration to which the entity expects to be entitled in exchange for those goods or services. The Company recognizes revenue using a five-step model resulting in revenue being recognized as performance obligations within a contract are satisfied. The steps within that model include: (i) identifying the existence of a contract with a customer; (ii) identifying the performance obligations within the contract; (iii) determining the contract’s transaction price; (iv) allocating the transaction price to the contract’s performance obligations; and (v) recognizing revenue as the contract’s performance obligations are satisfied. Judgment is required to apply the principles-based, five-step model for revenue recognition. Management is required to make certain estimates and assumptions about the Company’s contracts with its customers, including, among others, the nature and extent of its performance obligations, its transaction price amounts and any allocations thereof (including estimates of standalone selling prices), the events which constitute satisfaction of its performance obligations, and when control of any promised goods or services is transferred to its customers. The guidance also requires certain incremental costs incurred to obtain or fulfill a contract to be deferred and amortized on a systematic basis consistent with the transfer of goods or services to a customer.
The Company generates (i) product revenue from the sale of capsule endoscopy medical devices, (ii) service revenue from the provision of reading services for endoscopy videos, and (iii) product or service revenue depending on which video delivery option is selected by and arranged with the customer to download and view capsule endoscope video output. The customer’s video delivery options include (1) a capsule data access reading device shipped to the customer which works together with downloadable software installed locally on a customer’s computer (classified as product revenue), or (2) a software-as-a-service offering which involves the customer mailing capsule devices to an off-site Company-operated facility which processes uploads of video content to a cloud platform the customer can access via a web browser or a smart device application in order to view videos and generate reports (classified as service revenue).
F-11
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
The Company’s contracts with customers contain both fixed and variable consideration. For contracts with variable consideration, the Company uses the most likely amount method to estimate the transaction price. Variable consideration is constrained to the extent that it is deemed probable that a significant reversal of the amount of revenue recognized will not occur. Where products or services are not sold separately or experience a range of selling prices, the Company estimates standalone selling prices using a cost-plus-expected margin approach and utilizes the resultant standalone selling prices to allocate transaction prices at transaction inception on a relative standalone selling price basis. The Company recognizes revenue for its product and reading/interpretive service performance obligations at a point in time, that being when the performance obligations are satisfied and control is transferred to the customer (generally upon shipment for products, or conveyance of final deliverable for services). For the software-as-a-service video delivery offering performance obligation described in the foregoing paragraph, the associated revenue is recognized over time, typically less than one fiscal quarter, representing the estimate of the typical period of time in which customers derive utility from the cloud-based service.
The Company does not have any significant financing components as payment is received at or shortly after sale. Costs incurred to obtain or fulfill a contract are expensed as incurred when the amortization period is less than one year, which is the case for the Company. The Company considers all shipping and handling to be fulfillment activities and not a separate performance obligation. Shipping and handling costs are recorded as costs of revenue. The Company has elected an accounting policy to exclude sales taxes and other similar taxes from the measurement of the transaction price.
The Company has certain U.S. customers characterized as group purchasing organizations that function as procurement agents for their underlying medical practice, clinic, or hospital members; these group purchasing organizations typically charge the Company a fixed or variable (tiered based upon volume purchased during a period) fee in exchange for the right to do business with the group purchasing organization. Such fees, which do not provide a distinct separate benefit to the Company, are recorded as reductions to revenue in the same time period as the recognition of the related revenue.
The Company has historically experienced minimal product returns given the one-time-use nature of its capsule devices; estimated product returns (periodically adjusted to reflect actual experience), if warranted, are accounted for as reductions to revenue with offsets to a product return liability included within accrued expenses and other current liabilities in the balance sheet.
Contract Assets and Contract Liabilities; Accounts Receivable and Related Allowance
The Company records contract assets when it has completed performance prior to receiving consideration from the customer and where such amounts are unbilled; where billed, amounts are reflected as trade accounts receivable. The Company promptly invoices accounts receivable upon shipment or delivery; therefore, the Company’s contract assets were zero as of December 31, 2023.
Contract liabilities, portrayed as deferred revenue, reflect (i) obligations to provide goods for which the Company has already received consideration (generally arising from up-front payments received) and (ii) performance obligations to provide goods or services which are not yet satisfied in the context of a particular contract (principally consisting of video delivery obligations related to the Company’s off-premise cloud-based video delivery offering). The Company’s deferred revenue balance was $96,129 as of December 31, 2023.
Trade accounts receivable are recorded at invoiced amounts and do not bear interest. The Company grants trade credit to most of its customers in the normal course of business and generally does not require collateral. An allowance for current expected credit losses arises subsequent to the origination of a sale for estimated
F-12
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
uncollectible receivables based on the Company’s assessment of the collectability of customer accounts. A provision (or a reversal of the allowance) is recorded as a component of general and administrative expenses in the statement of operations. In determining the amount of the allowance, the Company considers aging of accounts, historical credit losses, customer-specific information, the current economic environment, supportable forecasts, and other relevant factors. Uncollectible receivables are written off against the allowance for credit losses when all attempts to collect the receivable have been exhausted.
Costs of Revenue
Costs of revenue include materials, direct labor, and manufacturing overhead costs related to sold products, as well as certain period costs such as non-allocated overhead, scrap, and outbound freight costs, fees paid to physicians for providing reading services, and the costs of operating the Company’s cloud-based software-as-a-service offering for video delivery such as shipping costs, processing costs, and data storage costs. All shipping and handling costs directly related to bringing products to their final point of sale are included in costs of revenue and were $63,280 for the year ended December 31, 2023.
Selling and Marketing Expenses
Selling and marketing-related expenses include costs directly attributable to actively marketing the Company’s products and services using both direct employees and outside contractors or vendors. These costs include, but are not limited to, salaries, bonuses, benefits, and stock-based compensation, sales commissions, travel costs and expense reimbursements, and the costs of sponsoring programs, events, and conferences. Advertising costs were nil for the year ended December 31, 2023.
Research and Development Expenses (Including Clinical Trial Expenses)
Research and development costs are expensed as incurred in accordance with ASC 730, Research and Development. Research and development expenses include costs directly attributable to the conduct of research and development programs, including salaries, bonuses, benefits, and stock-based compensation for Company employees focused on R&D or clinical trials, costs of independent contractors and outside vendors, costs of supplies consumed in or product inventory utilized in clinical trials, and the costs of the clinical trials themselves as charged by trial sites or vendors responsible for multiple trial sites.
The Company capitalizes prepayments for goods or services, including trial device inventory, that will be used, consumed, or rendered for future research and development activities and recognizes expense as the related goods are delivered or services are performed. The Company also records expenses and accruals for estimated costs of research and development activities, including third party provided services for clinical trials of medical devices. The Company bases its estimates on information available at the time. Costs for certain clinical trial activities are recognized based on the pattern of performance of the individual arrangements, which may differ from the pattern of billings incurred, and are reflected in the balance sheet as prepaid expenses or as accrued expenses.
During the year ended December 31, 2023 the Company incurred $2,363,673 of expenses related to ongoing clinical trial activities which are included within research and development expenses.
General and Administrative Expenses
General and administrative expenses consist primarily of salaries, bonuses, benefits, and stock-based compensation related to the Company’s executive, administrative, finance, human resources, and other
F-13
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
supporting functions. Also included are professional fees for legal services, consulting services, tax matters and audits, information technology, office expenses, rent, insurance, and foreign exchange gains (losses).
Net Loss Per Share
Basic net loss per common share is computed by dividing net loss by the weighted average number of common shares outstanding during the period without consideration of potentially dilutive securities. Diluted net loss per share reflects the potential dilution that could occur if options on common stock were exercised or convertible preferred stock shares were converted into common stock of the Company. Diluted net loss per share is the same as basic net loss per common share since the effect of the potentially dilutive securities are anti-dilutive. Potential dilutive common share equivalents consist of incremental common shares issuable upon exercise of vested stock options, common stock warrants, and the Company’s various series of convertible preferred stock.
Cash
Cash comprises demand deposits in the form of checking accounts and money market accounts (which may be deposited into and withdrawn at will with no restrictions as to investment or redemption) and includes cash balances denominated in currencies other than the U.S. Dollar (Euro and New Taiwan Dollar) for which fluctuations in period-end exchange rates result in gains or losses reported within other income (expense), net in the statement of operations.
Inventory
Inventories, consisting of materials, direct labor and manufacturing overhead, are subdivided into raw materials, work-in-process, and finished goods and are stated at the lower of cost (moving average cost) or net realizable value.
The Company evaluates inventories as to (i) net realizable value and (ii) circumstances or indicators of loss, damage, insufficient time to expiry (determined by battery life), or excess quantities, and provisions accordingly via direct charges against the carrying value of inventory. For the year ended December 31, 2023, the Company did not record any inventory provisions.
Long-Lived Assets
Property and equipment is recorded at historical cost, less accumulated depreciation. The Company capitalizes major improvements and expenses repairs and maintenance as incurred. Depreciation is recorded using the straight-line method based on the estimated useful lives of the depreciable property or, for leasehold improvements, the remaining term of the lease, whichever is shorter. The useful lives of the assets are as follows:
| Asset Class |
Estimated Useful Life | |
| Machinery & equipment | 5 years | |
| Computer equipment and office equipment | 3 to 5 years | |
| Purchased software | 3 to 5 years | |
| Leasehold improvements | Shorter of remaining lease term or useful life |
Upon disposal (retirement or sale) of property and equipment, as applicable, the cost and related accumulated depreciation are removed and the resulting gain or loss is reflected within operating expenses. There were no gains or losses for the year ended December 31, 2023.
F-14
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Long-lived assets are evaluated for impairment as warranted by triggering events related to changes in facts and circumstances. There have historically been no impairments and there were none for the year ended December 31, 2023.
Leases
The Company has operating leases for office and storage space in California and office space in Taiwan, Republic of China.
The Company determines if an arrangement is a lease at inception and classifies its leases at commencement. Operating leases are presented as right-of-use (“ROU”) assets and the corresponding lease liabilities are depicted in operating lease liabilities, current and operating lease liabilities, non-current in the balance sheet. ROU assets represent the right to use an underlying asset, and lease liabilities represent the obligation for lease payments in exchange for the ability to use the asset for the duration of the lease term.
ROU assets and lease liabilities are recognized at commencement date and determined using the present value of the future minimum lease payments over the lease term. The Company utilizes an incremental borrowing rate when assessing lease classification and measuring lease liabilities. The estimated incremental borrowing rate considers credit rating practices applied by well known statistical rating organizations to borrowers, Company-specific factors, and the actual lease term at commencement date. The lease term may include options to extend when it is reasonably certain that the Company will exercise that option. The Company recognizes operating lease expense on a straight-line basis over the lease term.
The Company has lease agreements which contain both lease and non-lease components, which it has elected to account for as a single lease component. Variable lease payments that are not dependent on an index or rate are not included in lease measurements and are accounted for as incurred.
Convertible Preferred Stock
The Company’s primary means of financing from start-up through development and into the commercial stage has been the issuance of convertible preferred stock in various series; these series share similar rights and obligations and contain similar qualifications as to the requirements an equity initial public offering would have to meet in order for the stock to convert to common stock. Convertible preferred stock is recorded at fair value on the date of issuance (historically, the same as the issue price), net of issuance costs. Application of the guidance contained in ASC 480-10-S99-3A, SEC Staff Announcement: Classification and Measurement of Redeemable Securities has led the Company to present each of the series of convertible preferred stock as mezzanine equity due to the similar features of the series concerning liquidation preferences and other rights and privileges inuring to the respective series’ holders, one or more of which may result in a settlement in cash not entirely within the Company’s control.
Stock-Based Compensation
The Company accounts for employee and nonemployee equity compensation awards in accordance with ASC 718, Compensation—Stock Compensation (“ASC 718”). In accordance with ASU 2018-07, the Company accounts for stock-based compensation for awards granted to nonemployees in a similar fashion to the way it accounts for stock-based compensation awards to employees. ASC 718 requires the recognition of stock-based compensation expense, using a fair-value based method, for costs related to all stock- based compensation awards. Stock-based compensation awards issued under the Company’s 2005 Stock Plan take the form of stock
F-15
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
options (non-qualified or incentive stock options) or restricted stock. The Black-Scholes option-pricing model is used to determine the fair value of options. A mixture of income and market approaches to valuation are used to determine the fair value of the underlying common stock. The Black-Scholes option-pricing model requires judgment to develop inputs, some of which are more subjective, including: (i) the fair value of the underlying common stock on the date of grant; (ii) the expected term of the award; (iii) the expected volatility of the underlying common stock; (iv) the risk-free interest rate; and (v) expected dividends. The following descriptions reflect the Company’s option-pricing model input determinations.
| • | Fair Value of Common Stock: Due to the absence of an active market for the common stock, the valuation methodologies are employed in accordance with ASC 820, Fair Value Measurement. The estimated fair value of common stock is determined using one or more, or a combination, of valuation techniques including (i) the income approach (based upon undiscounted free cash flows reduced to present value by the weighted average cost of capital) and (ii) various market approaches such as application of market multiples, as adjusted to the Company’s circumstances, derived from the guideline public company method, the comparable transactions method, or the application of the Black-Scholes option pricing model to the Company’s equity. The valuation of common stock qualifies as a Level 3 measurement due to the use of significant unobservable inputs which directly or indirectly reflect Company-specific assumptions, the Company’s own financial forecasts, its competitive position, and macroeconomic factors. Significant changes to the assumptions could result in different fair values of common stock. |
| • | Expected Term: The expected term represents the period that the options granted are expected to be outstanding and is determined using the simplified method (based on the mid-point between the vesting date and the end of the contractual term) for substantially all awards (and the contractual term for certain nonemployee awards). The Company, as a pre-public entity with a limited history of stock option exercises, is unable to conclude that such limited history is a good predictor of exercise behavior that would be expected as a public entity; moreover, the Company does not expect substantially different exercise or post-vesting termination behavior patterns among sub-groups of equity awardees. |
| • | Expected Volatility: The Company uses a volatility measurement correlated to the expected term based upon peer group historical stock price volatility calculated from a selected peer group of public companies of varying sizes within the medical devices sector that are believed to possess reasonably similarities, due to the absence of an active market for the Company’s common stock and the absence of traded options from which to imply volatility. |
| • | Risk-Free Interest Rate: The risk-free interest rate correlated to the expected term is based on the constant maturity rate of U.S. Treasury securities with similar maturities as of the date of the grant. |
| • | Expected Dividend: Expected dividends are zero based upon the Company’s current and expected future dividend-declaring behavior. |
The Company’s employee and nonemployee stock-based compensation awards contain only service conditions for vesting. The Company recognizes stock-based compensation cost as a component of the related functional expense (costs of revenue or category of operating expenses) on a straight-line basis over the requisite service period, which is explicitly stated in the case of awards issued to employees and implicitly understood to be the period over which services are provided for nonemployees. The Company recognizes forfeitures as they occur and reverses any previously recognized compensation cost associated with pre-vesting forfeitures.
Income Taxes
The Company accounts for income taxes in accordance with ASC 740, Income Taxes (“ASC 740”), which requires that the asset and liability method be used. Deferred tax assets and liabilities are determined based on the
F-16
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
differences between financial reporting (book basis) and the tax reporting (tax basis) amounts of assets and liabilities; these pre-tax differences are measured and reflected after-tax using the enacted tax rates and law that will be in effect when the differences are expected to reverse.
ASC 740 requires that deferred tax assets be reduced by a valuation allowance if it is more likely than not that some or all of the deferred tax assets will not be realized. The Company evaluates the realizability of its deferred tax assets at least annually and more frequently as warranted by changes in facts and circumstances and adjusts the amount of the valuation allowance accordingly. Factors considered include the presence of reversing taxable temporary differences as a source of taxable income, forecasts of future taxable income, and available tax planning strategies that could be implemented and are both prudent and feasible. For all historical periods and through calendar 2023, the Company has recorded a full valuation allowance with respect to all of its deferred tax assets based upon the significant negative evidence presented by its accumulated deficit position, its operating loss and negative operating cash flows, and the uncertainty associated with the timing and amount of future profits and therefore taxable income that enable realization. In future periods, if the Company generates book income and taxable income, changes in judgment may reduce or eliminate the valuation allowance.
The Company accounts for uncertain tax positions in accordance with a two-step model. Firstly, a recognition threshold must be met; a tax benefit from an uncertain tax position may only be recognized if the tax position would, more likely than not, be sustained by the taxing authority assuming an examination and the application of typical practices and precedents. Secondly, a measurement rule is applied; the amount of tax benefit that can be recognized is the portion that is greater than 50% likely of being realized upon a settlement. The determination as to whether a tax benefit will, more likely than not, be realized is based upon the technical merits of the tax position as well as consideration of the available facts and circumstances. As of December 31, 2023 the Company’s uncertain tax positions were principally related to the non-recognition of a portion of Federal and U.S. state research and development tax credits. The Company recognizes any interest and penalties associated with its income tax positions in income tax expense (benefit).
Accounting Standards Issued and Recently Adopted
In September 2016, the FASB issued ASU No. 2016-13, Financial Instruments—Credit Losses, which was subsequently amended by ASU 2018-19 and ASU 2019-10. This standard, among other things, requires the measurement of expected credit losses for financial instruments carried at amortized cost held at the reporting date based on historical experience, current conditions and reasonable forecasts. The Company adopted this standard on January 1, 2023, with no impact on its financial statements and related disclosures given the Company’s credit loss exposure is limited to trade accounts receivable for which, historically, uncollectible receivables written off have been, and the related allowance for credit losses was, at December 31, 2023, insignificant in amount.
In August 2020, the FASB issued ASU No. 2020-06, Debt—Debt with Conversion and Other Options (Subtopic 470-20) and Derivatives and Hedging—Contracts in Entity’s Own Equity (Subtopic 815-40): Accounting for Convertible Instruments and Contracts in an Entity’s Own Equity, which simplifies accounting for convertible instruments by removing major separation models required under current GAAP. The ASU removes certain settlement conditions that are required for equity contracts to qualify for the derivative scope exception, and it also simplifies the diluted earnings per share calculation in certain areas. Early adoption is permitted, but no earlier than January 1, 2021. The Company early adopted ASU 2020-06 with effect from January 1, 2022; however, such early adoption had no impact on the financial statements and related disclosures.
F-17
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
In November 2023, the FASB issued Accounting Standards Update 2023-07, Segment Reporting (Topic 280): Improvements to Reportable Segment Disclosures (“ASU 2023-07”). This guidance requires disclosure on an annual and interim basis of the following: (i) significant segment expenses regularly provided to the chief operating decision maker (“CODM”) and a measure of segment profit or loss; (ii) an amount for other segment items by reportable segment and a description of its composition; (iii) all annual disclosures about a reportable segment’s profit and loss and assets as currently required by Topic 280; (iv) clarifying if the CODM uses more than one measure of a segment’s profit or loss in assessing segment performance and deciding how to allocate resources; and (v) disclosing the title and position of the CODM and how the CODM uses the reported measures. Public entities with a single reportable segment are required to provide all the disclosures required by this amendment. ASU 2023-07 is effective for annual periods beginning after December 15, 2023, and for quarters in the years after December 15, 2024; early adoption is permitted. The Company, a single reportable segment entity, early adopted ASU 2023-07 with effect from January 1, 2023; the related disclosures are included in the Segments note.
Accounting Pronouncements Issued and Not Yet Adopted
In December 2023, the FASB issued Accounting Standards Update 2023-09, Income Taxes (Topic 740)—Improvements to Income Tax Disclosures (“ASU 2023-09”). The new guidance provides for disclosure on an annual basis of the following: (i) specific categories in the rate reconciliation, and (ii) additional information for reconciling items that meet a quantitative threshold of greater than 5% of the amount computed by multiplying pretax income (or loss) by the applicable statutory income tax rate. ASU 2023-09 is effective for public business entities for annual periods beginning after December 15, 2025; early adoption is permitted. The Company is currently evaluating the impact of ASU 2023-09 on its income tax disclosures.
| 4. | FAIR VALUE MEASUREMENTS |
The Company’s demand deposit accounts include money market demand deposits which constitute cash, not cash equivalents, given the lack of restrictions or redemption conditions and the ability to immediately withdraw funds. The carrying amounts of the Company’s financial assets (which include cash and accounts receivable) and liabilities (which include accounts payable and accrued expenses) approximate fair value due their insensitivity to interest rates and/or short time periods to maturity and qualify as Level 1 measurements. There were no transfers between Level 1, 2 or 3 during 2023.
| 5. | BALANCE SHEET COMPONENTS |
Inventory
| December 31, 2023 |
||||
| Finished goods |
$ | 247,511 | ||
| Work in process |
534,005 | |||
| Raw materials |
1,412,988 | |||
|
|
|
|||
| Inventory |
$ | 2,194,504 | ||
|
|
|
|||
During the year ended December 31, 2023 there were no provisions to write down inventories.
F-18
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Property and Equipment, Net
| December 31, 2023 |
||||
| Machinery and equipment |
$ | 805,031 | ||
| Computer equipment, office equipment, and purchased software |
91,904 | |||
| Leasehold improvements |
28,527 | |||
|
|
|
|||
| Accumulated depreciation and amortization |
(152,754 | ) | ||
|
|
|
|||
| Property and equipment, net |
$ | 772,708 | ||
|
|
|
|||
For the year ended December 31, 2023, depreciation and amortization expenses of $141,355 were recorded as components of costs of revenue and operating expense in the statement of operations. Property and equipment, net at December 31, 2023 was located in Taiwan, Republic of China in the amount of $631,709, with the remainder located in the U.S.
Prepaid Expenses and Other Current Assets; Other Long-Term Assets
At December 31, 2023 prepaid expenses and other current assets were as follows. Other long-term assets consisted of security deposits related to operating leases.
| December 31, 2023 |
||||
| Advance payments for inventory purchases |
$ | 210,905 | ||
| Other |
371,458 | |||
|
|
|
|||
| Prepaid expenses and other current assets |
$ | 582,363 | ||
|
|
|
|||
Accrued Expenses and Other Current Liabilities
Accrued expenses and other current liabilities comprise the following:
| December 31, 2023 |
||||
| Accrued compensation |
$ | 307,706 | ||
| Accrued clinical trial expenses |
216,169 | |||
| Accrued sales taxes |
89,109 | |||
| Other |
161,511 | |||
|
|
|
|||
| Accrued expenses and other current liabilities |
$ | 774,495 | ||
|
|
|
|||
F-19
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
| 6. | REVENUE AND DEFERRED REVENUE |
For the year ended December 31, 2023, the Company’s total revenue disaggregated by geographic source and type of customer was as follows. The Company’s service revenues were attributable to U.S. customers, with product revenues attributable to both U.S. customers and outside-of-U.S. customers.
| Revenue by Geography and Type of Customer | Year Ended December 31, 2023 |
|||
| United States—sales to medical practices and hospitals: |
||||
| Products (capsules and data access reading devices) |
$ | 6,611,961 | ||
| Services (cloud-based video access and reading services) |
626,516 | |||
| Outside United States—sales to distributors: |
||||
| Products (capsules and data access reading devices) |
2,514,373 | |||
|
|
|
|||
| Total revenue |
$ | 9,752,850 | ||
|
|
|
|||
For the year ended December 31, 2023, service revenue included $426,108 of revenue from the Company’s cloud-based software-as-a-service video delivery offering recognized over time (commencing with the shipment of capsule endoscopes) and $200,408 of revenue from the provision of video reading to customers (recognized at a point in time upon provision of deliverables); of these two amounts combined, $74,477 represented the recognition of previously recorded deferred revenue. During the year, and as of December 31, 2023, the Company’s contract liabilities for revenue, reported in deferred revenue—current in the balance sheet, principally related to the cloud-based software-as-a-service video delivery offering, for which the recognition period for revenue deferrals was approximately three months.
Fees paid to group purchasing organizations recorded as reductions to revenue were $107,736 for the year ended December 31, 2023.
| 7. | LEASES |
The Company has operating leases for office and storage space in California and office space in Taiwan, Republic of China, with remaining lease terms ranging from less than one year to 4 years as of December 31, 2023.
Leases with an initial term of twelve months or less are not recorded on the balance sheet; these are limited to minor leases in Taiwan, which are excluded from amounts presented in the following tables. The operating leases include certain variable payments related to operating expenses, which are billed by the landlord, as is customary with these types of charges for office space.
The Company’s existing leases contain escalation clauses and renewal options. The Company has evaluated several factors in assessing whether it is reasonably certain that the Company will exercise a contractual renewal option. For leases with renewal options that are reasonably certain to be exercised, the Company includes the renewal term in the total lease term used in deriving the right-of-use asset and lease liability. The Company’s main office lease includes a renewal option of three years, which was not included in the term pursuant to an evaluation of those factors.
F-20
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
The following table presents components of total lease expense recognized for the year ended December 31, 2023.
| Year Ended December 31, 2023 |
||||
| Operating lease expense |
$ | 408,973 | ||
| Short-term lease expense |
11,129 | |||
|
|
|
|||
| Total lease expense |
$ | 420,102 | ||
|
|
|
|||
Operating lease right-of-use assets and lease liabilities are reflected in the balance sheet as follows:
| December 31, 2023 |
||||
| Assets |
||||
| Operating lease right-of-use assets |
$ | 1,509,706 | ||
|
|
|
|||
| Total operating lease right-of-use assets |
$ | 1,509,706 | ||
|
|
|
|||
| Liabilities |
||||
| Operating lease liabilities—current |
$ | 276,333 | ||
| Operating lease liabilities—long-term |
1,237,504 | |||
|
|
|
|||
| Total operating lease liabilities |
$ | 1,513,837 | ||
|
|
|
|||
The weighted-average remaining lease term for operating leases was 4.0 years and the weighted-average incremental borrowing rate was 11.8% as of December 31, 2023. Variable lease payments were $26,039 for the year ended December 31, 2023.
Future minimum lease payments under non-cancellable leases as of December 31, 2023, are as follows:
| Operating Leases |
||||
| Year Ending December 31, 2024 |
$ | 437,412 | ||
| 2025 |
434,420 | |||
| 2026 |
488,130 | |||
| 2027 |
502,774 | |||
| 2028 and thereafter |
— | |||
|
|
|
|||
| Total minimum lease payments |
1,862,736 | |||
|
|
|
|||
| Less: interest |
(348,899 | ) | ||
|
|
|
|||
| Total operating lease liabilities |
$ | 1,513,837 | ||
|
|
|
|||
Lease Modifications and New Leases
In October 2023, the Company consummated a third amendment to its extant main office space lease in Saratoga, California with the landlord modifying the previous amended and restated lease terms. This third amendment adjusted the term of the extant lease and adjusted the lease rate per square foot. This third amendment effectively commenced in November 2023 and restated the original lease term to expire on December 31, 2027; it provided
F-21
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
for a three-year renewal thereafter with base rent to be adjusted to then-prevailing market rates. In conjunction with this lease modification, the Company re-evaluated relevant factors concerning renewal and concluded renewal was not reasonably certain. Accordingly, the lease modification, which did not change the classification of the extant lease as an operating lease, involved adjusting the lease term, the future minimum lease payments, and re-measuring the lease liability using a 11.8% incremental borrowing rate. This modification, given the overall shortening of lease term, resulted in a $16,837 reduction to the pre-existing right-of-use asset and lease liability.
In relation to the preceding modification, the Company also procured an additional office suite from the landlord, taking possession of the space in December 2023. This additional office space is also subject to an original term expiring on December 31, 2027 with a three-year renewal option for which renewal was not concluded to be reasonably certain. This additional rented space, classified as an operating lease, was accounted for as a new lease, with discounting of the future minimum lease payments at a 11.8% incremental borrowing rate. This new lease resulted in an additional right-of-use asset and associated lease liability of $185,484 at lease inception.
| 8. | COMMITMENTS AND CONTINGENCIES |
Contract Manufacturing Commitments
The Company, post-acquisition of raw materials, relies significantly upon third-party contract manufacturers located in Taiwan, Republic of China for sub-assembly manufacturing. Management believes that any present supply concentrations related to a focus on a small number of key suppliers can be adequately hedged against with the use of more than one supplier and a readiness to quickly adjust sources (and geographic source within Asia) of supply.
Indemnity Matters
In the ordinary course of business, the Company may provide indemnification of varying scope and terms to vendors and other parties with respect to certain matters including, but not limited to, losses arising out of breach of such agreements or from intellectual property infringement claims made by third parties. The maximum potential amount of future payments that the Company could be required to make under these indemnification agreements is, in many cases, unlimited. At December 31, 2023, the Company has not incurred any material costs as a result of such indemnifications and is not currently aware of any indemnification claims.
Litigation and Legal Matters
The Company may be the subject of adverse litigation as a defendant, arbitration, claims, or proceedings related to things such as intellectual property infringement or disputes, commercial contracts, employment, or other matters in the ordinary course of business. Such matters are subject to many uncertainties and outcomes may be unpredictable. Such matters may be fully or partially mitigated by the effect of insurance coverage in place or counter-party indemnitors. The Company provides for liabilities via a charge to operating expenses when it is both probable that a liability has been incurred and the amount of the loss can be reasonably estimated.
The Company is not a party to any material litigation, arbitration, claims or proceedings and therefore did not have any contingency-related liabilities as of December 31, 2023.
Receipt of Federal Government Assistance
During 2023, the Company received cash related to refundable payroll tax credits from the U.S. Federal government in the amount of $736,152 reflecting receipt of credit claims related to calendar 2020 and 2021
F-22
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
remitted to the U.S. Internal Revenue Service under the Employee Retention Credit program, a source of COVID-19 pandemic era Federal relief to businesses paying payroll taxes. This amount, due to the uncertainty of eventual receipt following claims made, was not recognized until receipt in 2023 and was reflected within general and administrative expenses given its relationship to payroll taxes reflected as operating expenses. There can be no assurance that the Company’s program benefit received may not be examined, and if examined, possibly challenged as to a portion or all of the benefit received which, if required to be returned, would have an adverse impact on the Company’s liquidity.
| 9. | CAPITAL STOCK |
Convertible Preferred Stock—Financing History
The Company has issued several Series of preferred stock to investors as its primary financing vehicle to fund research and development and commercial stage maturity.
From October 2006, with closure before the next series, the Company issued to various investors 17,962,675 shares of Series A Preferred Stock (“Series A”), par value $0.001 per share, for an original issue price of $0.27 per share, resulting in proceeds of $4,849,922 (net of issuance costs of zero).
From November 2008, with closure before the next series , the Company issued to various investors 6,000,000 shares of Series B Preferred Stock (“Series B”), par value $0.001 per share, for original issue prices of $0.70 to $0.79 per share, resulting in proceeds of $4,318,938 (net of issuance costs of zero).
From February 2010, with closure before the next series , the Company issued to various investors 2,931,022 shares of Series C Preferred Stock (“Series C”), par value $0.001 per share, for an original issue price of $0.87 per share, resulting in proceeds of $2,549,989 (net of issuance costs of zero).
From January 2011, with closure before the next series , the Company issued to various investors 2,825,835 shares of Series C-1 Preferred Stock (“Series C-1”), par value $0.001 per share, for an original issue price of $0.89 per share, resulting in proceeds of $2,514,993 (net of issuance costs of zero).
From December 2011, with closure before the next series, the Company issued to various investors 1,733,329 shares of Series D Preferred Stock (“Series D”), par value $0.001 per share, for an original issue price of $0.90 per share, resulting in proceeds of $1,559,996 (net of issuance costs of zero).
From June 2012, with closure before the next series, the Company issued to various investors 555,553 shares of Series D-1 Preferred Stock (“Series D-1”), par value $0.001 per share, for an original issue price of $0.90 per share, resulting in proceeds of $499,998 (net of issuance costs of zero).
From October 2012, with closure before the next series, the Company issued to various investors 9,725,761 shares of Series D-2 Preferred Stock (“Series D-2”), par value $0.001 per share, for an original issue price of $0.90 per share (with certain exceptions described in the next sentence), resulting in proceeds of $8,386,994 (net of issuance costs of zero). Included in these issued Series D-2 shares were a limited number issued at a lower price pursuant to previous pricing granted to a small number of earlier investors; given the then-existing fair value was indicated by the $0.90 per share issuance price, the excess of the per-share issuance price over the proceeds was charged to accumulated deficit as a deemed dividend in the amount of $366,191, resulting in a Series D-2 Preferred Stock carrying value of $8,753,185. Unlike all other Series of preferred stock, which have a liquidation preference equal to the original issuance price plus declared and unpaid dividends, Series D-2 has a liquidation preference equal to
F-23
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
two times the original issuance price plus declared and unpaid dividends; this particular liquidation preference reflected the negotiation results between the particular investors and the Company.
From January 2014, with closure before the next series, the Company issued to various investors 13,826,084 shares of Series E Preferred Stock (“Series E”), par value $0.001 per share, for an original issue price of $1.15 per share, resulting in proceeds of $15,899,997 (net of issuance costs of zero).
From March 2016, with closure before the next series, the Company issued to various investors 4,000,005 shares of Series F-1 Preferred Stock (“Series F-1”), par value $0.001 per share, for an original issue price of $1.15 per share, resulting in proceeds of $4,600,006 (net of issuance costs of zero).
From October 2016, with closure before the next series, the Company issued to various investors 8,320,000 shares of Series F-2 Preferred Stock (“Series F-2”), par value $0.001 per share, for an original issue price of $1.25 per share, resulting in proceeds of $10,400,000 (net of issuance costs of zero).
From January 2018 until final closing for the series, the Company issued to various investors 5,925,931 shares of Series G Preferred Stock (“Series G”), par value $0.001 per share, for an original issue price of $1.35 per share, resulting in proceeds of $8,000,007 (net of issuance costs of zero).
From August 2018, with closure before the next series, the Company issued to various investors 6,792,389 shares of Series G-1 Preferred Stock (“Series G”), par value $0.001 per share, for an original issue price of $1.45 per share, resulting in proceeds of $9,848,964 (net of issuance costs of zero).
From November 2019 until December 31, 2022, pursuant to the originally resolved financing round and subsequent extensions thereof, the Company issued to various investors 23,602,655 shares of Series H Preferred Stock (“Series H”), par value $0.001 per share, for an original issue price of $1.45 per share, resulting in proceeds of $34,473,836 (net of issuance costs of zero). Via further extension, the Company issued 14,210,319 shares of Series H Preferred Stock in 2023 for an original issue price of $1.45 per share, resulting in proceeds of $20,354,976 (net of issuance costs of zero). Subsequent to December 31, 2023, via further extension, the Company issued additional shares of Series H Preferred Stock (refer to Subsequent Events note).
All Series of preferred stock are collectively referred to as the “Preferred Stock”.
Convertible Preferred Stock—Conversion Rights
Preferred Stock shares may be converted into shares of common stock at a ratio determined by dividing the original issuance price by the conversion price (the “Conversion Rate”). The conversion price is initially set at the original issuance price and may be adjusted downward, or upward, thereby reciprocally adjusting the Conversion Rate, for capital stock events such as stock splits, stock dividends, reorganizations, or other events where the consideration received relative to the number of shares issued would imperil or dilute holders of Preferred Stock. Thus, the Conversion Rate is 1:1 at issuance; there have been no events that have altered the Conversion Rate from 1:1 for any Series of Preferred Stock.
Preferred Stock shares convert to shares of common stock (i) electively, upon the receipt by the Company of a written request consented to by holders of at least a majority of the Preferred Stock then outstanding at the effective date specified in the request (provided further that such conversion shall not occur for either of Series G-1 and Series H unless a majority of holders within each of those series so consents), or (ii) automatically, upon the closing of a firm commitment underwritten initial public offering pursuant to an effective registration
F-24
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
statement filed under the Securities Act of 1933, as amended, provided that the offering price per share of common stock is not less than $2.90 or $3.00, depending on the Series (as adjusted for recapitalizations) and the aggregate gross proceeds to Company are $25 million or greater.
Convertible Preferred Stock—Liquidation Preferences
Liquidation, for purposes of liquidation preferences that favor holders of Preferred Stock, means a liquidation, dissolution, or winding up of the Company, whether voluntary or involuntary, an acquisition of the Company by another entity (other than a transaction which does not result in a change in the holders of at least 50% of voting stock outstanding before and after the transaction), or the sale, lease, exclusive license, or conveyance of all or substantially all of the Company’s assets.
The liquidation preference for each series of Preferred Stock is the sum of (i) the original issuance price per share (as adjusted for recapitalizations)—that is, one times the original issue price, and (ii) all declared but unpaid dividends, with the exception of Series D-2, where the liquidation preference includes two times the original issue price per share.
Liquidation preferences are rank-ordered by priority group—first: Series E, F-1, F-2, G, G-1 and H; second: Series D-2; third: Series D and D-1; fourth and last: Series A, B, C, and C-1. Should distributable assets upon liquidation be insufficient to satisfy respective liquidation preferences within a priority group, the distributable assets are to be distributed to the stockholders within the priority group pro rata and in relationship to the proportion of available distributable assets to the full liquidation preferences that would otherwise be due. When distributable assets are sufficient for a priority group, the remaining distributable assets shall cascade to the next priority group, and so on and so forth in descending order of priority group. Should remaining distributable assets exist, after satisfying the liquidation preferences of all Preferred Stock, such remainder is distributable to the holders of the Preferred Stock and common stock in proportion to the number of shares of common stock held by them on a combined basis, with the shares of Preferred Stock being treated as if converted.
Convertible Preferred Stock—Dividends
Outstanding shares of Preferred Stock are entitled to receive dividends, when, as and if declared by the Board of Directors at a dividend rate of 8.00% on original issuance price (as adjusted for recapitalizations) payable in preference and priority to any declaration or payment of any distribution related to common stock. The dividends are not cumulative and Preferred Stockholders do not acquire rights to dividends because they are not declared or paid in any given year. After the payment or setting aside for payment of the dividends for all Series of Preferred Stock, any additional dividends declared or paid in any fiscal year shall be split pro rata among the holders of the Series G-1 Preferred Stock, Series H Preferred Stock (with only those two Series additionally participating) and common stock in proportion to the number of shares of common stock held by them on an as-converted basis, as applicable.
The Company has never declared or paid any dividends.
Convertible Preferred Stock—Voting Rights
The holders of Preferred Stock have the same voting rights as the holders of common stock, on an as-converted basis at the then-prevailing Conversion Rate.
F-25
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
As of December 31, 2023, the Board of Directors of the Company comprised eight members. For holders of the following types or series of capital stock, voting together within each group, the following director entitlements exist: Series A – one director; Series C – one director; Series D-2 – one director; Series E – one director; Series G-1 – one director; Series H – two directors, and common stock – one director. Any director elected pursuant to the preceding, once resigned or removed, may only be replaced by a successor elected by the holders of the same type or series of capital stock.
Convertible Preferred Stock—Protective Provisions
As long as any Preferred Stock is issued and outstanding, the Company shall not, without first obtaining the approval (by vote or written consent) of the holders of a majority of outstanding shares of Preferred Stock, voting together as a single class:
| • | Amend, alter or repeal any provision of the certificate of incorporation or bylaws (including pursuant to a merger) if such action would adversely affect the Preferred Stock; |
| • | Increase or decrease the authorized number of shares of Preferred Stock (including any series thereof) or common stock; |
| • | Authorize or create any new class or series of shares having rights, preferences or privileges senior to or on parity with any series of Preferred Stock; |
| • | Voluntarily liquidate or dissolve, or enter into any transaction or series of related transactions deemed to be a liquidation, dissolution or any disposition of the Company’s property; |
| • | Effect a firm commitment underwritten initial public offering pursuant to an effective registration statement filed under the Securities Act of 1933; |
| • | Enter into any transaction (other than customary service or employment agreements) with any officer or director of the Company, or any party related thereto or affiliated therewith, unless approved by a majority of the Board of Directors; |
| • | Issue any indebtedness that is convertible into any equity security, or incur any other indebtedness other than loans, leases and similar transactions in the ordinary course of business; and |
| • | Redeem, repurchase or otherwise acquire any shares of Preferred Stock or common stock, except for the repurchase of shares of common stock from employees, officers, directors, consultants or other persons performing services pursuant to agreements granting the Company option or repurchase rights upon the occurrence of certain events. |
Common Stock
Pursuant to the Company’s certificate of incorporation, as amended and restated several times since the Company’s inception (most recently amended in September 2023), at December 31, 2023 the Company had 170,000,000 shares of common stock authorized for issuance, par value $0.001 per share. Of the authorized shares, 6,117,333 shares were issued and outstanding as of December 31, 2023. Additional shares issued in 2023 were limited to those related to exercises of stock options and common stock warrants, as depicted in the statement of convertible preferred stock and stockholders’ deficit.
The excess of authorized shares over issued and outstanding shares is intended to provide full conversion capacity for existing shares of preferred stock. Additionally, the Company has reserved 10,500,000 shares of common stock under the auspices of the Amended and Restated 2005 Stock Plan related to the issuance of equity
F-26
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
awards to officers, directors, employees and consultants. Refer to the stock-based compensation footnote for further description of award issuances under said plan.
| 10. | STOCK-BASED COMPENSATION |
Amended and Restated 2005 Stock Plan
In 2005, as originally instituted, and as subsequently amended and restated, the Board of Directors adopted, and stockholders have approved, the Amended and Restated 2005 Stock Plan (“2005 Plan”) under which the Company may grant equity-based incentive awards to the Company’s employees, directors, and consultants.
The following types of equity incentive awards are authorized to be issued:
| • | Stock options; either Incentive Stock Options (“ISO” —employees only) or Non-Qualified Stock Options (“NQSO”); and |
| • | Restricted stock awards (“RSA”). |
Under the 2005 Plan, the Company has reserved 10,500,000 shares of common stock for issuance upon the granting or exercise of awards. Through December 31, 2023, the Company has issued RSAs, ISOs and NQSOs under the 2005 Plan.
All RSAs took the form of 2,411,000 restricted common stock shares issued to certain founding individuals of the Company at or shortly after the inception of the Company and all such RSAs have either (i) fully vested (stock released from restrictions) according to service conditions stipulated in the agreements (according to a schedule of 25% on the first anniversary of the award and 1/48th each month thereafter, for a total vesting term of four years) or (ii) been forfeited pre-vesting.
Options granted to employees and non-employees contain only service vesting conditions, with typical grants vesting over a four-year period (a schedule of 25% on the first anniversary of the award and 1/48th each month thereafter) but may be granted with different vesting profiles ranging from vested-at-grant (immediate vesting) to different numbers of years. Option grants may contain provisions regarding acceleration of vesting on the occurrence of a change in control or major corporate event. Option grants are priced, due to the presence of service conditions only, using the Black-Scholes option pricing model. Options granted to employees and non-employees expire no more than 10 years from the date of grant.
Common Stock Warrants
The Company, outside of the 2005 Plan, through December 31, 2022, issued warrants to purchase its common stock, which substantively are identical to stock options, to four non-employee service providers, with one of the providers being subject to a service vesting condition. Common stock shares of 300,000 were warranted. Of the warrants on common stock shares, 50,000 were forfeited pre-vesting prior to December 31, 2022, 100,000 expired unexercised in 2023, 100,000 were exercised with common stock issued in 2023 for proceeds of $145,000, and 50,000 warrants on common stock remain outstanding at December 31, 2023 with an exercise price of $1.45 per share.
These common stock warrants are virtually identical to the Company’s stock options, with the exception of the contractual term (which is the earlier of five years from grant or a change in control of the Company); being outside of the 2005 Plan, they are excluded in the stock option award activity and stock-based compensation
F-27
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
expense tables and descriptions which follow given the description of numbers of warrants in the preceding paragraph. The Company has recognized warrant-related expense upon the warrants’ grant dates given immediate vesting and has classified such expense within operating expenses identically to the manner in which cash payments to the service providers would be classified.
Stock Options
A rollforward of activity for the year ended December 31, 2023 follows:
| Options on Common Stock |
Weighted Average Exercise Price |
Weighted Average Remaining Contractual Life (years) |
Aggregate Intrinsic Value |
|||||||||||||
| Outstanding as of December 31, 2022 |
4,300,097 | $ | 0.09 | 5.56 | $ | 334,991 | ||||||||||
| Granted |
804,000 | 0.17 | ||||||||||||||
| Exercised |
(365,686 | ) | 0.08 | |||||||||||||
| Forfeited, canceled, or expired |
(182,523 | ) | 0.08 | |||||||||||||
|
|
|
|||||||||||||||
| Outstanding as of December 31, 2023 |
4,555,888 | 0.11 | 5.47 | 285,485 | ||||||||||||
|
|
|
|||||||||||||||
| Exercisable as of December 31, 2023 |
3,414,257 | 0.10 | 4.42 | 252,366 | ||||||||||||
|
|
|
|||||||||||||||
| Exercisable and expected to vest as of December 31, 2023 |
3,414,257 | $ | 0.10 | 4.42 | $ | 252,366 | ||||||||||
|
|
|
|||||||||||||||
The aggregate intrinsic values are calculated as the differences between the exercise prices of the underlying options and the fair value of the Company’s common stock for options that were in the money.
The Company had 1,141,631 unvested stock options outstanding as of December 31, 2023. The weighted-average fair value of options granted during the year ended December 31, 2023 was $0.11 per share.
During the year ended December 31, 2023, 365,686 stock options were exercised, resulting in cash proceeds of $29,342. The Company satisfied stock option exercises with newly-issued shares.
As of December 31, 2023, total unrecognized compensation cost of $91,179 related to unvested stock options is expected to be recognized as expense over a weighted average period of 1.3 years.
When utilizing the Black-Scholes option-pricing model to determine the grant date fair value of stock options granted to employees or non-employees, the Company used the following weighted average assumptions:
| Year Ended December 31, 2023 |
||||
| Risk-free interest rate |
3.72 | % | ||
| Expected term (in years) |
5.90 | |||
| Volatility of common stock |
66.6 | % | ||
| Expected dividend rate |
0 | % | ||
| Fair value of common stock |
$ | 0.17 | ||
F-28
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Stock-Based Compensation Expense
Stock-based compensation, measured at the grant date, is recognized ratably over the requisite service period, using the straight-line method of expense attribution. The Company recognized total stock-based compensation expense for employees and non-employees as components of the statement of operations as follows.
| Year Ended December 31, 2023 |
||||
| Costs of revenue |
$ | 708 | ||
| Selling and marketing |
9,209 | |||
| Research and development |
21,098 | |||
| General and administrative |
$ | 11,620 | ||
|
|
|
|||
| Total stock-based compensation expense |
$ | 42,635 | ||
|
|
|
|||
| 11. | SEGMENTS |
The Company constitutes a single operating (and reportable) segment known as the Capsule Endoscopy Segment; its chief operating decision maker is its President. The composition of this segment reflects the Company’s focus on capsule endoscope medical devices addressing different areas within the human gastrointestinal tract. Refer to Note 3, “Summary of Significant Accounting Policies” and Note 6, “Revenue and Deferred Revenue” for descriptions of products and services. Segment accounting policies are the same as those described in said note. The chief operating decision maker assesses performance for the Capsule Endoscopy Segment and decides how to allocate resources based upon net income (loss) as reported in the statement of operations; also considered is operating income (loss), measured on the same basis as that in the accompanying statement of operations. Net income (loss) and operating income (loss) are used in the process of evaluating expenses and determining where to prioritize resources.
Segment assets are reported on the balance sheet as total assets. Segment revenue is identical to that reported in the financial statements; significant segment expenses are identical to those identified in the accompanying statement of operations under the line items for selling and marketing expenses, research and development expenses, and general and administrative expenses. Other segment items are represented by non-operating income (expense), net (which includes insignificant interest income) and income tax expense (benefit) as reported in the accompanying statement of operations. Segment depreciation expense and capital expenditures are identical to the amounts depicted in the accompanying statement of cash flows.
| 12. | INCOME TAXES |
The domestic and foreign components of loss before income taxes are as follows:
| Year Ended December 31, 2023 |
||||
| Domestic |
$ | 10,719,489 | ||
| Foreign |
573,923 | |||
|
|
|
|||
| Loss before income taxes |
$ | 11,293,412 | ||
|
|
|
|||
F-29
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
The following table reconciles the federal statutory rate to the effective income tax rate (“ETR”):
| 2023 | ||||
| Federal statutory income tax rate |
21.0% | |||
| Permanent book-tax differences |
1.2% | |||
| State income taxes, net of federal benefit |
3.7% | |||
| Federal and State research and development tax credits |
2.3% | |||
| Change in valuation allowance |
(28.5)% | |||
| Other |
0.2% | |||
|
|
|
|||
| ETR |
(0.1)% | |||
|
|
|
|||
The components of the income tax provision for the year ended December 31, 2023 are as follows:
| Year Ended December 31, 2023 |
||||
| Current provision |
||||
| U.S. Federal |
$ | — | ||
| U.S. State |
3,549 | |||
| Foreign |
7,884 | |||
|
|
|
|||
| Total current provision |
11,433 | |||
| Deferred provision |
||||
| U.S. Federal |
— | |||
| U.S. State |
— | |||
| Foreign |
— | |||
|
|
|
|||
| Total deferred provision |
— | |||
|
|
|
|||
| Total income tax provision |
$ | 11,433 | ||
|
|
|
|||
Deferred income taxes are accounted for using the asset and liability method, which requires recognition of deferred tax assets and liabilities for the expected future consequences of temporary differences between the financial reporting basis and the tax basis of assets and liabilities, as measured by enacted state and federal tax rates.
F-30
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
The main components of deferred tax assets (liabilities) are as follows:
| December 31, 2023 |
||||
| Deferred tax assets: |
||||
| Net operating loss carryforwards |
$ | 23,990,365 | ||
| Capitalized R&D expenses |
3,279,751 | |||
| Research and development tax credits |
1,469,633 | |||
| Accrued expenses |
81,918 | |||
| Operating lease liabilities |
368,127 | |||
| Other |
12,026 | |||
|
|
|
|||
| Total deferred tax assets |
29,201,820 | |||
| Valuation allowance |
(28,684,376 | ) | ||
|
|
|
|||
| Net deferred tax assets |
517,444 | |||
|
|
|
|||
| Deferred tax liabilities: |
||||
| Depreciation |
(150,321 | ) | ||
| Operating lease right-of-use assets |
(367,123 | ) | ||
|
|
|
|||
| Total deferred tax liabilities |
(517,444 | ) | ||
|
|
|
|||
| Net deferred tax assets |
$ | — | ||
|
|
|
|||
Valuation Allowance Considerations
A valuation allowance against a deferred tax asset must be established when it is “more likely than not” that all, or a portion of, deferred tax assets will not be realized. A review of all available positive and negative evidence needs to be considered, including the scheduled reversal of deferred tax liabilities, projected future taxable income, and tax planning strategies. After consideration of all the information available, in particular the Company’s accumulated deficit and history of operating losses, it was concluded that uncertainty exists with respect to future realization of deferred tax assets. A valuation allowance fully reserving net deferred tax assets, established in previous periods, was increased during the year ended December 31, 2023 in the amount of $3,189,366 to ensure net deferred tax assets were fully offset by a valuation allowance as of December 31, 2023.
Net Operating Losses and Tax Credit Carryforwards
The Company has experienced net operating losses (“NOLs”) for tax purposes since inception. As of December 31, 2023, the Company has total available gross (pre-tax) Federal and U.S. state NOL carryforwards of $91,403,212 and $69,691,526, respectively. Under the Federal Tax Cuts and Jobs Act of 2017, NOLs incurred in tax years beginning on or after January 1, 2018, are carried forward indefinitely and subject to a usage limitation of 80% of taxable income; NOLs incurred in tax years prior to January 1, 2018, are subject to a twenty-year carryforward before expiring and not subject to a usage limitation of taxable income.
F-31
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
NOLs and tax credit carryforward values as of December 31, 2023 are as follows:
| Amount | Expiration Years |
|||||||
| Net Operating Losses (gross; pre-tax) |
||||||||
| NOLs—Federal (arising after December 31, 2017) |
$ | 43,283,022 | No expiry | |||||
| NOLs—Federal (arising before January 1, 2018) |
$ | 48,120,190 | 2027 | |||||
| NOLs—State—indefinite |
$ | 425,835 | No expiry | |||||
| NOLs—State—definite lived |
$ | 69,265,691 | 2028 | |||||
| R&D Tax Credits |
||||||||
| R&D tax credits—Federal and State |
$ | 1,469,633 | 2027 | |||||
Utilization of NOL carryforwards and research and development tax credit carryforwards may be subject to a substantial annual limitation under Section 382 of the Internal Revenue Code of 1986 as amended (“Section 382”), or, for states, state laws, due to ownership changes that have occurred previously or that could occur in the future. These ownership changes may limit the amount of carryforwards that can be utilized annually to offset future taxable income.
In general, an ownership change, as defined by Section 382, results from transactions increasing the ownership of certain stockholders or public groups in the stock of a corporation by more than 50% over a three-year period. The Company has not conducted a study to assess whether a change of control has occurred or whether there have been multiple changes of control since inception. If the Company has experienced a change of control, as defined by Section 382, at any time since inception, utilization of the NOL carryforwards or research and development tax credit carryforwards would be subject to an annual limitation under Section 382. Any limitation may result in expiration of a portion of the NOL carryforwards or research and development tax credit carryforwards before utilization.
Section 174 Capitalization
The Tax Cuts and Jobs Act of 2017 made a significant change to Section 174 of the Internal Revenue Code of 1986 as amended that went into effect for taxable years beginning after December 31, 2021. The change eliminated the ability to currently deduct R&D expenses. Instead, taxpayers must now capitalize and amortize these costs for tax purposes. Capitalized Section 174 costs must be amortized over 5 years (15 years for expenditures attributable to foreign research) beginning with the midpoint of the tax year in which the expenditures are paid or incurred.
The Company had $8,167,664 and $1,146,304 of domestic and foreign R&D expenses for the tax year ended December 31, 2023, respectively, which are being capitalized and amortized over the respective periods noted above. These amounts are reflected in gross deferred tax assets (together with previous periods’ amounts).
F-32
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Uncertain Tax Positions
A reconciliation of the beginning and ending balance of total unrecognized tax benefits on a tax-effected basis for the year ended December 31, 2023 follows; these relate principally to research and development credit carryforwards.
| Unrecognized tax benefits, beginning of year |
$ | 1,353,751 | ||
| Additions for current tax positions |
273,749 | |||
| Changes for previous tax positions |
6,927 | |||
|
|
|
|||
| Net unrecognized tax benefits, end of year |
$ | 1,634,427 | ||
|
|
|
The Company has not yet conducted a study of its research and development credit carryforwards. This study may result in an adjustment to the Company’s research and development credit carryforwards; however, until a study is completed and any adjustment is known, the tax assets associated with the research and development credit carryforwards have been fully offset with a reserve. A full valuation allowance has been provided against the Company’s net deferred tax assets, and if an adjustment to the net unrecognized tax benefits were to occur, this adjustment would be offset by an equal adjustment to the valuation allowance.
The Company has recognized no interest and penalties associated with substantially all of its unrecognized tax benefits. There are no tax positions for which it is reasonably possible that the total amounts of unrecognized tax benefits will significantly increase or decrease within twelve months of the reporting date.
The Company files U.S. Federal and State income tax returns in jurisdictions with varying statutes of limitations. In the normal course of business, the Company is subject to examination by taxing authorities throughout the U.S. These examinations could include examining the timing and amount of deductions, the allocation of income among various tax jurisdictions and compliance with laws.
The Company’s remaining open tax years subject to examination by federal and state tax authorities include the years ended December 31, 2018, through 2023 in the U.S and December 31, 2019 through 2023 in the Company’s foreign operations. However, for tax years from inception through the present, federal and state taxing authorities may examine and adjust loss carryforwards in the years in which those loss carryforwards are ultimately utilized. There are currently no pending income tax examinations.
| 13. | NET LOSS PER SHARE |
| Year Ended December 31, 2023 |
||||
| Numerator: |
||||
| Net loss—basic and diluted |
$ | (11,304,845 | ) | |
| Denominator: |
||||
| Weighted-average number of shares of common stock outstanding—basic and diluted |
5,862,935 | |||
| Net loss per share—basic and diluted |
$ | (1.93 | ) | |
Basic net loss per share is the same as diluted net loss per share as the inclusion of all potential common shares outstanding would have been anti-dilutive. Certain series of convertible preferred stock, as described in the foregoing note regarding capital stock (section “Convertible Preferred Stock – Dividends”), are participating securities; however, as a result of the net loss position, they are excluded from the calculation of basic net loss per share.
F-33
Table of Contents
CAPSOVISION, INC.
NOTES TO FINANCIAL STATEMENTS
DECEMBER 31, 2023
Potentially dilutive securities that were not included in the preceding calculations because they would be anti-dilutive were as follows:
| As of December 31, 2023 |
||||
| Convertible preferred stock (all series) |
118,411,557 | |||
| Warrants on common stock |
50,000 | |||
| Options on common stock |
4,555,888 | |||
|
|
|
|||
| Total |
123,017,445 | |||
|
|
|
|||
Subsequent to December 31, 2023, additional shares of Series H Preferred Stock were issued as further described in Note 15, “Subsequent Events”.
| 14. | RELATED PARTIES |
For the year ended December 31, 2023, the Company’s material transactions with related parties were limited to certain R&D expenses (approximately $1 million) incurred with a R&D and manufacturing services vendor that holds an insignificant percentage of the Company’s capital stock. There were no amounts due from or to related parties at December 31, 2023.
| 15. | SUBSEQUENT EVENTS |
The following information reflects an evaluation of subsequent events undertaken through December 16, 2024, the date of issuance of these financial statements.
On November 13, 2024 the Company’s Board of Directors approved the pursuit of an equity initial public offering of the Company, with proceeds to be used for general corporate purposes (and further research and development); future timing is dependent upon macroeconomic conditions. There is no assurance that such an offering will be successfully completed, or, if completed, completed with desired terms.
Additional Series H Preferred Stock Financing
Through October 24, 2024, amendments to the Company’s certificate of incorporation pursuant to Delaware General Corporation Law were approved which provided for (i) an increase in the authorized number of shares of common stock by 10,000,000 shares, to 180,000,000 shares, and (ii) an increase in the authorized number of shares of Series H Preferred Stock to 48,157,821 shares, facilitating fundraising from additional Series H Preferred Stock issuances.
Through the date of issuance of these financial statements, the Company issued to various investors 10,344,835 shares of Series H Preferred Stock, par value $0.001 per share, for an original issue price of $1.45 per share, resulting in proceeds of $15,000,011 (net of issuance costs of zero).
Equity Incentive Plan Amendment
Contemporaneous with the approvals of the additional financings described in the foregoing paragraphs, the number of shares of common stock reserved for issuance under the Company’s Amended and Restated 2005 Stock Plan was increased by 5,162,500 shares to 15,662,500 shares.
F-34
Table of Contents
Shares
Common Stock
CapsoVision, Inc.
Prospectus
The Benchmark Company
Through and including , 2025 (the 25th day after the date of this prospectus), all dealers effecting transactions in these securities, whether or not participating in this offering, may be required to deliver a prospectus. This is in addition to a dealer’s obligation to deliver a prospectus when acting as an underwriter and with respect to an unsold allotment or subscription.
Table of Contents
PART II
INFORMATION NOT REQUIRED IN PROSPECTUS
| Item 13. | Other Expenses of Issuance and Distribution. |
The following table sets forth the estimated costs and expenses to be incurred in connection with the issuance and distribution of the shares of common stock being registered hereby, other than underwriting discounts and commissions. All amounts are estimates except the SEC registration fee and the Financial Industry Regulatory Authority, Inc. (“FINRA”) filing fee and the stock exchange listing fee.
The following expenses will be borne solely by the registrant:
| Amount | ||||
| SEC registration fee |
$ | |||
| FINRA filing fee |
||||
| Nasdaq listing fee |
||||
| Printing and engraving expenses |
||||
| Legal fees and expenses |
||||
| Accounting fees and expenses |
||||
| Transfer agent and registrar fees and expenses |
||||
| Miscellaneous fees and expenses |
||||
|
|
|
|||
| Total |
||||
|
|
|
|||
| Item 14. | Indemnification of Directors and Officers. |
Pursuant to Section 145 of the Delaware General Corporation Law (the “DGCL”), a corporation shall have the power to indemnify any person who was or is a party or is threatened to be made a party to any threatened, pending or completed action, suit or proceeding, whether civil, criminal, administrative or investigative (other than a derivative action by or in the right of such corporation) by reason of the fact that such person is or was a director, officer, employee or agent of such corporation, or serving at the request of such corporation in such capacity for another corporation, partnership, joint venture, trust or other enterprise, against expenses (including attorneys’ fees), judgments, fines and amounts paid in settlement actually and reasonably incurred in connection with such action, suit or proceeding, if such person acted in good faith and in a manner he or she reasonably believed to be in or not opposed to the best interests of such corporation, and, with respect to any criminal action or proceeding, had no reasonable cause to believe his or her conduct was unlawful.
The DGCL also permits indemnification by a corporation under similar circumstances for expenses (including attorneys’ fees) actually and reasonably incurred by such persons in connection with the defense or settlement of a derivative action or suit, except that no indemnification shall be made in respect of any claim, issue or matter as to which such person shall have been adjudged to be liable to such corporation unless the Delaware Court of Chancery or the court in which such action or suit was brought shall determine upon application that such person is fairly and reasonably entitled to indemnity for such expenses which such court shall deem proper.
To the extent a present or former director or officer is successful in the defense of such an action, suit or proceeding referenced above, or in defense of any claim, issue or matter therein, a corporation is required by the DGCL to indemnify such person for actual and reasonable expenses incurred in connection therewith. Expenses (including attorneys’ fees) incurred by such persons in defending any action, suit or proceeding may be paid in advance of the final disposition of such action, suit or proceeding upon in the case of a current officer or director, receipt of an undertaking by or on behalf of such person to repay such amount if it is ultimately determined that such person is not entitled to be so indemnified.
II-1
Table of Contents
The DGCL provides that the indemnification described above shall not be deemed exclusive of other indemnification that may be granted by a corporation pursuant to its bylaws, disinterested directors’ vote, stockholders’ vote and agreement or otherwise.
Section 102(b)(7) of the DGCL enables a corporation, in its certificate of incorporation or an amendment thereto, to eliminate or limit the personal liability of a director to the corporation or its stockholders for monetary damages for violations of the directors’ fiduciary duty, except (i) for any breach of the director’s duty of loyalty to the corporation or its stockholders, (ii) for acts or omissions not in good faith or which involve intentional misconduct or a knowing violation of law, (iii) pursuant to Section 174 of the DGCL (providing for liability of directors for unlawful payment of dividends or unlawful stock purchases or redemptions) or (iv) for any transaction from which a director derived an improper personal benefit. Our certificate of incorporation provides for such limitations on liability for its directors.
The DGCL also provides corporations with the power to purchase and maintain insurance on behalf of any person who is or was a director, officer, employee or agent of such corporation, or is or was serving at the request of such corporation in a similar capacity for another corporation, partnership, joint venture, trust or other enterprise, against any liability asserted against him or her in any such capacity or arising out of his or her status as such, whether or not the corporation would have the power to indemnify him or her against such liability as described above. In connection with this offering, the registrant will obtain liability insurance for its directors and officers. Such insurance would be available to its directors and officers in accordance with its terms.
Our certificate of incorporation requires us to indemnify and hold harmless, to the fullest extent permitted by applicable law as it presently exists or may hereafter be amended, any person (a “covered person”) who was or is made or is threatened to be made a party or is otherwise involved in any threatened, pending or completed action, suit or proceeding, whether civil, criminal, administrative or investigative (a “proceeding”) by reason of the fact that he or she is or was a director, officer or member of a committee, or, while a director or officer, is or was serving at our request as a director or officer, employee or agent of another corporation, partnership, joint venture, trust or other enterprise or non-profit entity, including service with respect to employee benefit plans, against all liability and loss suffered and expenses (including attorneys’ fees), judgment, fines and amounts paid in settlement actually and reasonably incurred by such person in connection with a proceeding.
In addition, under our certificate of incorporation, in certain circumstances, we are required to pay the expenses (including attorneys’ fees) incurred by a covered person in defending a proceeding in advance of the final disposition of such proceeding; provided, however, that we are not required to advance any expenses to a person against whom we directly bring an action, suit or proceeding alleging that such person (1) committed an act or omission not in good faith or (2) committed an act of intentional misconduct or a knowing violation of law. Additionally, an advancement of expenses incurred by a covered person shall be made only upon delivery to us of an undertaking, by or on behalf of such covered person, to repay all amounts so advanced if it shall ultimately be determined by final judicial decision from which there is no further right to appeal or otherwise in accordance with Delaware law that such covered person is not entitled to be indemnified for such expenses.
In addition, we plan to enter into indemnification agreements with our directors and executive officers that provide for additional indemnification protections, which form of agreement has been filed as an exhibit to this registration statement.
| Item 15. | Recent Sales of Unregistered Securities. |
Set forth below is information regarding shares of capital stock issued by us within the past three years. Also included is the consideration received by us for such shares and information relating to the section of the Securities Act, or rule of the Securities and Exchange Commission, under which exemption from registration was claimed.
II-2
Table of Contents
Equity Plan-Related Issuances
Since January 1, 2022, the registrant has granted to its employees and non-employees options to purchase an aggregate of 424,000 shares of its common stock under its 2005 Plan, at an exercise price of $0.11 per share, in connection with services provided to the registrant by such parties.
Since January 1, 2022, the registrant has granted to its employees and non-employees options to purchase an aggregate of 4,832,000 shares of its common stock under its 2005 Plan, at an exercise price of $0.17 per share, in connection with services provided to the registrant by such parties.
Since January 1, 2022, the registrant has issued to its employees and non-employees an aggregate of 1,602,889 shares of its common stock upon the exercise of stock options under its 2005 Plan, for an aggregate cash amount of approximately $0.1 million.
The issuances of such stock options and the shares of common stock issuable upon the exercise of such options were issued pursuant to written compensatory plans or arrangements with the registrant’s employees, directors and consultants, in reliance on the exemption provided by Rule 701 promulgated under the Securities Act, or pursuant to Section 4(a)(2) under the Securities Act, relative to transactions by an issuer not involving any public offering, to the extent an exemption from such registration was required.
Issuance of Preferred Stock
Since January 1, 2022, the registrant has issued an aggregate of 33,120,656 shares of its Series H preferred stock to 88 accredited investors at a purchase price of $1.45 per share, for an aggregate purchase price of $48.0 million. Each issuance of preferred stock was made pursuant to Section 4(a)(2) of the Securities Act and/or Rule 506 as a transaction not involving a public offering.
| Item 16. | Exhibits and Financial Statement Schedules. |
See the Exhibit Index attached to this registration statement, which Exhibit Index is incorporated herein by reference.
Schedules not listed above have been omitted because the information required to be set forth therein is not applicable or is shown in the financial statements or notes thereto.
| Item 17. | Undertakings. |
Insofar as indemnification for liabilities arising under the Securities Act may be permitted to directors, officers, and controlling persons of the registrant pursuant to the foregoing provisions, or otherwise, the registrant has been advised that in the opinion of the SEC such indemnification is against public policy as expressed in the Securities Act, and is, therefore, unenforceable. In the event that a claim for indemnification against such liabilities (other than the payment by the registrant of expenses incurred or paid by a director, officer, or controlling person of the registrant in the successful defense of any action, suit, or proceeding) is asserted by such director, officer, or controlling person in connection with the securities being registered, the registrant will, unless in the opinion of its counsel the matter has been settled by controlling precedent, submit to a court of appropriate jurisdiction the question of whether such indemnification by it is against public policy as expressed in the Securities Act and will be governed by the final adjudication of such issue.
II-3
Table of Contents
The undersigned registrant hereby undertakes that:
| 1. | For purposes of determining any liability under the Securities Act, the information omitted from the form of prospectus filed as part of this Registration Statement in reliance upon Rule 430A and contained in a form of prospectus filed by the Registrant pursuant to Rule 424(b)(1) or (4) or 497(h) under the Securities Act shall be deemed to be part of this Registration Statement as of the time it was declared effective. |
| 2. | For the purpose of determining any liability under the Securities Act, each post-effective amendment that contains a form of prospectus shall be deemed to be a new registration statement relating to the securities offered therein, and the offering of such securities at that time shall be deemed to be the initial bona fide offering thereof. |
II-4
Table of Contents
Exhibit Index
| Exhibit Number |
Exhibit Description | |
| 1.1* | Form of Underwriting Agreement. | |
| 3.1* | Amended and Restated Certificate of Incorporation (currently in effect). | |
| 3.2* | Form of Amended and Restated Certificate of Incorporation, to be in effect immediately prior to the completion of this offering. | |
| 3.3* | Bylaws (currently in effect). | |
| 3.4* | Form of Amended and Restated Bylaws, to be in effect immediately prior to the completion of this offering. | |
| 4.01* | Form of Common Stock Certificate. | |
| 4.02* | Amended and Restated Investors’ Rights Agreement, dated November 21, 2019, by and among CapsoVision, Inc. and the investors listed therein. | |
| 4.03* | Warrant to Purchase Shares of Common Stock, dated January 12, 2021, issued to the Nai-Yu Pai and Helen M Pai Trust. | |
| 5.1* | Opinion of O’Melveny & Myers LLP | |
| 10.01†* | Memorandum of Understanding, dated May 14, 2008, by and between Largan Precision Company, Ltd. and CapsoVision, Inc. | |
| 10.02†* | Amendment to Memorandum of Understanding, dated January 1, 2010, by and between Largan Precision Company, Ltd. and CapsoVision, Inc. | |
| 10.03†* | Memorandum of Understanding, dated October 28, 2022, by and between Largan Precision Company, Ltd. and CapsoVision, Inc. | |
| 10.04†* | Development Agreement, dated August 8, 2013, by and between Toshiba Corporation and CapsoVision, Inc. | |
| 10.05†* | Development and Manufacturing Agreement dated June 3, 2014, by and between Moai Electronics Corporation and CapsoVision, Inc., as amended on March 30, 2015. | |
| 10.06#* | Supplement to Development and Manufacturing Agreement dated February 10, 2020, by and among Moai Electronics Corporation, SpeedBridge Technology Corp. and CapsoVision, Inc. | |
| 10.07#* | 2005 Stock Incentive Plan. | |
| 10.08#* | Form Agreements under 2005 Stock Incentive Plan. | |
| 10.09#* | 2025 Equity Incentive Plan. | |
| 10.10#* | Form Agreements under 2025 Equity Incentive Plan. | |
| 10.11#* | Form of Indemnification Agreement for Directors and Officers. | |
| 10.12#* | Form of Employment Agreement for Executive Officers | |
| 23.1* | Consent of Baker Tilly US, LLP, independent registered public accounting firm. | |
| 23.2* | Consent of O’Melveny & Myers LLP (included in Exhibit 5.1). | |
| 24.1* | Power of Attorney (reference is made to the signature page to the Registration Statement). | |
| 107* | Filing Fee Table. | |
II-5
Table of Contents
* To be filed by amendment.
# Indicates management contract or compensatory plan.
† Portions of the exhibit, marked by brackets, have been omitted because the omitted information is (i) not material and (ii) the type of information that the registrant customarily and actually treats as private or confidential. The confidential portions of this exhibit have been omitted and filed separately with the Securities and Exchange Commission.
II-6
Table of Contents
SIGNATURES
Pursuant to the requirements of the Securities Act, the registrant has duly caused this registration statement to be signed on its behalf by the undersigned, thereunto duly authorized, in the city of Saratoga, California, on , 2025.
| CAPSOVISION, INC. | ||
| By: |
||
| Kang-Huai (Johnny) Wang | ||
| President and Chief Executive Officer | ||
II-7
Table of Contents
POWER OF ATTORNEY
KNOW ALL PERSONS BY THESE PRESENTS, that each person whose signature appears below hereby constitutes and appoints Kang-Huai (Johnny) Wang as attorney-in-fact and agent, with full power of substitution and re-substitution, to sign on his or her behalf, individually and in any and all capacities, including the capacities stated below, any and all amendments (including post-effective amendments) to this registration statement and any registration statements filed by the registrant pursuant to Rule 462 of the Securities Act of 1933, as amended, relating thereto and to file the same, with all exhibits thereto, and other documents in connection therewith, with the Securities and Exchange Commission, granting to said attorney-in-fact and agent, full power and authority to do and perform each and every act and thing requisite and necessary to be done in and about the premises, as fully to all intents and purposes as he or she might or could do in person, hereby ratifying and confirming all that said attorney-in-fact and agent, or his or her substitute, may lawfully do or cause to be done by virtue hereof.
Pursuant to the requirements of the Securities Act of 1933, as amended, this registration statement has been signed below by the following persons in the capacities and on the dates indicated:
| Signature |
Title |
Date | ||
|
Kang-Huai (Johnny) Wang |
Director, President and Chief Executive Officer (Principal Executive Officer) | , 2025 | ||
|
Kevin Lundquist |
Chief Financial Officer (Principal Financial Officer) | , 2025 | ||
|
Eliyahou Harari |
Director | , 2025 | ||
|
Chen Lung Tsai |
Director | , 2025 | ||
II-8